
使用原生质谱和/交换质谱测试高阶蛋白质复合物的探测:使用新鲜和商业血红蛋白样本的案例研究
T-Y Lui1, Xiangfeng Chen1,2, Danna Hu1
1Department of Chemistry, The Chinese University of Hong Kong, Hong Kong SAR 999077, P. R. China.

概括
原生质谱 (MS) 和/交换MS (HDX-MS) 揭示了蛋白质复合物的完整性. 新鲜的血红蛋白保留了它的结构,而商业样本显示变质,突出了结构验证的需要.
科学领域:
- 生物物理化学 生物物理化学
- 蛋白质组学是指蛋白质组学.
- 结构生物学 结构生物学
背景情况:
- 蛋白质复合体的原生质谱 (MS) 对矩阵效应敏感,通常需要缓冲区交换.
- 为本地MS优化缓冲区交换协议具有挑战性,对于准确的结果至关重要.
- 血红蛋白 (Hb) 作为一个模型系统来研究缓冲区交换对蛋白质复合体分析的影响.
研究的目的:
- 评估缓冲区交换协议对原生MS蛋白质组合分析的影响.
- 为了比较新鲜和商业血红蛋白样本的结构完整性,使用本地MS和HDX-MS.
- 为了证明本地MS和HDX-MS在评估高阶蛋白质结构方面的互补性质.
主要方法:
- 原生质谱法 (MS) 用于分析血红蛋白复合体.
- 用/交换质谱法 (HDX-MS) 进行结构比较研究.
- 使用血红蛋白作为模型,研究了缓冲区交换优化.
主要成果:
- 蛋白质组合的本地MS光谱高度依赖缓冲区交换协议.
- HDX-MS数据与缓冲器交换样本的本地MS分析有很好的相关性.
- 新鲜的Hb样本保持了他们的四度结构,而商业的Hb样本在很大程度上被变质为二度形式.
结论:
- 原生MS和HDX-MS是分析高阶蛋白质复合物的互补技术.
- 缓冲区交换协议显著影响本土成员国的结果.
- 在下游生物医学应用之前,对蛋白质高阶结构完整性的验证至关重要.
相关概念视频
 01:21
Mass Spectrometry: Complex Analysis
740
Mass spectrometry is an important technique for the identification of pure compounds. However, it has some limitations for the analysis of complex mixtures, often due to excessive fragmentation making the spectrum too complicated to decipher. Mass spectrometry can be combined with suitable separation methods in sequence, forming hyphenated methods, which are useful in the analysis of complex mixtures.
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
740
 01:33
Peptide Identification Using Tandem Mass Spectrometry
6.4K
Tandem mass spectrometry, also known as MS/MS or MS2, is an analytical technique that employs two mass analyzers. Essentially it is a series of mass spectrometers that helps isolate a particular biomolecule and then helps study its chemical properties.
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
6.4K
 01:15
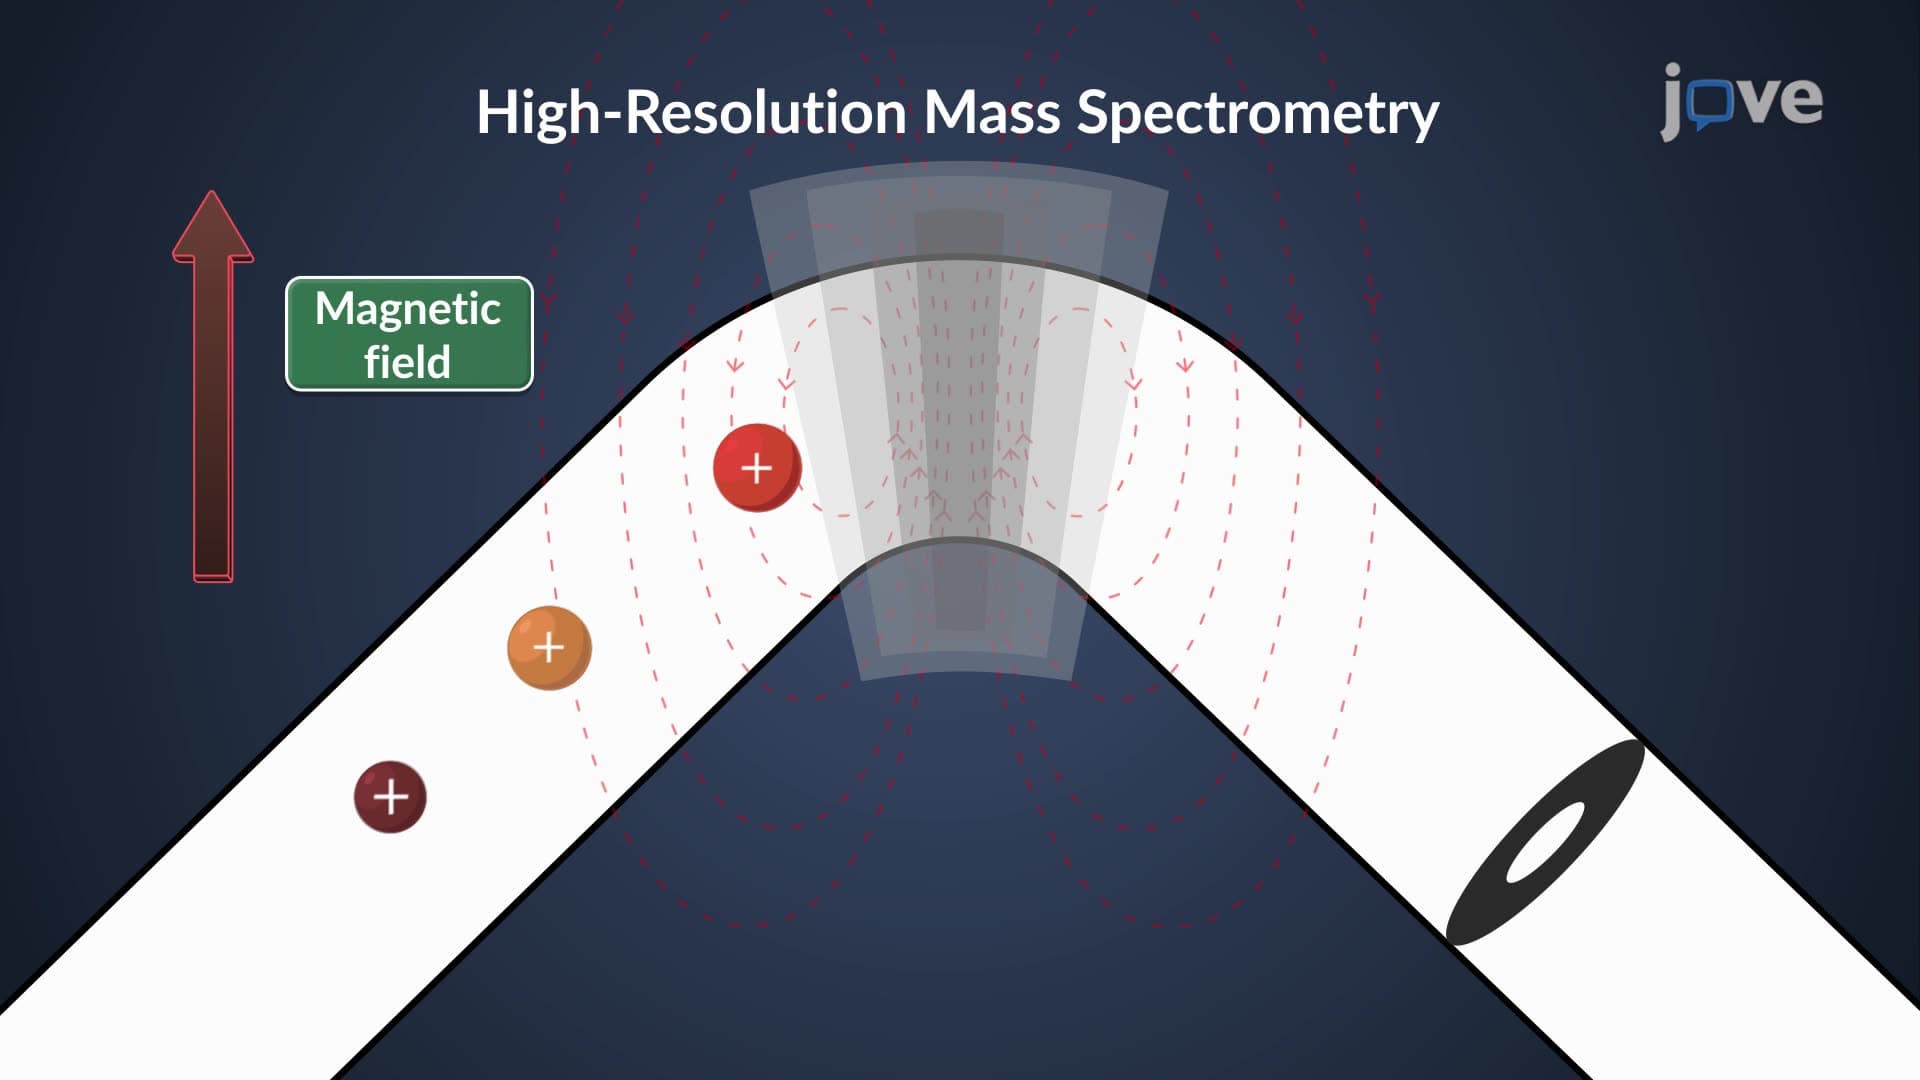
High-Resolution Mass Spectrometry (HRMS)
1.3K
The resolution of a mass spectrometer depends on the efficiency of separating ions with different ion masses. The mass of an atom is approximated to the sum of the masses of protons and neutrons inside, considering the masses of protons and neutrons as equal. However, the masses of the proton (1.6726 × 10⁻²⁴ g) and neutron (1.6749 × 10⁻²⁴ g) are not truly equal. There is a minor error in the expression of atomic masses relative to the simplest atom of hydrogen. For...
1.3K