
在下一代测序时代的重复:从历史位置到人口数据库
Kevin Uguen1,2,3, Jacques L Michaud4,5,6, Emmanuelle Génin7
1Univ Brest, Inserm, EFS, UMR 1078, GGB, Brest, France. kevin.uguen@univ-brest.fr.
European journal of human genetics : EJHG
|July 9, 2024

概括
短串联重复 (STR) 是重要的基因组标记. 先进的测序技术改善了对遗传多样性,疾病和法医应用的STR分析,需要全面的全球数据库.
科学领域:
- 基因组学就是基因组学.
- 分子生物学分子生物学
- 人口遗传学 人口遗传学
背景情况:
- 短串联重复 (STR) 构成了人类基因组的重要组成部分,在遗传多样性,疾病病原和进化研究中发挥着关键作用.
- STRs是理解人类变异的基本标记,并在从法医科学到医学遗传学等领域有应用.
研究的目的:
- 探索当前人类基因组中短串重复 (STR) 分析的现状.
- 审查用于检测基因组变异的不断发展的技术,特别是STR.
- 突出STRs在医学遗传学,法医科学和人口遗传学中的意义和影响.
主要方法:
- 审查STR分析的当前方法,包括高通量测序.
- 对解释人口特异性STR变异的挑战进行分析.
- 检查STRs在遗传疾病中的作用及其临床应用.
主要成果:
- 高通量测序显著提升了STRs的准确映射和分析.
- STR变异在全球不同的人口中存在解释方面的挑战.
- 人们越来越认识到STR在临床诊断和遗传研究中的重要性.
结论:
- 迫切需要建立全面的STR数据库,以捕捉全球遗传多样性.
- 准确的STR数据解释对于推进临床诊断,遗传研究和法医应用至关重要.
- 将STR分析整合到常规遗传研究和临床实践中,对于未来的发现至关重要.
相关概念视频
 03:00
Next-generation Sequencing
88.6K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
88.6K
 01:57
Sanger Sequencing
754.0K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
754.0K
 02:51
Multi-species Conserved Sequences
3.9K
Next-generation sequencing technologies have created large genomic databases of a variety of animals and plants. Ever since the human genome project was completed, scientists studied the genome of primates, mammals, and other phylogenetically distant living beings. Such large-scale studies have provided new insights into the evolutionary relationship between organisms.
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
3.9K
 03:21
RNA-seq
9.9K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.9K
 01:05
Maxam-Gilbert Sequencing
11.1K
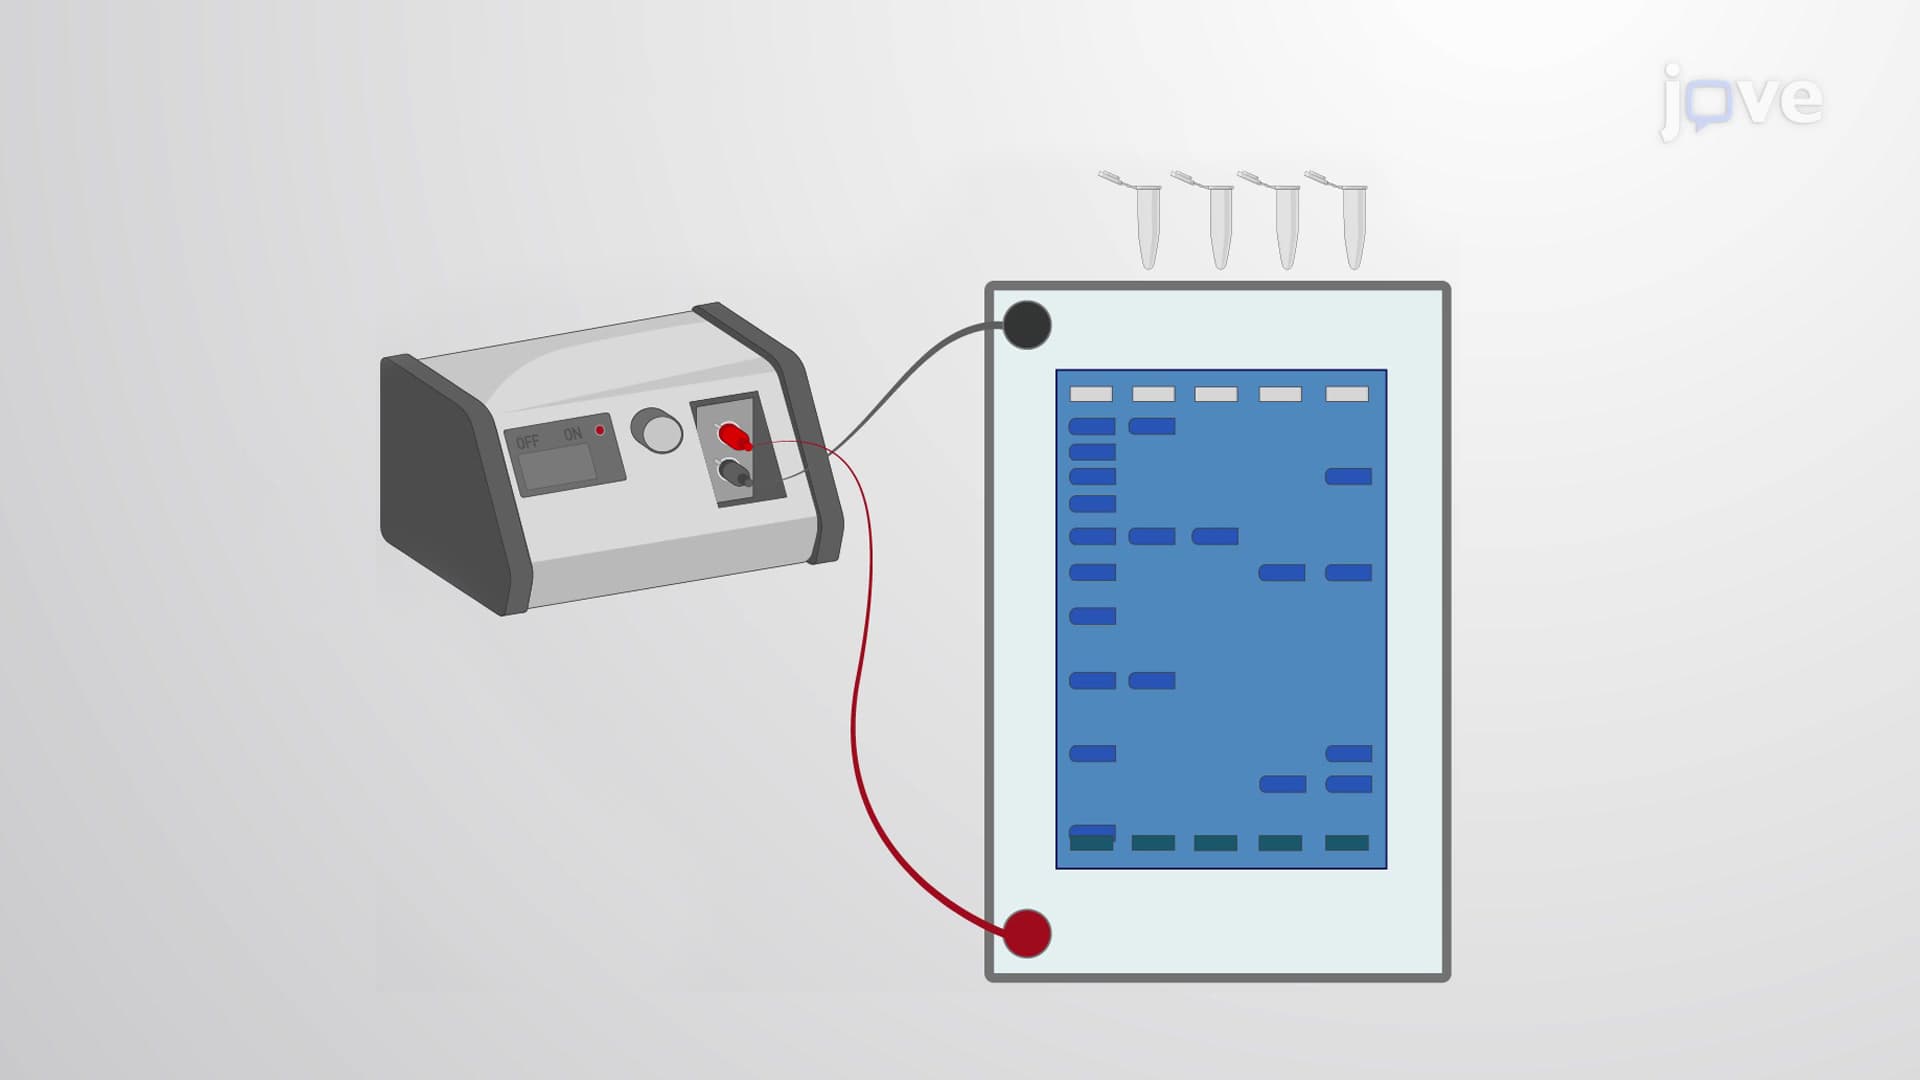
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
11.1K
 01:11
Genome-wide Association Studies-GWAS
13.3K
Genome-wide association studies or GWAS are used to identify whether common SNPs are associated with certain diseases. Suppose specific SNPs are more frequently observed in individuals with a particular disease than those without the disease. In that case, those SNPs are said to be associated with the disease. Chi-square analysis is performed to check the probability of the allele likely to be associated with the disease.
GWAS does not require the identification of the target gene involved in...
GWAS does not require the identification of the target gene involved in...
13.3K