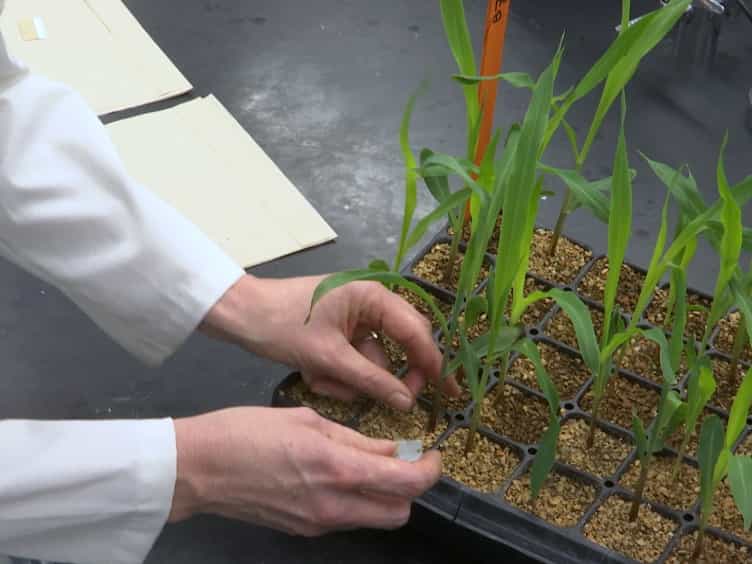

通过高通量测序解开涉及骨昏迷发展的lncRNA-miRNA-mRNA调控网络
Zihe Song1, Chenghao Zhang1, Guotao Song1
1State Key Laboratory of Tree Genetics and Breeding, Co-Innovation Center for Sustainable Forestry in Southern China, Nanjing Forestry University, Nanjing 210037, China.
International journal of molecular sciences
|July 13, 2024

概括
研究人员调查了种子昏迷的发展,确定了关键的分子参与者. 这项研究揭示了长非编码RNA (lncRNA) 网络调节昏迷形成,为控制这种植物分散机制提供了洞察力.
科学领域:
- 植物生物学 植物生物学
- 分子遗传学 分子遗传学
- 文字转录学 (Transcriptomics) 是一个学科.
背景情况:
- 种子昏迷 (毛的附属物) 促进长距离的分散,但导致环境问题.
- 了解昏迷发展机制对于管理其负面影响至关重要.
研究的目的:
- 阐明分子机制,特别是涉及长非编码RNAs (lncRNAs),调节种子昏迷的启动和发展.
- 构建参与昏迷形成,细胞骨安排和细胞壁重塑的调节网络.
主要方法:
- 在骨昏迷的三个发育阶段进行链特异性RNA测序 (RNA-seq).
- 对lncRNA和信使RNA (mRNA) 表达特征的分析.
- 通过共同定位和共同表达来预测 lncRNA 基因.
- 构建 lncRNA-miRNA-TF 和 lncRNA-miRNA-mRNA 调控网络.
主要成果:
- 在昏迷发育期间鉴定了1888个lncRNA和52810个mRNA.
- 构建了用于昏迷启动,细胞骨组织 (素,素) 和细胞壁合成 (纤维素合成酶,糖糖合成酶,扩张) 的调节网络.
- 揭示了lncRNAs在调节昏迷发育期间关键细胞过程中的作用.
结论:
- 这项研究提供了对IncRNA参与种子昏迷形成的全面了解.
- 对昏迷发展的分子基础的新见解为未来的研究和管理提供了潜在的途径.
更多相关视频
相关概念视频
 02:39
lncRNA - Long Non-coding RNAs
8.5K
In humans, more than 80% of the genome gets transcribed. However, only around 2% of the genome codes for proteins. The remaining part produces non-coding RNAs which includes ribosomal RNAs, transfer RNAs, telomerase RNAs, and regulatory RNAs, among other types. A large number of regulatory non-coding RNAs have been classified into two groups depending upon their length – small non-coding RNAs, such as microRNA, which are less than 200 nucleotides in length, and long non-coding RNA...
8.5K
 03:21
RNA-seq
9.9K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.9K