相关实验视频
Updated: Jun 20, 2025
09:58

Light-driven Enzymatic Decarboxylation
Published on: May 22, 2016
10.7K
识别和工程弗拉文依赖的化酶用于选择性生物催化剂
1Department of Chemistry, Indiana University, Bloomington, Indiana 47405, United States.
Accounts of chemical research
|July 22, 2024

概括
工程制造的黄素依赖化酶 (FDHs) 能够选择性化各种非原生化合物. 定向进化和基因组挖掘扩大了FDH用于生物催化合成的实用性,包括enantioselective反应.
科学领域:
- 生物催化和酶工程 生物催化和酶工程
- 有机化学 有机化学
- 合成生物学 合成生物学
背景情况:
- 有机化合物在制药和农业化学品中至关重要,但它们的合成通常需要恶劣的条件和多步骤的过程.
- 黄素依赖化酶 (FDHs) 为选择性化提供了一种以自然为灵感的解决方案,利用FADH2和分子氧.
- 以往通过现场定向突变发生的FDH工程工作在基质范围和对非本地基质的选择性方面取得了有限的改善.
研究的目的:
- 克服FDH工程中的局限性,用于非本土基质的制备性化.
- 扩大FDHs用于各种生物催化化反应的实用性,超出简单的芳香化.
- 开发FDH选择性的预测模型,并识别具有独特催化能力的新型FDH.
主要方法:
- 优化FDH RebH及其还原酶RebF的表达条件.
- 针对RebH的进化以提高稳定性,基质范围和位点选择性.
- 进行X射线晶体学,分子动力学模拟和全家族基因组挖掘,以识别和描述多样化的FDHs.
- 工程FDHs在enantioselective脱对称化,atroposelective化,环化和由单元FDH/FRed AetF催化反应中的工程FDHs的探索.
主要成果:
- 工程 RebH 证明了非原生基质的准备性化,具有催化剂控制的选择性.
- 定向进化显著提高了FDH稳定性,基质范围和地点选择性,达到合成有用的水平.
- 基因组挖掘发现了各种FDH,具有新的基质范围和复杂化合物的互补区域选择性.
- 已经证明,FDHs可催化异构选择性反应,包括脱对称化,异构选择性化和环化.
- 单个组件的FDH/FRed AetF表现出独特的基质化能力和高反选择性,包括选择性芳香和反选择性乙化.
结论:
- 通过定向进化和基因组挖掘开发的工程FDHs代表了生物催化化强大的工具.
- 这些酶提供了增强的稳定性,更广泛的基质范围,与野生类型对应物相比,提高了选择性.
- 在复杂的合成应用中证明的实用性,包括enantioselective转换,突显了FDHs对未来化学合成的重大承诺.
相关概念视频
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
3.3K
Catalytic hydrogenation of alkenes is a transition-metal catalyzed reduction of the double bond using molecular hydrogen to give alkanes. The mode of hydrogen addition follows syn stereochemistry.
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
3.3K
 01:13
Electrophilic Aromatic Substitution: Fluorination and Iodination of Benzene
5.9K
Bromination and chlorination of aromatic rings by electrophilic aromatic substitution reactions are easily achieved, but fluorination and iodination are difficult to achieve. Fluorine is so reactive that its reaction with benzene is difficult to control, resulting in poor yields of monofluoroaromatic products. To address this, Selectfluor reagent is used as a fluorine source in which a fluorine atom is bonded to a positively charged nitrogen.
5.9K
 00:51
Base-Promoted α-Halogenation of Aldehydes and Ketones
3.4K
α-Halogenation of aldehydes and ketones is a reaction involving the substitution of α hydrogens with halogens in the presence of a base. The reaction begins with the abstraction of α hydrogen by the base to produce a nucleophilic enolate ion. This intermediate undergoes a subsequent nucleophilic substitution with the halogen to produce a monohalogenated carbonyl compound. If the starting substrate has more than one α hydrogen, it is difficult to stop the reaction...
3.4K
 01:24
ortho–para-Directing Deactivators: Halogens
5.5K
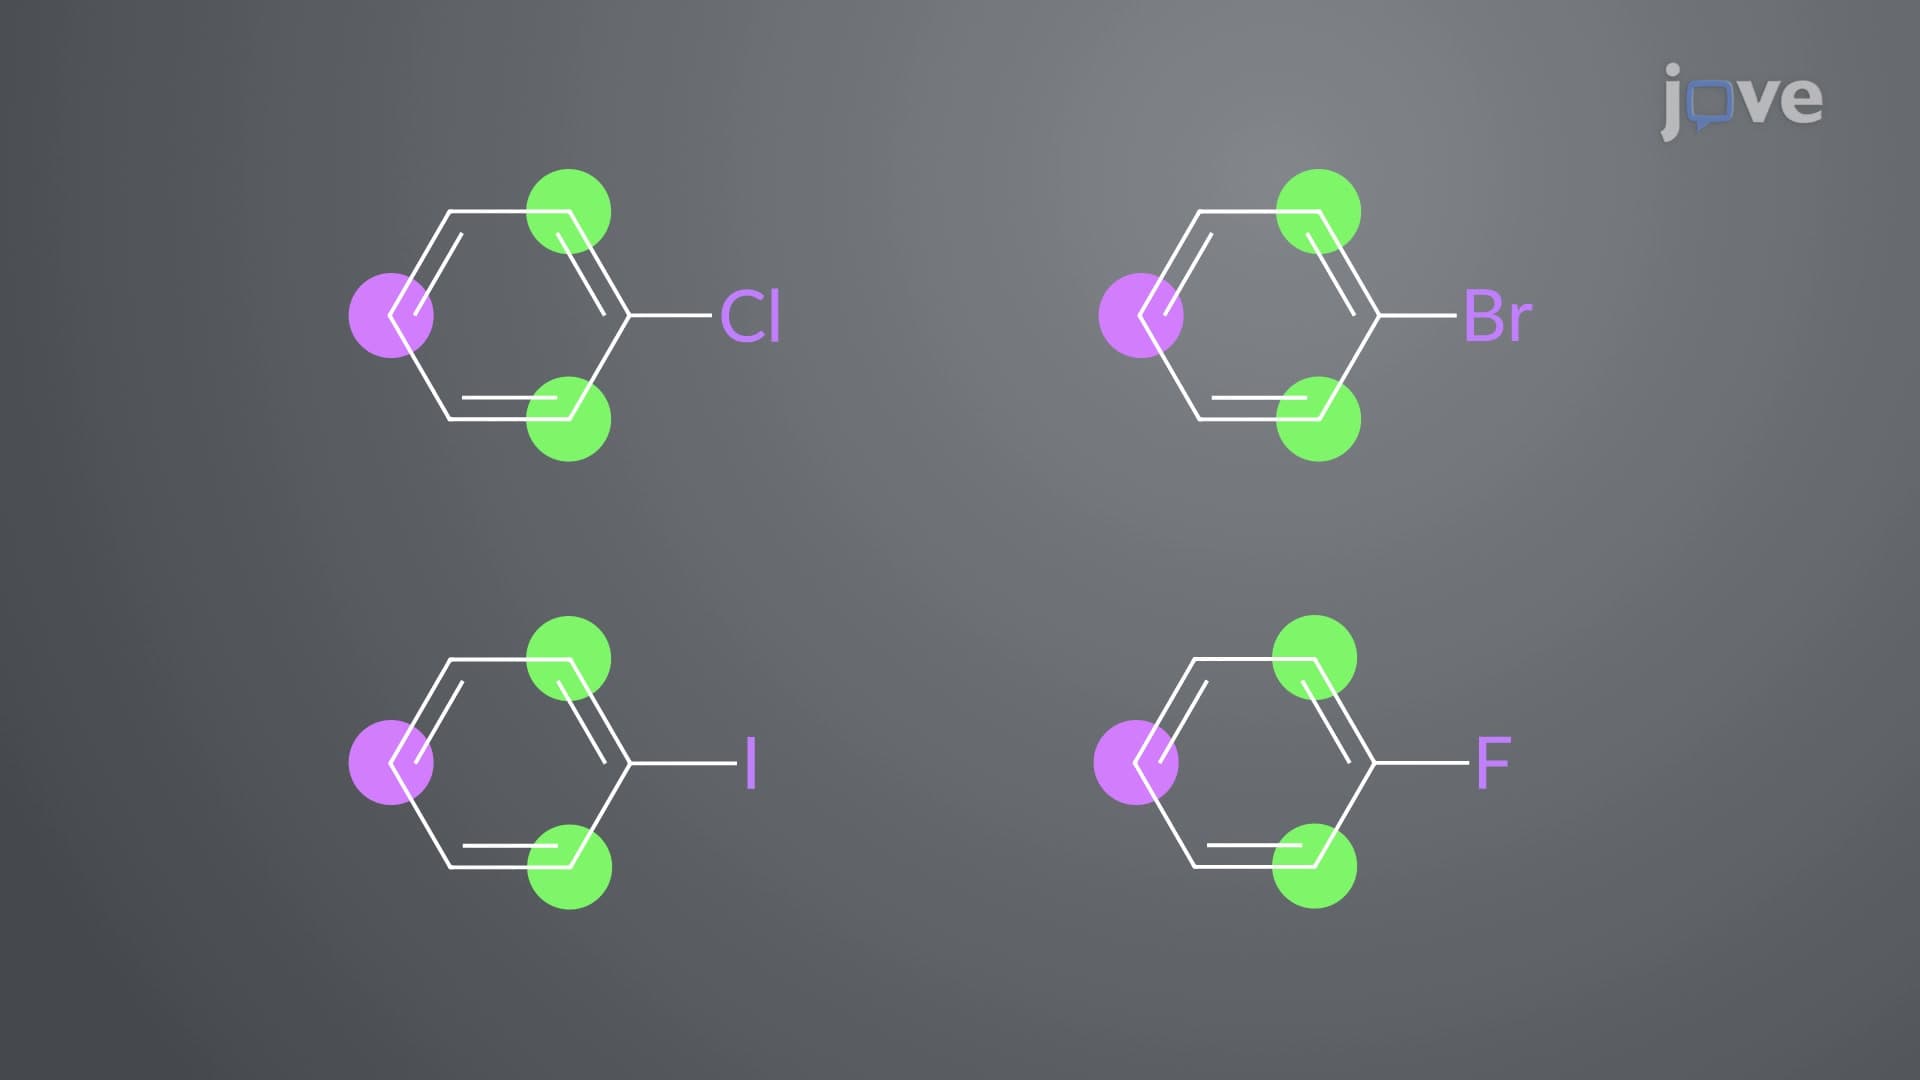
Halogens are ortho–para directors. They are more electronegative than carbon. Therefore, as ring substituents, they can withdraw electrons through the inductive effect and deactivate the aromatic ring towards electrophilic substitution. Halogens also have an electron-donating resonance effect on the ring, which influences the orientation of the incoming electrophile. If an electrophile attacks at the ortho or the para position, the halogen donates electrons and stabilizes the intermediate...
5.5K
 01:21
Acid-Catalyzed α-Halogenation of Aldehydes and Ketones
3.7K
By replacing an α-hydrogen with a halogen, acid-catalyzed α-halogenation of aldehydes or ketones yields a monohalogenated product
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...
3.7K
 02:17
Regioselectivity of Electrophilic Additions to Alkenes: Markovnikov's Rule
14.1K
If a set of reactants can yield multiple constitutional isomers, but one of the isomers is obtained as the major product, the reaction is said to be regioselective. In such reactions, bond formation or breaking is favored at one reaction site over others.
The hydrohalogenation of an unsymmetrical alkene can yield two haloalkane products, depending on which vinylic carbon takes up the halogen. However, one product usually predominates, where hydrogen adds to the vinylic carbon bearing the...
The hydrohalogenation of an unsymmetrical alkene can yield two haloalkane products, depending on which vinylic carbon takes up the halogen. However, one product usually predominates, where hydrogen adds to the vinylic carbon bearing the...
14.1K