
在P2X7受体的高亲和激动作用是由三种残留物介导的,这些残留物位于orthosteric口袋外
Adam C Oken1, Nicolas E Lisi1, Ipsita Krishnamurthy1
1Department of Chemical Physiology & Biochemistry, Oregon Health & Science University, Portland, OR, 97239, USA.
Nature communications
|August 6, 2024

概括
研究人员发现了高亲和激动剂如何激活P2X7受体,这对于癌症治疗至关重要. 这项研究揭示了开发向P2X7癌症治疗的关键残留物和结构细节.
科学领域:
- 分子生物学分子生物学
- 结构生物学 结构生物学
- 生物物理学的生物物理.
背景情况:
- P2X受体是调节细胞信号的ATP离子通道.
- P2X7受体激活与癌症生物学有关,通过亡途径.
- 了解P2X7高亲和度激活机制对于治疗开发至关重要.
研究的目的:
- 阐明P2X7受体高亲和激活的基础结构机制.
- 定义P2X7受体在封闭状态下孔结构的分子细节.
- 为了确定由BzATP负责高亲和激素的特定残留物.
主要方法:
- 高分辨率冷电子显微镜 (cryo-EM) 用于确定受体结构.
- 电生理学记录以评估受体功能.
- 直接结合试验用于量化激素相互作用.
主要成果:
- 确定野生型大鼠P2X7的冷-EM结构,具有BzATP和apo状态.
- 揭示了孔隙结构的分子细节和封闭状态下的Na+离子相互作用.
- 在ATP结合部位之外确定了三种特定的残留物,这些残留物对于BzATP的高亲和激动作用至关重要.
结论:
- 这项研究提供了有关高亲和度激动剂对P2X7受体激活的原子层次见解.
- 结构和功能数据为开发特定亚型P2X7激动剂铺平了道路.
- 这些发现支持P2X7受体调节在癌症治疗中的潜力.
更多相关视频
相关概念视频
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
2.6K
Adrenergic agonists' structure-activity relationship (SAR) determines their selectivity and efficacy. These agonists comprise a phenylethylamine moiety with an aromatic ring and an ethylamine side chain.
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
2.6K
 01:25
Drug-Receptor Interaction: Agonist
2.4K
Agonists are drugs that interact with specific receptors in the body to produce a biological response. When an agonist binds to a receptor, it activates or enhances the receptor's function, leading to physiological effects. The interaction between agonist drugs and receptors is crucial for their therapeutic action in various medical treatments.
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous...
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous...
2.4K
 01:29
The Two-State Receptor Model
1.9K
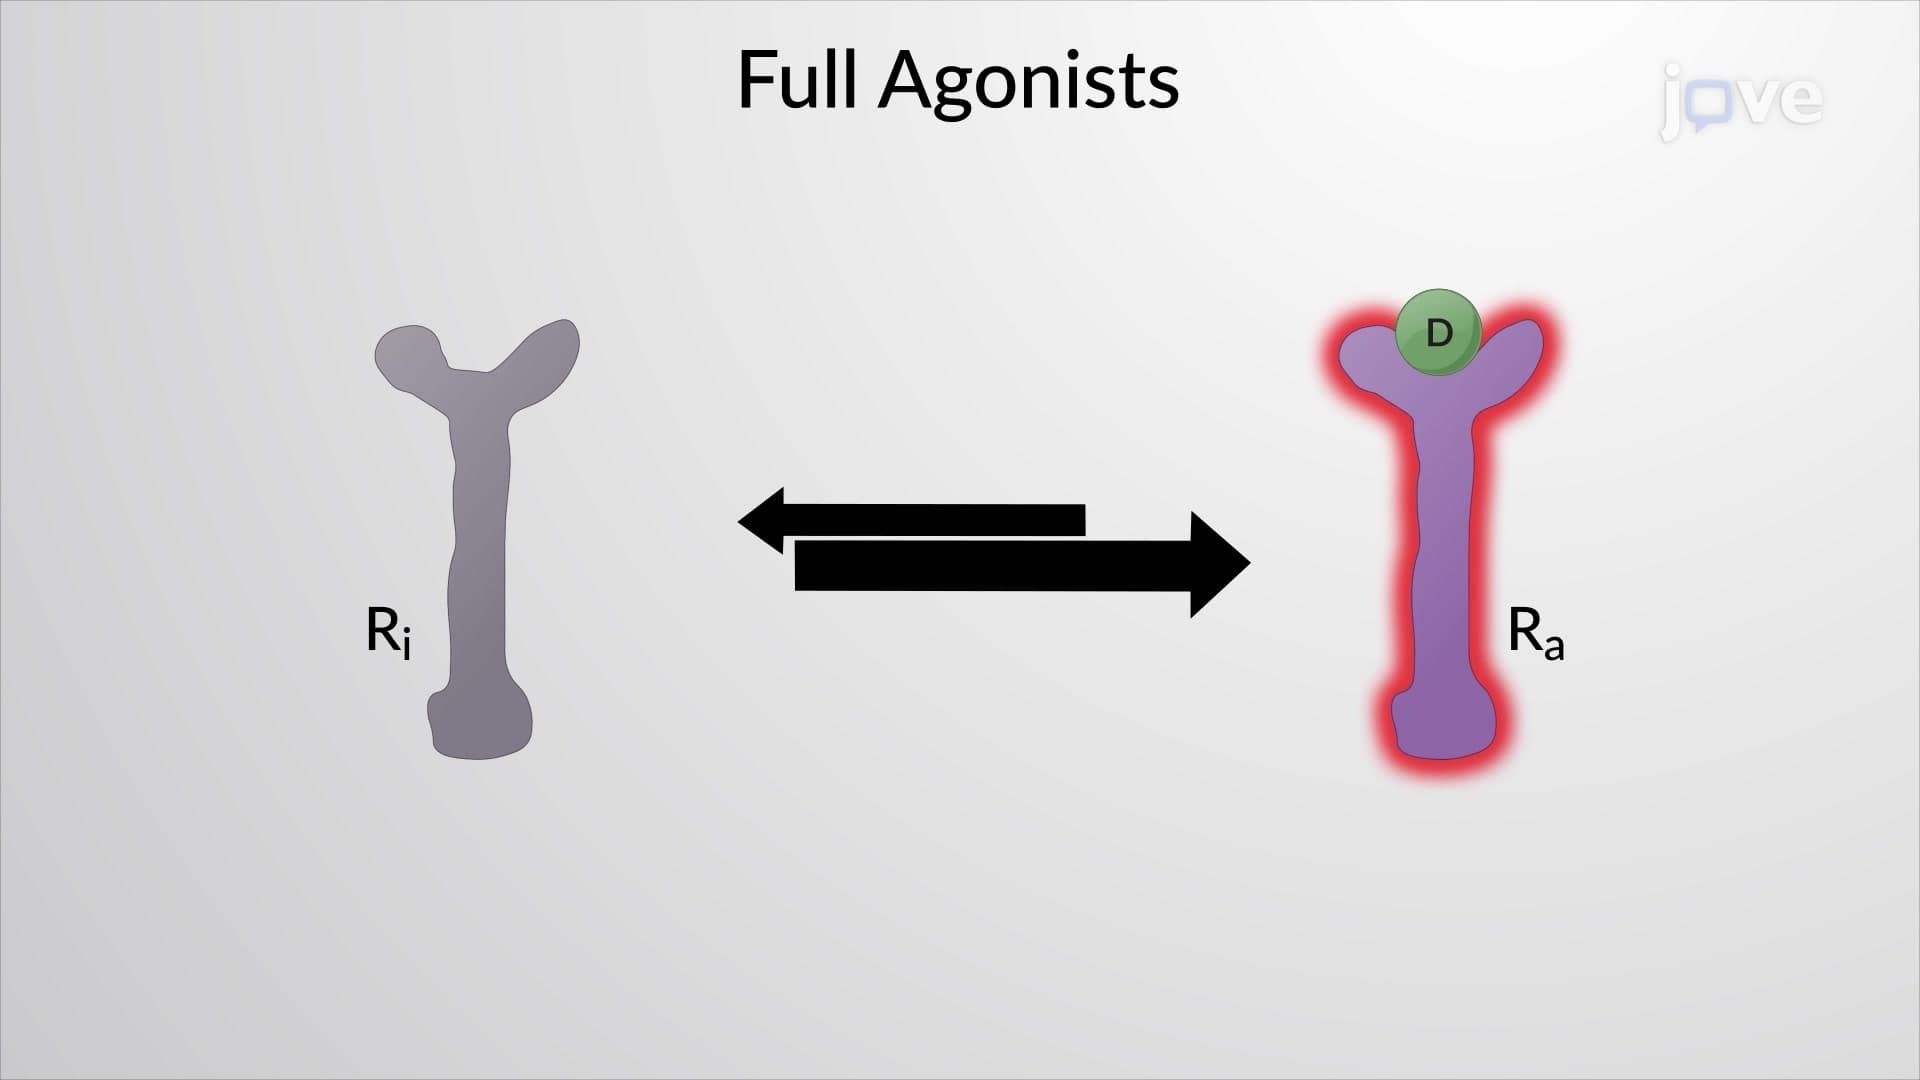
The two-state receptor model explains a drug's interaction with receptors, such as G protein-coupled receptors and ligand-gated ion channels, to induce or inhibit a biological response. When no natural ligands are present, a receptor exists in an equilibrium of inactive (Ri) and active (Ra) conformations. The inactive form does not produce a response, while the active form generates a basal effect known as constitutive activity.
The binding affinity of a drug determines its interaction with...
The binding affinity of a drug determines its interaction with...
1.9K
 01:29
Drug-Receptor Interactions
5.1K
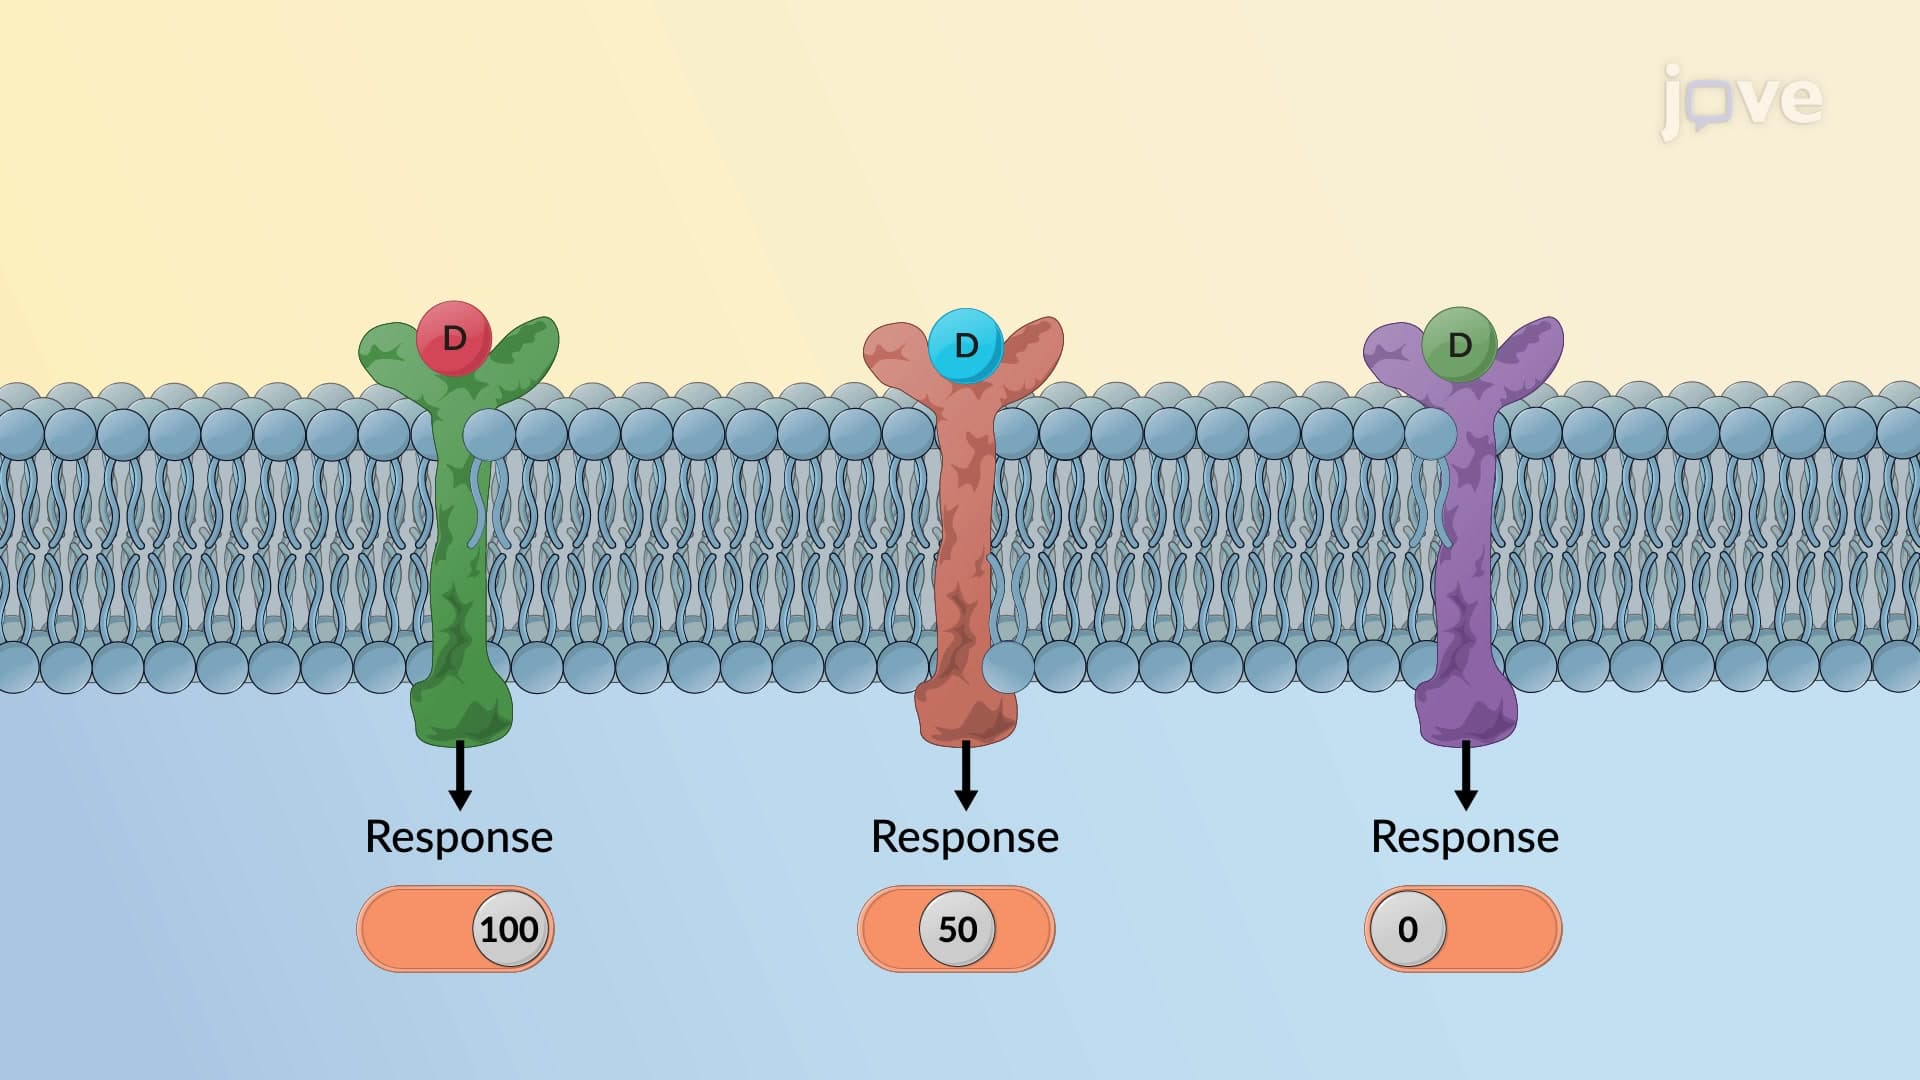
Drug-receptor interaction describes the binding of receptors by drugs, but not all drug-receptor interactions result in activation and tissue response. For instance, the binding of agonists activates the receptor to generate a cellular reaction, while antagonists bind to receptors without causing their activation.
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
5.1K
 01:12
GPCR Desensitization
5.9K
G protein-coupled receptor (GPCR) signaling plays a crucial role in cell functioning. GPCR desensitization is an equally essential process. It allows cells to respond to changing environments and regain sensitivity to new stimuli while preventing unnecessary stimulation when no longer needed. Prolonged exposure to stimuli leads to GPCR desensitization. It involves blocking the receptors from binding and activating additional G proteins. This inhibits activation of downstream effectors, thereby...
5.9K
 01:31
Adrenergic Receptors: ɑ Subtype
1.5K
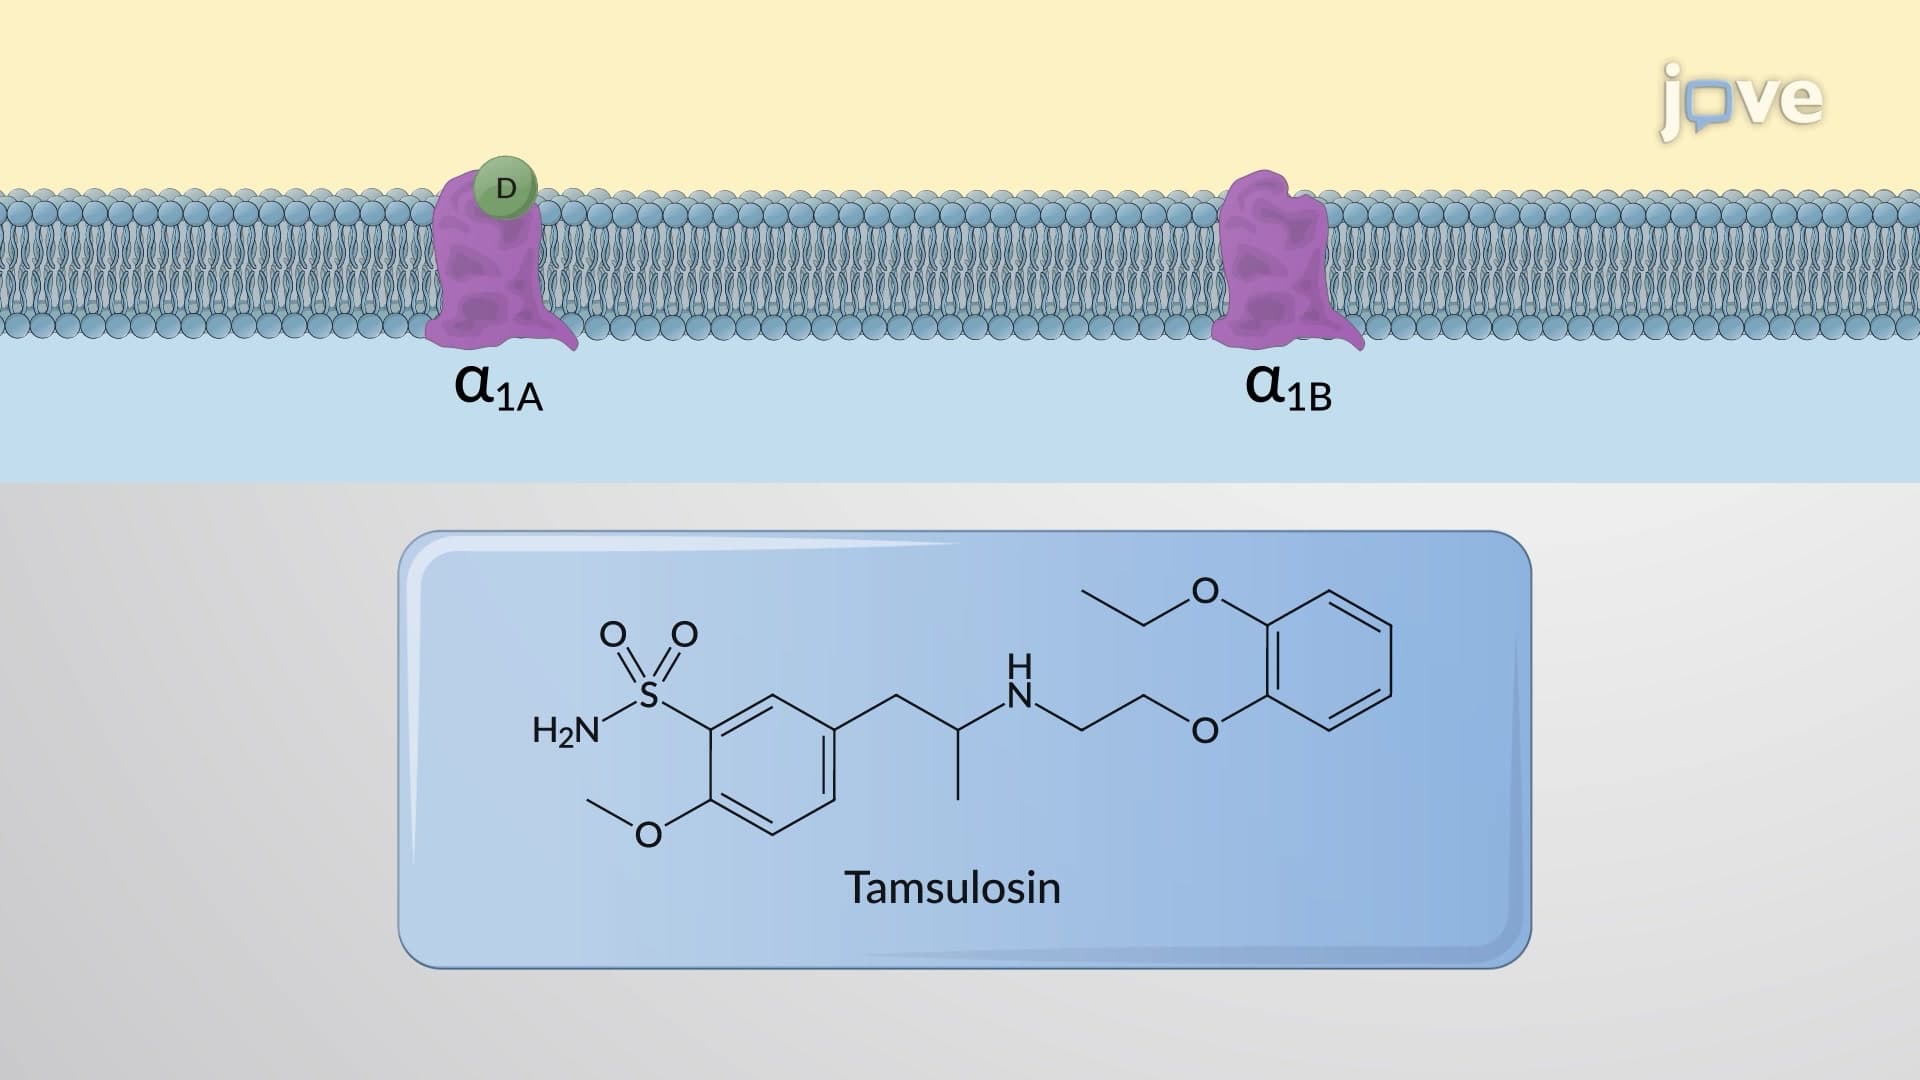
Adrenoceptors are classified into α and ꞵ classes based on their potencies to catecholamine agonists. α-adrenoceptors show the following order of catecholamine potency:
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase...
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase...
1.5K