

在密度函数理论上放置错误条
Simuck F Yuk1, Irmak Sargin2, Noah Meyer3
1Department of Chemistry and Life Science, United States Military Academy, West Point, NY, 10996, USA.
Scientific reports
|August 30, 2024

概括
在密度函数理论 (DFT) 计算中预测错误对于材料科学至关重要. 这项研究使用材料信息学来估计DFT错误,为功能选择提供"错误条",并加速新材料的发现.
科学领域:
- 计算材料科学科学 计算材料科学
- 量子化学 是一个量子化学.
- 材料 信息学 信息学
背景情况:
- 使用密度函数理论 (DFT) 准确预测材料特性,严重依赖于交换相关性 (XC) 函数的选择.
- 估计与不同XC函数相关的先验错误具有挑战性,阻碍了高通量选的高效性.
- 了解功能特定错误对于开发更准确的预测模型至关重要.
研究的目的:
- 开发一种材料信息学方法,用于预测来自不同XC函数的DFT计算中的错误.
- 分析二元氧化物和三元氧化物的常见XC函数 (LDA,PBE-GGA,PBEsol,VDW-DF) 的系统错误.
- 提供定量错误估计 ("错误条") 以指导高通量材料发现的功能选择.
主要方法:
- 用四个不同的XC函数计算二元和三元氧化物的结构和弹性特性.
- 材料信息技术的应用,以分析和预测与每个XC功能相关的系统错误.
- 预测的DFT错误与内在的功能性质的相关性,如电子密度和杂交.
主要成果:
- 预测LDA,PBE-GGA,PBEsol和VDW-DF (具有C09交换) 函数的系统错误.
- 预测的错误被成功地用于改进DFT计算的格子参数.
- 计算错误与函数在描述电子密度和杂交中的表现之间建立了明确的联系.
结论:
- 材料信息学为估计 DFT XC 功能错误提供了一个强大的框架.
- 这些发现为在高通量材料选中选择合适的功能提供了实用的"错误条".
- 这项工作为开发改进的XC功能铺平了道路,并加速了新材料的发现.
更多相关视频

相关概念视频
 02:58
Crystal Field Theory - Octahedral Complexes
26.2K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
26.2K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
41.8K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
41.8K
 01:12
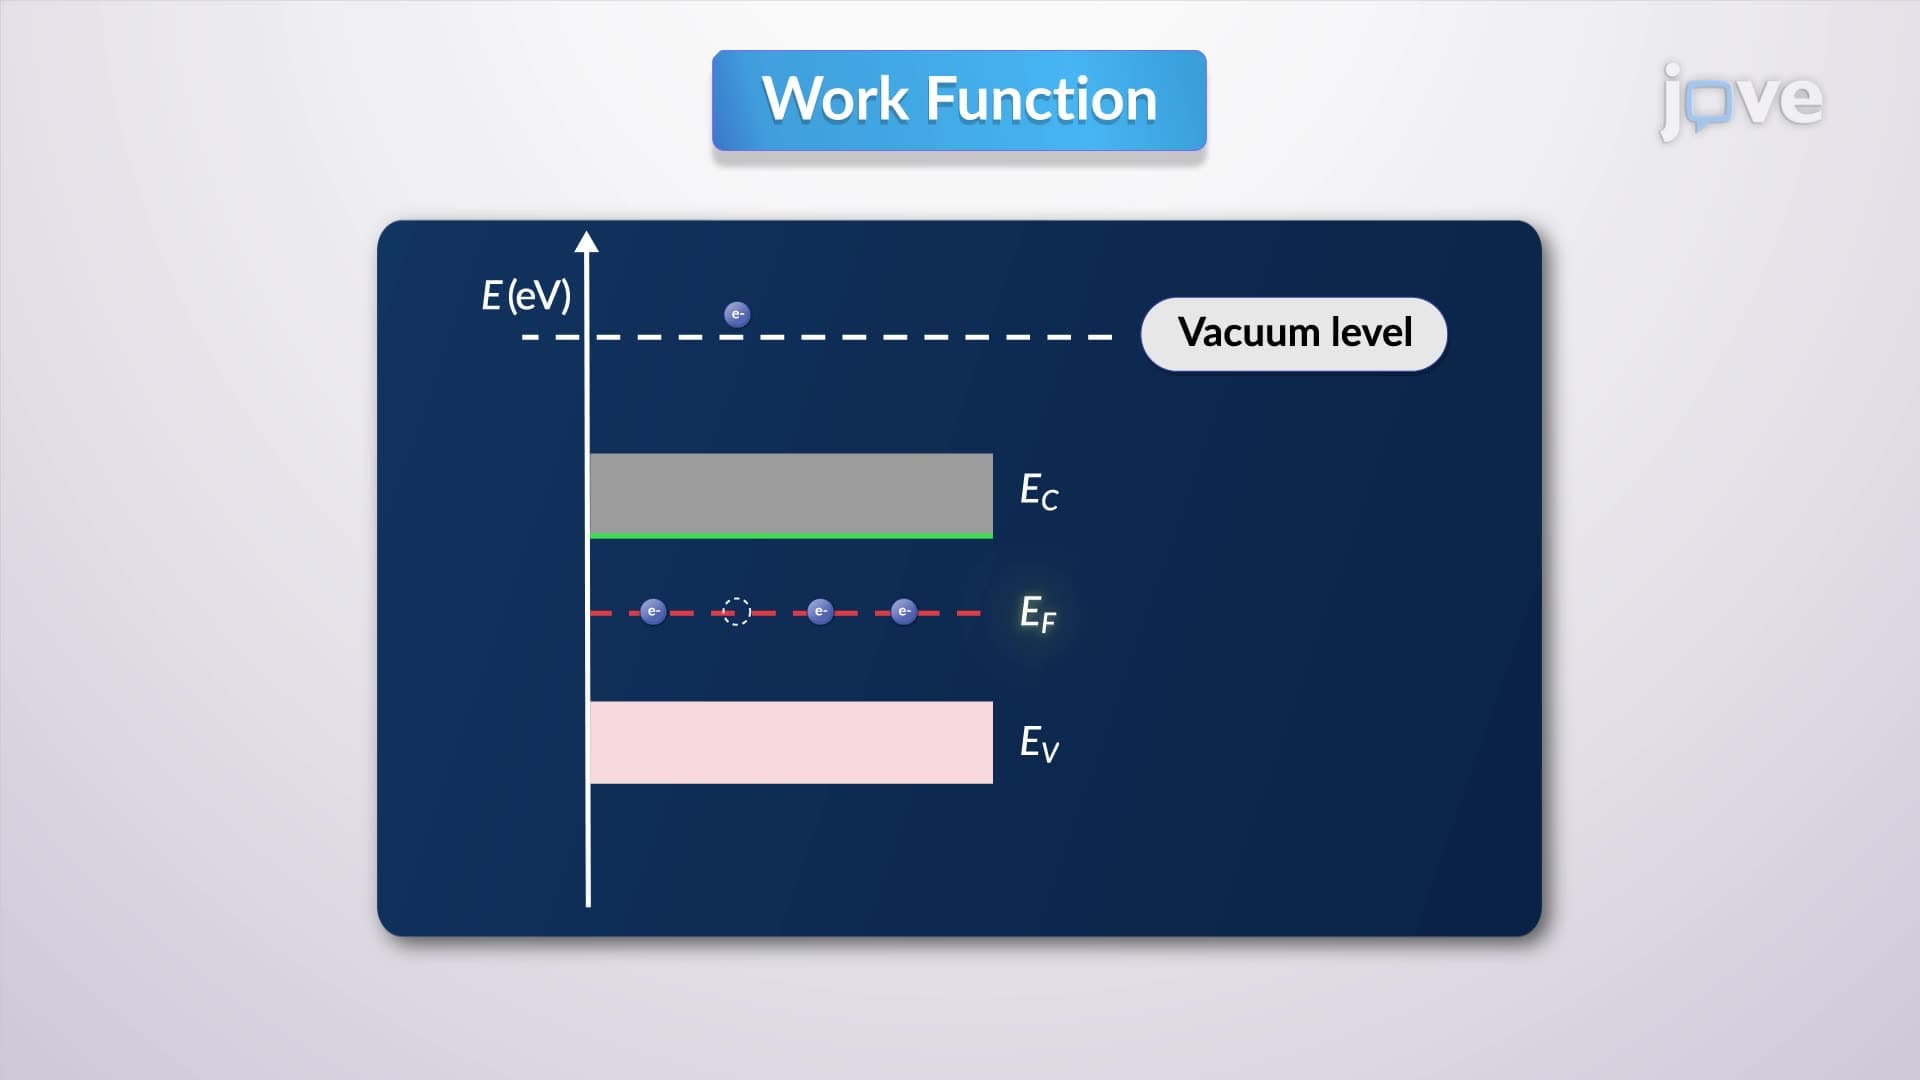
Fermi Level Dynamics
228
The vacuum level denotes the energy threshold required for an electron to escape from a material surface. It is usually positioned above the conduction band of a semiconductor and acts as a benchmark for comparing electron energies within various materials.
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
228
 02:35
Molecular Orbital Theory I
31.8K
Overview of Molecular Orbital Theory
31.8K
 03:06
The Pauli Exclusion Principle
35.7K
The arrangement of electrons in the orbitals of an atom is called its electron configuration. We describe an electron configuration with a symbol that contains three pieces of information:
35.7K
 03:51
Molecular Orbital Theory II
19.0K
Molecular Orbital Energy Diagrams
19.0K