
Ab initio处理分子Coster-Kronig衰变使用复杂尺度的运动方程合集群理论
Jan Philipp Drennhaus1, Anthuan Ferino-Pérez1, Florian Matz1
1Department of Chemistry, KU Leuven, Celestijnenlaan 200F, B-3001 Leuven, Belgium. thomas.jagau@kuleuven.be.
Physical chemistry chemical physics : PCCP
|September 4, 2024

概括
这项研究将分子奥格尔光谱方法扩展到科斯特-克罗尼格衰变,揭示了自旋轨道合对和硫化衰变分支比的显著影响.
科学领域:
- 原子和分子物理 原子和分子物理
- 量子化学 是一个量子化学.
- 频谱学是一种光谱学.
背景情况:
- 科斯特-克罗尼格衰变涉及L1外空缺的非辐射填充,发射低能电子.
- 这个过程类似于奥格衰变,但在电子运动能量方面是不同的.
- 了解这些衰变对于原子和分子电子结构至关重要.
研究的目的:
- 扩展分子奥格尔光谱的计算方法到科斯特-克罗尼格衰变.
- 在和硫化中计算2s-1状态的电离能和衰变宽度.
- 构建和分析L1L2,3M科斯特-克罗尼格和L1MM奥格尔光谱.
主要方法:
- 使用复杂尺度的运动方程合集群理论.
- 扩展以前开发的Auger光谱构造方法.
- 计算电离能,总和部分衰变宽度.
主要成果:
- 成功计算了L1L2,3M科斯特-克罗尼格和L1MM奥格尔工艺的光谱.
- 对整个光谱的现有实验和理论数据取得了良好的一致性.
- 在分支比率中发现了实质性的分歧,特别是关于旋转轨道合效应.
结论:
- 开发的方法对于模拟科斯特-克罗尼格衰变光谱是有效的.
- 旋转轨道合在科斯特-克罗尼格衰变中起着重要作用,即使对于较轻的元素.
- 进一步调查旋转轨道效应是准确预测的必要条件.
更多相关视频
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
6.2K
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
8.4K
相关概念视频
 02:35
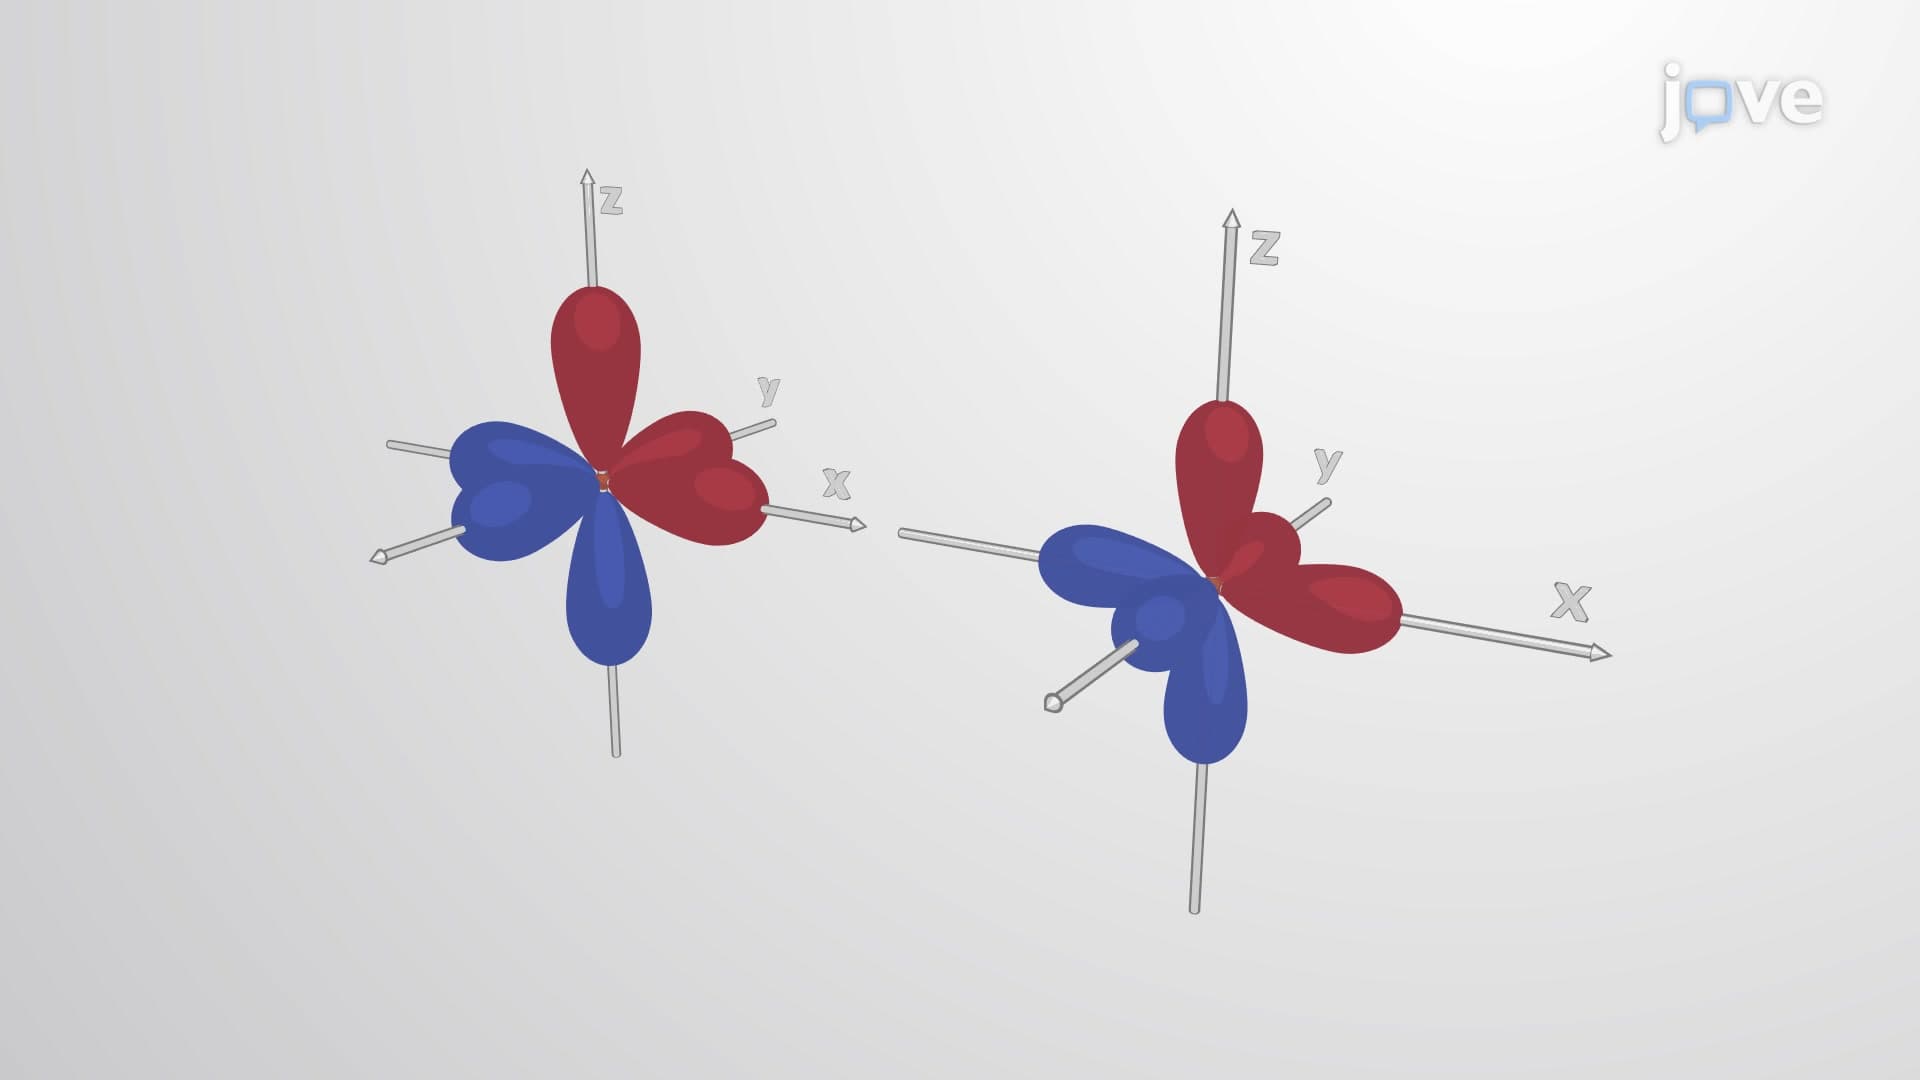
Molecular Orbital Theory I
31.8K
Overview of Molecular Orbital Theory
31.8K
 03:51
Molecular Orbital Theory II
19.0K
Molecular Orbital Energy Diagrams
19.0K
 03:24
Hybridization of Atomic Orbitals I
46.7K
The mathematical expression known as the wave function, ψ, contains information about each orbital and the wavelike properties of electrons in an isolated atom. When atoms are bound together in a molecule, the wave functions combine to produce new mathematical descriptions that have different shapes. This process of combining the wave functions for atomic orbitals is called hybridization and is mathematically accomplished by the linear combination of atomic orbitals. The new orbitals that...
46.7K
 03:35
Hybridization of Atomic Orbitals II
31.9K
sp3d and sp3d 2 Hybridization
31.9K
 01:27
π Electron Effects on Chemical Shift: Overview
1.1K
An applied magnetic field causes loosely bound π-electrons in organic molecules to circulate, producing a local or induced diamagnetic field over a large spatial volume. As the molecules tumble in solution, the field generated by π-electrons in spherical substituents results in a zero net field. However, the net field generated by π-electrons in non-spherical substituents is not zero. The effect of this induced field depends on the orientation of the molecule with respect to B0,...
1.1K
 02:40
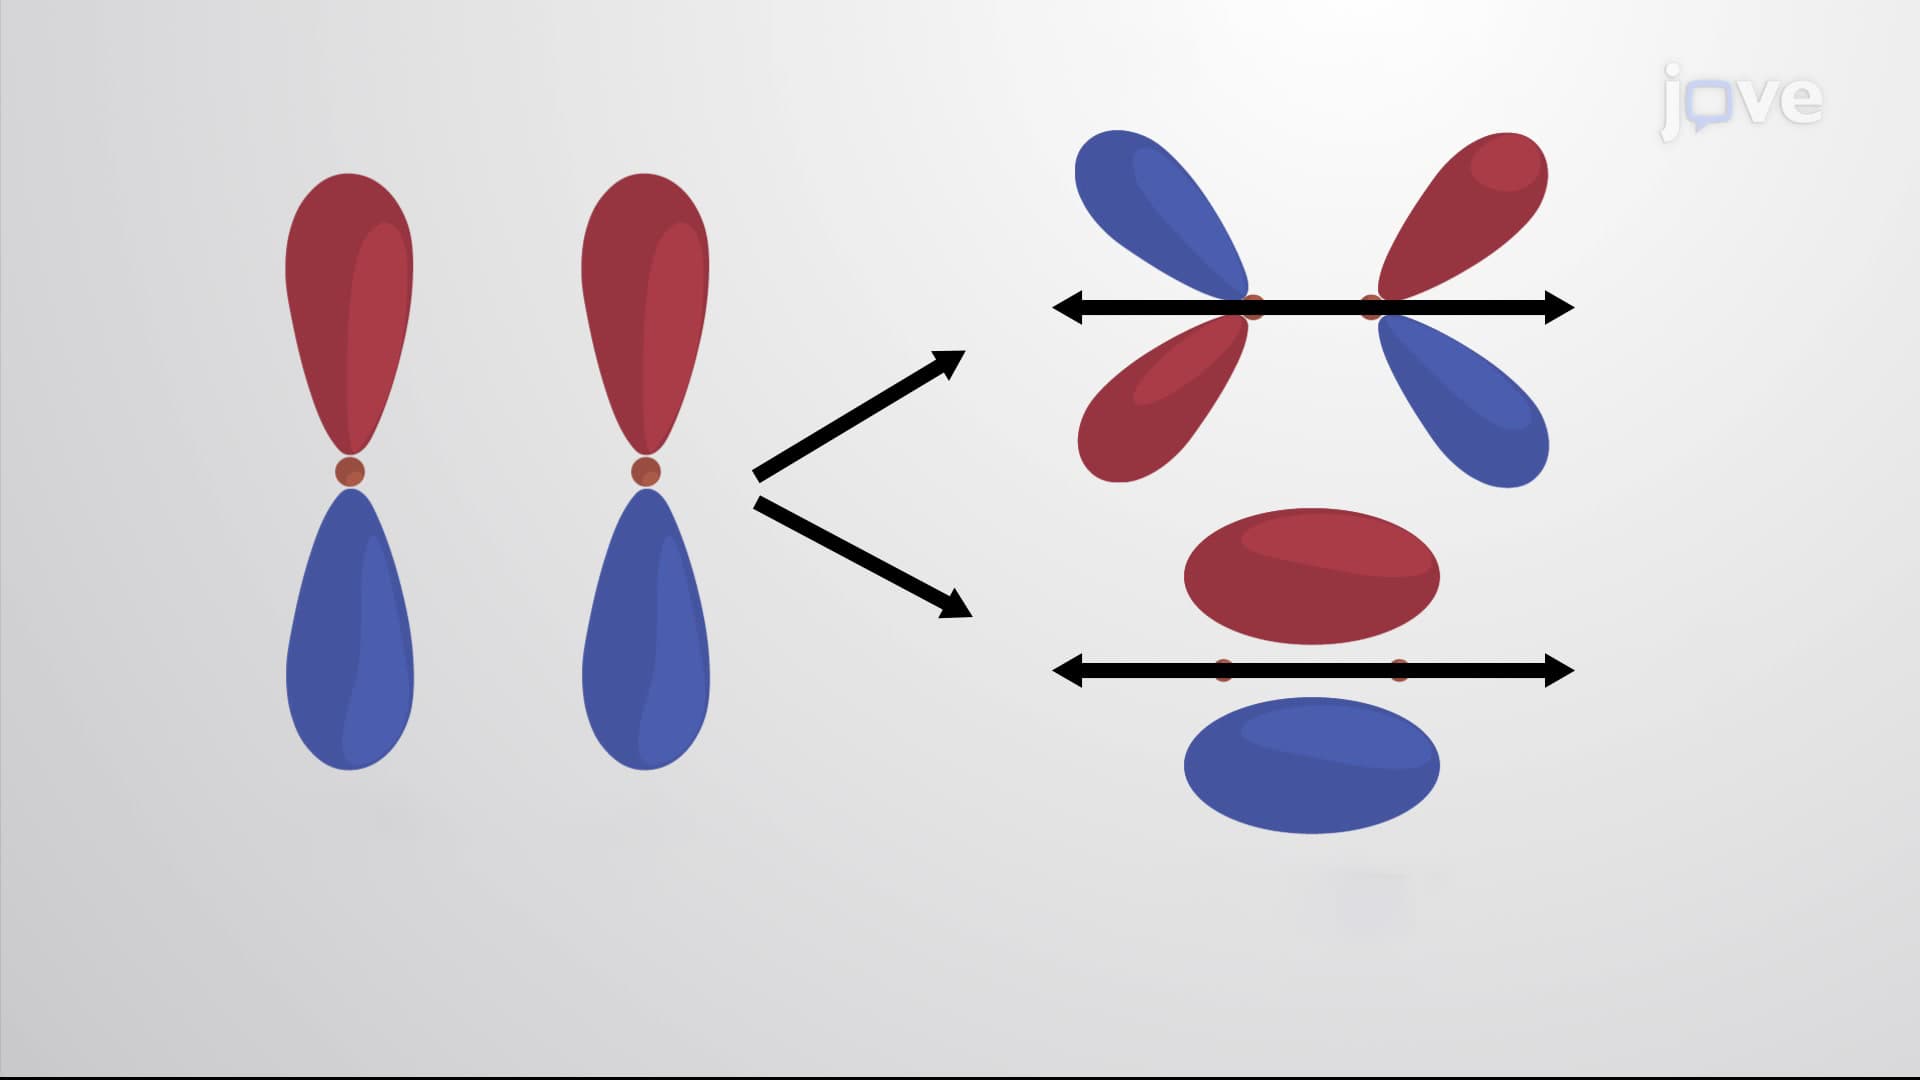
MO Theory and Covalent Bonding
10.3K
The molecular orbital theory describes the distribution of electrons in molecules in a manner similar to the distribution of electrons in atomic orbitals. The region of space in which a valence electron in a molecule is likely to be found is called a molecular orbital. Mathematically, the linear combination of atomic orbitals (LCAO) generates molecular orbitals. Combinations of in-phase atomic orbital wave functions result in regions with a high probability of electron density, while...
10.3K