相关实验视频
Updated: Jun 13, 2025
09:40

Novel Sequence Discovery by Subtractive Genomics
Published on: January 25, 2019
8.6K
大木的基因组序列,Luzula sylvatica (胡德斯) 高丁 高丁
Zoë A Goodwin1, David Bell1, Michelle L Hart1
1Royal Botanic Garden Edinburgh, Edinburgh, Scotland, UK.
Wellcome open research
|September 9, 2024

概括
我们介绍了大木草 (Luzula sylvatica) 的基因组组合,这是了解植物基因组学的重要一步. 这种组合为未来在Juncaceae家族的遗传研究提供了基础资源.
科学领域:
- 基因组学就是基因组学.
- 植物科学 植物科学
- 生物信息学是一种生物信息学.
背景情况:
- 科 (Juncaceae) 家族,包括科 (Luzula sylvatica),是具有多种生态作用的单植物的一个重要群体.
- 基因组测序为植物进化,适应和遗传多样性提供了关键的见解.
- 高质量的基因组组合对于比较基因组学和功能研究至关重要.
研究的目的:
- 为了生成Luzula sylvatica (大木草) 的高质量基因组组.
- 为Juncaceae家族提供染色体级的基因组资源.
- 为了促进未来研究Luzula sylvatica的遗传构成和进化历史.
主要方法:
- 一个个体的Luzula sylvatica标本的全基因组测序.
- 基因组序列的生物信息组装和支架.
- 核,线粒体和塑体基因组的分析和表征.
主要成果:
- 产生了一个跨越444.5兆基的基因组组件.
- 该组件在很大程度上被支架成6种染色体伪分子.
- 线粒体和塑体基因组的组合长度分别为633.36千基和201.32千基.
结论:
- 提出的基因组组合代表了Luzula sylvatica的宝贵基因组资源.
- 这次集会将有助于理解波尔氏体内遗传结构和进化关系.
- 这种基因组序列的可用性支持未来的功能基因组学和大木草的保护工作.
相关概念视频
 02:54
Evolutionary Relationships through Genome Comparisons
5.7K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
5.7K
 00:58
Genomic DNA in Eukaryotes
46.8K
Eukaryotes have large genomes compared to prokaryotes. To fit their genomes into a cell, eukaryotic DNA is packaged extraordinarily tightly inside the nucleus. To achieve this, DNA is tightly wound around proteins called histones, which are packaged into nucleosomes that are joined by linker DNA and coil into chromatin fibers. Additional fibrous proteins further compact the chromatin, which is recognizable as chromosomes during certain phases of cell division.
46.8K
 01:51
Lampbrush Chromosomes
2.4K
2.4K
 01:05
Maxam-Gilbert Sequencing
11.1K
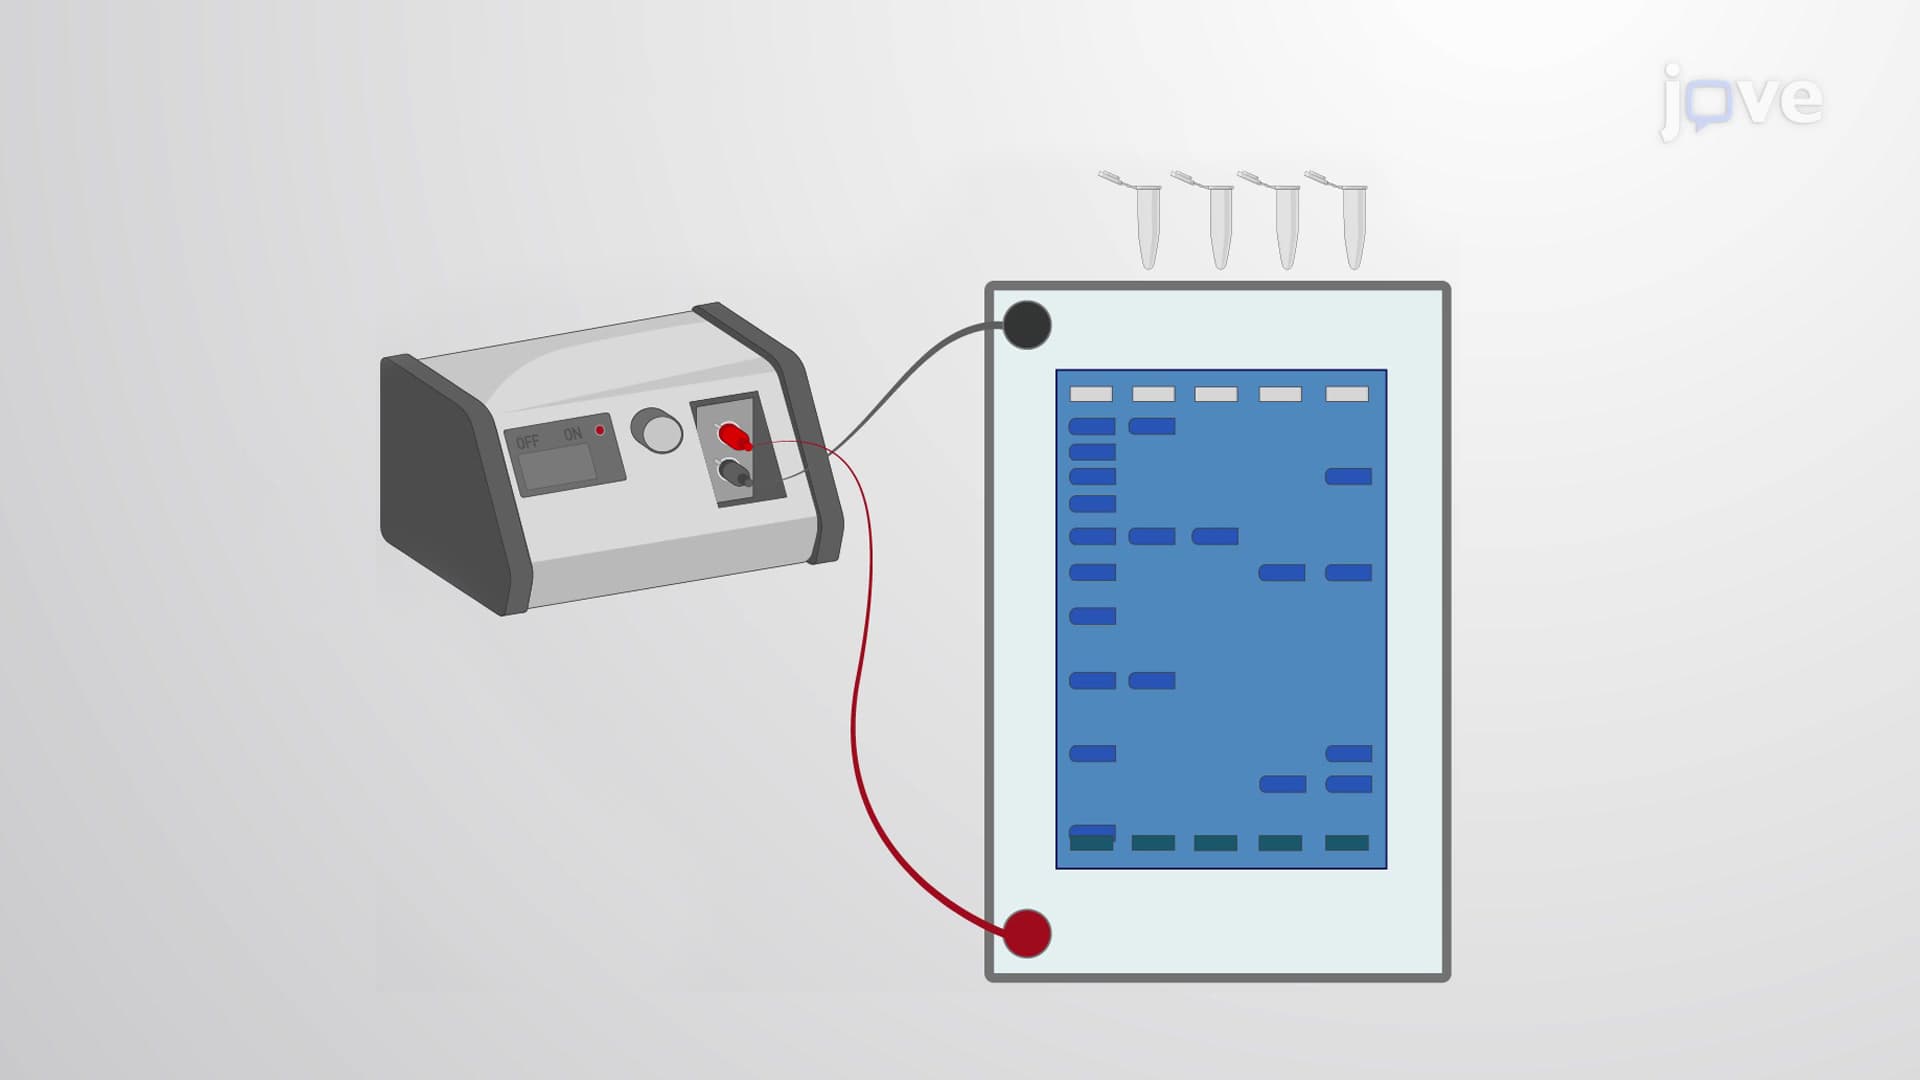
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
11.1K
 02:51
Multi-species Conserved Sequences
3.9K
Next-generation sequencing technologies have created large genomic databases of a variety of animals and plants. Ever since the human genome project was completed, scientists studied the genome of primates, mammals, and other phylogenetically distant living beings. Such large-scale studies have provided new insights into the evolutionary relationship between organisms.
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
3.9K
 02:35
RACE - Rapid Amplification of cDNA Ends
6.3K
Rapid Amplification of cDNA Ends, or RACE, is one of the most effective methods to obtain a full-length cDNA from an mRNA sequence between a known internal region to the unknown sequence at the 5’ or 3’ end. The unknown region is cloned in the cDNA by a gene-specific primer that binds the known end, and a hybrid primer that attaches a predefined anchor sequence to the unknown end of the cDNA. The sequence in between is amplified by PCR with an anchor primer and a gene-specific...
6.3K