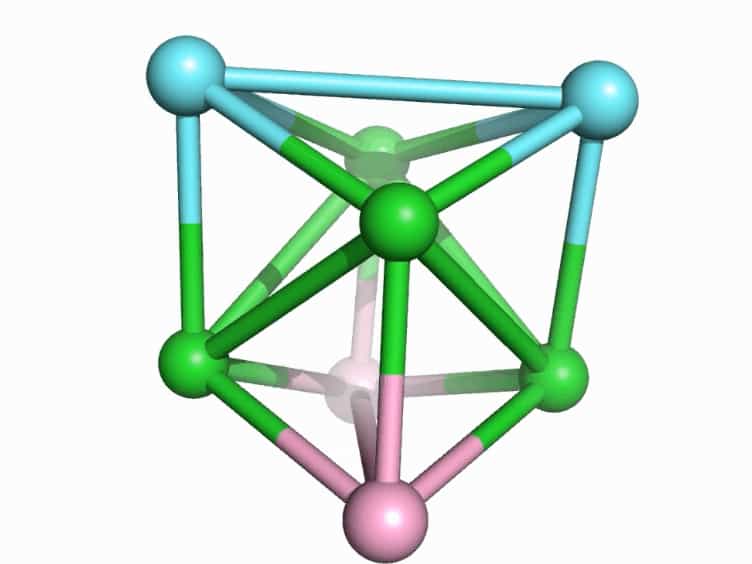

一个DFT计算研究I型酸盐A8Sn46-x (A = Cs或NH4,x = 0或2) 的DFT计算研究
Nikolaos Kelaidis1, Emmanuel Klontzas1, Andreas Kaltzoglou1
1Theoretical and Physical Chemistry Institute, National Hellenic Research Foundation, 11635 Athens, Greece.
Materials (Basel, Switzerland)
|September 28, 2024

概括
计算研究揭示了锡 (Sn) 酸盐的缺陷显著改变了电气性能,使得可调节的半导体行为. 首选的缺陷配置导致适合热电应用的窄带间隙材料.
科学领域:
- 材料科学 材料科学 材料科学
- 固态物理 固态物理
- 计算化学的计算化学
背景情况:
- 半导体聚合物对热电材料来说是有前途的.
- 了解结构属性关系对于优化热电性能至关重要.
研究的目的:
- 研究I型酸盐的晶体结构,形成度和物理性质.
- 探索缺陷对-锡和-锡克拉特酸盐的热电性质的影响.
主要方法:
- 开始的VASP计算用于几何优化和属性计算.
- 分析电子和声密度的状态.
- 电子定位函数用于化学键的比较.
主要成果:
- 在Sn框架中缺陷的存在和分布显著改变了形成能量和电特性 (从金属到半导体).
- 特定的缺陷配置 (每六个成员的Sn环中的一个) 是热力学上首选的,产生狭窄带间隙半导体行为.
- 酸在腔中的旋转可能会降低晶格导热率.
结论:
- 锡酸盐的缺陷工程为调整热电特性提供了一条途径.
- 确定了首选的缺陷结构和的客体动力学,突出了先进热电材料的潜力.
相关概念视频
 02:42
Ionic Crystal Structures
14.2K
Ionic crystals consist of two or more different kinds of ions that usually have different sizes. The packing of these ions into a crystal structure is more complex than the packing of metal atoms that are the same size.
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
14.2K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
41.7K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
41.7K
 02:58
Crystal Field Theory - Octahedral Complexes
26.2K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
26.2K
 02:37
Metallic Solids
18.3K
Metallic solids such as crystals of copper, aluminum, and iron are formed by metal atoms. The structure of metallic crystals is often described as a uniform distribution of atomic nuclei within a “sea” of delocalized electrons. The atoms within such a metallic solid are held together by a unique force known as metallic bonding that gives rise to many useful and varied bulk properties.
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
18.3K
 02:57
Coordination Number and Geometry
15.6K
For transition metal complexes, the coordination number determines the geometry around the central metal ion. Table 1 compares coordination numbers to molecular geometry. The most common structures of the complexes in coordination compounds are octahedral, tetrahedral, and square planar.
15.6K
 02:42
Valence Bond Theory
8.5K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
8.5K