
萨法里:使用氨酸/氨酸编码的古代DNA的泛基因组对齐
Joshua Rubin1, Jan van Waaij1, Louis Kraft1
1Department of Health Technology, Section for Bioinformatics, Technical University of Denmark, Kongens Lyngby, Denmark.
bioRxiv : the preprint server for biology
|October 17, 2024

概括
由于损伤和短序,调整古代DNA是很困难的. 我们的新RYmer指数和SAFARI工具提高了化石的对齐精度,增强了遗传洞察力.
科学领域:
- 古遗传学是古遗传学的一部分.
- 生物信息学是一种生物信息学.
- 计算生物学 计算生物学
背景情况:
- 由于短序列,化学损伤 (C→T,G→A替代) 和参考偏差,古代DNA (aDNA) 的对齐具有挑战性.
- 泛基因组图减轻了一些偏差,但仍然受到aDNA特定替代模式的影响.
研究的目的:
- 开发一种用于强大的古老DNA序列对齐的新方法.
- 为了提高古代DNA分析和下游应用的准确性.
主要方法:
- 引入RYmer指数,这是一个修改后的最小化指数,代表纯氨酸 (R) 和氨酸 (Y),以抵抗化学损伤.
- 实施SAFARI,这是使用RYmers.mers的vg长鹿基因组对齐器的古老DNA损伤感知版本.
- 评估SAFARI的性能与使用古代DNA序列的当前对齐方法的对比.
主要成果:
- 与现有的方法相比,SAFARI可以从古老的DNA序列中产生更准确的对齐.
- 瑞默指数有效地挽救了受脱氧化影响的对齐,这是DNA损伤的常见形式.
- 该算法改善了对古代DNA损伤率的估计,特别是在高度降解的样本中.
结论:
- 莱默指数和SAFARI在古代DNA对齐方面取得了重大进展.
- 改进的对齐精度转化为从古生物学数据中获得更好的洞察力.
- 萨法里可以与现有的 pangenome 工具无集成,扩大其应用范围.
相关概念视频
 02:05
DNA as a Genetic Template
21.8K
Two structural features of the DNA molecule provide a basis for the mechanisms of heredity: the four nucleotide bases and its double-stranded nature. The Watson-Crick model of double-helical DNA structure, proposed in 1952, drew heavily upon the X-ray crystallography work of researchers Rosalind Franklin and Maurice Wilkins. Watson, Crick, and Wilkins jointly received the Nobel Prize in Physiology or Medicine for their work in 1962. Franklin was, controversially, excluded from the prize for...
21.8K
 02:27
DNA Base Pairing
27.1K
Erwin Chargaff’s rules on DNA equivalence paved the way for the discovery of base pairing in DNA. Chargaff’s rules state that in a double-stranded DNA molecule,
27.1K
 02:54
Evolutionary Relationships through Genome Comparisons
5.7K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
5.7K
 01:57
Sanger Sequencing
753.7K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
753.7K
 01:05
Maxam-Gilbert Sequencing
11.1K
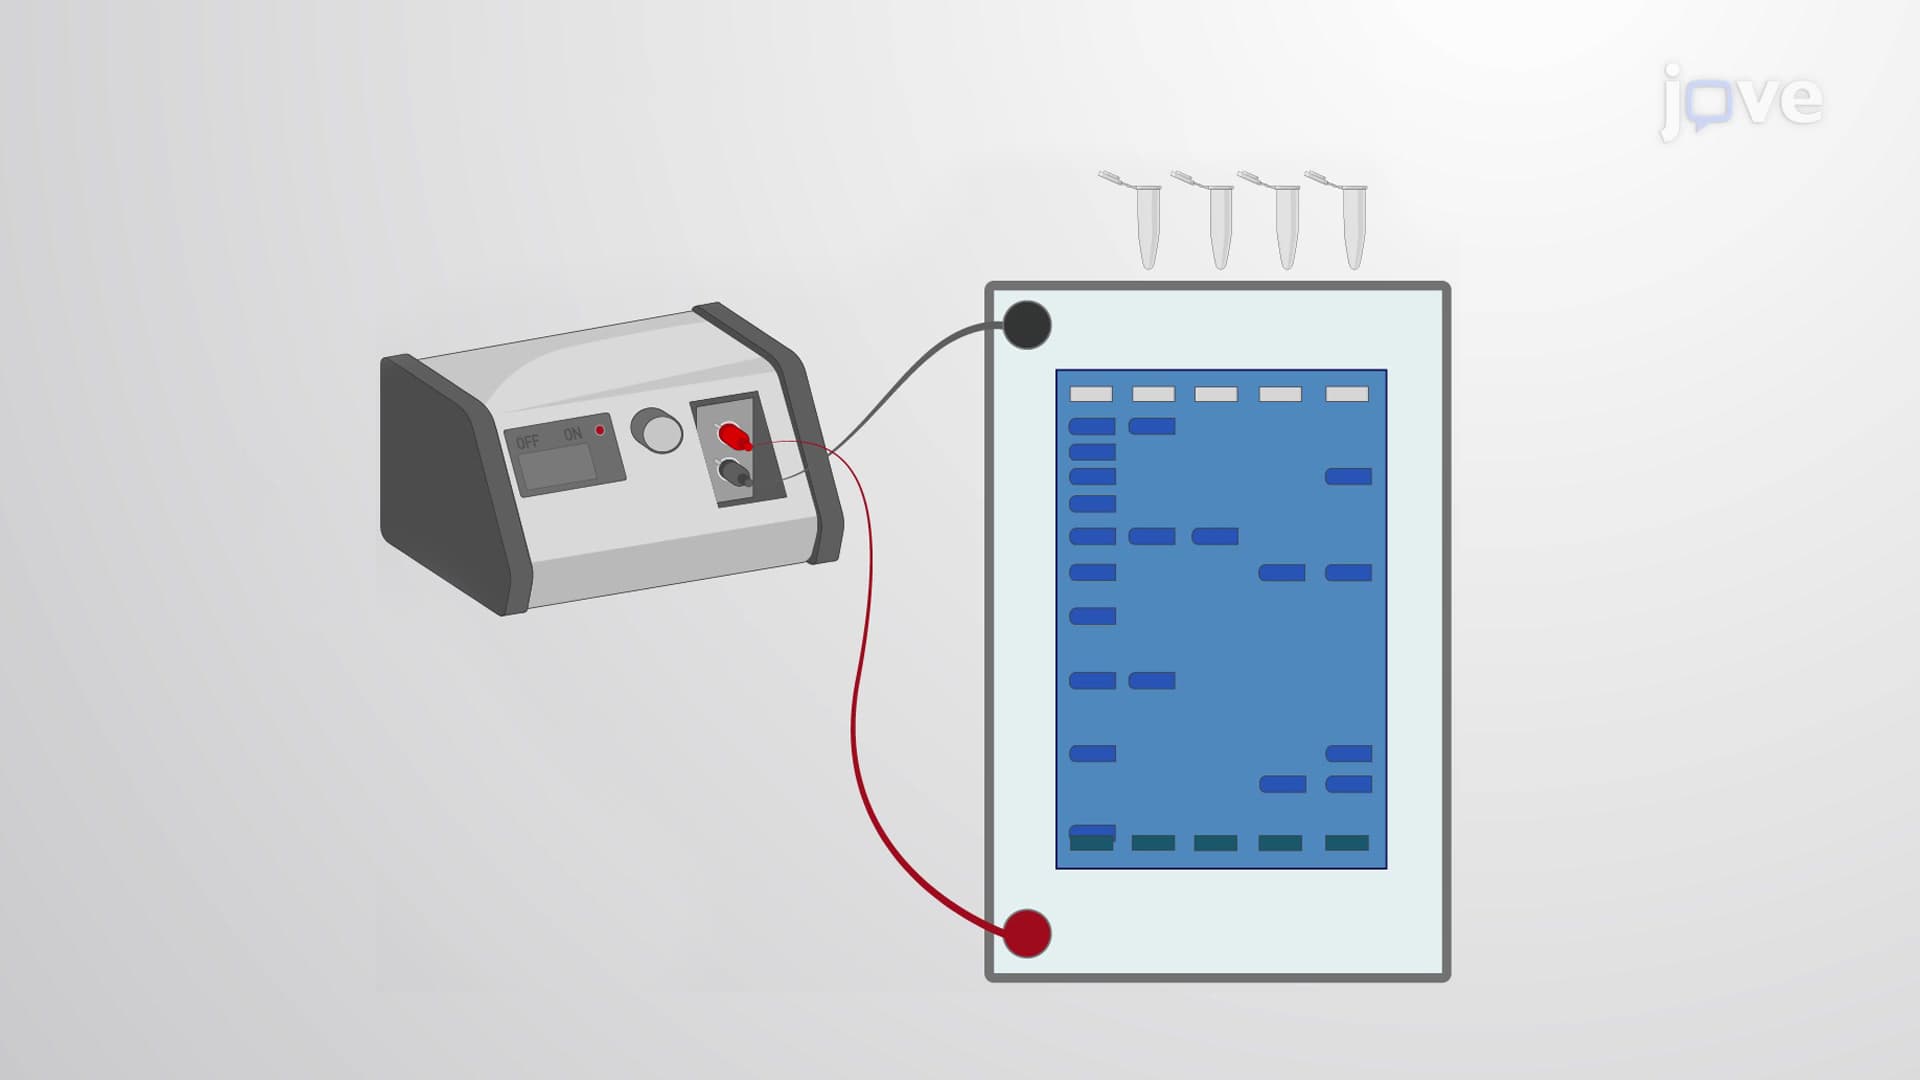
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
11.1K
 03:00
Next-generation Sequencing
88.3K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
88.3K