
在高精度热化学的背景下,降级CCSD的前景 (T)
Tingting Zhao1, James H Thorpe1, Devin A Matthews1
1Department of Chemistry, Southern Methodist University, Dallas, Texas 75275, USA.

对分子进行精确的反应能量计算是具有挑战性的. 结合集群单双三倍 (CCSD(T)) 计算的新近似方法显著降低了计算成本,同时保持了高精度.
科学领域:
- 量子化学是一种量子化学.
- 计算化学是一种计算化学.
- 热化学建模 热化学建模
背景情况:
- 对于中型气相分子中的反应能量,达到亚化学精度 (1 kJ mol-1) 仍然是一个重大挑战.
- 扰动三倍校正 (通常标记为 (T)) 到合集群单双三倍校正 (CCSD ((T)) 对于高精度的热化学建模至关重要,但由于计算成本高 (O ((N7) 缩放) 而受到影响.
研究的目的:
- 开发和评估 (T) 校正的新近似方法,以降低计算成本.
- 评估中型分子 (T) 校正的五种不同近似的准确性.
- 确定保持高准确度,同时显著降低计算需求的近似值.
主要方法:
- 对 (T) 校正的五种近似方法的评估.
- 将这些方法应用于W4-17数据集中的分子子集.
- 将近的 (T) 贡献与正规的,密度相适应的 (T) 结果进行比较.
主要成果:
- 所有经过测试的近似方法与使用最小数量的投影仪的正规密度 (T) 贡献相比,都实现了低于0.1kJ mol-1的精度.
- Z̃T近似证明了计算成本和准确性之间的最佳平衡.
- 确定了高精度模型化学的CCSD (T) 组件计算成本的潜在数量级降低.
结论:
- 对于 (T) 校正的近似方法可以实现对反应能量的高精度.
- 对于加速高精度的热化学计算,Z̃T近似显示出显著的希望.
- 这些发现可以使更大的分子系统更有效地计算反应能量.
更多相关视频
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
相关概念视频
 01:16
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in...
 01:10
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 02:55
Thermochemical Equations
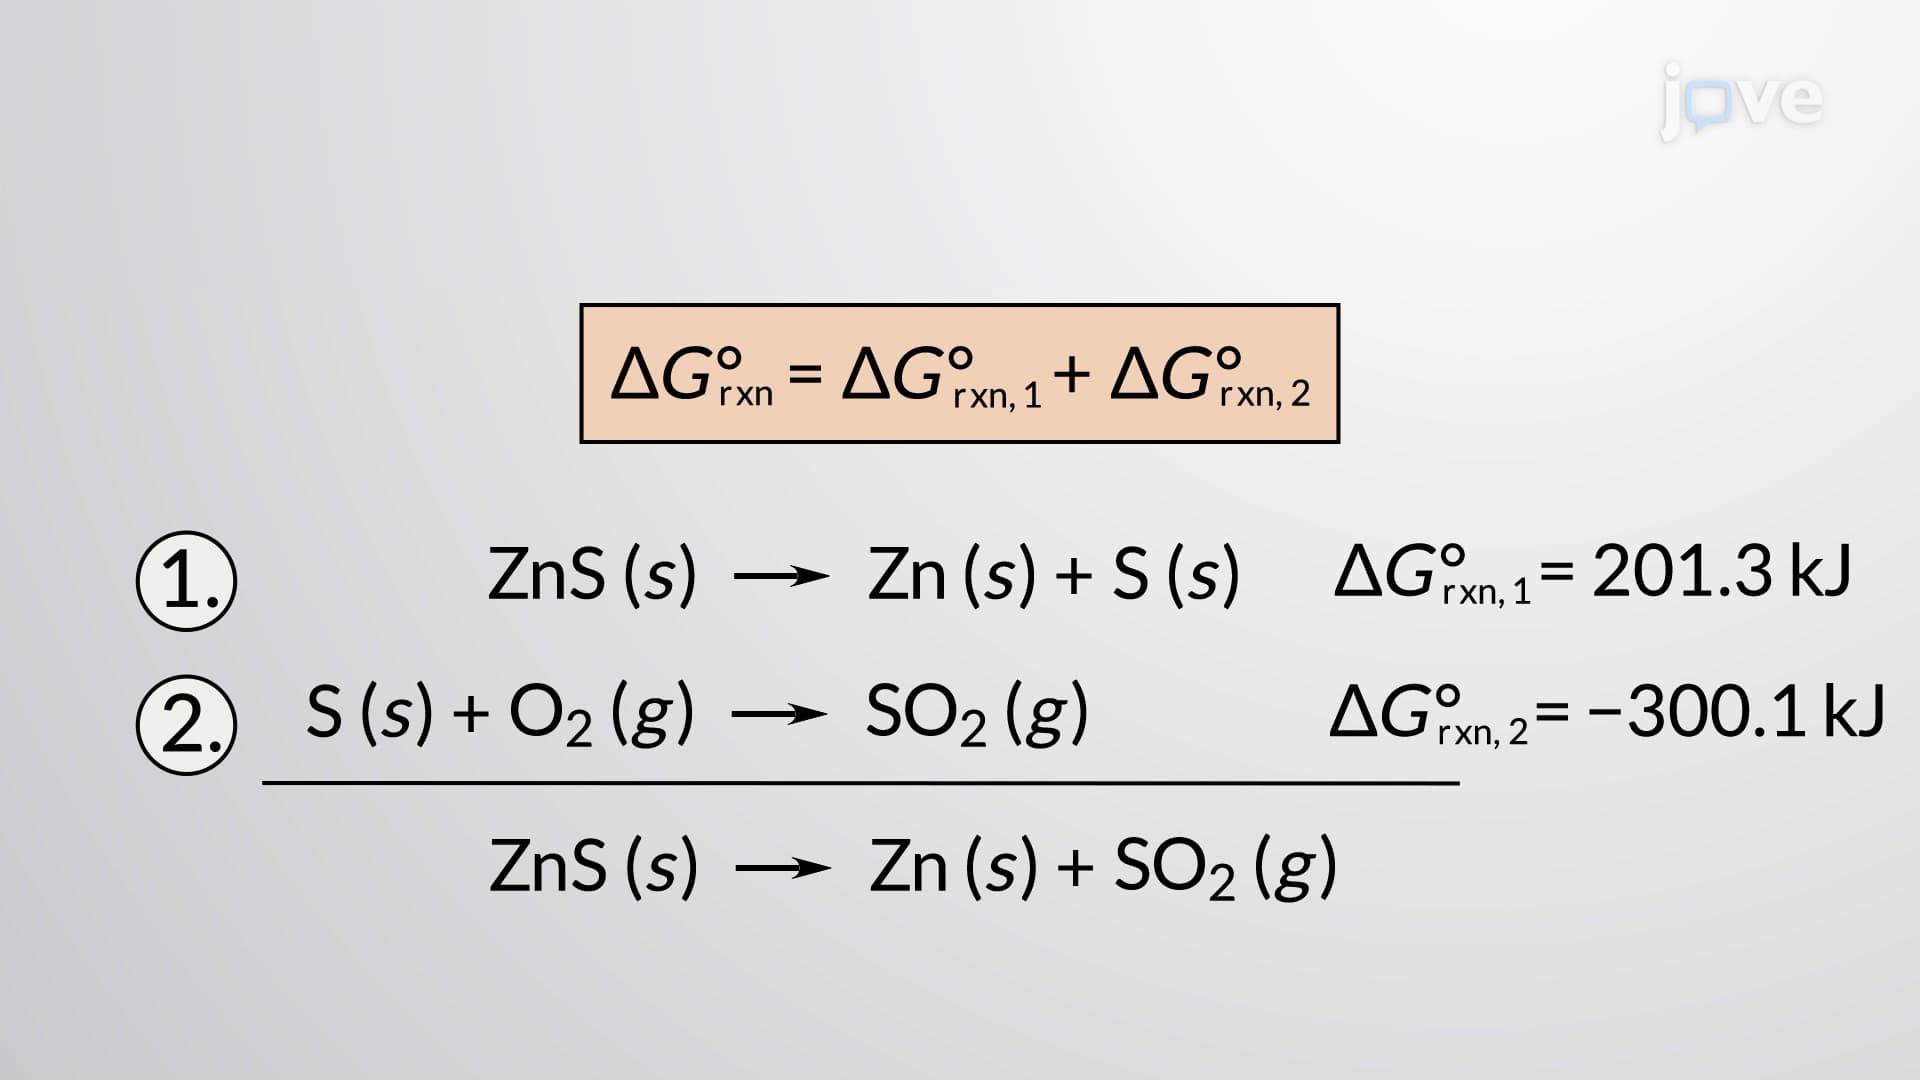 02:49
Calculating Standard Free Energy Changes
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:27
Predicting Molecular Geometry