

由于伪电位和交换相关函数不兼容,DFT计算的固体机械和结构性质的不确定性
1Institute of Fundamental Technological Research, Polish Academy of Sciences, Pawińskiego 5B, 02-106 Warsaw, Poland.
Journal of chemical theory and computation
|October 25, 2024

概括
常见的伪电位可能会在晶体属性计算中引入错误. 本研究评估了密度函数理论 (DFT) 计算中流行的交换相关性 (XC) 函数的准确性.
科学领域:
- 计算材料科学科学 计算材料科学
- 固态物理 固态物理
- 量子化学 是一个量子化学.
背景情况:
- 伪潜在的产生是有限的,大多数可用于LDA和PBE功能.
- 存在各种各样的交换相关性 (XC) 函数,但并非所有函数都有相应的伪潜值.
- 不一致的伪潜在使用可能导致物质性质预测的不准确性.
研究的目的:
- 为了量化机械和结构性质的误差,由于DFT计算中的伪潜在不一致性.
- 为了评估常用的XC函数 (LDA,PBE,PBEsol,SCAN) 的性能,用于晶体属性计算.
主要方法:
- 密度函数理论 (DFT) 的计算.
- 对晶体属性的系统评估,包括晶格常数,凝聚能,表面能,弹性常数和散装模量.
- 不同XC函数的比较分析.
主要成果:
- 在机械和结构性质中识别和量化错误,这些错误来自伪潜力不匹配.
- 提供了LDA,PBE,PBEsol和SCAN功能的一致性性能评估.
- 强调了 XC 功能选择对 DFT 预测准确性的影响.
结论:
- 伪潜在的可用性显著影响DFT计算的材料属性的可靠性.
- 选择XC功能对于准确预测晶体机械和结构特征至关重要.
- 为选择适当的XC函数用于DFT研究提供了建议.
更多相关视频
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
7.6K
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
12.7K
相关概念视频
 01:13
Force and Potential Energy in One Dimension
5.3K
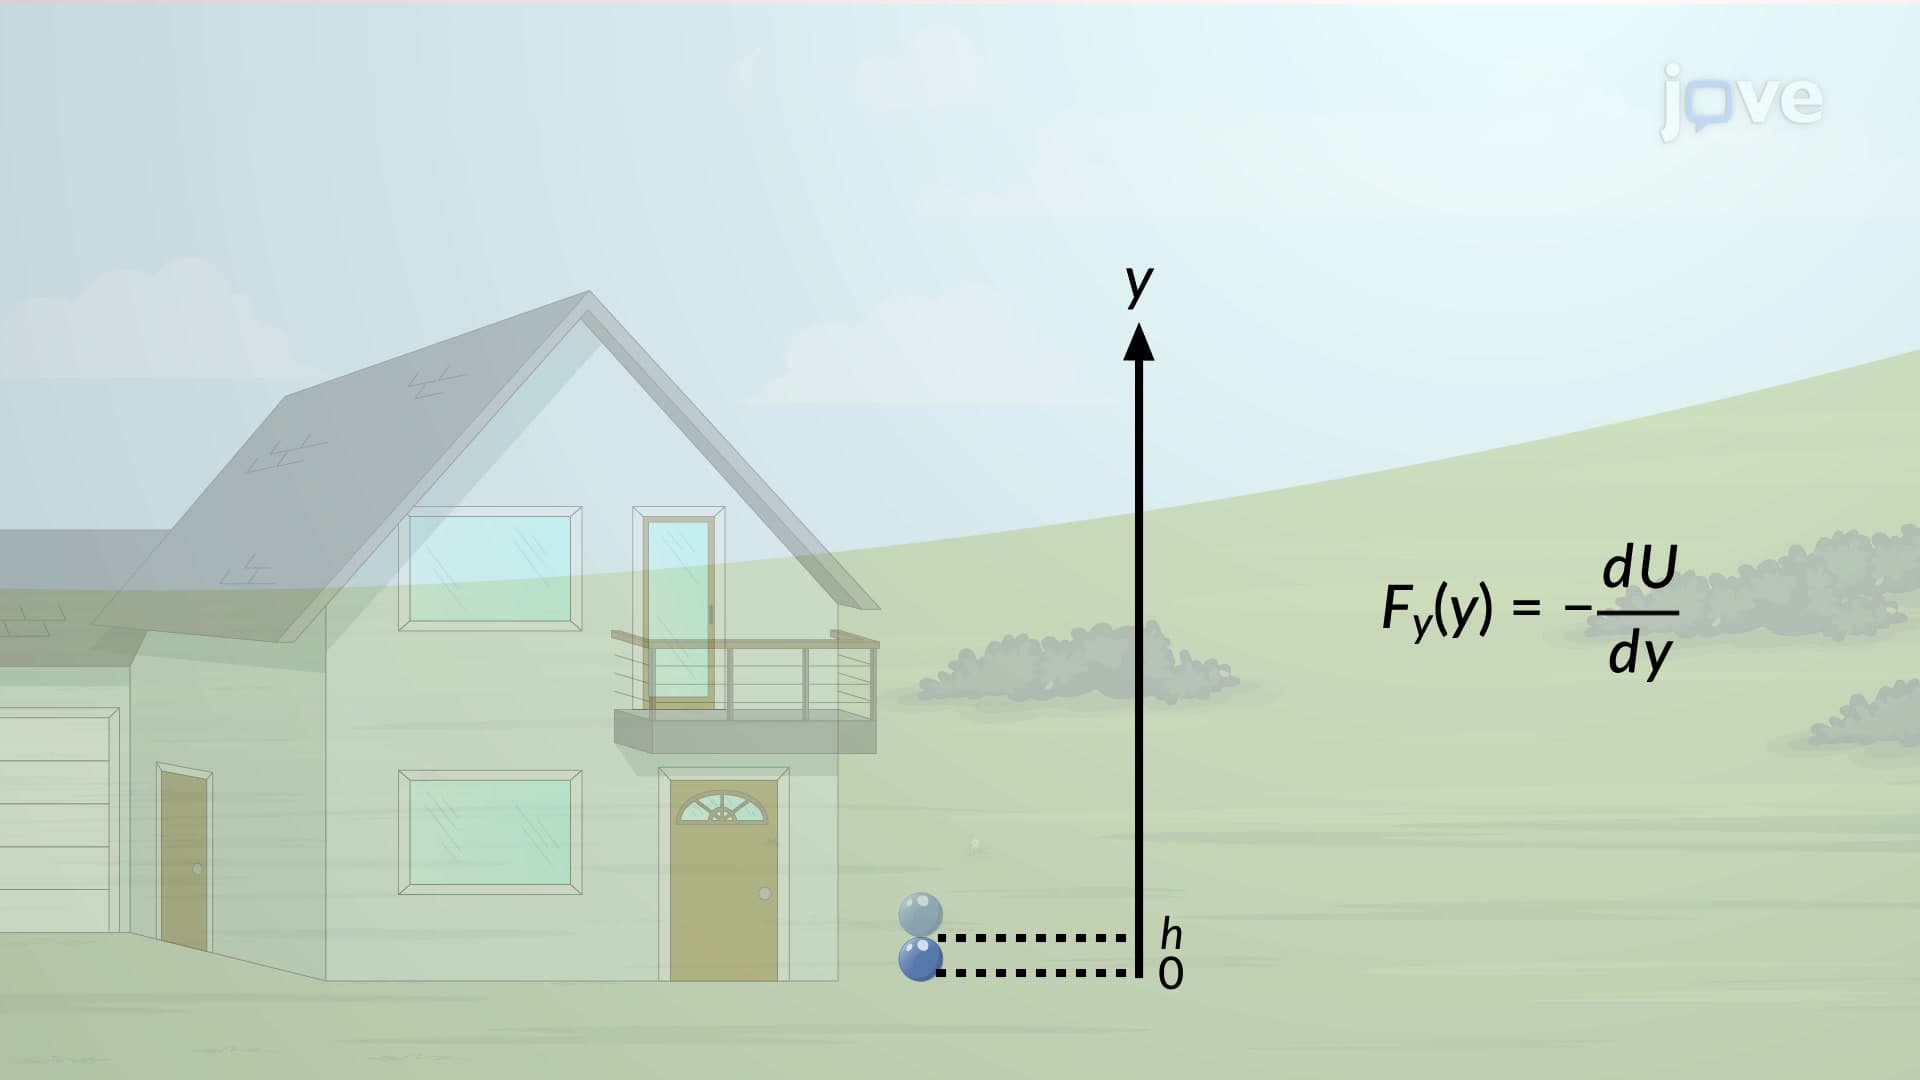
Force can be calculated from the expression for potential energy, which is a function of position. The component of a conservative force, in a particular direction, equals the negative of the derivative of the corresponding potential energy with respect to the displacement in that direction. For regions where potential energy changes rapidly with displacement, the work done and force is maximum. Also, when force is applied along the positive coordinate axis, the potential energy decreases with...
5.3K
 02:58
Crystal Field Theory - Octahedral Complexes
26.2K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
26.2K
 04:08
The Uncertainty Principle
23.1K
Werner Heisenberg considered the limits of how accurately one can measure properties of an electron or other microscopic particles. He determined that there is a fundamental limit to how accurately one can measure both a particle’s position and its momentum simultaneously. The more accurate the measurement of the momentum of a particle is known, the less accurate the position at that time is known and vice versa. This is what is now called the Heisenberg uncertainty principle. He...
23.1K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
41.6K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
41.6K
 02:40
MO Theory and Covalent Bonding
10.3K
The molecular orbital theory describes the distribution of electrons in molecules in a manner similar to the distribution of electrons in atomic orbitals. The region of space in which a valence electron in a molecule is likely to be found is called a molecular orbital. Mathematically, the linear combination of atomic orbitals (LCAO) generates molecular orbitals. Combinations of in-phase atomic orbital wave functions result in regions with a high probability of electron density, while...
10.3K
 01:10
Van der Waals Equation
4.0K
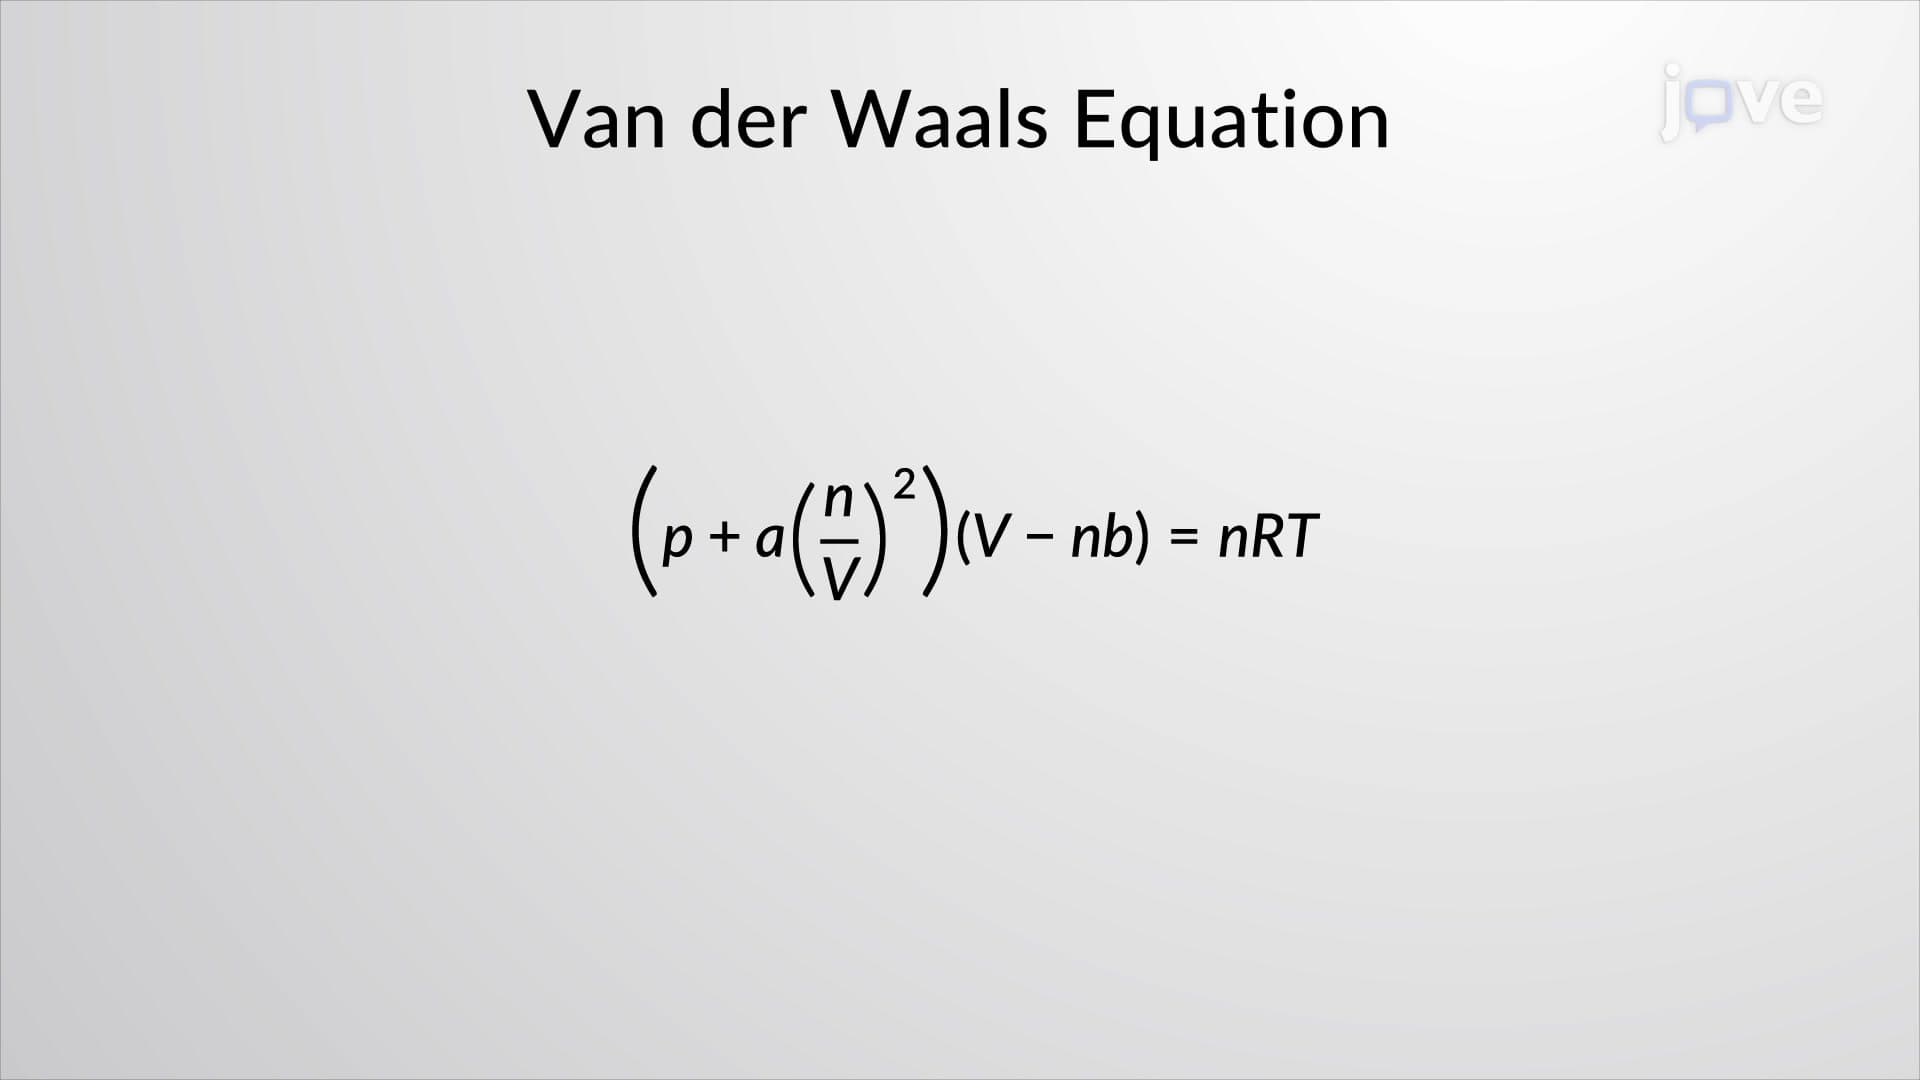
The ideal gas law is an approximation that works well at high temperatures and low pressures. The van der Waals equation of state (named after the Dutch physicist Johannes van der Waals, 1837−1923) improves it by considering two factors.
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
4.0K