替代对磁化能力的影响:密度函数理论和基准合集群计算的见解
Terri E Field-Theodore1,2, Peter R Taylor3, David J D Wilson4
1Scuola Superiore Meridionale, Largo San Marcellino 10, 80138 Naples, Italy.
The journal of physical chemistry. A
|October 25, 2024

概括
计算分子磁性,特别是对于化化合物,是复杂的. 这项研究对替代对磁性效应进行了基准测试,提供了对量子化学挑战的见解,并完善了理论模型.
科学领域:
- 量子化学 是一个量子化学.
- 计算化学计算化学
- 分子光谱学 分子光谱学
背景情况:
- 计算含分子的磁化张量在量子化学中提出了重大挑战.
- 替代对分子磁化性的影响需要详细的研究.
研究的目的:
- 进行一项基准研究,研究替代物对小型封闭外分子中的分子磁性 (同otropic和异otropic) 的影响.
- 为了确定磁化计算的准确理论极限,并评估密度函数理论 (DFT) 的近似值.
- 重新评估Flygare关于替代和磁化性异构性的假设.
主要方法:
- 结合集群 (CCSD(T)) 理论与大基数集和伦敦原子轨道.
- 完整的基础集 (CBS) 外推,以获得准确的理论极限.
- 理论结果与实验气相数据的比较,包括零点振动校正.
- 评估各种密度函数理论 (DFT) 的近似与基准CCSD (T) /CBS结果.
主要成果:
- 为24个含分子的磁化性生成了一个基准数据集.
- 评估了几个DFT近似计算磁化性的性能.
- 使用结合集群 (CCSD(T)) 理论建立了完整的基础集 (CBS) 极限.
- 鉴定了Flygare关于磁化性异性变化由于替代而导致的异性变化的假设的例外情况,并根据碳杂交进行了合理化.
结论:
- 这项研究为含有的分子提供了可靠的磁化数据,这对计算化学至关重要.
- 这些发现提供了对用于磁化性计算的DFT方法的批判性评估.
- 对Flygare假设偏差的合理化有助于我们更好地理解分子磁化中的结构属性关系.
相关概念视频
 03:07
Electron Affinity
35.3K
The electron affinity (EA) is the energy change for adding an electron to a gaseous atom to form an anion (negative ion).
35.3K
 01:16
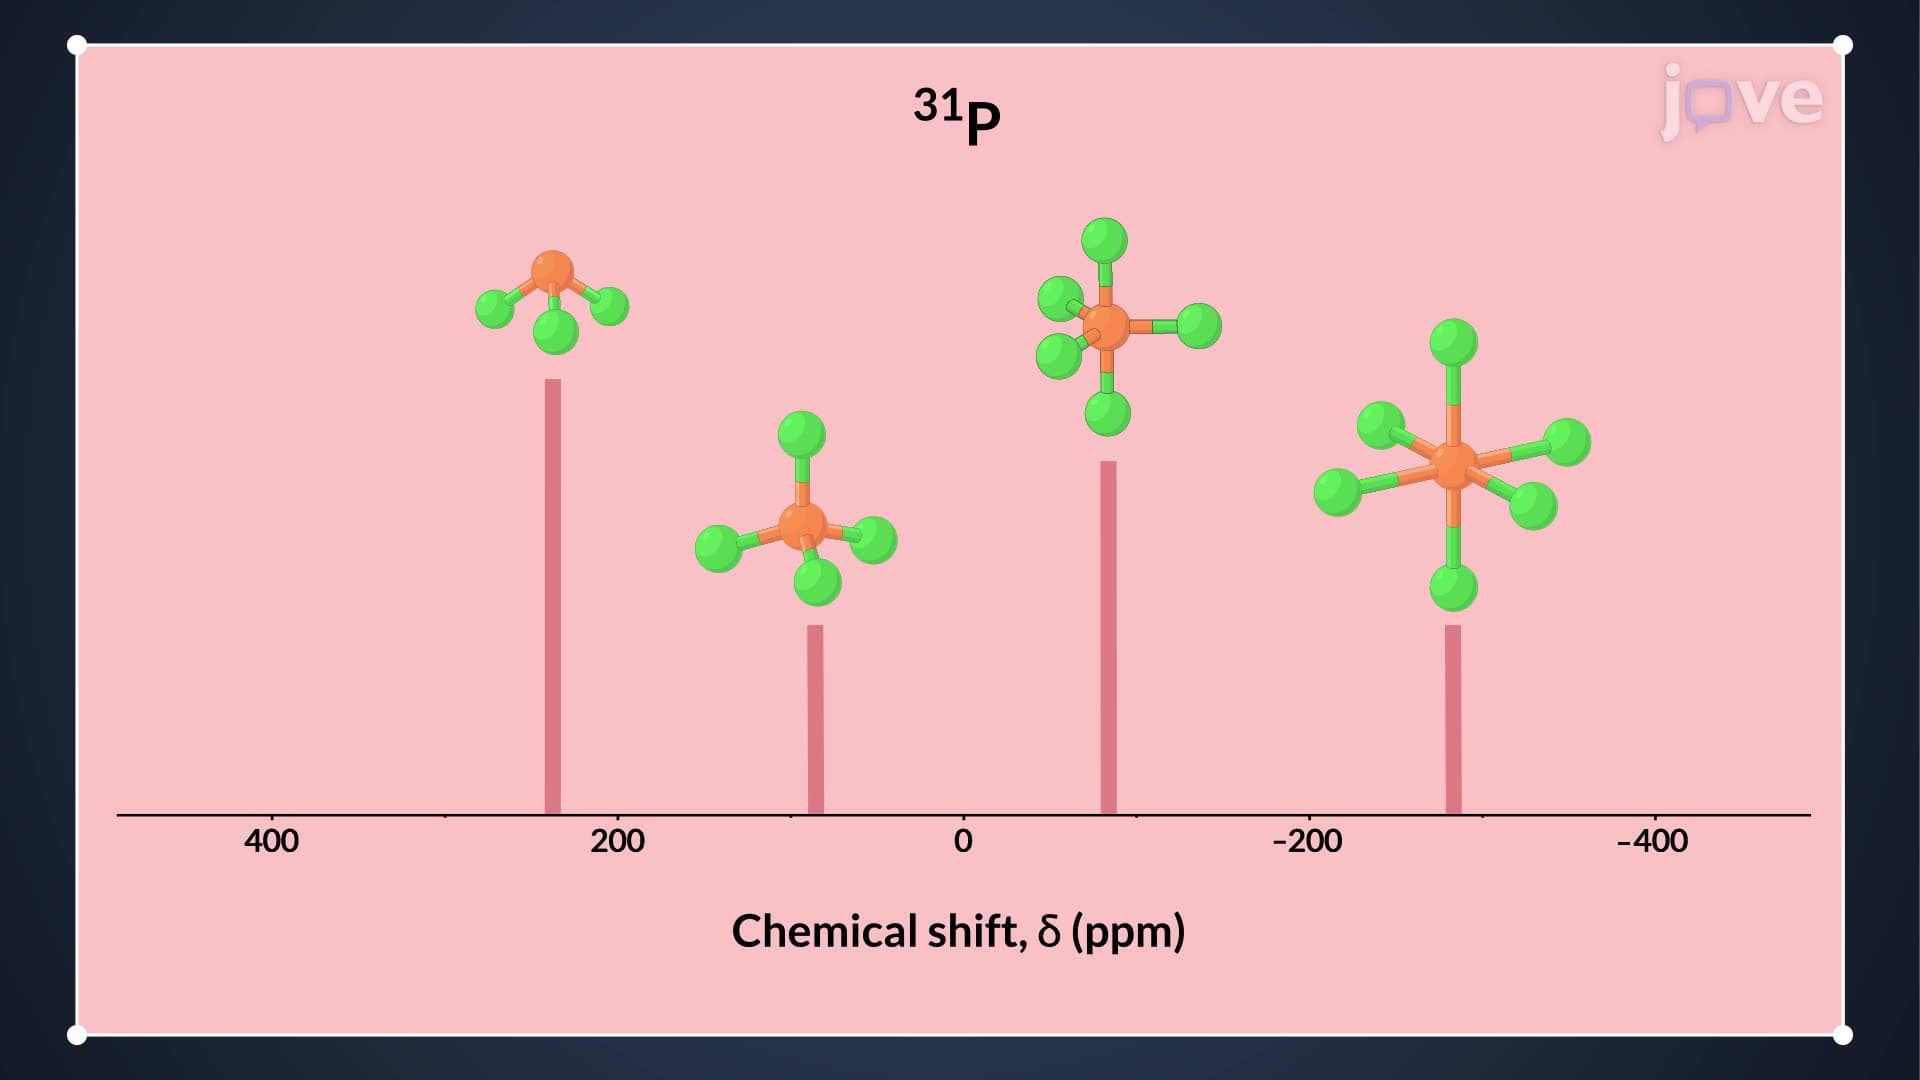
Other Nuclides: 31P, 19F, 15N NMR
368
Many organic, inorganic, and biological molecules contain spin-half nuclei such as nitrogen-15, fluorine-19, and phosphorus-31. As a result, NMR studies of these nuclei have found extensive applications in chemical and biological research.
While fluorine-19 and phosphorous-31 have high natural abundances (100%) and positive gyromagnetic ratios, nitrogen-15 has a low natural abundance and a negative gyromagnetic ratio. However, nitrogen-15 is still preferred over nitrogen-14 (which has a...
While fluorine-19 and phosphorous-31 have high natural abundances (100%) and positive gyromagnetic ratios, nitrogen-15 has a low natural abundance and a negative gyromagnetic ratio. However, nitrogen-15 is still preferred over nitrogen-14 (which has a...
368
 02:42
Valence Bond Theory
8.5K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
8.5K
 03:02
Colors and Magnetism
11.6K
Color in Coordination Complexes
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
11.6K
 03:02
The Aufbau Principle and Hund's Rule
46.6K
To determine the electron configuration for any particular atom, we can build the structures in the order of atomic numbers. Beginning with hydrogen, and continuing across the periods of the periodic table, we add one proton at a time to the nucleus and one electron to the proper subshell until we have described the electron configurations of all the elements. This procedure is called the aufbau principle, from the German word aufbau (“to build up”). Each added electron occupies the...
46.6K
 02:44
The Born-Haber Cycle
21.7K
Lattice Energy
21.7K