

一个基于多极的反应力场,用于碳化合物
Junben Weng1,2, Hongqiang Cui1,2, Da Zheng1,2
1Laboratory of Molecular Modeling and Design, State Key Laboratory of Molecular Reaction Dynamics, Dalian Institute of Chemical Physics, Chinese Academy of Sciences, Dalian 116023, China.

基于原子多极的新型OPERATOR反应力场简化了化学反应的建模. 这种基于物理的方法准确地捕捉了反应期间的静电变化,优于现有的方法,如ReaxFF用于碳化合物分解模拟.
科学领域:
- 计算化学的计算化学
- 材料科学 材料科学 材料科学
- 化学物理 化学物理
背景情况:
- 量子化学方法对于大规模的反应系统来说在计算上是昂贵的.
- 现有的反应力场依赖于复杂的,基于知识的纠正,阻碍了发展.
- 需要准确有效的模型来模拟化学反应的分子动力学模拟.
研究的目的:
- 介绍一个新的基于原子多极的无约束反应模型 (操作器).
- 在AMOEBA框架内开发一个有物理动机的简化力场.
- 改进反应过程中静电相互作用和化学环境的描述.
主要方法:
- 在AMOEBA框架内使用物理动机模型构建了OPERATOR力场.
- 动态生成的原子多极体,以捕捉反应期间的环境变化.
- 集成的电荷透,极化,电荷转移,范德瓦尔斯电荷和三体电位.
- 优化使用DFT衍生的潜在能量表面 (PESs) 的碳化合物跨越数以百万计的形状.
主要成果:
- 操作员力场准确地复制了原子断和能量.
- 与ReaxFF反应力场相比,具有可比或优越的性能.
- 对n-heptane分解的分子动力学模拟成功地重现了实验产品和反应.
结论:
- 运算器力场提供了一个简化,基于物理的方法来建模化学反应.
- 它捕捉动态静电变化的能力对于反应模拟至关重要.
- 预计将促进未来对各种元素复杂化学反应机制的研究.
更多相关视频
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 05:57
Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
相关概念视频
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 02:14
Combustion Energy: A Measure of Stability in Alkanes and Cycloalkanes
Alkanes undergo combustion in the presence of excess oxygen and high-temperature conditions to give carbon dioxide and water. A combustion reaction is the energy source in natural gas, liquified...
 02:33
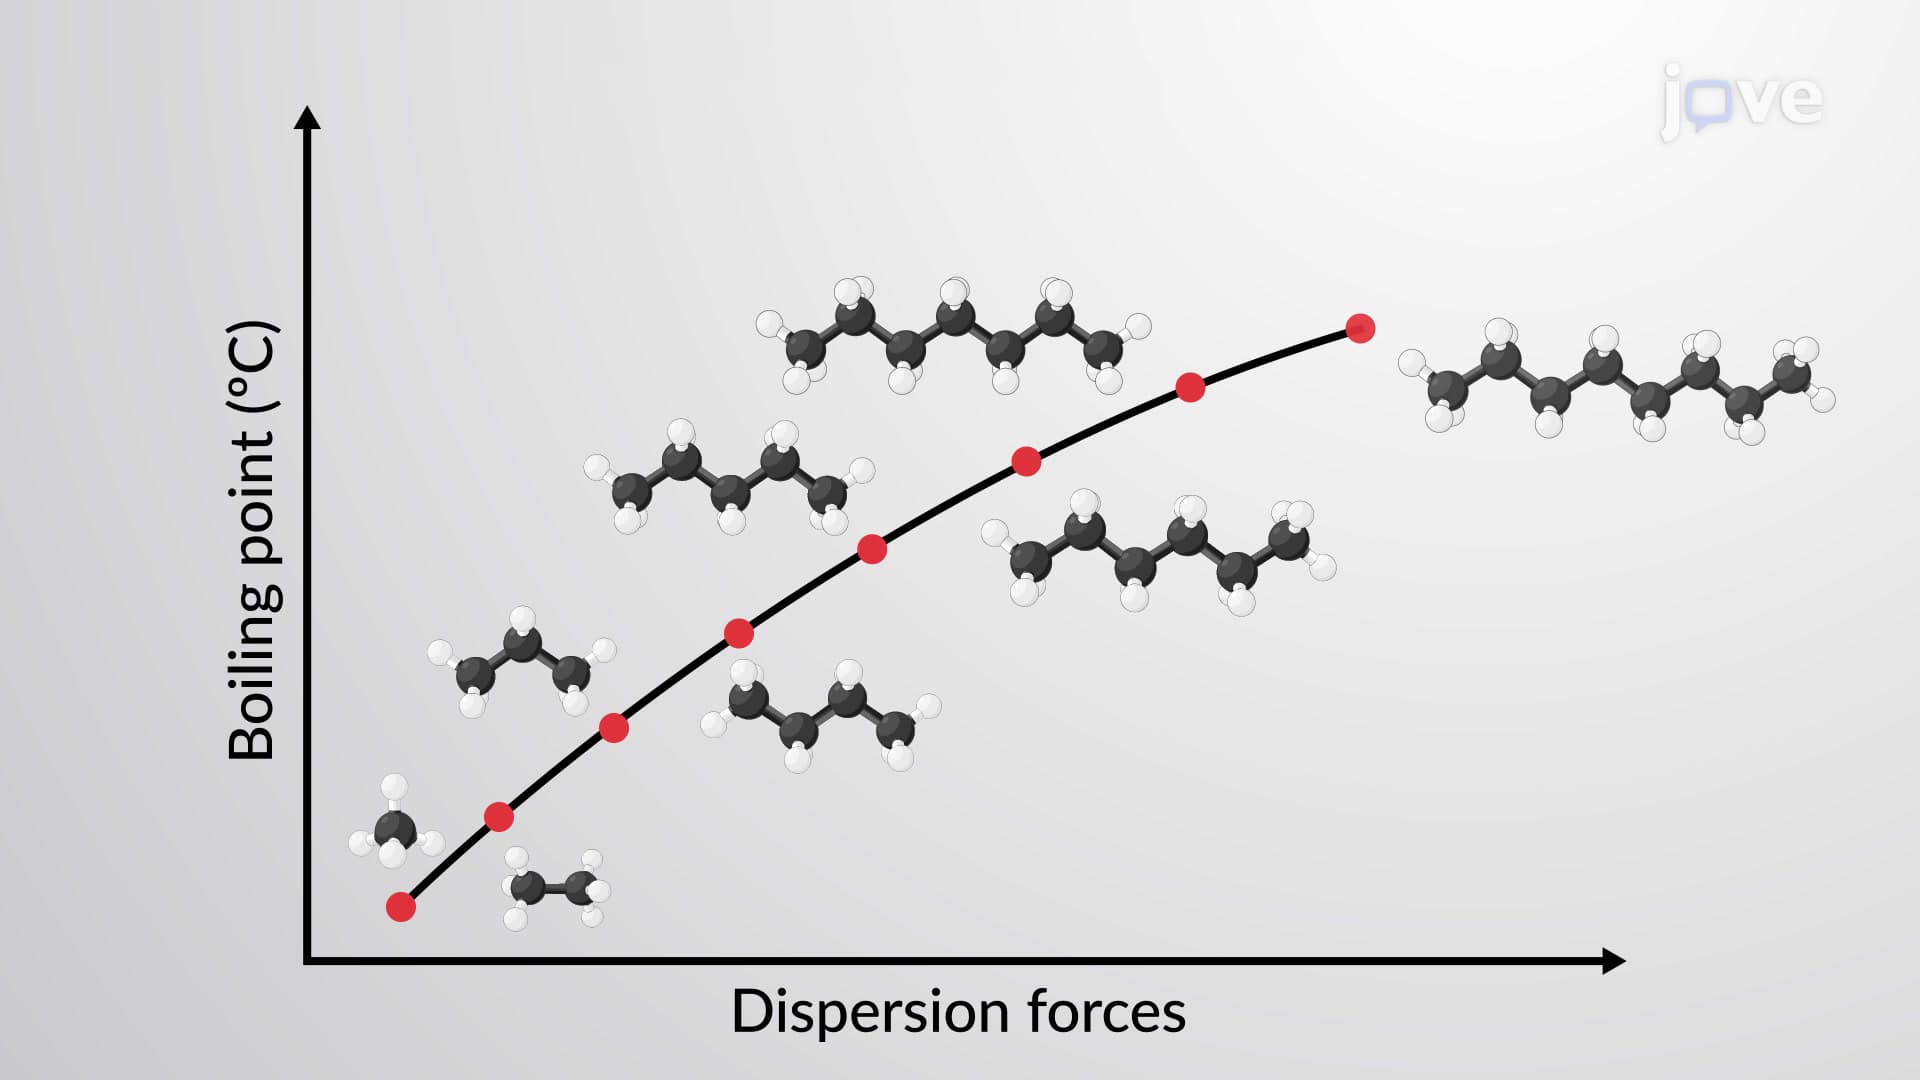
Physical Properties of Alkanes
 01:13
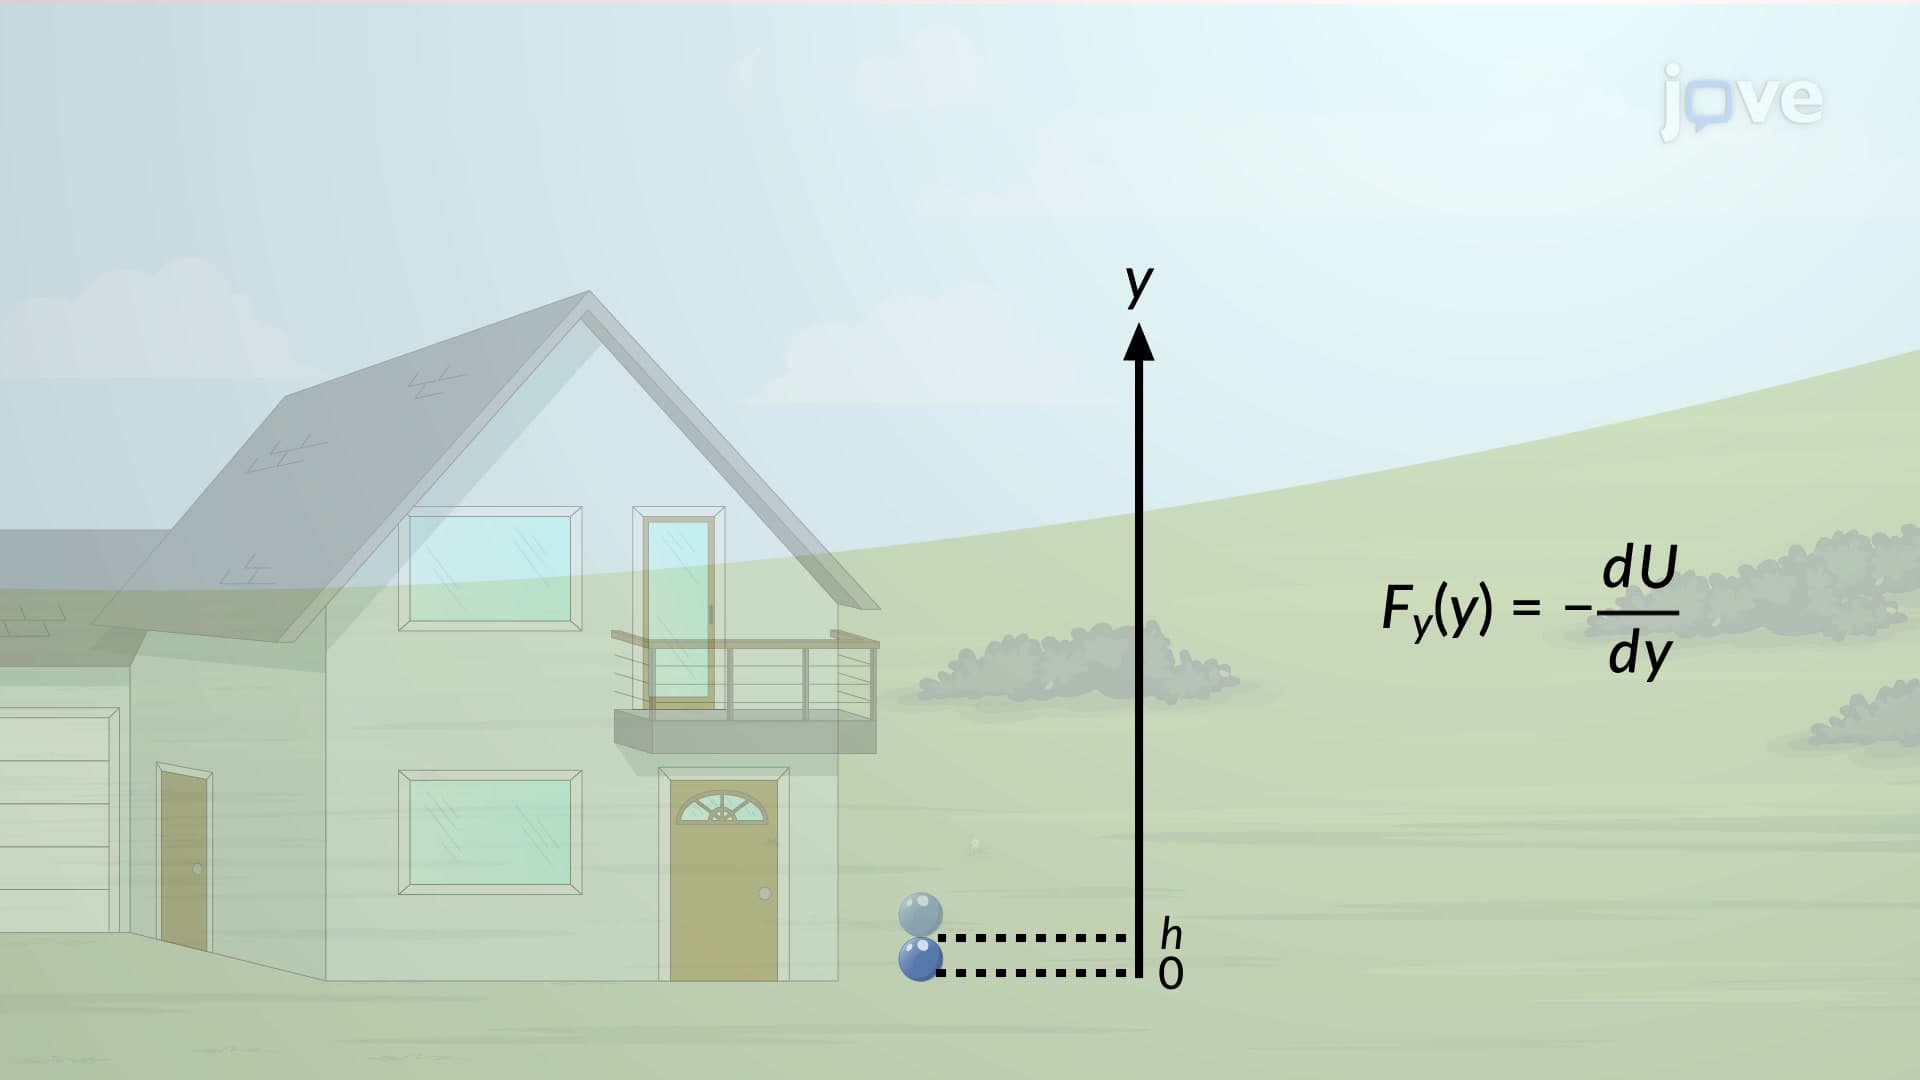
Force and Potential Energy in One Dimension
 01:27
π Electron Effects on Chemical Shift: Overview
 01:20
Two-Dimensional Force System