

表面跳跃中的非adiabatic合与紧密结合密度的功能理论:分子电机的案例
Gonzalo Díaz Mirón1, Carlos R Lien-Medrano2, Debarshi Banerjee1,3
1Condensed Matter and Statistical Physics, The Abdus Salam International Centre for Theoretical Physics, 34151 Trieste, Italy.

本研究介绍了一种有效的计算方法,用于模拟分子光相互作用,使用轨迹表面跳跃与密度功能紧密结合 (DFTB). 经过验证的方法准确地模拟了分子电机等复杂系统中的光物理过程.
科学领域:
- 计算化学的计算化学
- 物理化学 物理化学
- 量子力学就是量子力学.
背景情况:
- 非协同分子动力学 (NAMD) 对于理解分子光相互作用至关重要.
- 准确的NAMD需要先进的电子结构方法,这些方法在计算上昂贵.
- 像密度功能紧密结合 (DFTB) 这样的近似方法用于更大的系统,但需要验证.
研究的目的:
- 开发和验证NAMD的新实现,使用轨迹表面跳跃与DFTB相结合.
- 通过将其与更高层次的电子结构计算进行比较来评估该方法的准确性.
- 将验证的方法应用于复杂的系统,如分子电机,以深入了解它们的光物理行为.
主要方法:
- 实现轨迹表面跳跃 (TSH) 算法.
- 将TSH与密度功能紧密结合 (DFTB) 电子结构方法集成.
- 计算非adiabatic合向量,以提高准确性.
- 验证使用甲酸和 furan 系统.
- 应用于复杂的分子电机系统.
主要成果:
- 开发的NAMD-DFTB方法准确地捕捉了关键的光物理机制,包括光吸收后的二面旋转.
- 该模拟成功地重现了在时间依赖光实验中观察到的从明亮到黑暗状态的过渡.
- 该方法提供了一个计算可行的方法来研究复杂的分子系统.
结论:
- 经过验证的NAMD-DFTB方法为研究分子系统中的光物理放松提供了一个计算效率高,准确的工具.
- 这种方法为分子电机和其他复杂系统的行为提供了宝贵的见解.
- 该研究解决了对近似NAMD技术进行全面验证的需求.
更多相关视频
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
相关概念视频
 01:22
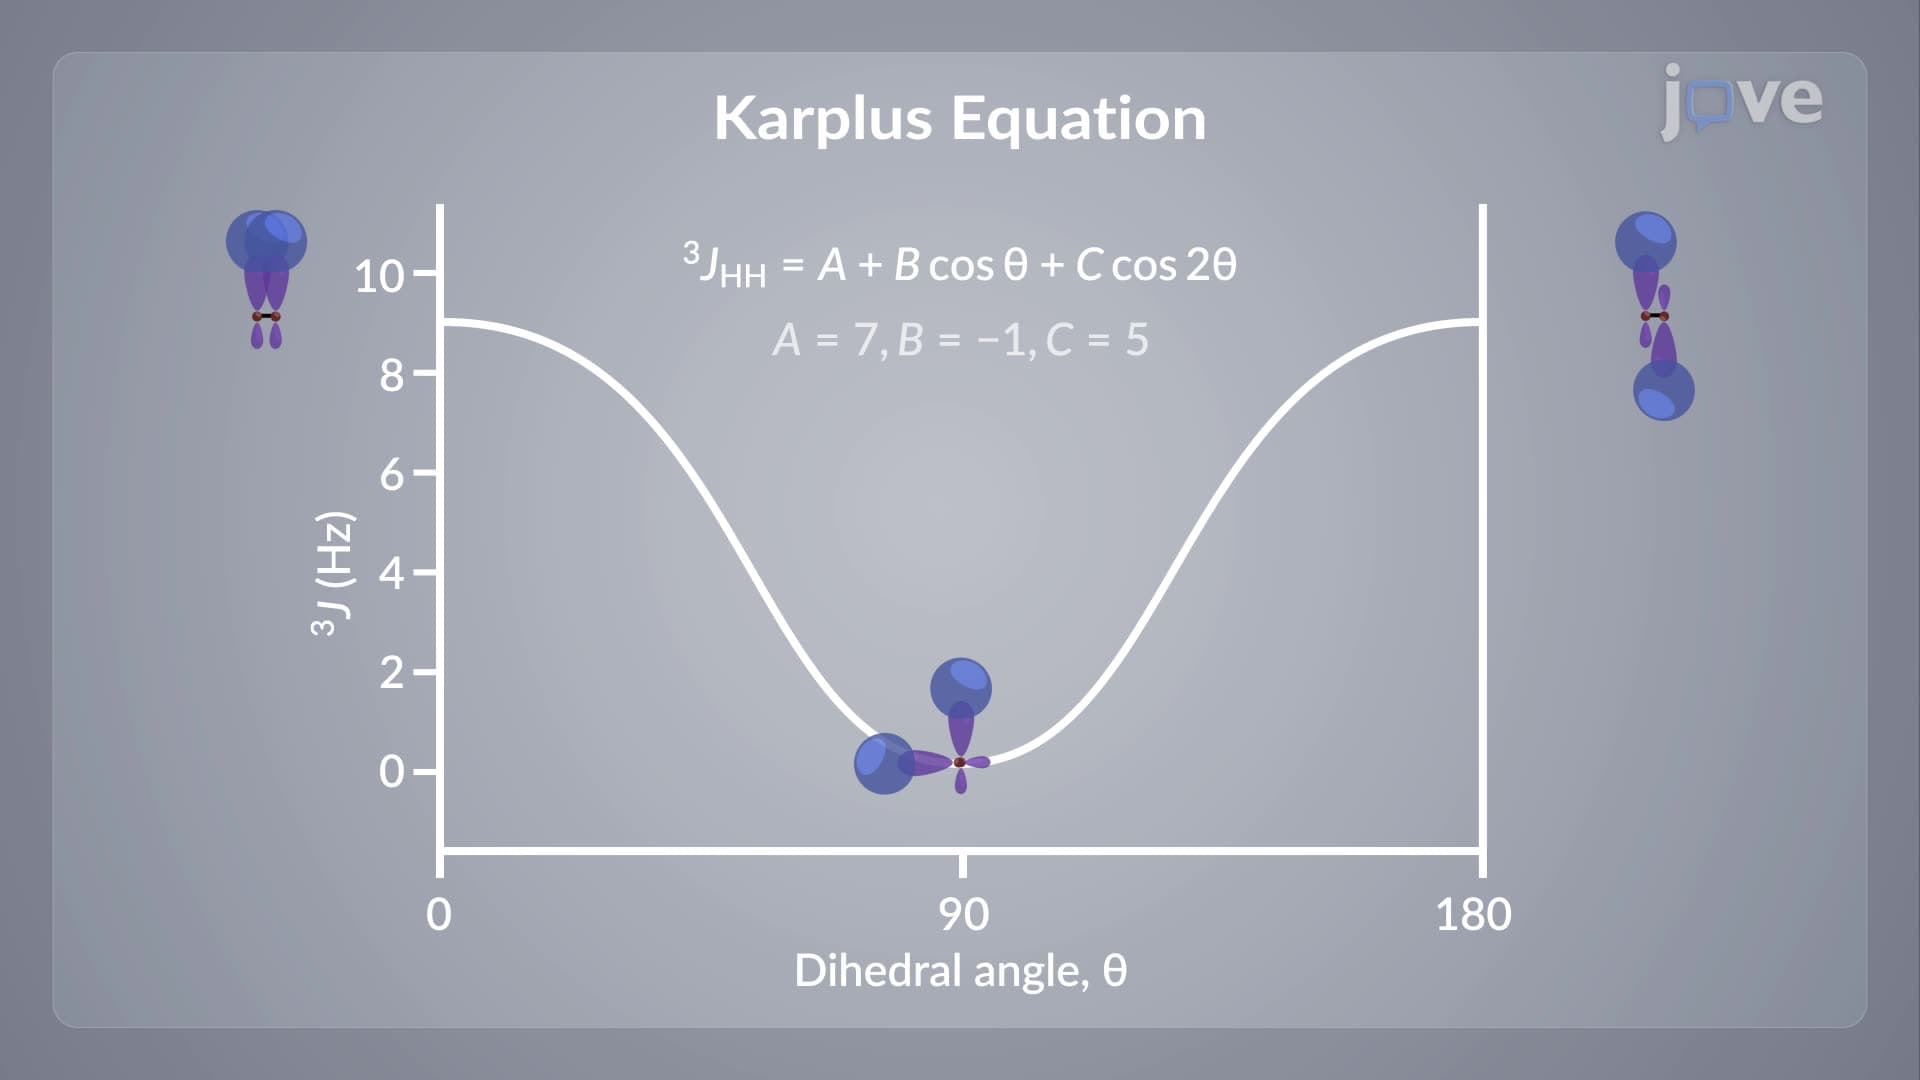
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...
 01:17
Spin–Spin Coupling: One-Bond Coupling
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...
 01:10
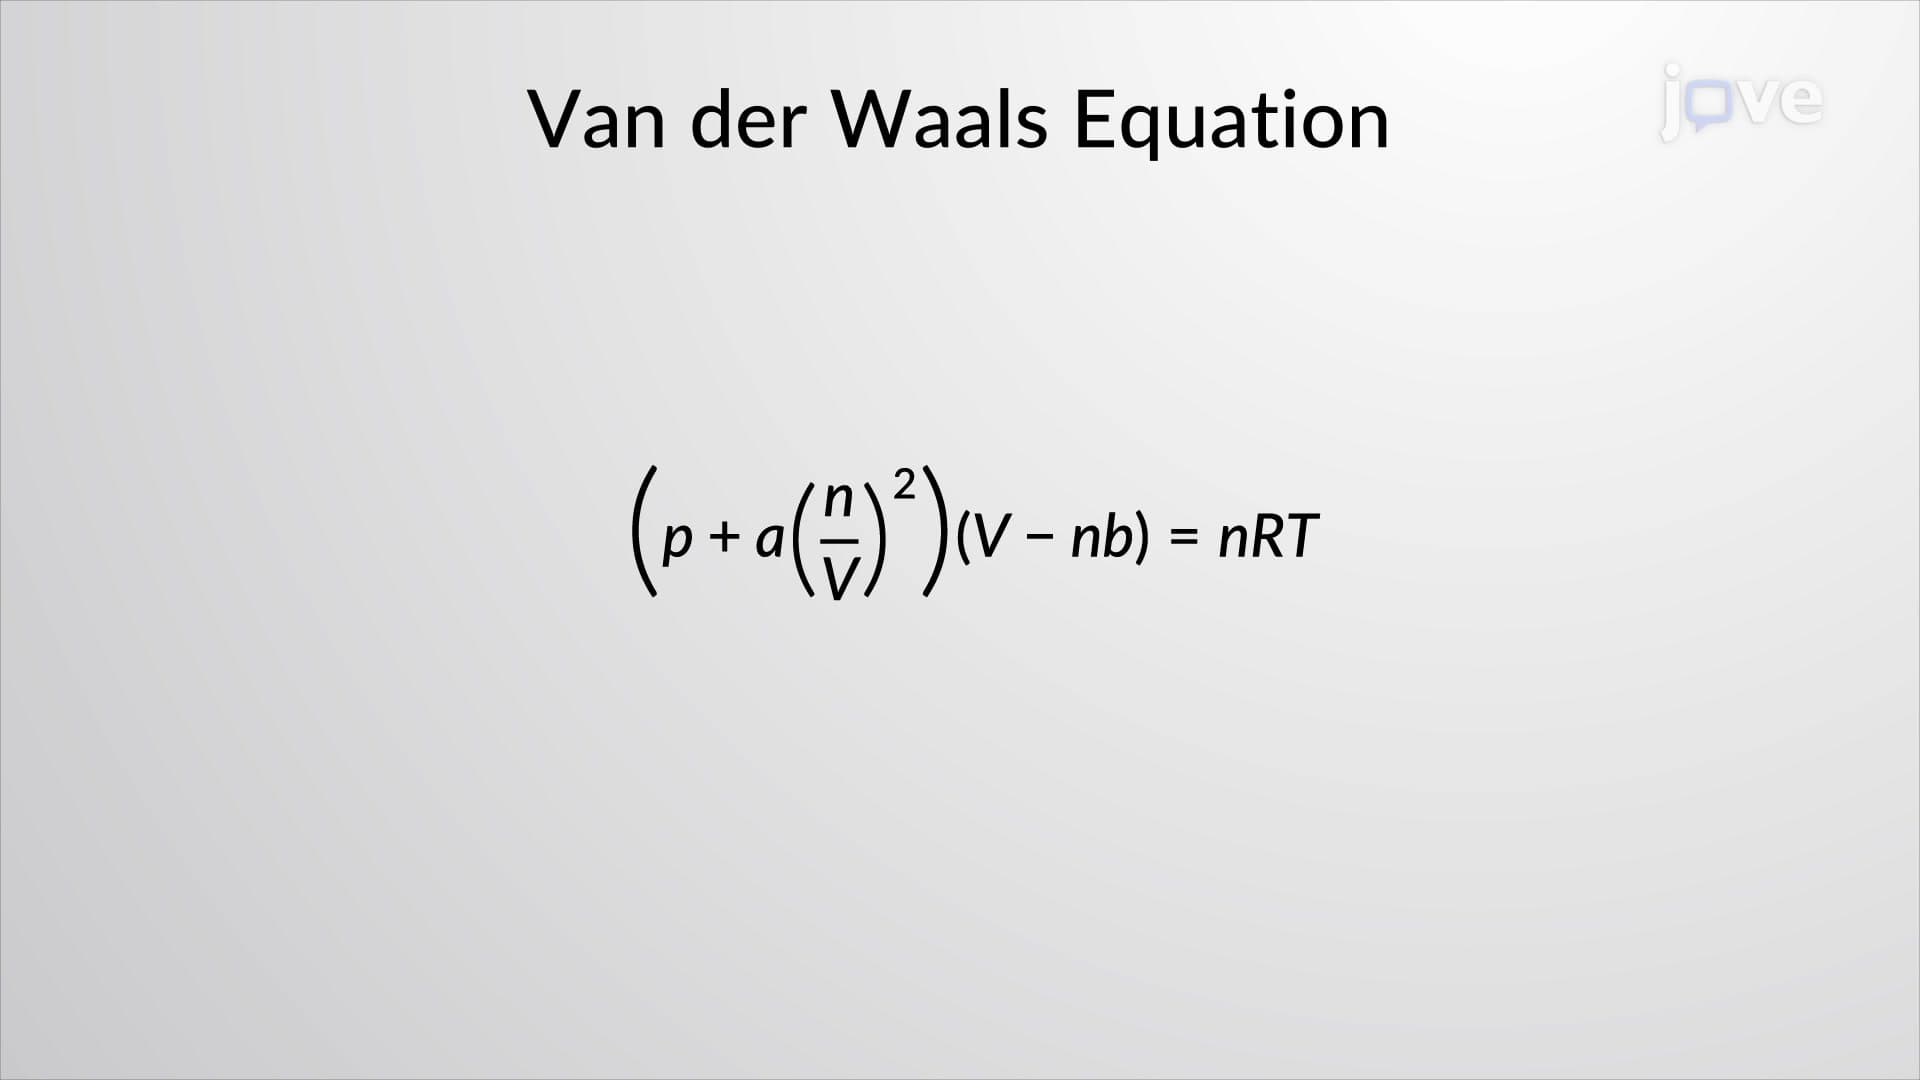
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:35
Hybridization of Atomic Orbitals II