确定GTPase-GEF复合体对界面药物结合特异性的静电贡献:蛋白质-药物-蛋白质复合物的案例研究
Frank A Jermusek1, Lauren J Webb1,2
1Interdisciplinary Life Sciences Graduate Program, The University of Texas at Austin, Austin, Texas 78712, United States.
Biochemistry
|November 26, 2024

概括
小分子药物设计需要了解蛋白质-蛋白质相互作用. 这项研究揭示了Arf1-GEF复合体中的静电环境和接口刚性如何影响brefeldin A (BFA) 的结合特异性,指导未来的药物开发.
科学领域:
- 生物化学 生化学
- 结构生物学 结构生物学
- 药物发现 药物发现 药物发现
背景情况:
- 蛋白质与蛋白质的相互作用对于细胞功能和药物设计至关重要.
- 小分子与蛋白质接口结合的特异性受到结构和静电互补性的影响.
- 对蛋白质 - 药物 - 蛋白质特异性的静电贡献比结构因素了解得更少.
研究的目的:
- 研究蛋白质-蛋白质接口的静电环境如何决定小分子结合的特异性.
- 为了比较BFA敏感和BFA耐药的Arf1-GEF复合体,以了解布雷菲尔丁A (BFA) 特异性的决定因素.
- 探索接口刚度在蛋白质-蛋白质相互作用中小分子破坏中的作用.
主要方法:
- 使用具有对BFA敏感或耐药的关氨酸核酸交换因子 (GEF) 的Arf1复合体.
- 采用Arf1的蓝类固醇标记和烯吸附频率测量,通过线性斯特克方程绘制界面电场的地图.
- 分析了烯振动光谱的温度依赖性,以评估结环境和接口刚性.
主要成果:
- 在Arf1表面的界面电场因被绑定的GEF而有显著的变化.
- Arf1-ARNOwt和Arf1-ARNO4M复合体在电场中表现出最显著的差异,这表明BFA与Arf1-ARNO结合需要更大的电场变化.
- 接口刚度与BFA灵敏度有很强的相关性,这表明灵活的接口更容易受到小分子的破坏.
结论:
- 静电环境为Arf1结合提供了质量一致的框架,但微妙的差异决定了BFA的特异性.
- 接口刚性是影响对orthosteric小分子结合敏感性的关键因素.
- 结果为设计小分子提供了洞察力,以提高特异性准蛋白质-蛋白质接口.
相关概念视频
 02:14
GTPases and their Regulation
8.3K
Guanine nucleotide-binding proteins (G-proteins), also known as GTPases, are a superfamily of proteins that regulate many cellular processes, such as cell signaling, vesicular transport, and the regulation of cell shape and motility. Mutation or dysfunction of these proteins can lead to disease. There are around 40,000 known G-proteins that can broadly be classified into two groups ‒ small G-proteins consisting of a single domain and large multi-domain G-proteins.
Large G-proteins,...
Large G-proteins,...
8.3K
 01:18
Factors Affecting Protein-Drug Binding: Drug-Related Factors
83
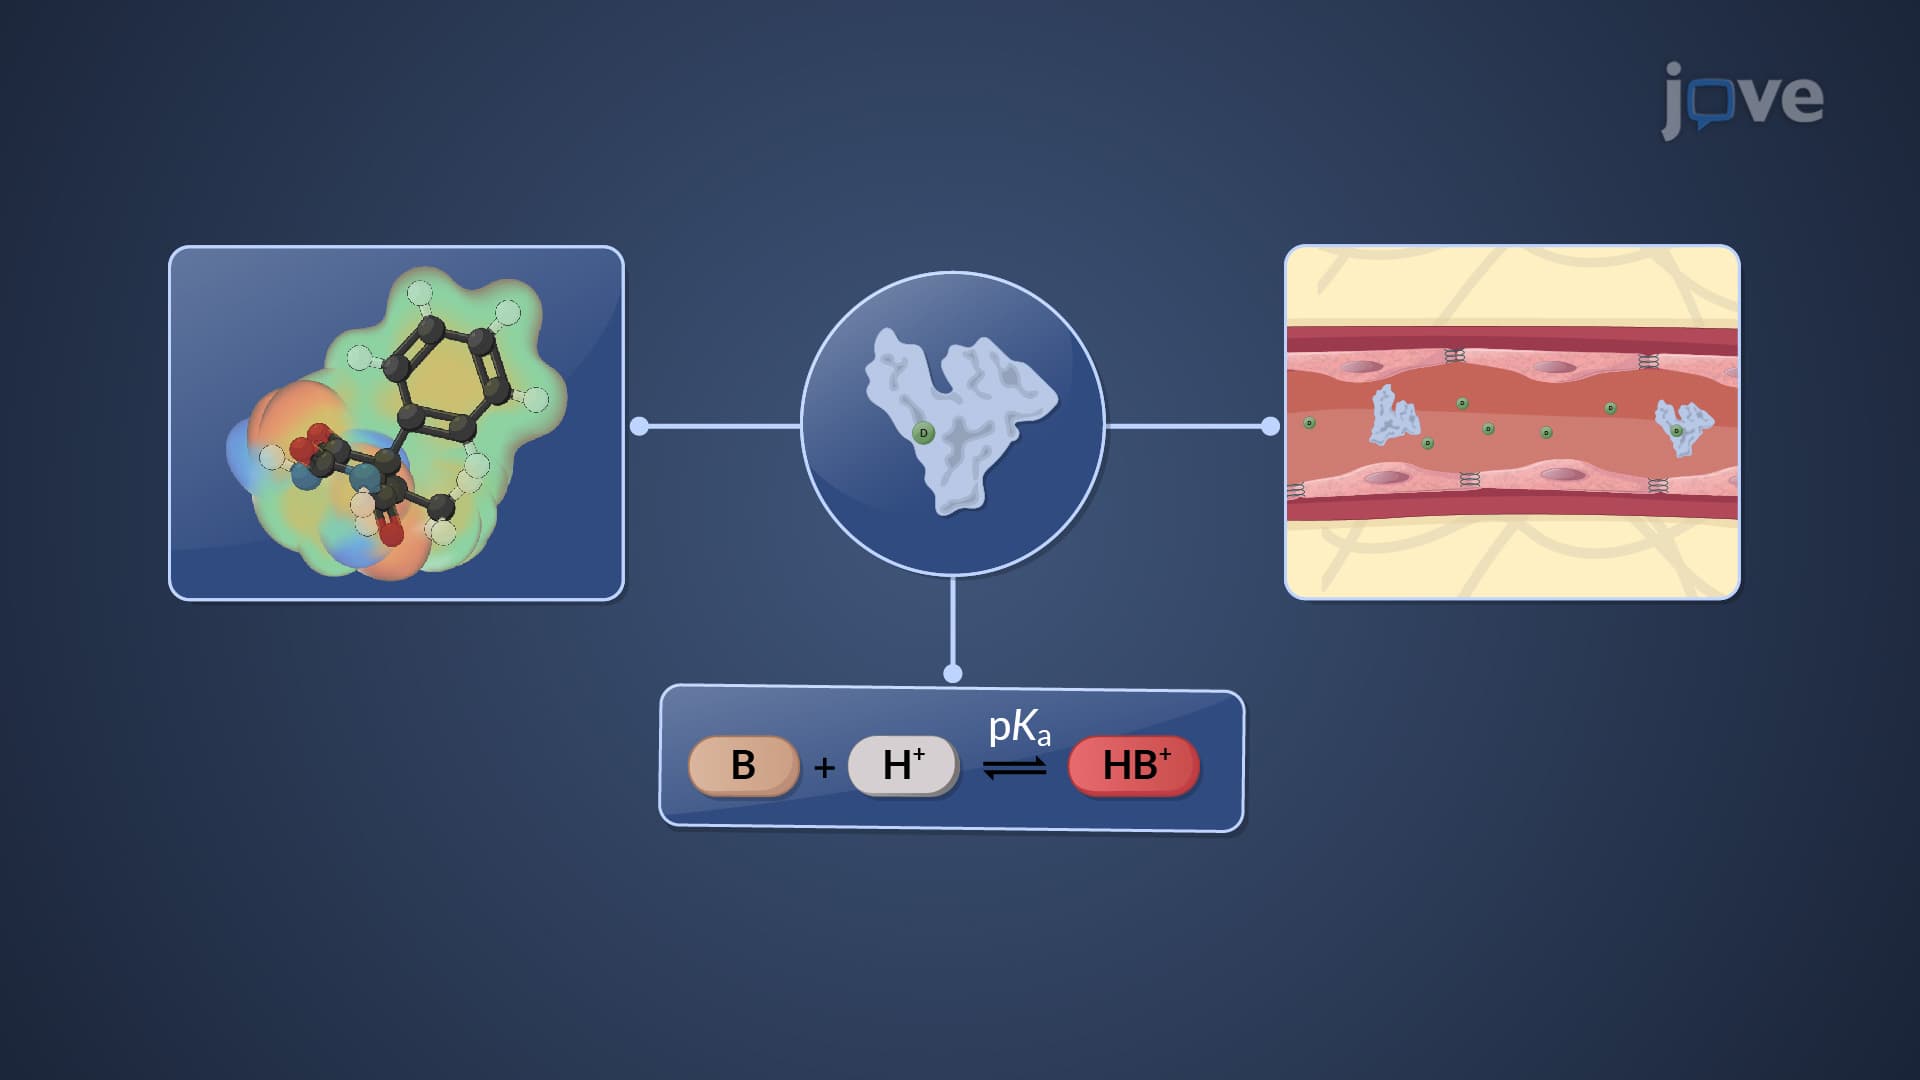
Drug binding to proteins is a complex phenomenon influenced by various drug-related factors, each playing a significant role in the interaction between drugs and proteins within the body.
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...
83
 01:22
Activation and Inactivation of G Proteins
6.8K
Heterotrimeric G proteins are guanine nucleotide-binding proteins. As the name suggests, heterotrimeric G proteins are composed of three subunits: alpha, beta, and gamma. They remain GDP-bound or GTP-bound inside the cells and switch between inactive/active states. The Gα subunit possesses the nucleotide-binding pocket that binds guanine nucleotides and switches between GDP or GTP-bound states. In contrast, the Gꞵ and Gγ subunits are always bound together with high...
6.8K
 02:04
Protein-protein Interfaces
12.5K
Many proteins form complexes to carry out their functions, making protein-protein interactions (PPIs) essential for an organism's survival. Most PPIs are stabilized by numerous weak noncovalent chemical forces. The physical shape of the interfaces determines the way two proteins interact. Many globular proteins have closely-matching shapes on their surfaces, which form a large number of weak bonds. Additionally, many PPIs occur between two helices or between a surface cleft and a...
12.5K
 01:24
Small GTPases - Ras and Rho
3.9K
Ras and Rho are small monomeric GTPases that act downstream of receptor tyrosine kinase (RTK) and regulate various cellular processes. These GTPases switch between active and inactive states by binding to guanine nucleotides.
Three regulatory proteins control their activity:
Three regulatory proteins control their activity:
3.9K
 01:25
Drug-Receptor Bonds
2.7K
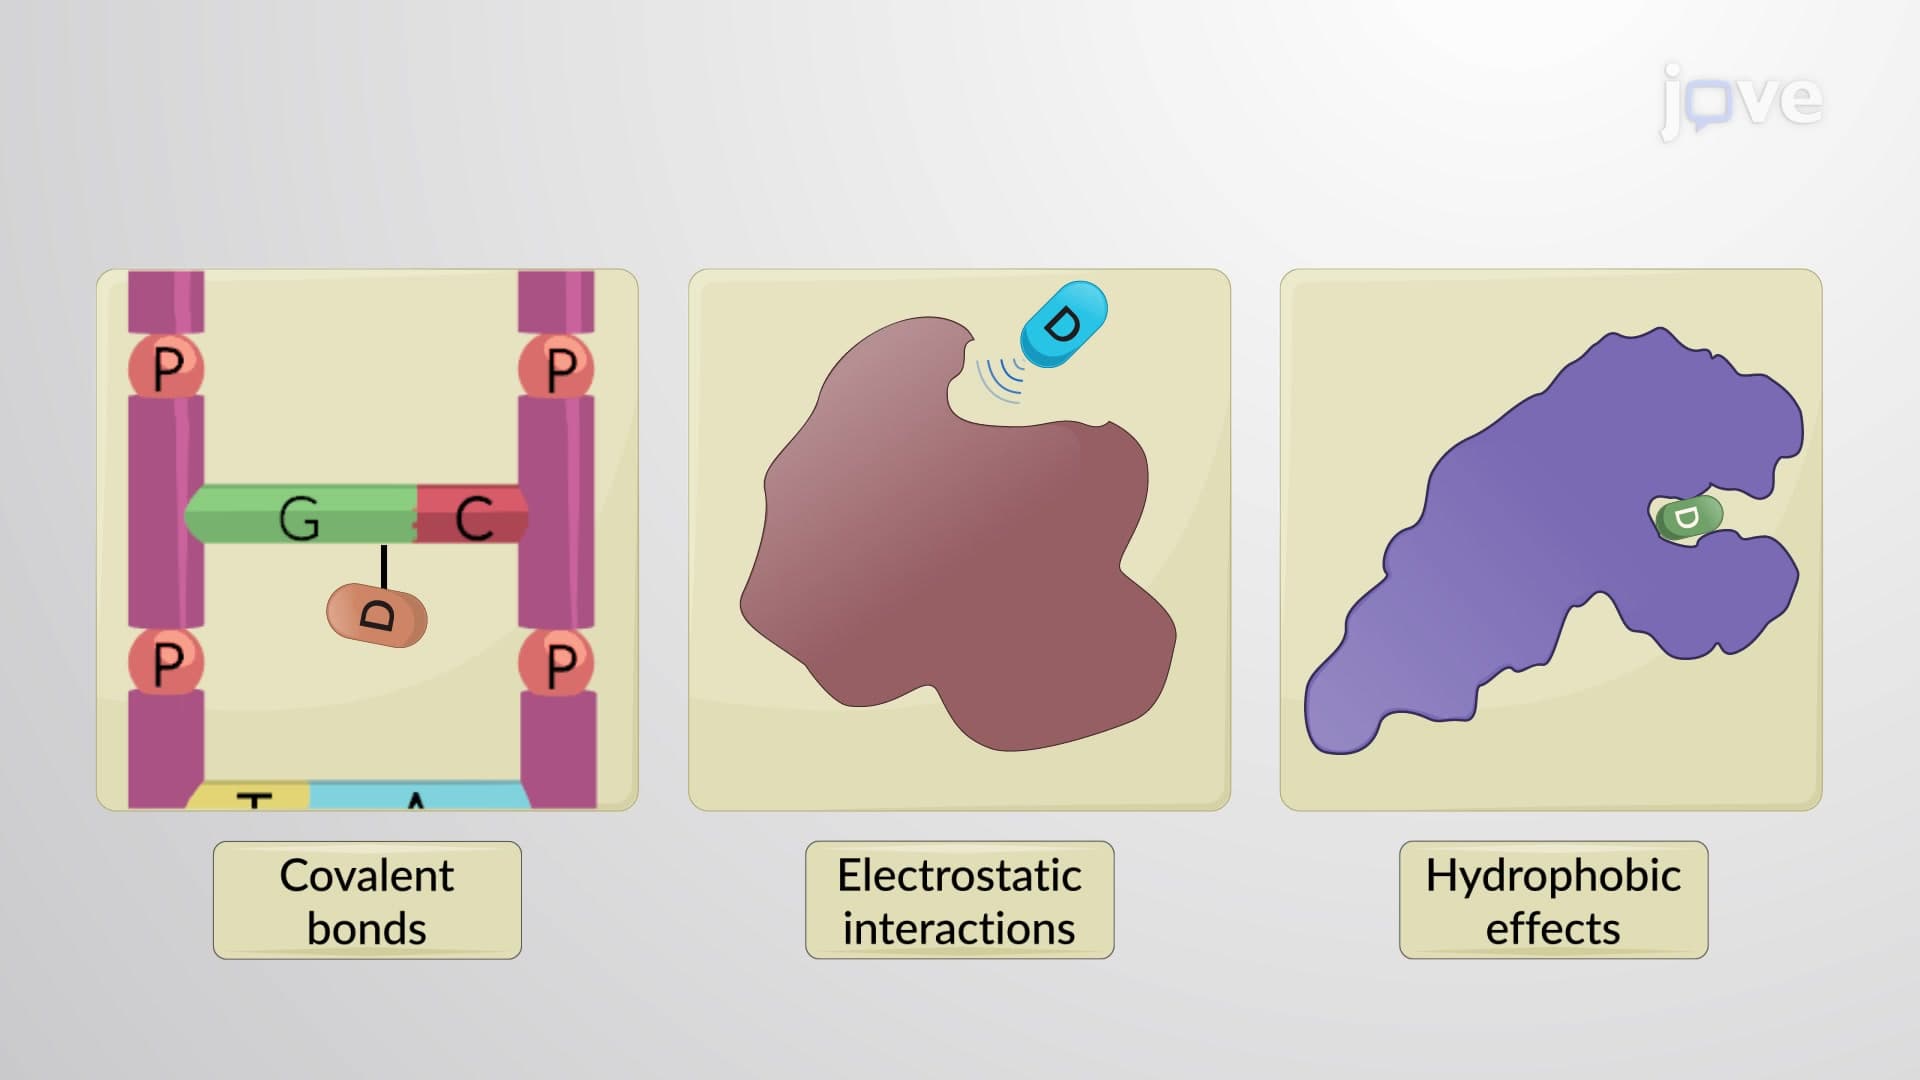
Drug-receptor bonds are formed through various chemical forces when drugs interact with target cells. Covalent bonds, strong and irreversible, are exemplified by DNA-alkylating anticancer agents that inhibit cell division. However, such irreversible drug binding lacks selectivity and can modify the DNA of the surrounding healthy cells. Covalent binding often contributes to tissue toxicity, as seen with chloroform and paracetamol metabolites binding to the liver, causing hepatotoxicity.
In...
In...
2.7K