在固体电解质NaAlCl4中探索阴离子替代与密度函数理论
Michael Häfner1,2, Matteo Bianchini1,2
1Faculty of Biology, Chemistry and Earth Sciences, Universität Bayreuth, Universitätsstrasse 30, 95447 Bayreuth, Germany.

研究人员探索了NaAlCl4中的替代元素,以改善室温电池的导率. 他们发现,银和的替代物是有希望的,而为新的导体提供了最好的潜力.
科学领域:
- 材料科学 材料科学 材料科学
- 电化学 电化学 电化学
- 计算材料科学科学 计算材料科学
背景情况:
- 化 (NaAlCl4) 是一种已知用于高温基电池的固体电解质.
- 它的离子导电性不足以有效的室温电池应用.
- 发现新的导体对于先进的能量储存至关重要.
研究的目的:
- 评估NaAlCl4中各种元素替代物的有效性,以提高离子导电性.
- 为了确定室温导体的有前途的候选人.
- 探索热力学特性在发现新的固体电解质中的作用.
主要方法:
- 使用密度函数理论 (DFT) 与热力学修正相结合.
- 在飞行中使用机器学习的潜能来加快语音计算.
- 在NaAlCl4系统中研究了同和异替代.
主要成果:
- (Na) 取代 (K) 和 (Ga) 取代 (Al) 的同价替代是热力学上有利的.
- 替代 (Na) 的白银 (Ag) 也显示出有希望的结果.
- 最有前途的替代涉及 (Zn) 在NaAlCl4Na2ZnCl4 tieline上,创建具有离子导电空缺的分层结构.
结论:
- 元素替代,特别是有价替代,可以显著提高NaAlCl4的离子导电性质.
- 已识别的替代物,特别是涉及,为室温电池提供了新的固体电解质的途径.
- 将热力学特性纳入计算发现加速了可靠导体的开发.
更多相关视频
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 06:44
From Molecules to Materials: Engineering New Ionic Liquid Crystals Through Halogen Bonding
Published on: March 24, 2018
相关概念视频
 02:48
Ionic Bonding and Electron Transfer
 02:42
Ionic Crystal Structures
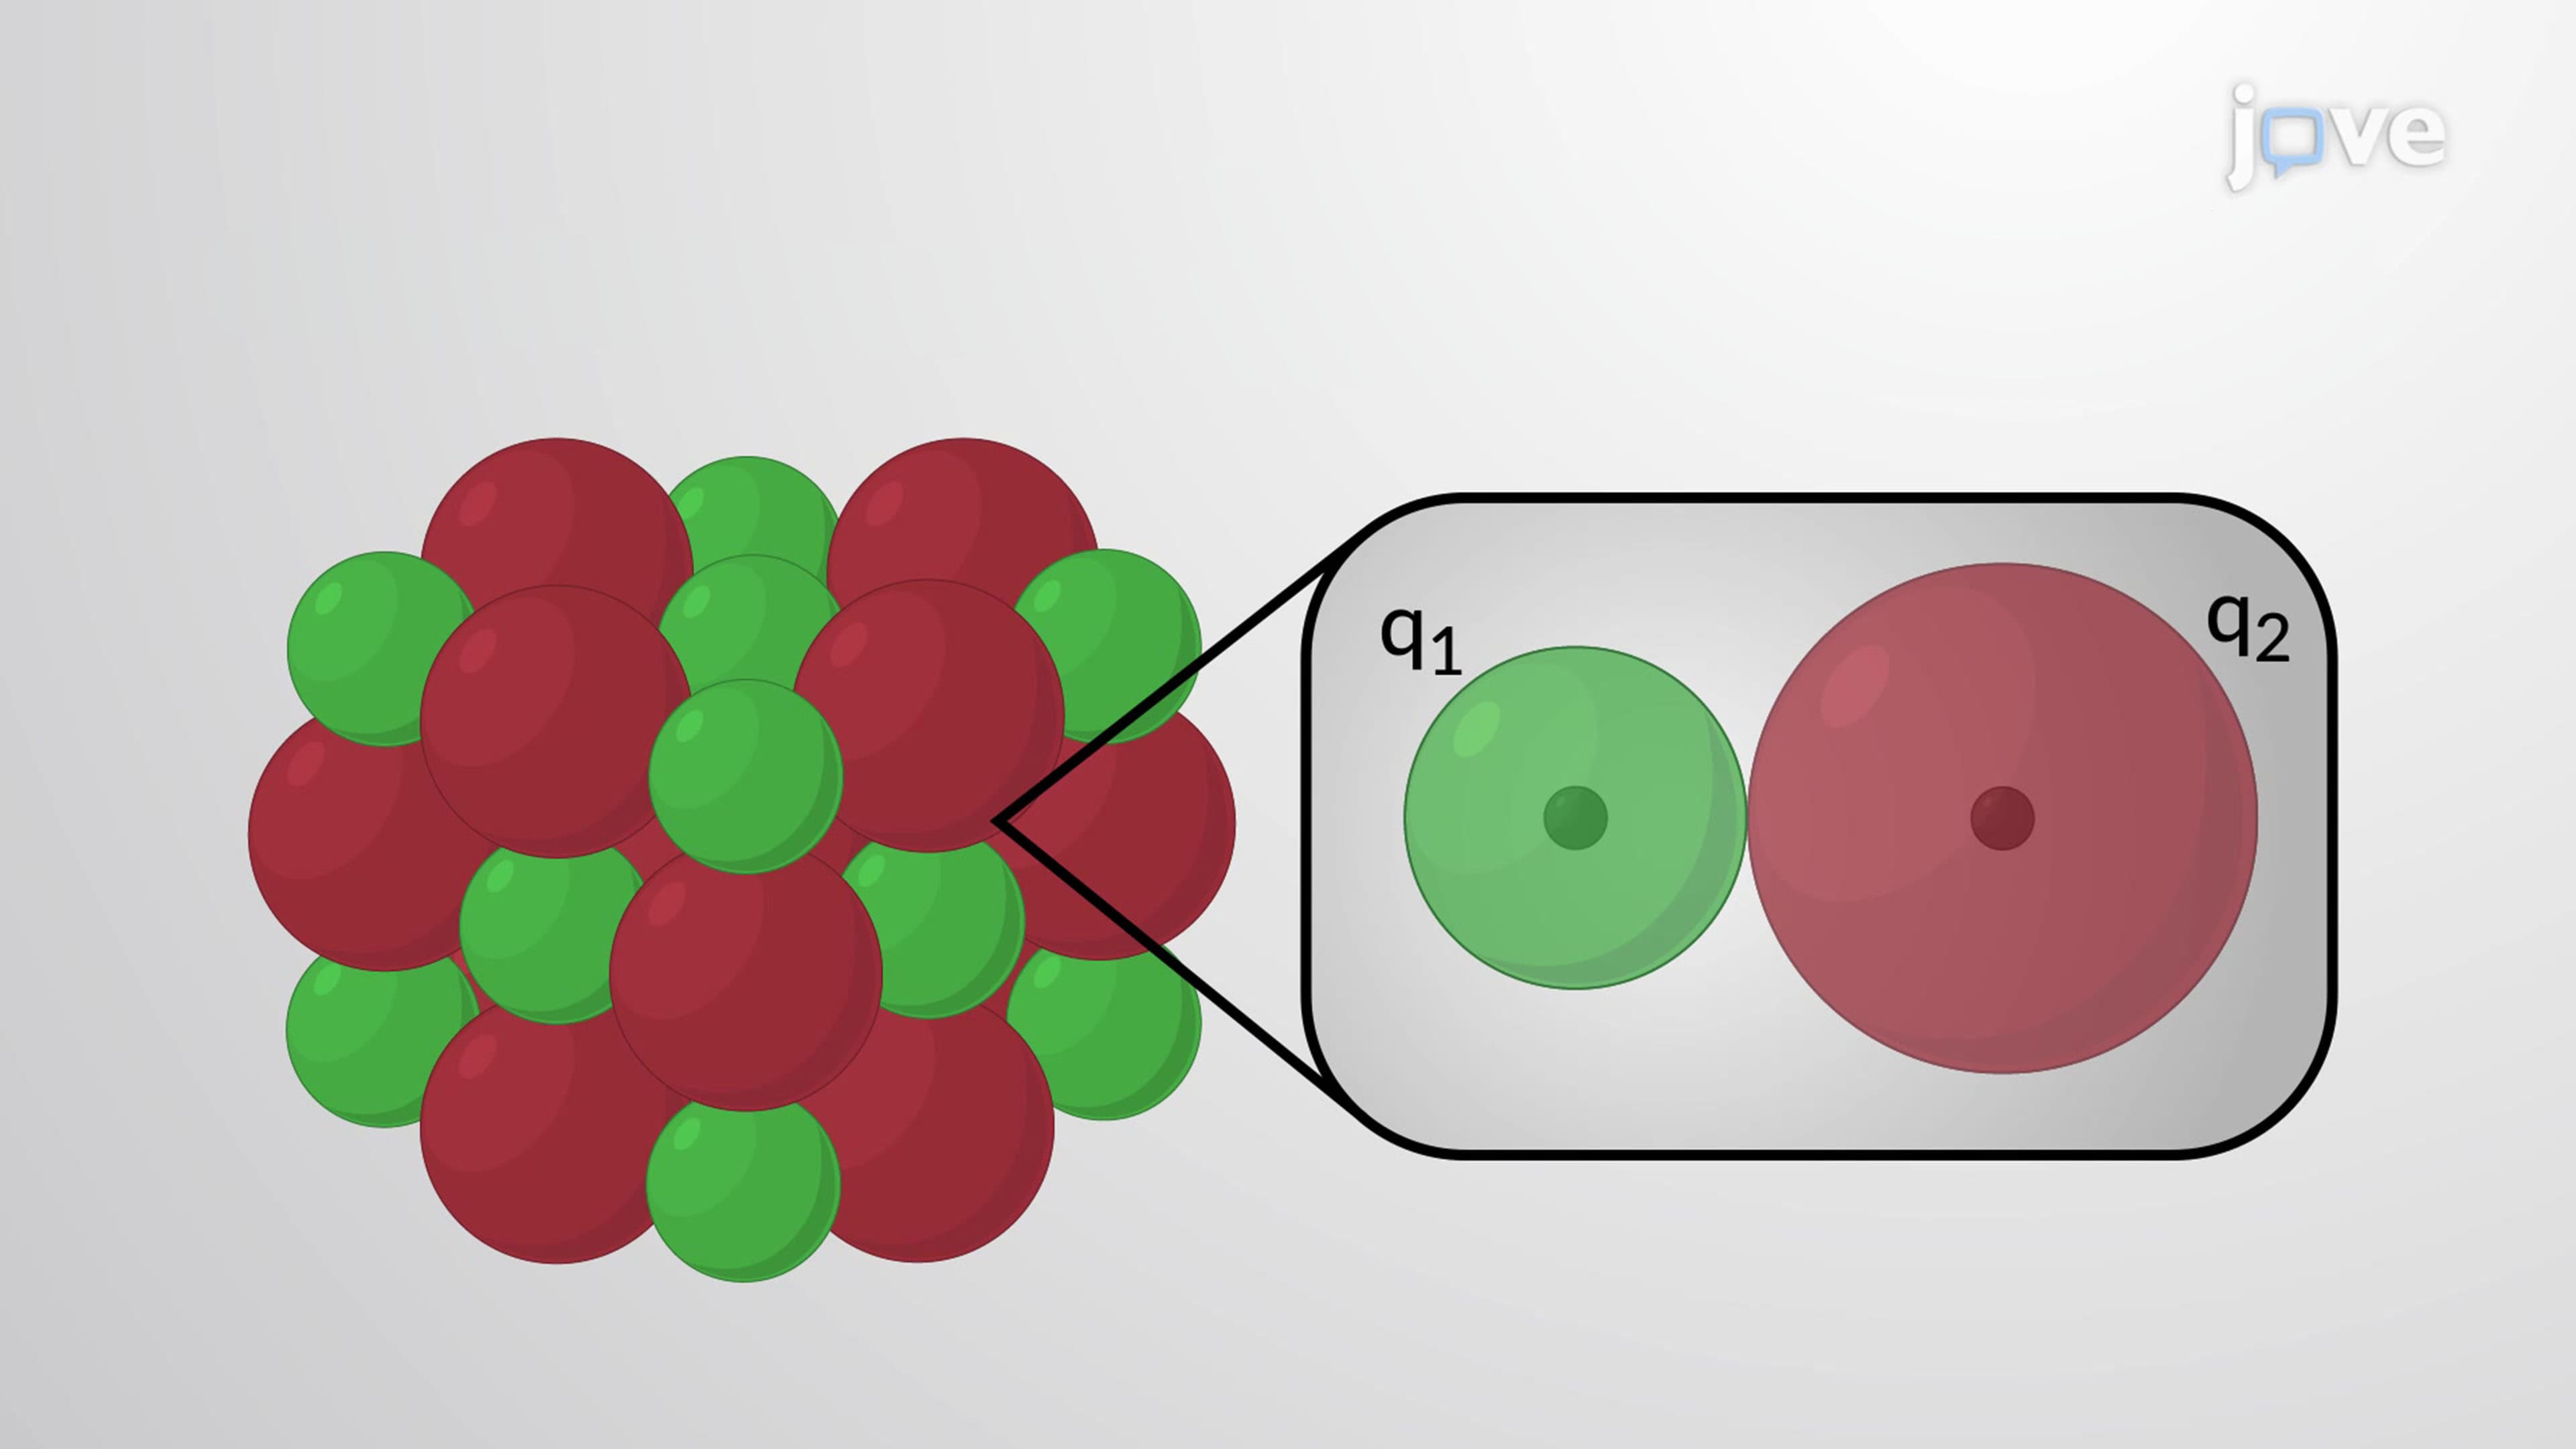
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 03:03
Chemical Reactions in Aqueous Solutions
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:54
Trends in Lattice Energy: Ion Size and Charge