基于未结合的万科米辛度监测的万科米辛的临床有效性和安全性
Fefei Ren1, Shan Li, Yixin Liu
1The Second Hospital of Hebei Medical University, Shijiazhuang, China.
Therapeutic drug monitoring
|December 11, 2024

概括
监测万科米辛的总低度 (Cmin_total) 和未结合的低度 (Cmin_free) 是至关重要的. 这两种度的较高水平与急性损伤 (AKI) 的风险增加有关.
科学领域:
- 药理动力学和药理动力学
- 临床化学 临床化学
- 腎臟病學 (nephrology) 是一種醫學專業.
背景情况:
- 范科米辛是治疗严重的格拉姆阳性感染的关键抗生素.
- 监测万科米辛度对于优化疗效和最大限度地降低毒性至关重要.
- 了解影响万科米辛水平及其临床影响的因素至关重要.
研究的目的:
- 为了监测万科米辛的总低度 (Cmin_total) 和未结合的低度 (Cmin_free).
- 分析影响万科米辛Cmin_total和Cmin_free.free的因素.
- 评估万科米辛度与临床疗效和急性损伤 (AKI) 的相关性.
主要方法:
- 血样本使用蛋白质沉和空腔纤维离心超处理.
- 使用高性能液体染色学 (HPLC) 来确定Cmin_total和Cmin_free.
- 收集了临床数据,以调查与疗效和毒性的相关性.
主要成果:
- 范科米Cmin_total和Cmin_free分别在0.62-56.08 mcg·mL-1和0.61-38.51 mcg·mL-1之间.
- Cmin_total和Cmin_free是相关的 (r=0.8899) 并且受到基底肌素和囊素C的影响.
- 较高的Cmin_free (>8.6mcg·mL-1) 和同时服用的毒药物是与胺相关的AKI的危险因素.
结论:
- 范科米辛Cmin_total和Cmin_free是相关的.
- 较高的Cmin_total和Cmin_free度可能会增加AKI的风险.
- 治疗药物监测万科米对于患者的安全至关重要.
相关概念视频
 01:15
Drug Concentration Versus Time Correlation
617
The plasma drug concentration-time curve is a crucial tool in pharmacokinetics, representing the drug's concentration in plasma at different time intervals post-administration. This curve illustrates the drug's journey from absorption into the systemic circulation, distribution to body tissues, and eventual elimination through excretion or biotransformation.
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the...
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the...
617
 01:23
Drug Concentrations: Measurements
326
Drug concentration is the quantity of a drug present in a biological sample. Measuring drug amounts in biological samples allows the clinician to understand how a drug is absorbed, distributed, metabolized, and excreted. Samples can be obtained through invasive or non-invasive methods. Invasive techniques involve surgical or parenteral interventions to gather blood, cerebrospinal fluid, or tissue biopsy. Conversely, non-invasive approaches provide samples like urine, feces, and saliva.
Plasma...
Plasma...
326
 01:14
Time Course of Drug Effect
2.0K
The progression of a drug's impact can be analyzed by examining both the concentration-time course and the effect-time course. The concentration-time course is determined by the drug's half-life and is influenced by factors such as its pharmacokinetics, including absorption, distribution, metabolism, and elimination. The effect of the drug is often related to its concentration in the plasma and is calculated using the maximum drug effect and the plasma concentration that generates 50...
2.0K
 01:24
One-Compartment Open Model: Wagner-Nelson and Loo Riegelman Method for ka Estimation
387
This lesson introduces two critical methods in pharmacokinetics, the Wagner-Nelson and Loo-Riegelman methods, used for estimating the absorption rate constant (ka) for drugs administered via non-intravenous routes. The Wagner-Nelson method relates ka to the plasma concentration derived from the slope of a semilog percent unabsorbed time plot. However, it is limited to drugs with one-compartment kinetics and can be impacted by factors like gastrointestinal motility or enzymatic degradation.
On...
On...
387
 01:25
Clearance Models: Compartment Models
48
Clearance measures drug elimination from the central compartment, including plasma and highly perfused organs like kidneys and liver. Its calculation varies depending on pharmacokinetic models and administration routes. The one-compartment model, for instance, portrays the pharmacokinetics of polar drugs such as aminoglycoside antibiotics administered intravenously and readily excreted in urine. In this case, clearance is influenced by the terminal rate constant (λz) and the total volume...
48
 01:22
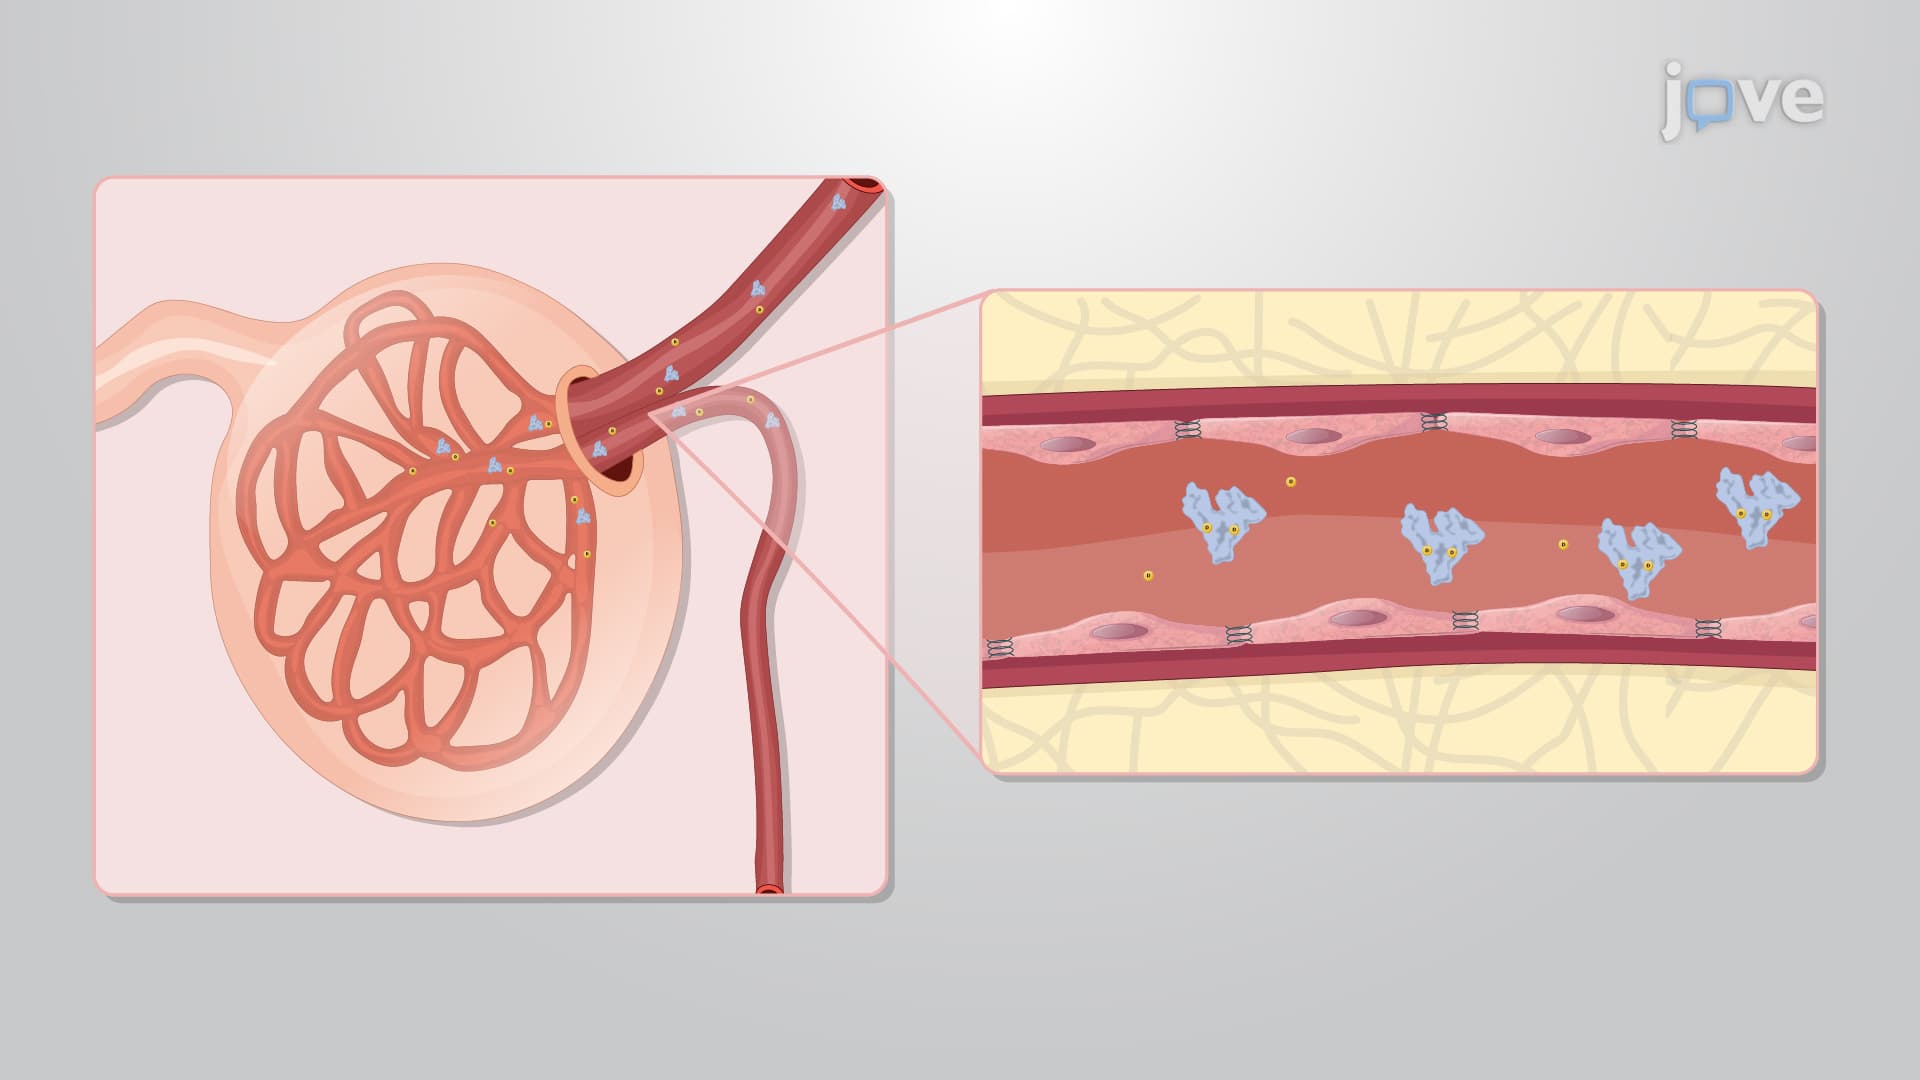
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
117
When a drug follows nonlinear pharmacokinetics, its bioavailability, the amount of the drug that reaches the systemic circulation, can change with different doses. This is due to the presence of a saturable pathway. The pathway becomes saturated as the drug concentration increases, decreasing the absorption rate. Consequently, the drug's bioavailability may be lower than expected at higher doses.
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
117