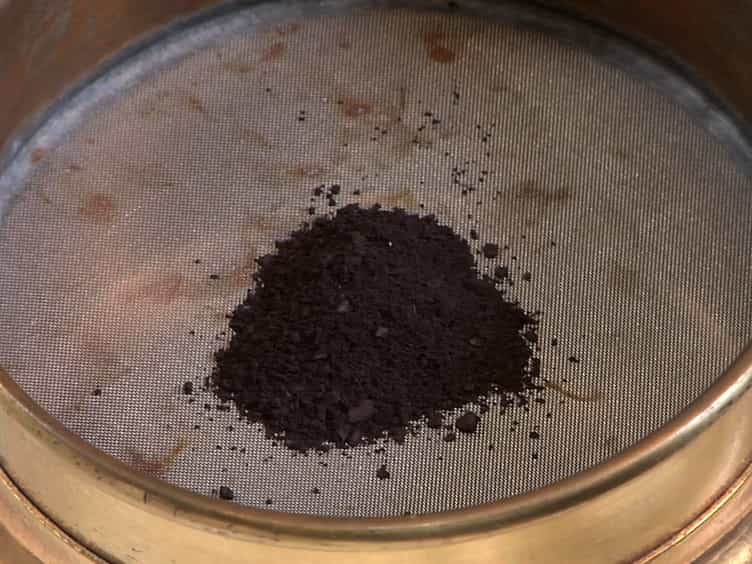
空位和低能量的3p-轨道内置Na4Fe3(PO4) 2(P2O7) 阴极具有优越的储存动力学
Ning Jiang1,2, Cheng Yang1, Yichao Wang1,2
1Shanghai Institute of Ceramics, Chinese Academy of Sciences, Shanghai, 200050, China.
Small (Weinheim an der Bergstrasse, Germany)
|January 6, 2025

概括
这项研究通过引入过渡金属空缺来增强用于离子电池 (SIB) 的铁基酸盐阴极. 这提高了导电性,并使高能量密度和特殊的循环稳定性用于实际应用.
科学领域:
- 材料科学 材料科学 材料科学
- 电化学 电化学 电化学
- 固态化学 固态化学
背景情况:
- 离子电池 (SIBs) 是有前途的储能解决方案,因为大量的.
- 铁基酸Na4Fe3(PO4)2(P2O7) (NFPP) 是SIBs的经济高效和环保的阴极材料.
- 核电发电厂本身具有较低的离子和电子导电性,这阻碍了其实际应用.
研究的目的:
- 研究低能3p轨道和过渡金属空缺在提高NFPP阴极性能方面的作用.
- 通过晶体学和电子相互作用探索NFPP的修饰机制.
- 为高性能SIB设计和合成一个增强的NFPP阴极.
主要方法:
- 结晶学分析分析的方法
- 电子相互作用研究的研究.
- 有各种各样的表征技术.
- 理论上的计算理论上的计算.
- 合成的Na4Fe2.85Al0.1(PO4)2(P2O7) (NF2.85A0.1PP) 的合成方法
主要成果:
- 该研究阐明了过渡金属空缺和3p轨道相互作用如何促进电荷重组和重建离子扩散路径.
- 设计的NF2.85A0.1PP阴极具有高能量密度,约为350Wh kg-1.1.
- 实现了卓越的电化学性能,包括卓越的动力学 (62 mAh g-1在10 A g-1),优异的功率密度 (23 kW kg-1,143 Wh kg-1),以及超过10,000个周期的显著循环稳定性.
结论:
- 引入过渡金属空缺是一个可行的策略,以克服聚离子阴极的导电性限制.
- 这项研究为设计下一代离子电池的超耐用,高速聚离子阴极提供了新的视角.
- 这些发现为更高效,更稳定的SIB铺平了道路.
更多相关视频
相关概念视频
 01:19
Ionic Strength: Effects on Chemical Equilibria
1.3K
The addition of an inert ionic compound increases the solubility of a sparingly soluble salt. For example, adding potassium nitrate to a saturated solution of calcium sulfate significantly enhances the solubility of calcium sulfate. Le Châtelier's principle cannot predict this shift in the equilibrium. Instead, this could be explained in terms of changes in the effective concentration of the ions in solution in the presence of added inert salt.
In this solution, the primary...
In this solution, the primary...
1.3K
 02:42
Valence Bond Theory
8.4K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
8.4K
 02:58
Crystal Field Theory - Octahedral Complexes
26.1K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
26.1K
 02:42
Ionic Crystal Structures
14.1K
Ionic crystals consist of two or more different kinds of ions that usually have different sizes. The packing of these ions into a crystal structure is more complex than the packing of metal atoms that are the same size.
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
14.1K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
41.4K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
41.4K
 03:26
Electron Configuration of Multielectron Atoms
38.8K
The alkali metal sodium (atomic number 11) has one more electron than the neon atom. This electron must go into the lowest-energy subshell available, the 3s orbital, giving a 1s22s22p63s1 configuration. The electrons occupying the outermost shell orbital(s) (highest value of n) are called valence electrons, and those occupying the inner shell orbitals are called core electrons. Since the core electron shells correspond to noble gas electron configurations, we can abbreviate electron...
38.8K