
多态过渡金属碳烯结合超出最小能量路径:旋转轨道相互作用的非局部性
Daisuke Yoshida1,2, Kaito Takahashi1,3
1Institute of Atomic and Molecular Sciences, Academia Sinica, Taipei 10617, Taiwan.
Inorganic chemistry
|January 13, 2025

概括
旋转轨道效应影响过渡金属碳和二复合体的形成. 计算显示,在 - 合体关联路径上存在旋转屏障,影响反应速率.
科学领域:
- 无机化学 无机化学
- 计算化学的计算化学
- 物理化学 物理化学
背景情况:
- 过渡金属碳和二复合物在生物学和催化中至关重要.
- 对过渡金属的联结合诱导了自旋状态的变化.
- 对这些关联路径的旋转轨道效应仍然未被评估.
研究的目的:
- 为了研究旋转轨道 (SO) 对过渡金属-碳基 (TM-CO) 和过渡金属-二 (TM-N2) 键的关联途径的影响.
- 计算 (Ni) 与一氧化碳 (CO) 和二氧化 (N2) 反应的关联潜在能量曲线.
主要方法:
- 使用了包含旋转轨道 (SO) 相互作用的多参考计算.
- 计算了11个电子状态的关联潜在能量曲线.
- 分析的旋转交叉反应:Ni + CO → NiCO和Ni + N2 → NiN2.
主要成果:
- 最小能量通路 (MEP) 提供了准确的异位能量,但不准确的中间键长图.
- 旋转轨道相互作用在2.3-2.5 Å的纽带长度区域加剧,在交叉点之前.
- 在Ni-adsorbate结合路径上发现了0.15 eV的自旋屏障.
结论:
- 旋转轨道相互作用在TM-连接体协会期间的旋转交叉机制中起着至关重要的作用.
- 已识别的旋转障碍对关联率有显著的影响,可能是数量级.
- 这项研究为转变金属化学中旋转轨道效应的重要性提供了新的视角.
更多相关视频
 06:53
Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
1.9K
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
8.3K
相关概念视频
 02:40
MO Theory and Covalent Bonding
10.3K
The molecular orbital theory describes the distribution of electrons in molecules in a manner similar to the distribution of electrons in atomic orbitals. The region of space in which a valence electron in a molecule is likely to be found is called a molecular orbital. Mathematically, the linear combination of atomic orbitals (LCAO) generates molecular orbitals. Combinations of in-phase atomic orbital wave functions result in regions with a high probability of electron density, while...
10.3K
 01:17
Spin–Spin Coupling: One-Bond Coupling
931
Coupling interactions are strongest between NMR-active nuclei bonded to each other, where spin information can be transmitted directly through the pair of bonding electrons. While nuclei polarize their electrons to the opposite spins, the bonding electron pair has opposite spins. Configurations with antiparallel nuclear spins are expected to be lower in energy. When coupling makes antiparallel states more favorable, J is considered to have a positive value. The one-bond coupling constant, 1J,...
931
 02:42
Valence Bond Theory
8.4K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
8.4K
 02:32
Bonding in Metals
46.8K
Metallic bonds are formed between two metal atoms. A simplified model to describe metallic bonding has been developed by Paul Drüde called the “Electron Sea Model”.
46.8K
 03:51
Molecular Orbital Theory II
19.0K
Molecular Orbital Energy Diagrams
19.0K
 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
1.0K
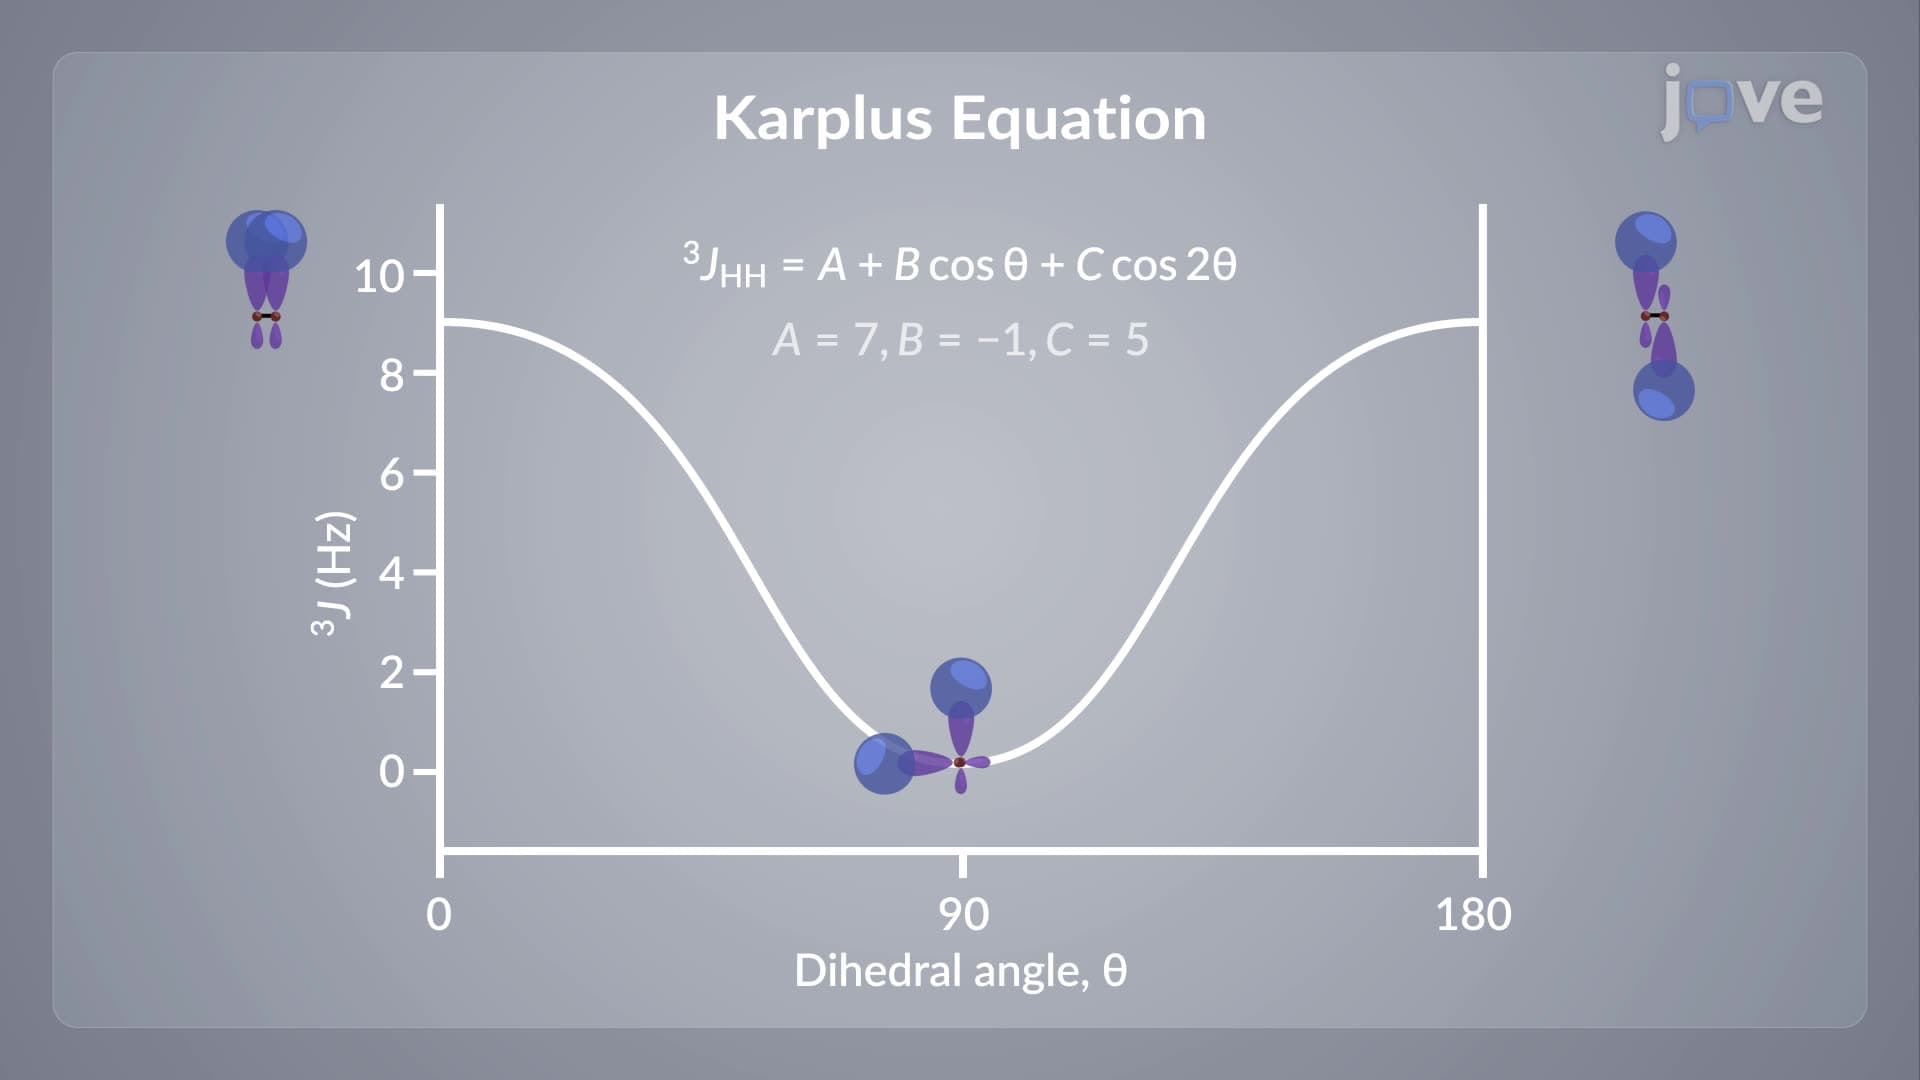
Vicinal or three-bond coupling is commonly observed between protons attached to adjacent carbons. Here, nuclear spin information is primarily transferred via electron spin interactions between adjacent C‑H bond orbitals. This generally favors the antiparallel arrangement of spins, so 3J values are usually positive.
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
1.0K