
在大鼠血中使用givinostat的UPLC-MS/MS方法:开发,验证,体内药理动力学研究和体内代谢稳定性研究
Ruanjuan Zhan1, Yanan Liu1,2, Jun Wu1,2
1The First Affiliated Hospital of Wenzhou Medical University, Zhejiang, People's Republic of China.
Drug design, development and therapy
|January 20, 2025

概括
吉维诺斯塔特是一种有前途的白血病HDAC抑制剂,在老鼠中被迅速吸收并缓慢清除. 这项研究开发了一种用于测量givinostat的快速分析方法,有助于其临床前和临床发展.
科学领域:
- 药理学 药理学是指药理学的学科.
- 分析化学 分析化学
- 生物化学 生物化学
背景情况:
- 吉维诺斯塔特是一种强效的基因素脱乙酶 (HDAC) 抑制剂,具有治疗复发性白血病和骨髓瘤的治疗潜力.
- 了解givinostat的药理动力学和代谢特征对于其临床应用至关重要.
研究的目的:
- 开发和验证用于量化givinostat度的快速试验.
- 在临床前模型中建立givinostat的初步药理动力学概况.
- 为了评估givinostat的体外代谢稳定性.
主要方法:
- 超高性能液态染色体协同质谱法 (UPLC-MS/MS) 用于givinostat测量.
- 埃利格斯图特 (Eliglustat) 作为检测量量的内部标准 (IS).
- 用蛋白沉进行样本制备,然后在C18柱上进行梯度化.
主要成果:
- 在UPLC-MS/MS试验中,在广泛的度范围 (24000 ng/mL) 上显示出优异的线性 (r2=0.998).
- 该方法表现出高精度 (95.8%-108.6%) 和精度 (RSD% <15%),超过90%的回收率和最小的矩阵效应.
- 鼠类的药理动力学研究显示,吉维诺斯塔特的吸收速度快,清除速度缓慢,体外半衰期为92.87分钟,内在清除速度缓慢.
结论:
- 吉维诺斯塔特在体内被快速吸收并缓慢排出,这与体外代谢稳定性研究结果一致.
- 开发的UPLC-MS/MS测定适用于givinostat的临床前药理动力学研究.
- 这项研究提供了有价值的数据,以支持givinostat的正在进行的临床开发.
更多相关视频
相关概念视频
 01:09
Methods for Studying Drug Absorption: In situ
In situ experiments, such as the Doluisio method and Single-Pass Perfusion technique, provide critical insights into drug uptake by simulating in vivo conditions for drug absorption.
The Doluisio method involves perfusing a prepared segment of a rat's small intestine with a solution of radiolabeled drug and a non-absorbable marker. This helps to differentiate between absorbed and non-absorbed drug concentrations. The intestinal segment is connected at both ends using tubing and syringes,...
The Doluisio method involves perfusing a prepared segment of a rat's small intestine with a solution of radiolabeled drug and a non-absorbable marker. This helps to differentiate between absorbed and non-absorbed drug concentrations. The intestinal segment is connected at both ends using tubing and syringes,...
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods
Pharmacokinetics is a vital branch of pharmacology that examines how drugs are absorbed, distributed, metabolized, and excreted by the body. Two key methodologies in pharmacokinetics are plasma drug concentration studies and urinary drug excretion analyses, both of which provide critical insights into a drug's therapeutic efficacy and bioavailability.Plasma Drug Concentration-Time StudiesPlasma drug concentration-time studies involve analyzing blood samples at specific intervals to quantify...
 01:20
Measurement of Bioavailability: Pharmacodynamic Methods
Pharmacodynamic methods provide insights into a drug's effects on physiological processes over time and play a crucial role in understanding bioavailability and therapeutic efficacy. These methods can be broadly classified into acute pharmacological and therapeutic response approaches, each with distinct mechanisms and applications.The acute pharmacological response method directly correlates a drug's physiological effects, such as ECG or pupil diameter changes, to its time course in the body.
 01:10
In Vitro Drug Release Testing: Overview, Development and Validation
In vitro dissolution and drug release tests assess how quickly and how much of a drug is released from its dosage form into an aqueous medium under standardized laboratory conditions. These tests are essential tools in pharmaceutical development and quality assurance, offering insight into the drug's performance before clinical use.During formulation development, dissolution testing identifies incomplete or inconsistent drug release issues. It also supports decisions on selecting the optimal...
 01:30
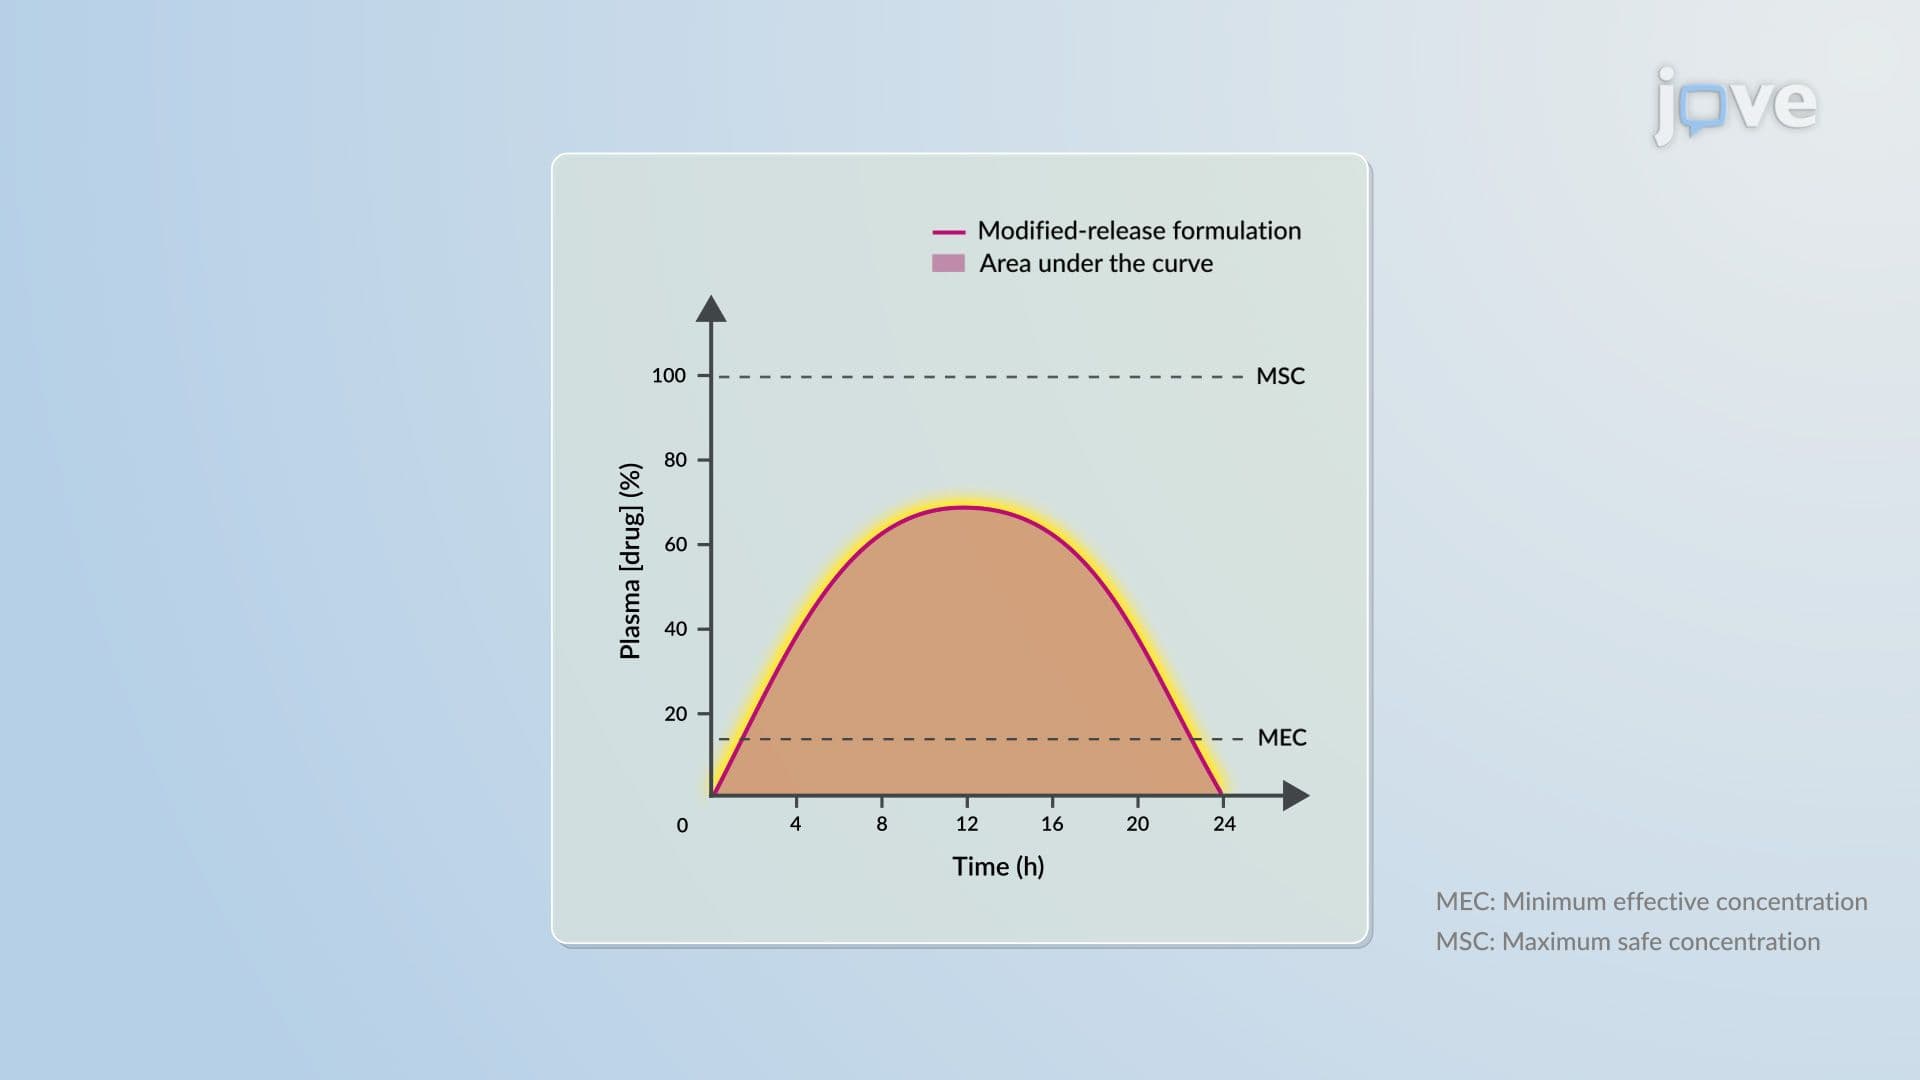
Modified-Release Drug Delivery Systems: Bioavailability
Modified-release (MR) dosage forms are designed to extend drug release over time, thereby maintaining stable plasma concentrations and reducing dosing frequency. However, their bioavailability is typically below 100% due to incomplete drug release and presystemic metabolism, and limitations in drug permeability across the gastrointestinal epithelium, all of which can restrict the fraction of the drug reaching systemic circulation. Consequently, studying the in vivo bioavailability of MR...