将密度纠正密度函数理论扩展到大型分子系统

Youngsam Kim1, Mingyu Sim1, Minhyeok Lee1
1Department of Chemistry, Yonsei University, 50 Yonsei-ro, Seodaemun-gu, Seoul 03722, Korea.

密度校正密度函数理论 (DC-DFT) 的计算使用双基法使得更高效. 这种方法加速了大型分子系统的哈特里-福克 (HF) 密度估计,增强了计算化学研究.
科学领域:
- 计算化学是一种计算化学.
- 量子化学是一种量子化学.
- 材料科学是一种材料科学.
背景情况:
- 实际的密度校正密度函数理论 (DC-DFT) 计算通常依赖于计算密集的哈特里-福克 (HF) 密度,限制它们对大型系统的应用.
- 对于超过一百个原子的系统,估计高频密度是一个重要的计算瓶.
研究的目的:
- 为了提高Hartree-Fock密度校正密度函数理论 (HF-DC-DFT) 对大分子系统的适用性.
- 引入和验证用于加速高频密度计算的双基法.
主要方法:
- 采用双基法,利用较小的基数集的密度矩阵,在较大的基数集上近似计算HF解决方案.
- 该方法在各种系统上进行了基准测试,包括GMTKN55数据库 (主要组化学) 和L7/S6L数据集 (大型分子系统).
- 详细介绍了HF-r2SCAN-DC4最近的重定型,评估其性能影响.
主要成果:
- 双基法在加速大型系统的高频密度估计方面表现出显著的有效性.
- 基准证实了该方法在各种化学系统中的准确性和可靠性.
- 对DNA和艾滋病毒系统的应用显示了与现有文献方法可比的结果.
结论:
- 双基法有效地将HF-DC-DFT计算的实际范围扩展到更大,更复杂的分子系统.
- 这种计算加速为研究复杂的生物和化学结构开辟了新的途径.
- 在HF-r2SCAN-DC4重定量化保持性能,确保在DC-DFT应用程序的持续准确性.
更多相关视频
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
相关概念视频
 04:01
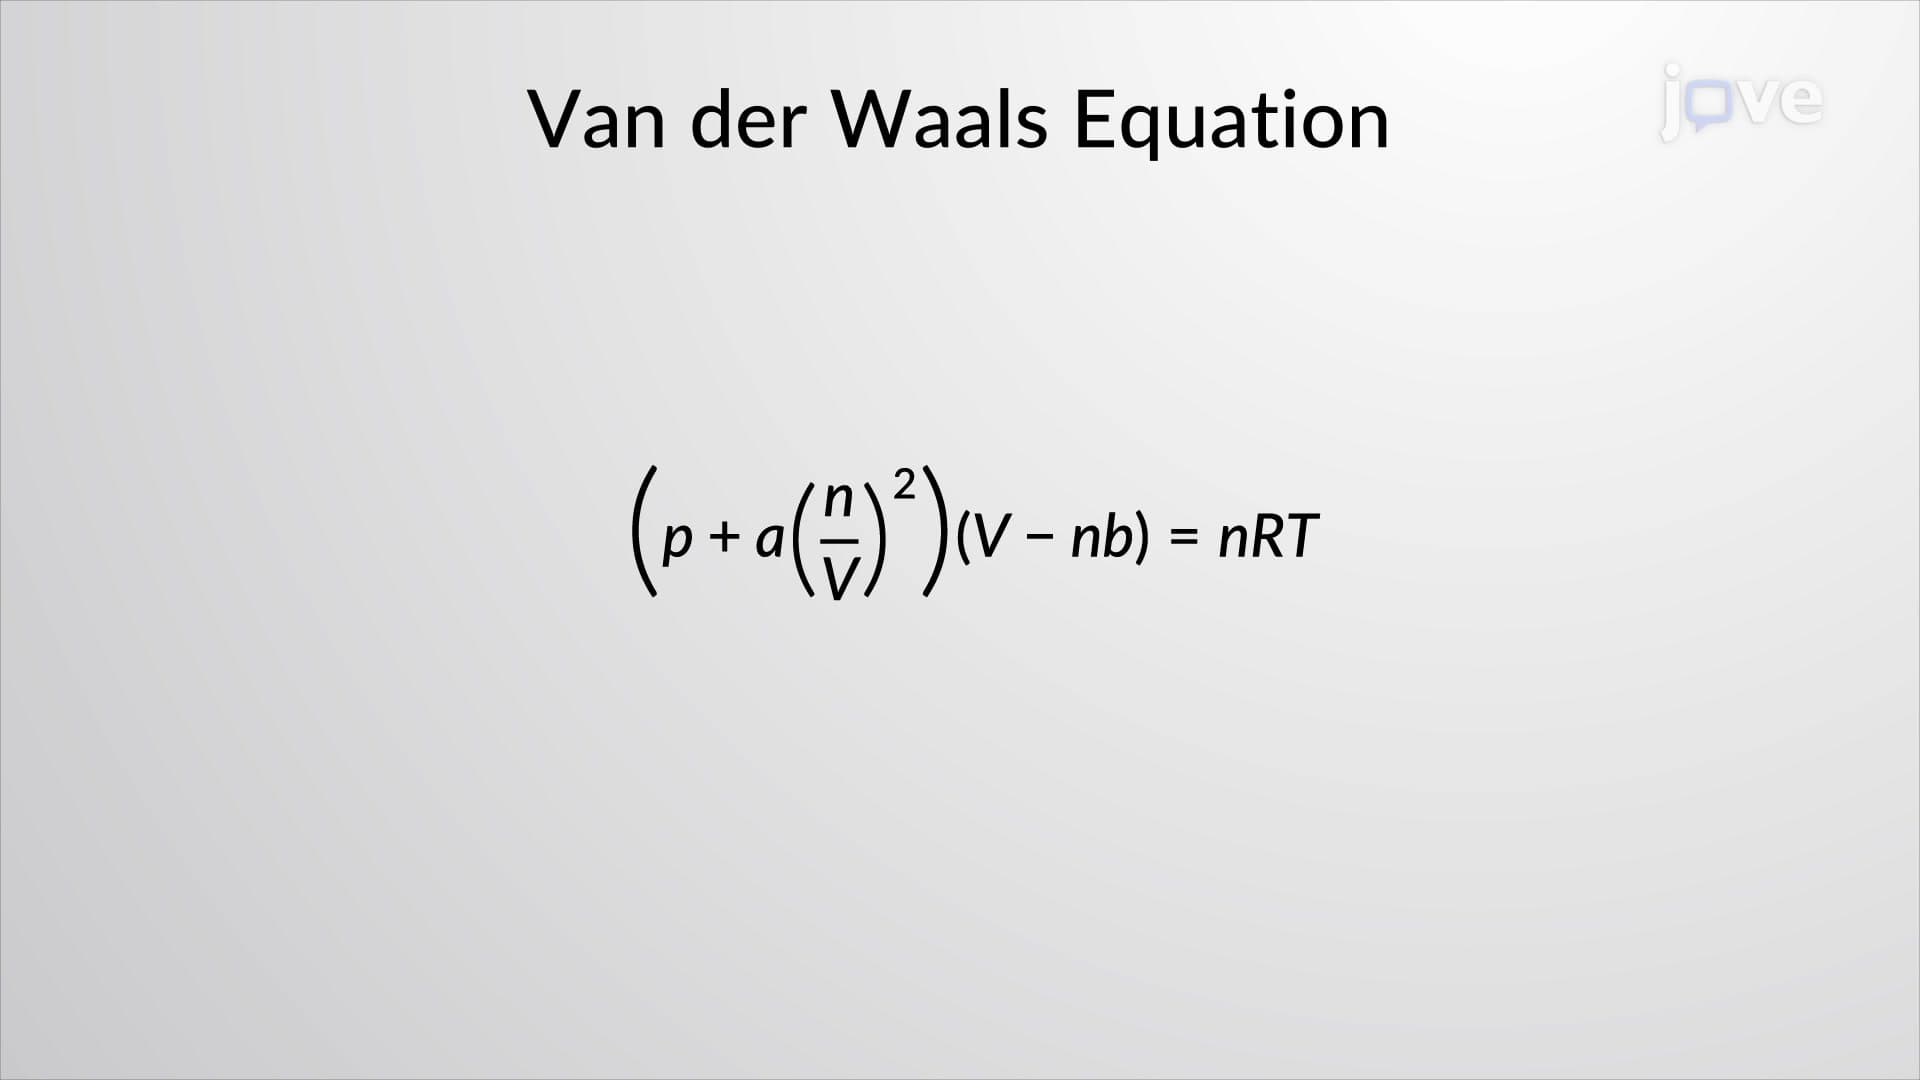
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:10
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 01:20
Maxwell-Boltzmann Distribution: Problem Solving
This distribution function f(v) is defined by saying that the expected number N (v1,v2) of particles with speeds between v1 and v2 is given by
 01:24
Van der Waals Interactions