

一个有用的可改变价值的光学探头,用于根据理论计算预测材料特性
Linghui Zhang1,2,3, Guoqiang Peng2, Yujiang Wang1
1College of Physics and Electronic Engineering, Northwest Normal University, Lanzhou, 730070, China. hanlili.06@163.com.
Physical chemistry chemical physics : PCCP
|January 23, 2025

概括
理论计算揭示了BaSnSi3O9.9中Ba和Sn位点的不同电子特性. 引入欧离子表明了这些位点如何影响材料特性,指导功能性材料设计.
科学领域:
- 材料科学 材料科学 材料科学
- 计算材料科学科学 计算材料科学
- 固态化学 固态化学
背景情况:
- 材料功能性质的可控制设计依赖于理论计算和实验验证.
- 了解电离位点的电子特征对于预测材料的行为至关重要.
- 第一原理计算提供了对材料属性的洞察,例如光学和介电行为.
研究的目的:
- 在BaSnSi3O9 (BSSO) 中研究Ba和Sn位点的独特电子特征,使用第一原则计算.
- 探索这些不同的位点对内置的功能离子,特别是欧 (Eu) 的特性的影响.
- 为设计具有定制功能性质的材料提供理论指导.
主要方法:
- 对无机化合物进行了第一原理计算,重点是BaSnSi3O9.
- 在Ba和Sn电位点分析了电子特征和电荷密度分布.
- 使用欧 (Eu) 作为光学探测器,以实验验证有关特定地点属性的理论预测.
主要成果:
- 在BSSO中确定了两个不同的阴离子位点:具有局部电子密度的Ba位点和具有强烈环境相互作用的Sn位点.
- 在Ba位点的欧离子表现出Eu3+和Eu2+发光的共存,发光灭度更高.
- 在Sn位点的欧离子仅显示了Eu3+发光和较低的发光灭度,这是由于氧相互作用造成的.
结论:
- 关于Ba和Sn站点独特电子特征的理论预测被实验验证,使用Eu作为探针.
- Ba和Sn位点的不同电子环境显著影响了内置离子的价值状态和光学特性.
- 这项研究强调了理论计算的潜力,以指导功能无机材料的合理设计和性能调制.
相关概念视频
 01:32
UV–Vis Spectroscopy of Conjugated Systems
6.8K
Organic compounds with conjugated double bonds show strong absorption features in the UV–visible region of the electromagnetic spectrum attributed to π → π* electronic excitations. Generally, a UV–vis absorption spectrum is recorded as a plot of absorbance vs wavelength. The wavelength of maximum absorbance, which manifests as a peak in the absorption spectrum, is denoted as λmax.
One of the factors influencing λmax is the extent...
One of the factors influencing λmax is the extent...
6.8K
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
23.7K
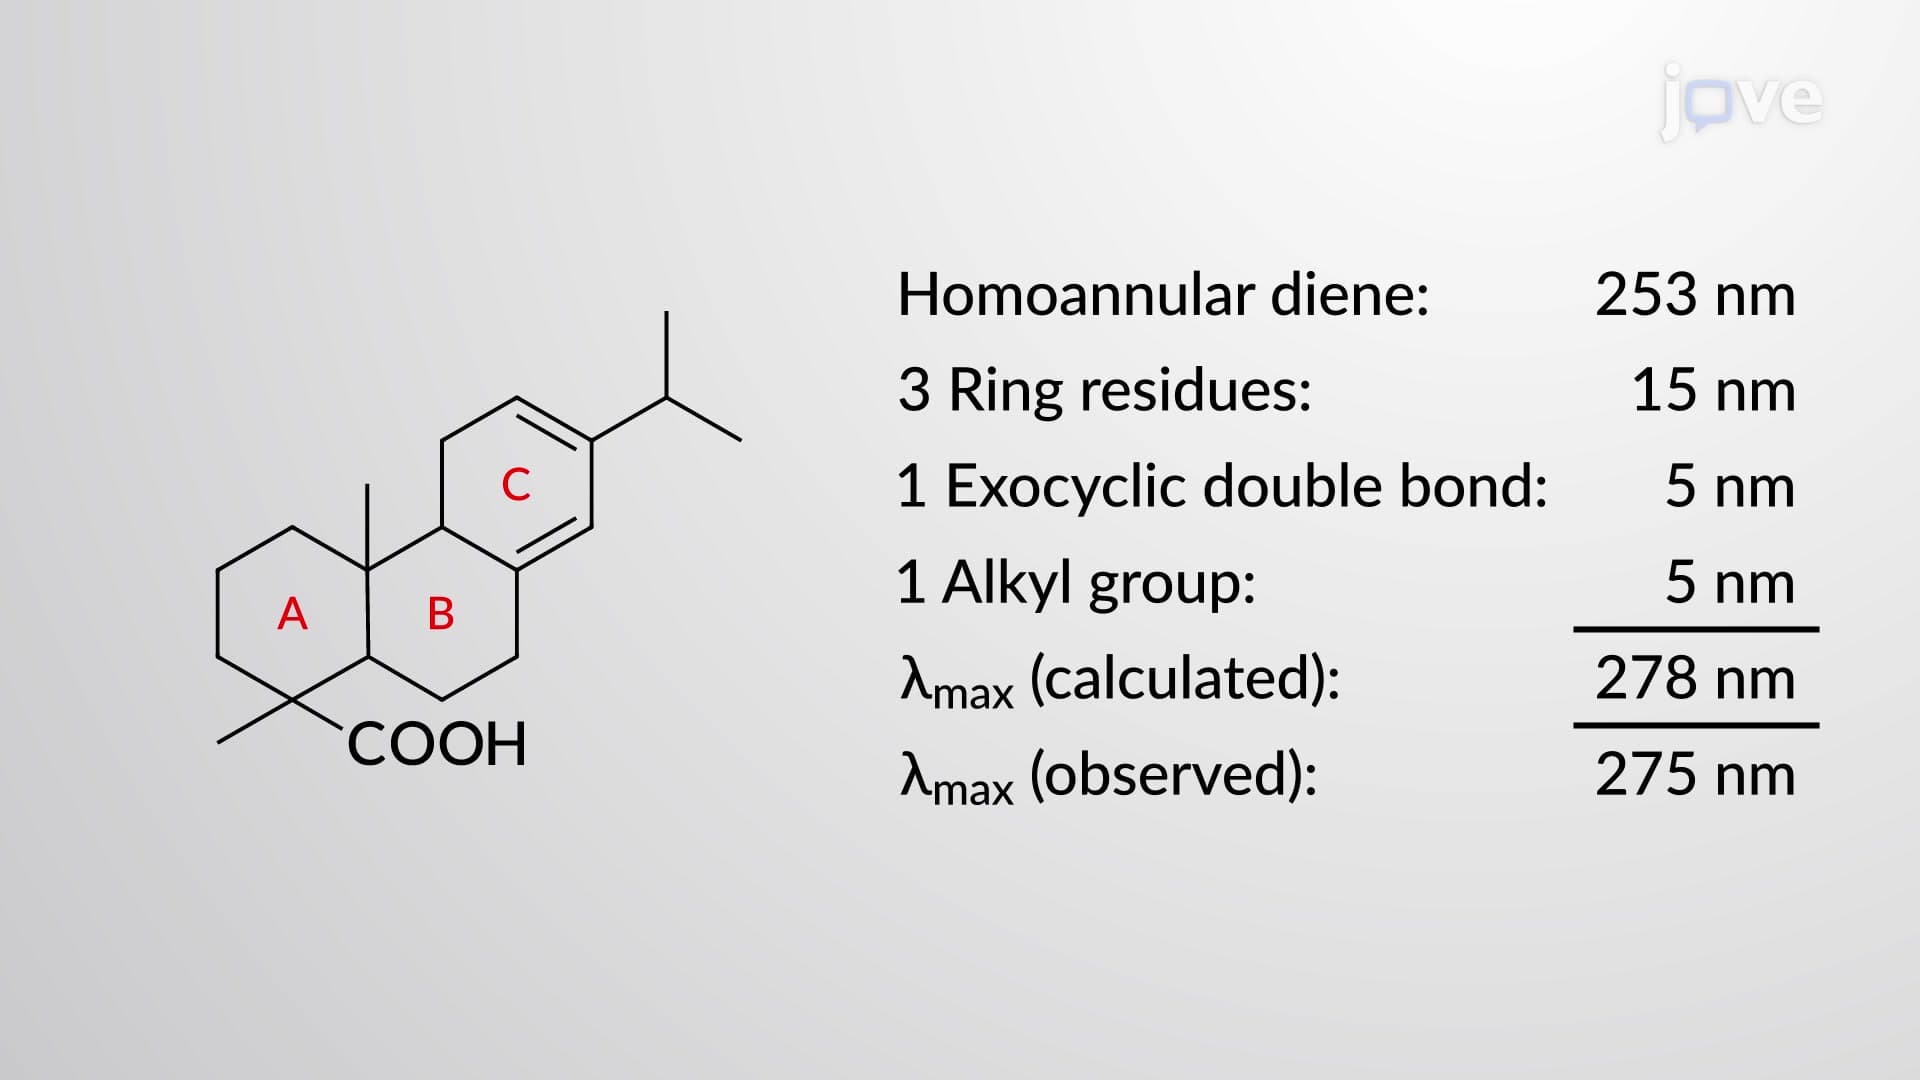
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given...
23.7K
 02:27
Predicting Molecular Geometry
34.0K
VSEPR Theory for Determination of Electron Pair Geometries
34.0K
 01:27
π Electron Effects on Chemical Shift: Overview
1.0K
An applied magnetic field causes loosely bound π-electrons in organic molecules to circulate, producing a local or induced diamagnetic field over a large spatial volume. As the molecules tumble in solution, the field generated by π-electrons in spherical substituents results in a zero net field. However, the net field generated by π-electrons in non-spherical substituents is not zero. The effect of this induced field depends on the orientation of the molecule with respect to B0,...
1.0K
 01:10
Van der Waals Equation
3.9K
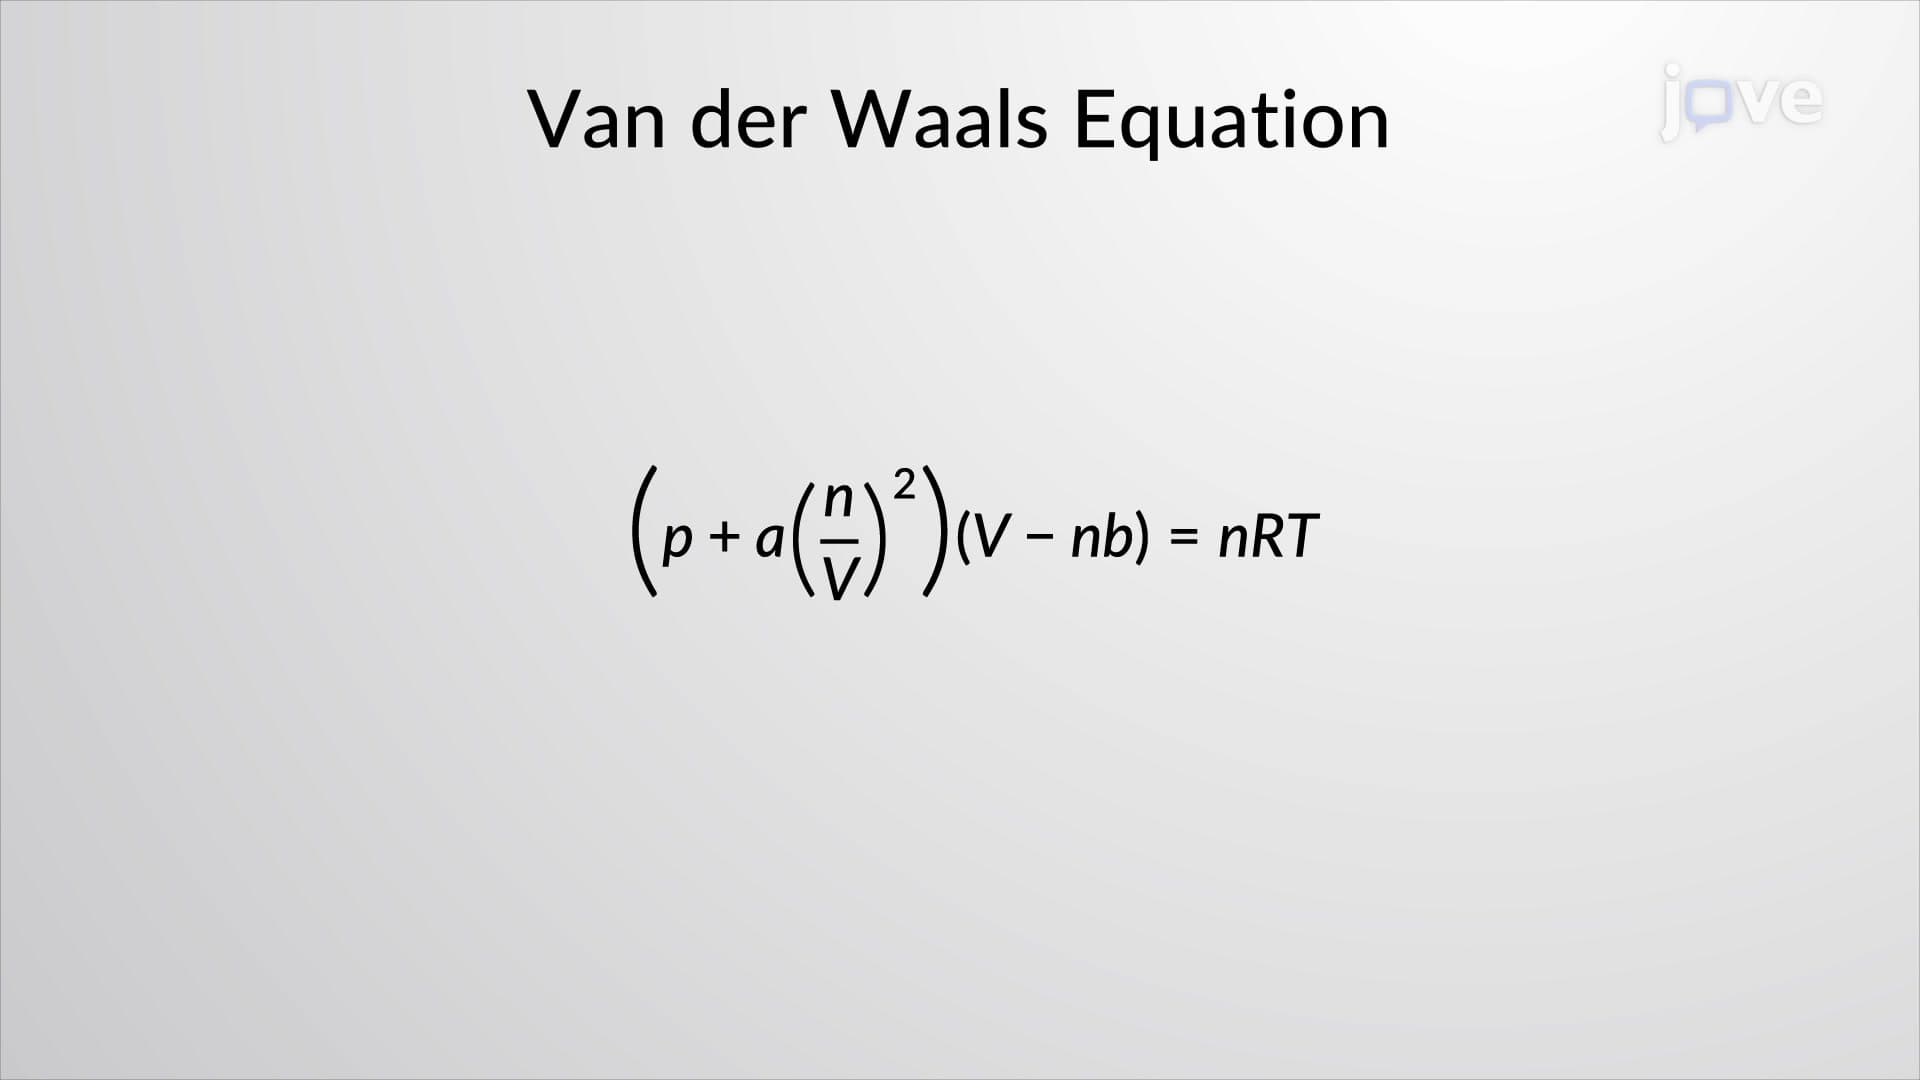
The ideal gas law is an approximation that works well at high temperatures and low pressures. The van der Waals equation of state (named after the Dutch physicist Johannes van der Waals, 1837−1923) improves it by considering two factors.
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
3.9K
 02:35
Molecular Orbital Theory I
31.7K
Overview of Molecular Orbital Theory
31.7K