相关实验视频
Updated: Jun 19, 2026
13:33

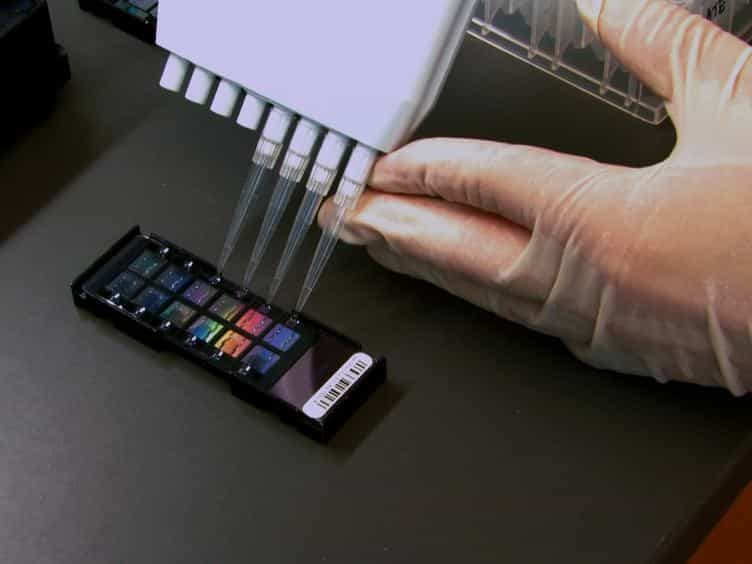
Infinium Assay for Large-scale SNP Genotyping Applications
Published on: November 19, 2013
38.9K
在双亲种群中,从杂的低覆盖度测序数据中快速准确地归纳基因型
Cécile Triay1, Alice Boizet2, Christopher Fragoso3,4
1DIADE, IRD, Cirad, University of Montpellier, Montpellier, France.
PloS one
|January 30, 2025

概括
一种名为NOISYmputer的新算法,使用低覆盖率的下一代测序 (LC-NGS) 数据,准确地对个体进行基因型鉴定. 它提高了遗传地图的精度,减少了错误,超过了现有的遗传分析方法.
科学领域:
- 基因组学就是基因组学.
- 生物信息学是一种生物信息学.
- 人口遗传学 人口遗传学
背景情况:
- 低覆盖下一代测序 (LC-NGS) 能够为创建遗传地图提供具有成本效益的基因定型.
- 挑战包括异构位的性能,缺失的数据,映射错误和技术NGS错误.
- 现有的归算方法与杂的LC-NGS数据作斗争.
研究的目的:
- 开发一种新的算法来归纳LC-NGS数据,以解决数据噪声并提高准确性.
- 为了准确地类型异质合体区域和估计交叉位置.
- 为使用LC-NGS数据进行基因分析提供一个强大而有效的工具.
主要方法:
- 开发了一种基于最大概率估计的算法 (NOISYmputer) 用于基因型和再组合断点赋值.
- 算法设计用于处理无需大量预先过的噪音数据.
- 将NOISYmputer与Tassel-FSFHap和LB-Impute进行比较,使用模拟和真实数据集.
主要成果:
- NOISYmputer在模拟的Illumina数据上实现了99.9%的断点精度和99.6%的回忆.
- 在处理杂数据方面,其表现优于Tassel-FSFHap和LB-Impute.
- 在真实数据集上提供了准确的遗传地图大小估计,与具有重大错误的替代工具不同.
结论:
- NOISYmputer有效地归因LC-NGS数据,纠正错误并提高基因型的准确性.
- 该算法在精度,回忆和计算效率方面提供了显著的优势.
- NOISYmputer是构建高密度遗传图和执行定量特征位置分析的宝贵工具.

相关概念视频
 02:05
Gene Evolution - Fast or Slow?
The genomes of eukaryotes are punctuated by long stretches of sequence which do not code for proteins or RNAs. Although some of these regions do contain crucial regulatory sequences, the vast majority of this DNA serves no known function. Typically, these regions of the genome are the ones in which the fastest change, in evolutionary terms, is observed, because there is typically little to no selection pressure acting on these regions to preserve their sequences.
In contrast, regions which code...
In contrast, regions which code...
 02:54
Evolutionary Relationships through Genome Comparisons
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
 02:26
Comparing Copy Number Variations and SNPs
Sequencing of the human genome has opened up several best-kept secrets of the genome. Scientists have identified thousands of genome variations that exist within a population. These variations can be a single nucleotide or a larger chromosomal variation.
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 01:05
Single Nucleotide Polymorphisms-SNPs
A single nucleotide polymorphism or SNP is a single nucleotide variation at a specific genomic position in a large population. It is the most prevalent type of sequence variation found in the human genome. Point mutations that occur in more than 1% of the population qualify as SNPs. These are present once every 1000 nucleotides on an average in the human genome. Replacement of a purine with another purine (A/G) or a pyrimidine with another pyrimidine (C/T) is known as a transition. In contrast,...
 01:29
Modern Molecular Taxonomy
Advancements in molecular biology have revolutionized the identification and characterization of bacteria, with multiple methods leveraging DNA sequencing for enhanced precision. As sequencing technologies improve and costs decline, these approaches are increasingly used in clinical, environmental, and evolutionary studies.Multilocus Sequence Typing (MLST) examines several housekeeping genes, essential chromosomal genes encoding cellular functions, to distinguish strains. Approximately...