通过全基因组分析检测非线性依赖
Wonuola A Akingbuwa1,2, Michel G Nivard3,4
1Department of Biological Psychology, Vrije Universiteit Amsterdam, Amsterdam, the Netherlands.
bioRxiv : the preprint server for biology
|February 24, 2025

概括
这项研究引入了一种新的统计方法,以揭示特征之间的复杂,非线性遗传关系. 研究结果揭示了BMI,睡眠和精神疾病之间的非线性遗传联系,挑战了遗传流行病学的传统假设.
科学领域:
- 遗传学 遗传学 是一个
- 生物统计学 生物统计学
- 精神病学流行病学 精神病学流行病学
背景情况:
- 传统的遗传流行病学通常假定特征之间的线性关系.
- 像遗传相关性这样的全球估计器可能会掩盖复杂的,非线性遗传关联.
- 了解非线性遗传关系对于有针对性的干预至关重要.
研究的目的:
- 引入一种新的统计方法来推断非线性双变基因关系.
- 研究身体质量指数 (BMI),睡眠时间,身高和精神疾病 (ADHD,神经性厌食症,抑郁症) 之间的遗传关系.
- 挑战遗传流行病学的线性假设.
主要方法:
- 开发了三角统计统计方法来推断非线性双变遗传关系.
- 在特征分布中利用细分的全基因组关联研究 (GWAS).
- 分析了特征细分的GWAS与第二个特征之间的遗传相关性.
- 将该方法应用于英国生物银行数据 (约. 450K个体). 这是一个很好的例子.
主要成果:
- 在特定假设下成功检索了非线性遗传关系的形状.
- 确定了BMI和抑郁症,BMI和厌食症,睡眠时间和抑郁症,睡眠时间和ADHD之间的非线性遗传关系.
- 在身高和精神特征之间的遗传关系中没有发现显著的非线性.
- 证明全球遗传估计器不足以捕捉潜在的复杂性.
结论:
- 使用新的统计方法,可以检测特征之间的非线性遗传关系.
- 在遗传流行病学的线性假设受到挑战.
- 两变的遗传关联在表型谱中各不相同,影响干预开发.
相关概念视频
 02:54
Evolutionary Relationships through Genome Comparisons
5.7K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
5.7K
 01:11
Genome-wide Association Studies-GWAS
12.3K
Genome-wide association studies or GWAS are used to identify whether common SNPs are associated with certain diseases. Suppose specific SNPs are more frequently observed in individuals with a particular disease than those without the disease. In that case, those SNPs are said to be associated with the disease. Chi-square analysis is performed to check the probability of the allele likely to be associated with the disease.
GWAS does not require the identification of the target gene involved in...
GWAS does not require the identification of the target gene involved in...
12.3K
 02:34
DNA Microarrays
17.1K
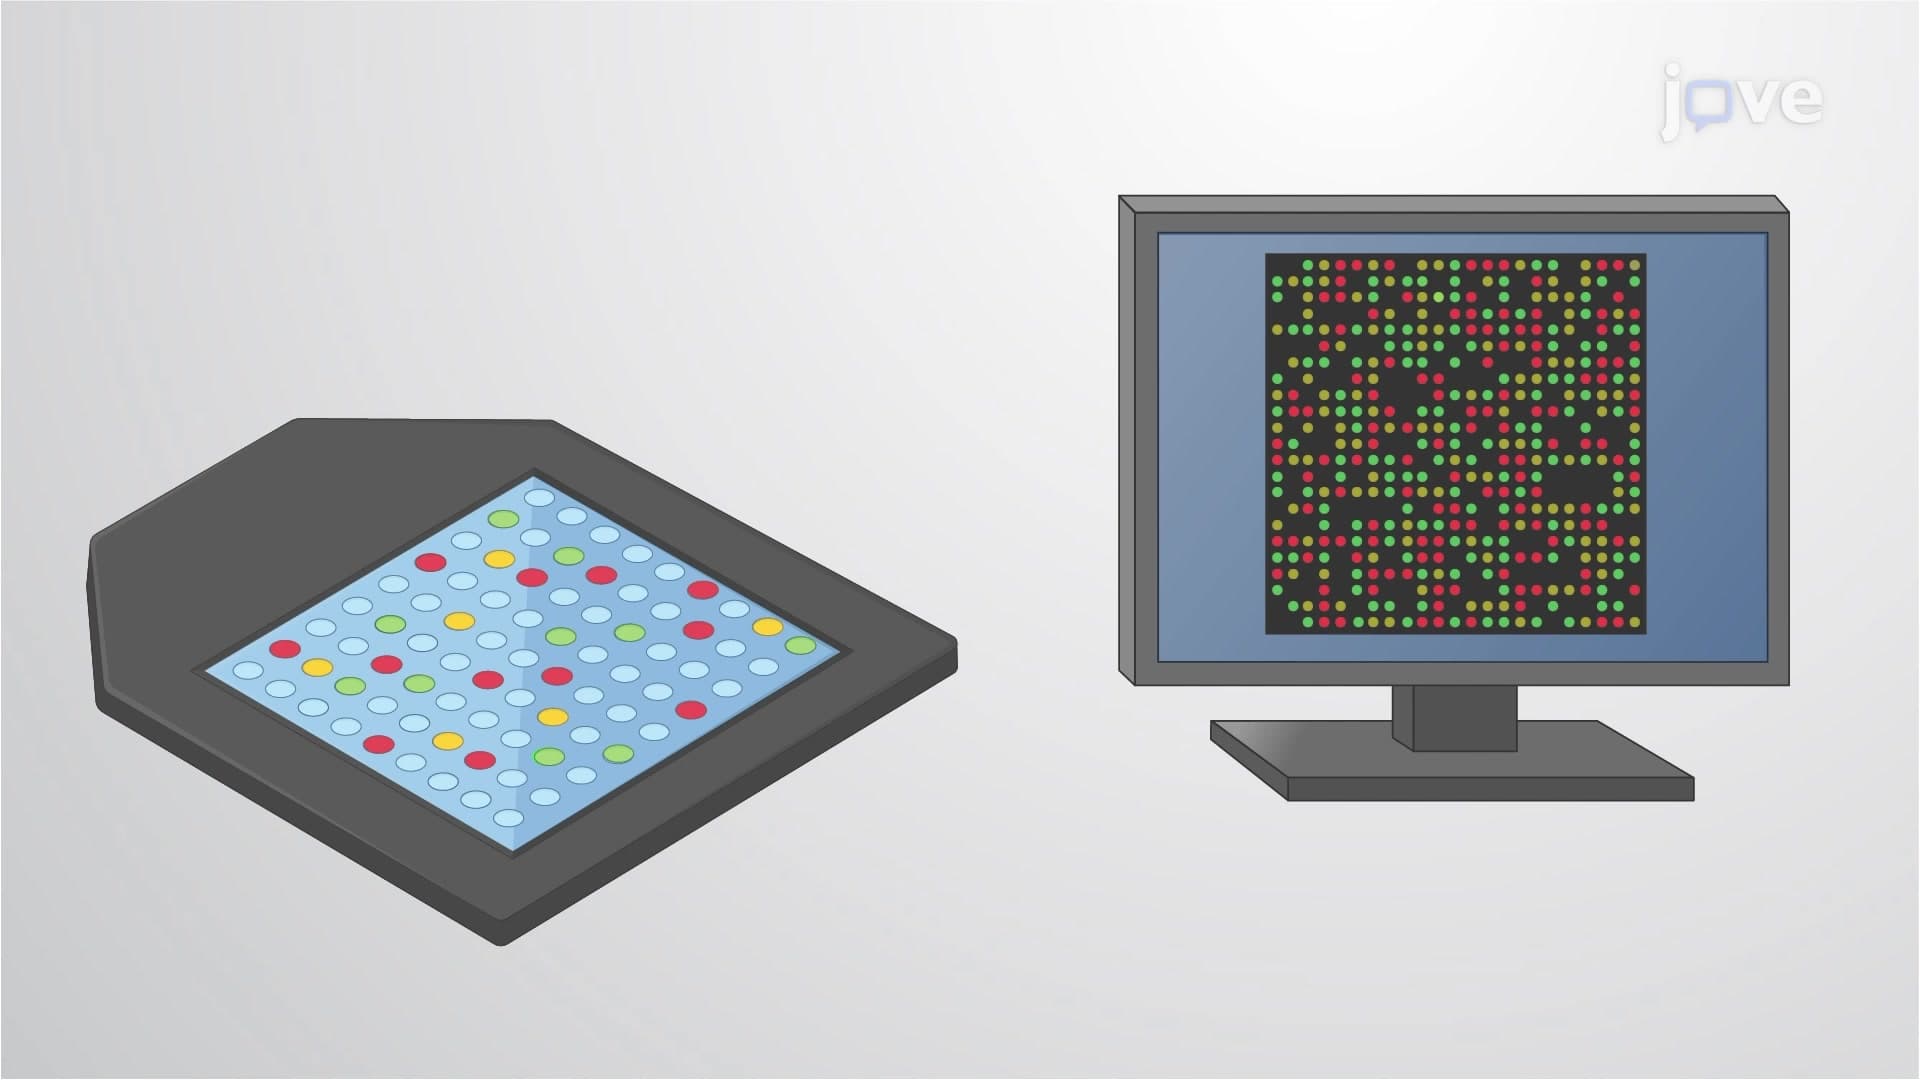
Microarrays are high-throughput and relatively inexpensive assays that can be automated to analyze large quantities of data at a time. They are used in genome-wide studies to compare gene or protein expression under two varied conditions, such as healthy and diseased states. Microarrays consist of glass or silica slides on which probe molecules are covalently attached through surface functionalization. Most commonly, the slides are prepared through the chemisorption of silanes to silica...
17.1K
 02:05
Gene Evolution - Fast or Slow?
7.0K
The genomes of eukaryotes are punctuated by long stretches of sequence which do not code for proteins or RNAs. Although some of these regions do contain crucial regulatory sequences, the vast majority of this DNA serves no known function. Typically, these regions of the genome are the ones in which the fastest change, in evolutionary terms, is observed, because there is typically little to no selection pressure acting on these regions to preserve their sequences.
In contrast, regions which code...
In contrast, regions which code...
7.0K
 02:26
Comparing Copy Number Variations and SNPs
17.0K
Sequencing of the human genome has opened up several best-kept secrets of the genome. Scientists have identified thousands of genome variations that exist within a population. These variations can be a single nucleotide or a larger chromosomal variation.
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
17.0K