
使用PaSiMap在Jalview中的序列聚类协议
Thomas Morell1, James Procter2, Geoffrey J Barton2
1Department of Biology, University of Konstanz, 78457 Konstanz, Germany.
STAR protocols
|February 28, 2025

概括
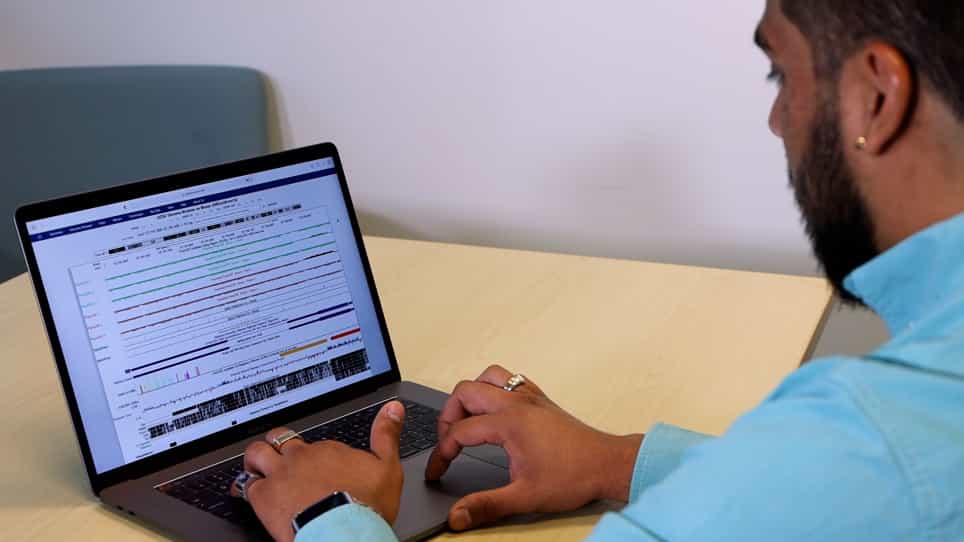
在 Jalview 中,双向相似度映射 (PaSiMap) 提供了一种新的蛋白质序列分析方法,不需要多个序列对齐. 本协议详细介绍了PaSiMap,用于有效的序列聚类和可视化.
科学领域:
- 生物信息学是一种生物信息学.
- 计算生物学 计算生物学
- 结构生物信息学 结构生物信息学
背景情况:
- 主要成分分析 (PCA) 常用于分析蛋白质序列关系.
- PCA通常需要预先计算的多个序列对齐,这可能是一个限制.
- 在序列数据中区分系统和随机差异对于准确的分析至关重要.
研究的目的:
- 在 Jalview.com 中使用对对相似度映射 (PaSiMap) 进行序列聚类的详细协议.
- 为分析蛋白质序列关系提供一种用户友好的方法.
- 为PCA提供一种不需要多个序列对齐的替代方案.
主要方法:
- 在 Jalview 软件平台中实现 PaSiMap.
- 一步一步的指南涵盖安装,序列数据导入和PaSiMap分析执行.
- 使用RStudio进行下游数据可视化和PaSiMap结果的解释.
主要成果:
- PaSiMap有效地区分了蛋白质序列数据集中的系统变化和随机变化.
- 该协议能够简化对蛋白质序列关系的可视化和分析.
- 成功应用PaSiMap用于序列聚类和关系映射.
结论:
- 在 Jalview 中的 PaSiMap 为蛋白质序列分析提供了一个强大且易于使用的工具.
- 该协议促进了蛋白质序列数据的有效聚类和解释.
- 该方法为研究蛋白质序列演变和功能的研究人员提供了有价值的替代方案.
相关概念视频
 01:05
Maxam-Gilbert Sequencing
10.9K
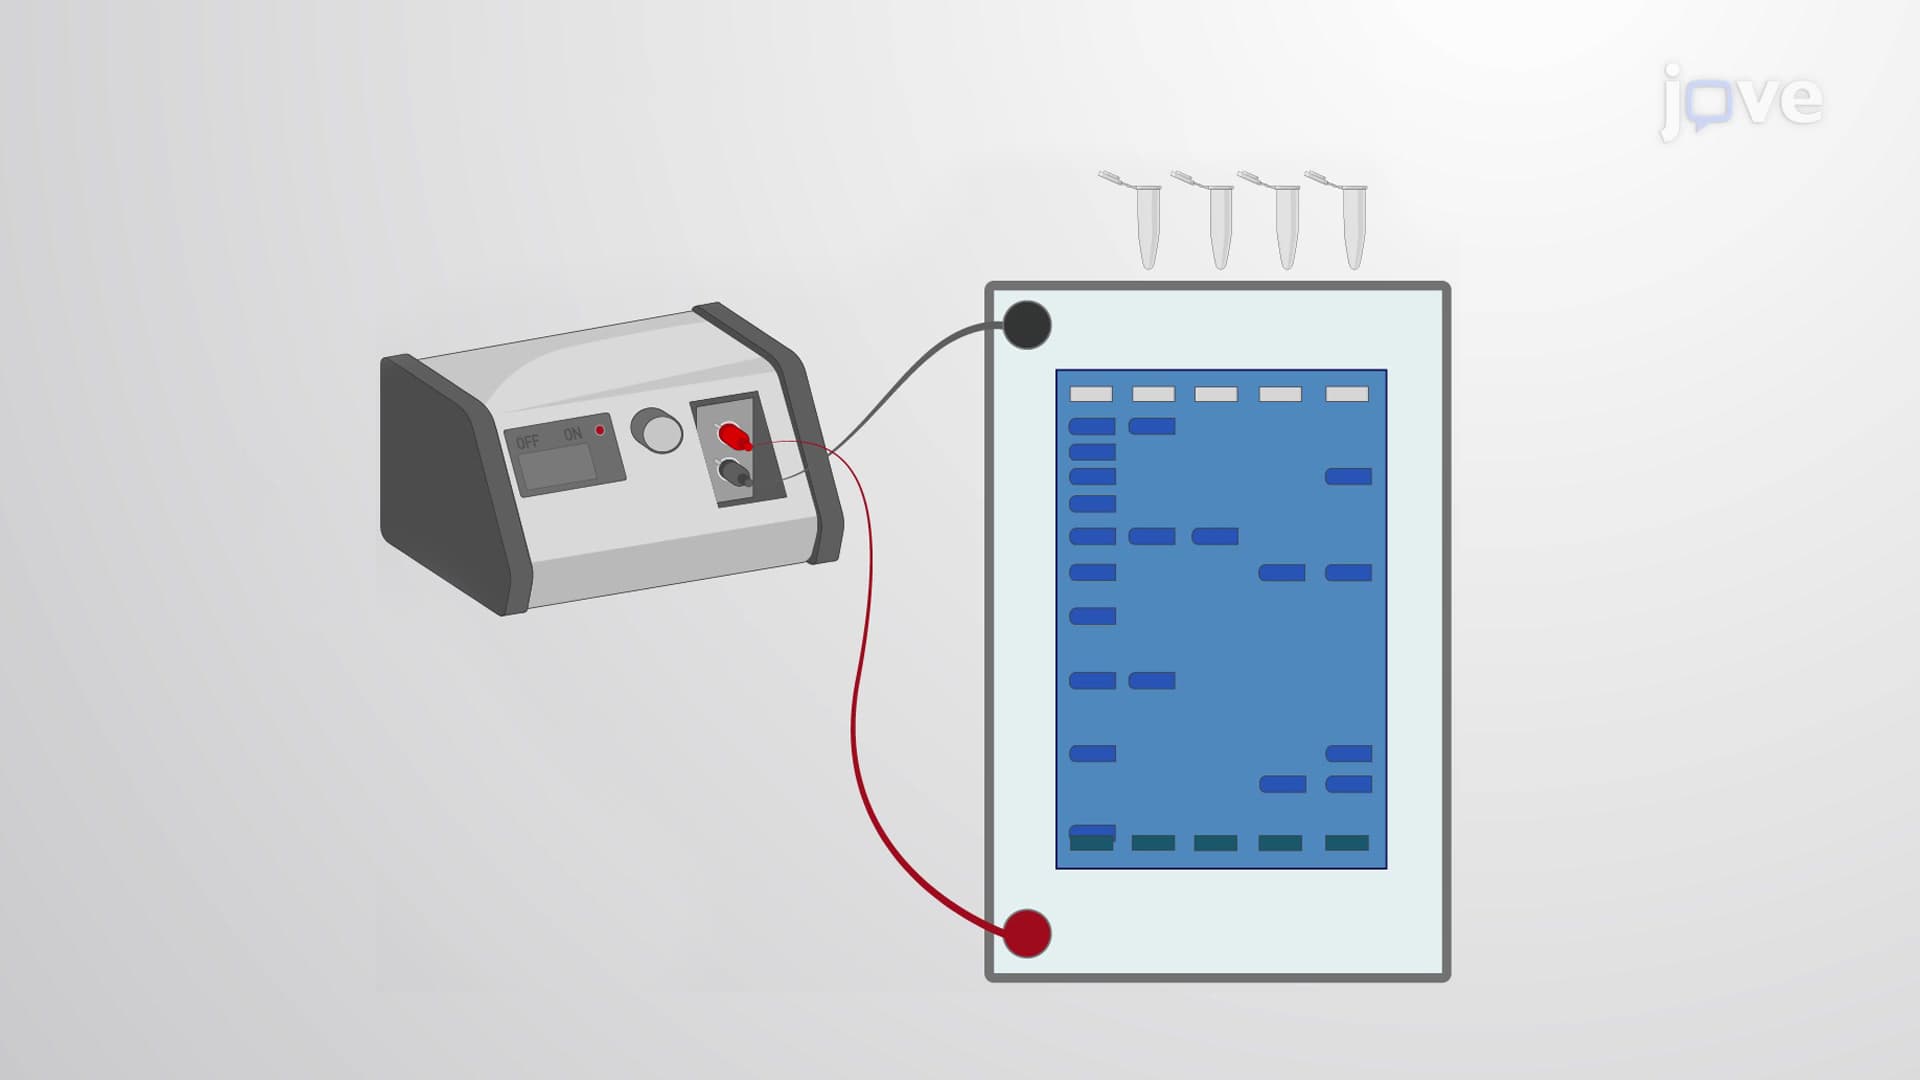
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
10.9K
 02:51
Multi-species Conserved Sequences
3.9K
Next-generation sequencing technologies have created large genomic databases of a variety of animals and plants. Ever since the human genome project was completed, scientists studied the genome of primates, mammals, and other phylogenetically distant living beings. Such large-scale studies have provided new insights into the evolutionary relationship between organisms.
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
3.9K
 01:57
Sanger Sequencing
752.2K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
752.2K
 01:26
Per-Unit Sequence Models
67
An ideal Y-Y transformer, grounded through neutral impedances, displays per-unit sequence networks akin to those of a single-phase ideal transformer when subjected to balanced positive- or negative-sequence currents. These currents do not produce neutral currents, and their associated voltage drops.
Zero-sequence currents, which are identical in magnitude and phase, generate a neutral current, resulting in voltage drops across the neutral impedance and the low-voltage winding. If the...
Zero-sequence currents, which are identical in magnitude and phase, generate a neutral current, resulting in voltage drops across the neutral impedance and the low-voltage winding. If the...
67
 01:20
Cluster Sampling Method
11.6K
Appropriate sampling methods ensure that samples are drawn without bias and accurately represent the population. Because measuring the entire population in a study is not practical, researchers use samples to represent the population of interest.
To choose a cluster sample, divide the population into clusters (groups) and then randomly select some of the clusters. All the members from these clusters are in the cluster sample. For example, if you randomly sample four departments from your...
To choose a cluster sample, divide the population into clusters (groups) and then randomly select some of the clusters. All the members from these clusters are in the cluster sample. For example, if you randomly sample four departments from your...
11.6K
 03:21
RNA-seq
9.8K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.8K