
通过完善的全长 ABO 哈普洛型测序,全面说明完整的 ABO 基因并解析 ABO 变异
Yanling Ying1, Jingjing Zhang1, Xiaozhen Hong1
1Blood Center of Zhejiang Province, Blood Transfusion Medicine Research Institute, Hangzhou, Zhejiang, People's Republic of China.
Clinical chemistry
|March 6, 2025

概括
全长度的 ABO 哈普洛型测序现在涵盖了整个基因,从5'到3'未翻译区域 (UTR). 这一进步改善了血液组的基因定型和对ABO基因变异的分析.
科学领域:
- 遗传学 遗传学 是一个
- 基因组学就是基因组学.
- 分子生物学分子生物学
背景情况:
- 准确的 ABO 基因基因型和注释对于理解其变体至关重要.
- 现有的方法对于全长的 ABO 单元类型缺乏全面的注释,特别是 5' 和 3' 未翻译区域 (UTR).
研究的目的:
- 开发和验证一种用于全长ABO单双型测序的方法.
- 为 ABO 杂型提供全面的注释,包括 3' UTR.
主要方法:
- 使用改善的一步超长距离PCR与抑制原始体来放大全长ABO基因 (5' UTR到3' UTR).
- 执行了单分子实时 (SMRT) 库构建和长读序列.
- 进行了全面的数据分析,包括基础调用,对齐,变体调用,聚类和注释.
主要成果:
- 产生了26.1kb的安普利康,代表了迄今为止最完整的ABO基因序列.
- 在中国人群中获得了完整的 ABO 哈普洛型序列和分类的 ABO 序列模式.
- 成功解决了具有结构变异 (SVs) 的 ABO 变异,例如删除,反转和模拟.
结论:
- 全长度的 ABO 哈普洛型测序成功地解决了 3' UTR 序列注释中的缺口.
- 这项技术推进了血液组基因组分析,ABO基因功能研究,进化研究和ABO变异的分辨率.
相关概念视频
 03:36
Genome Annotation and Assembly
18.8K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
18.8K
 01:11
Genome-wide Association Studies-GWAS
12.3K
Genome-wide association studies or GWAS are used to identify whether common SNPs are associated with certain diseases. Suppose specific SNPs are more frequently observed in individuals with a particular disease than those without the disease. In that case, those SNPs are said to be associated with the disease. Chi-square analysis is performed to check the probability of the allele likely to be associated with the disease.
GWAS does not require the identification of the target gene involved in...
GWAS does not require the identification of the target gene involved in...
12.3K
 03:21
RNA-seq
9.8K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.8K
 02:20
Blood Types
17.3K
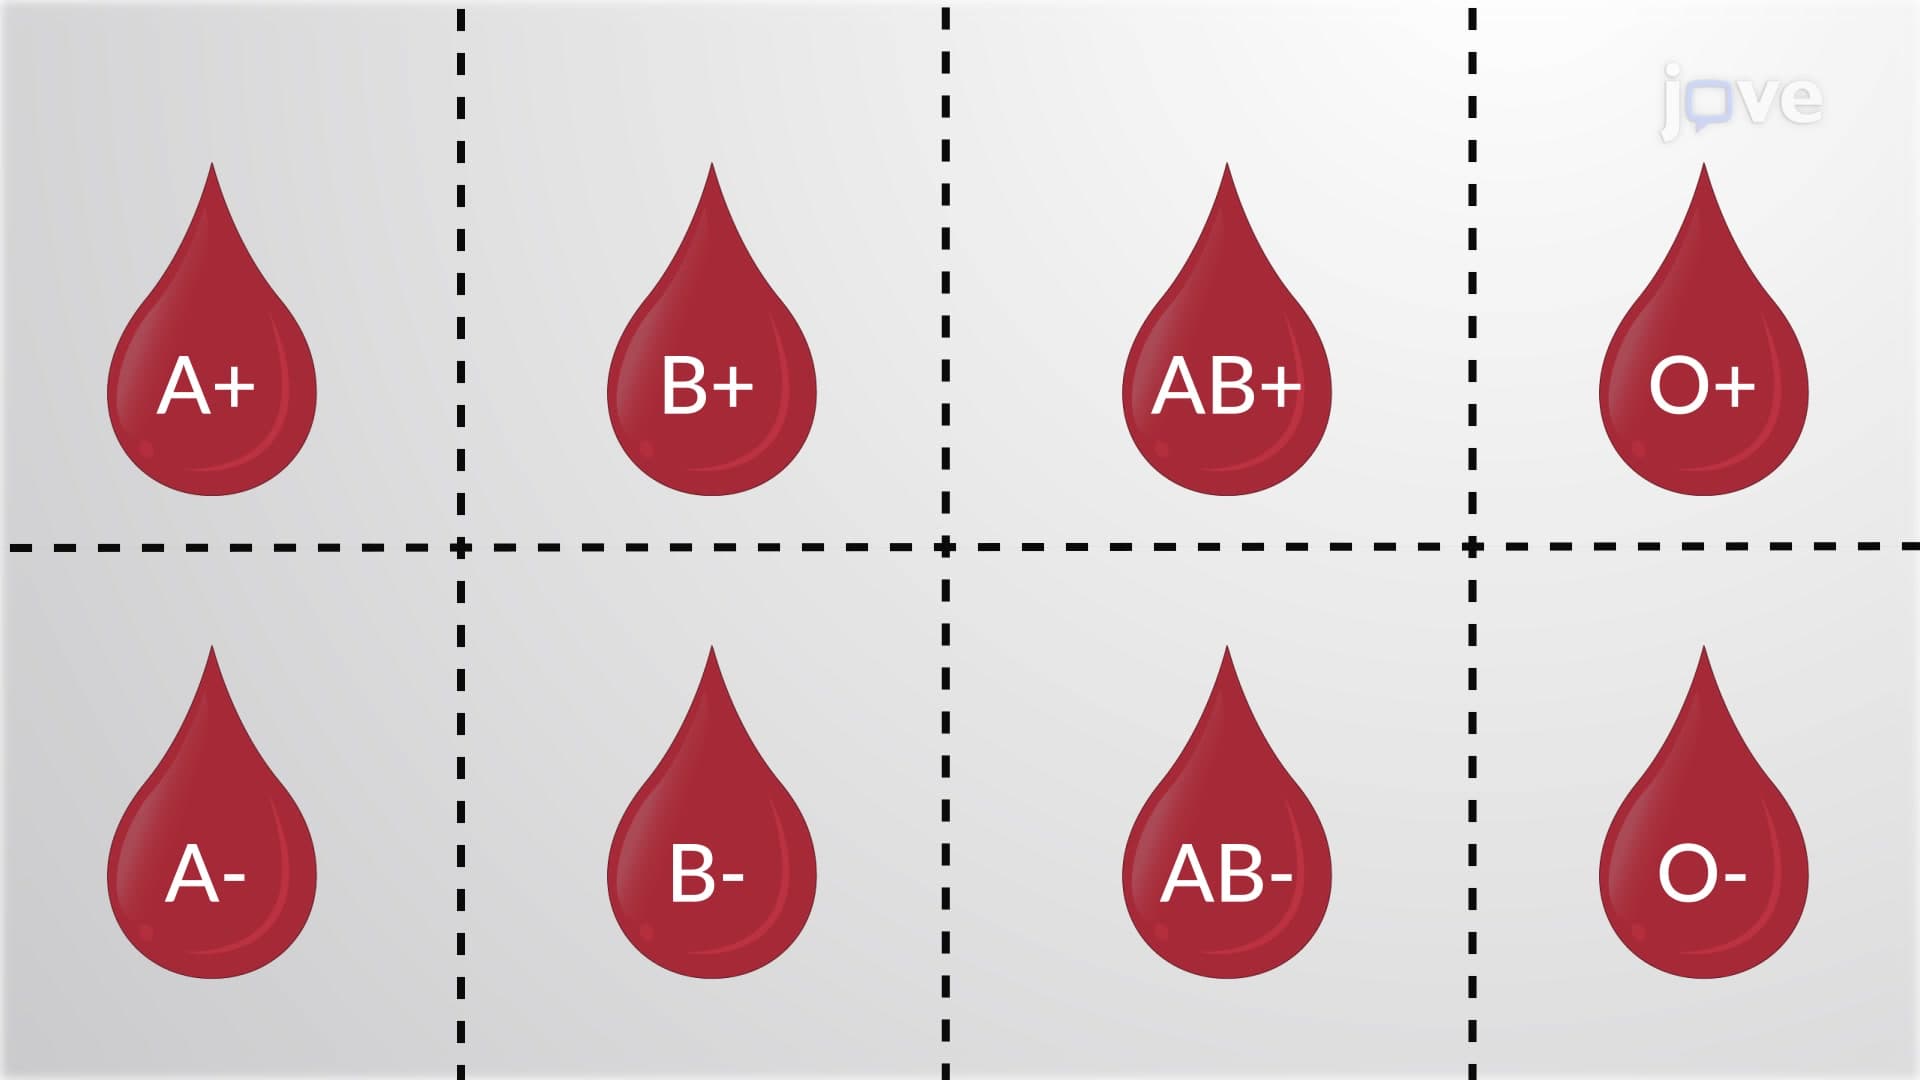
Human blood is classified into different types based on the presence of antigens on the red blood cell's surface and antibodies in the plasma. Proper identification of blood type is essential for successful blood transfusion. The International Society of Blood Transfusion has identified 38 human blood types based on the surface antigens on the red blood cells. The most common types are ABO, Rh, and MNS blood types.
ABO blood group
ABO antigens are glycoproteins encoded by genes present on...
ABO blood group
ABO antigens are glycoproteins encoded by genes present on...
17.3K
 03:00
Next-generation Sequencing
86.8K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
86.8K
 01:57
Sanger Sequencing
752.1K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
752.1K