相关实验视频
Updated: May 20, 2025
10:34
Ultra-long Read Sequencing for Whole Genomic DNA Analysis
Published on: March 15, 2019
22.6K
全基因组测序Flammulina filiformis和多奥米克分析,以应对低温
Xinmin Liang1, Jing Han1, Yuqin Cui2
1Sichuan Institute of Edible Fungi, Sichuan Academy of Agricultural Sciences, Chengdu 610066, China.
Journal of fungi (Basel, Switzerland)
|March 26, 2025

概括
状虫需要低温生长. 这项研究揭示了特定的碳水化合物活性酶 (CAZyme) 基因对于F. filiformis mycelia的寒冷适应至关重要.
科学领域:
- 菌类学 菌类学是指菌类学.
- 基因组学就是基因组学.
- 分子生物学分子生物学
背景情况:
- Flammulina filiformis的生长和原始形成是由低温的线索触发的.
- 了解寒冷应激反应的分子基础对于培养F. filiformis至关重要.
研究的目的:
- 阐明F. filiformis对寒冷压力的反应背后的分子机制.
- 为了确定关键的基因和途径参与低温适应.
主要方法:
- 使用PacBio HiFi和Hi-C.进行F. filiformis的新基因组测序.
- 多omics分析包括转录基因组和代谢基因组在不同温度下的菌体,原始体和果实体的概况.
- 识别和表达分析冷反应 (COR) 基因和碳水化合物活性酶 (CAZymes).
主要成果:
- 一个高质量的基因组序列 36.3 Mb 跨 12 个染色体与 11,886 个蛋白质编码基因组装.
- 确定了25个冷反应基因 (COR) 和520个适应寒冷的酶.
- 在低温 (10°C) 时丰富细胞中的碳水化合物代谢通路,515个CAZyme基因中,有58个基因被特别上调.
结论:
- 碳水化合物活性酶 (CAZyme) 基因在F. filiformis.低温适应方面发挥着重要作用.
- 在寒冷条件下,细胞中特定的CAZyme基因的升级是关键的反应机制.
- 这些发现支持针对F. filiformis改善的有针对性的育种策略.
相关概念视频
 02:02
Genomics
35.4K
Genomics is the science of genomes: it is the study of all the genetic material of an organism. In humans, the genome consists of information carried in 23 pairs of chromosomes in the nucleus, as well as mitochondrial DNA. In genomics, both coding and non-coding DNA is sequenced and analyzed. Genomics allows a better understanding of all living things, their evolution, and their diversity. It has a myriad of uses: for example, to build phylogenetic trees, to improve productivity and...
35.4K
 03:00
Next-generation Sequencing
86.6K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
86.6K
 01:57
Sanger Sequencing
751.9K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
751.9K
 03:21
RNA-seq
9.7K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.7K
 01:05
Maxam-Gilbert Sequencing
10.8K
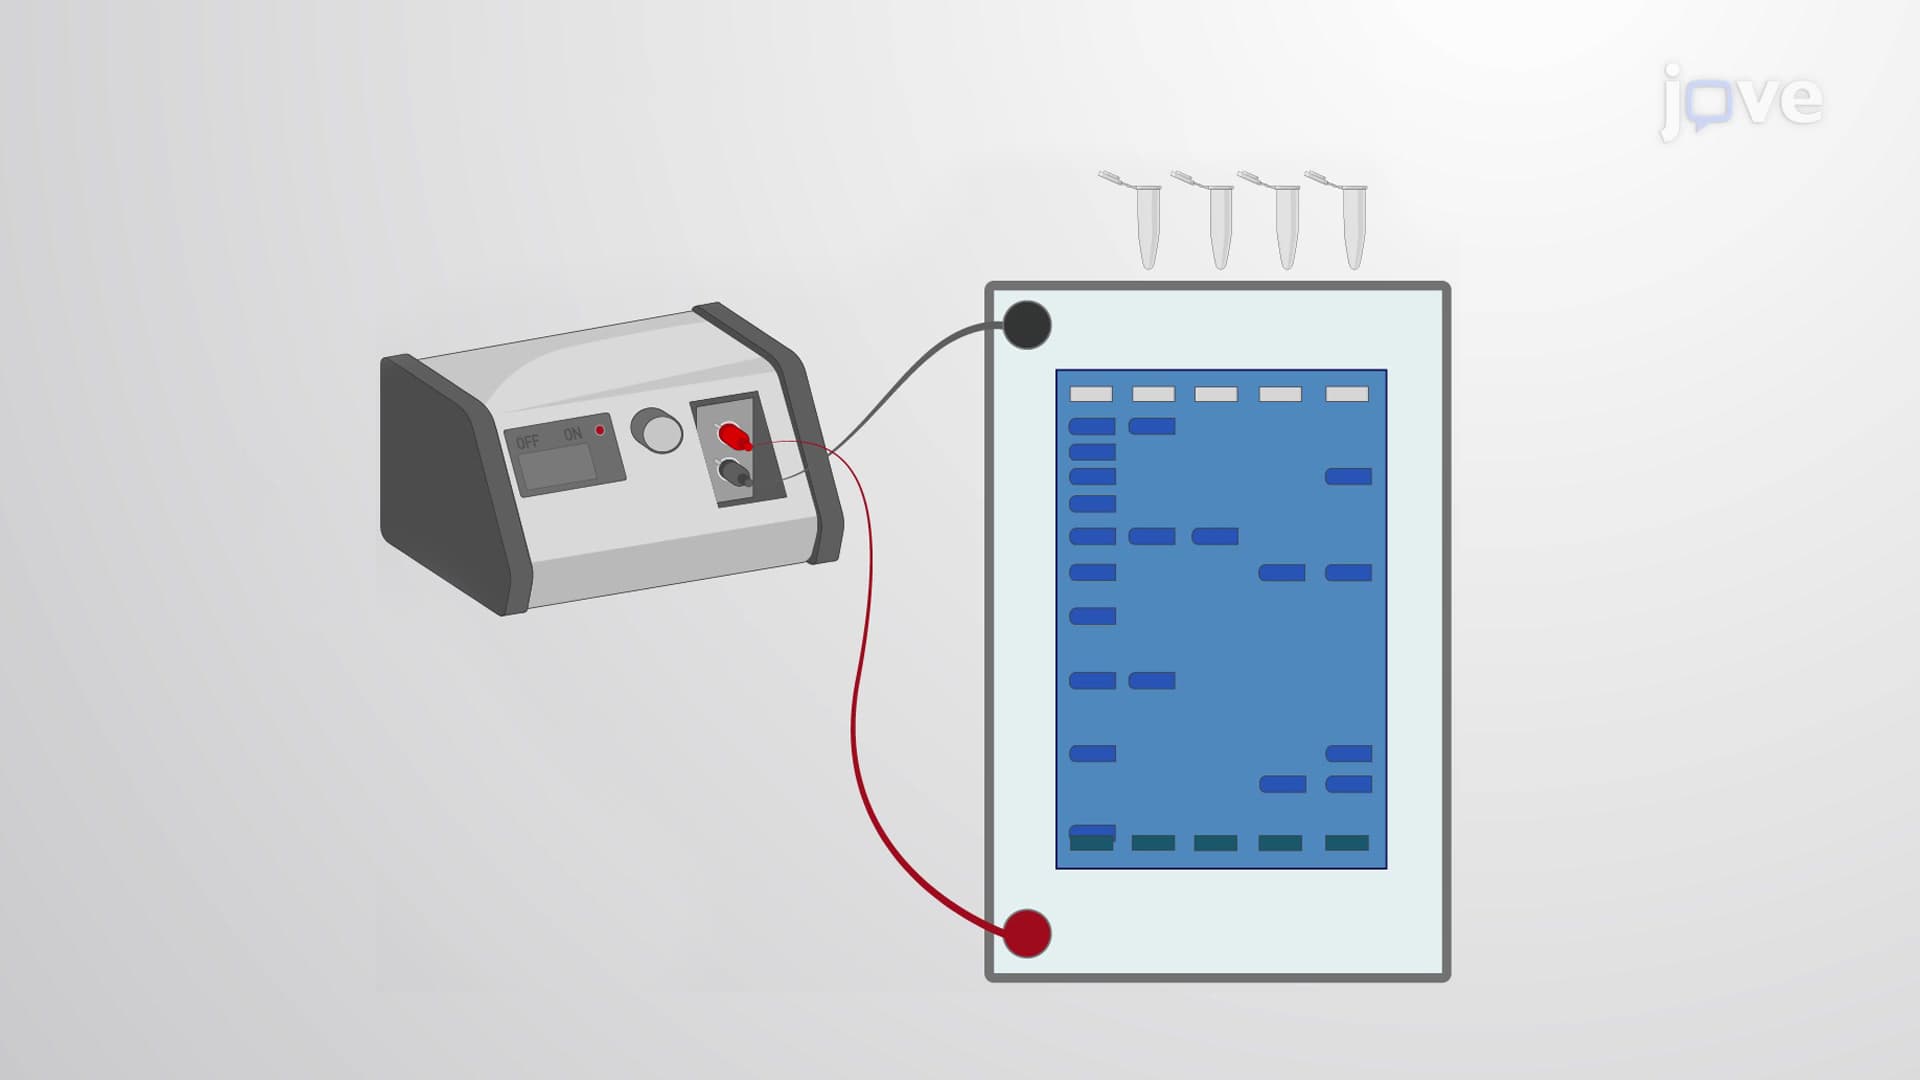
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
10.8K
 03:36
Genome Annotation and Assembly
18.7K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
18.7K