突发序列中的特定核酸替代增强了alphabaculoviruses中的多面体表达:改善了baculovirus表达载体
Susumu Katsuma1, Koshi Fukaura1, Noriko Matsuda-Imai1
1Department of Agricultural and Environmental Biology, Graduate School of Agricultural and Life Sciences, The University of Tokyo, Tokyo, Japan.
Applied and environmental microbiology
|April 7, 2025

概括
在baculovirus多面体 (POLH) 基因中的特定突变.
科学领域:
- 分子生物学分子生物学
- 病毒学 病毒学
- 生物技术是生物技术.
背景情况:
- 众所周知,αbaculoviruses产生了许多主要由多面体 (POLH) 组成的闭塞体 (OB).
- 波尔赫基因的高水平转录对OB产生至关重要,并且已被用于baculovirus表达载体系统 (BEVS).
- 波尔赫基因内的突破序列对于其超转录至关重要,但其精确的调节元素尚未完全理解.
研究的目的:
- 为了研究在POLH基因爆发序列中的A丰富区域的作用.
- 通过使用BEVS.来识别可以增强POLH表达的特定突变,以改善外来蛋白质的产生.
主要方法:
- 在爆发序列的A丰富区域中产生具有A-到-T替代的Bombyx mori核多重体病毒 (BmNPV) 突变.
- 对突变病毒中POLH促销器驱动的记者基因表达的分析.
- 在Autographa californica多重核聚合体病毒 (AcMNPV) 载体系统中对突变的评估.
- 在受感染细胞中量化POLH mRNA和蛋白质水平.
主要成果:
- 在POLH上游区域的位置 -16到 -14的三重A到T突变 (TTT) 显著增加了记者基因表达的4到5倍.
- 在这些位置的单个和双个A-to-T突变显示出较小但显著的影响.
- 在BmNPV和AcMNPV系统中,TTT突变增强了POLH表达.
- 这种突变还增加了OB产生的病毒中POLH mRNA和蛋白质的积累.
结论:
- 在baculovirus爆发序列中的特定核酸替代可以显著增强POLH基因转录.
- 在POLH上游区域的 -16至 -14位置的三重TTT突变特别有效地促进表达.
- 这些发现提供了一种简单的方法来改善baculovirus表达载体系统中的外来蛋白质产量.
相关概念视频
 01:35
Mutations
32.9K
Mutations are changes in the sequence of DNA. These changes can occur spontaneously or they can be induced by exposure to environmental factors. Mutations can be characterized in a number of different ways: whether and how they alter the amino acid sequence of the protein, whether they occur over a small or large area of DNA, and whether they occur in somatic cells or germline cells.
Chromosomal Alterations Are Large-Scale Mutations
While point mutations are changes in a single nucleotide in...
Chromosomal Alterations Are Large-Scale Mutations
While point mutations are changes in a single nucleotide in...
32.9K
 01:02
Long-patch Base Excision Repair
6.9K
Since the discovery of the two BER pathways, there has been a debate about how a cell chooses one pathway over the other and the factors determining this selection. Numerous in vitro experiments have pointed out multiple determinants for the sub-pathway selection. These are:
6.9K
 02:28
Leaky Scanning
5.1K
During most eukaryotic translation processes, the small 40S ribosome subunit scans an mRNA from its 5' end until it encounters the first start AUG codon. The large 60S ribosomal subunit then joins the smaller one to initiate protein synthesis. The location of the translation initiation is largely determined by the nucleotides near the start codon as there may be multiple translation initiation sites present on the mRNA. Marilyn Kozak discovered that the sequence RCCAUGG (where R...
5.1K
 02:46
Genome Copying Errors
4.1K
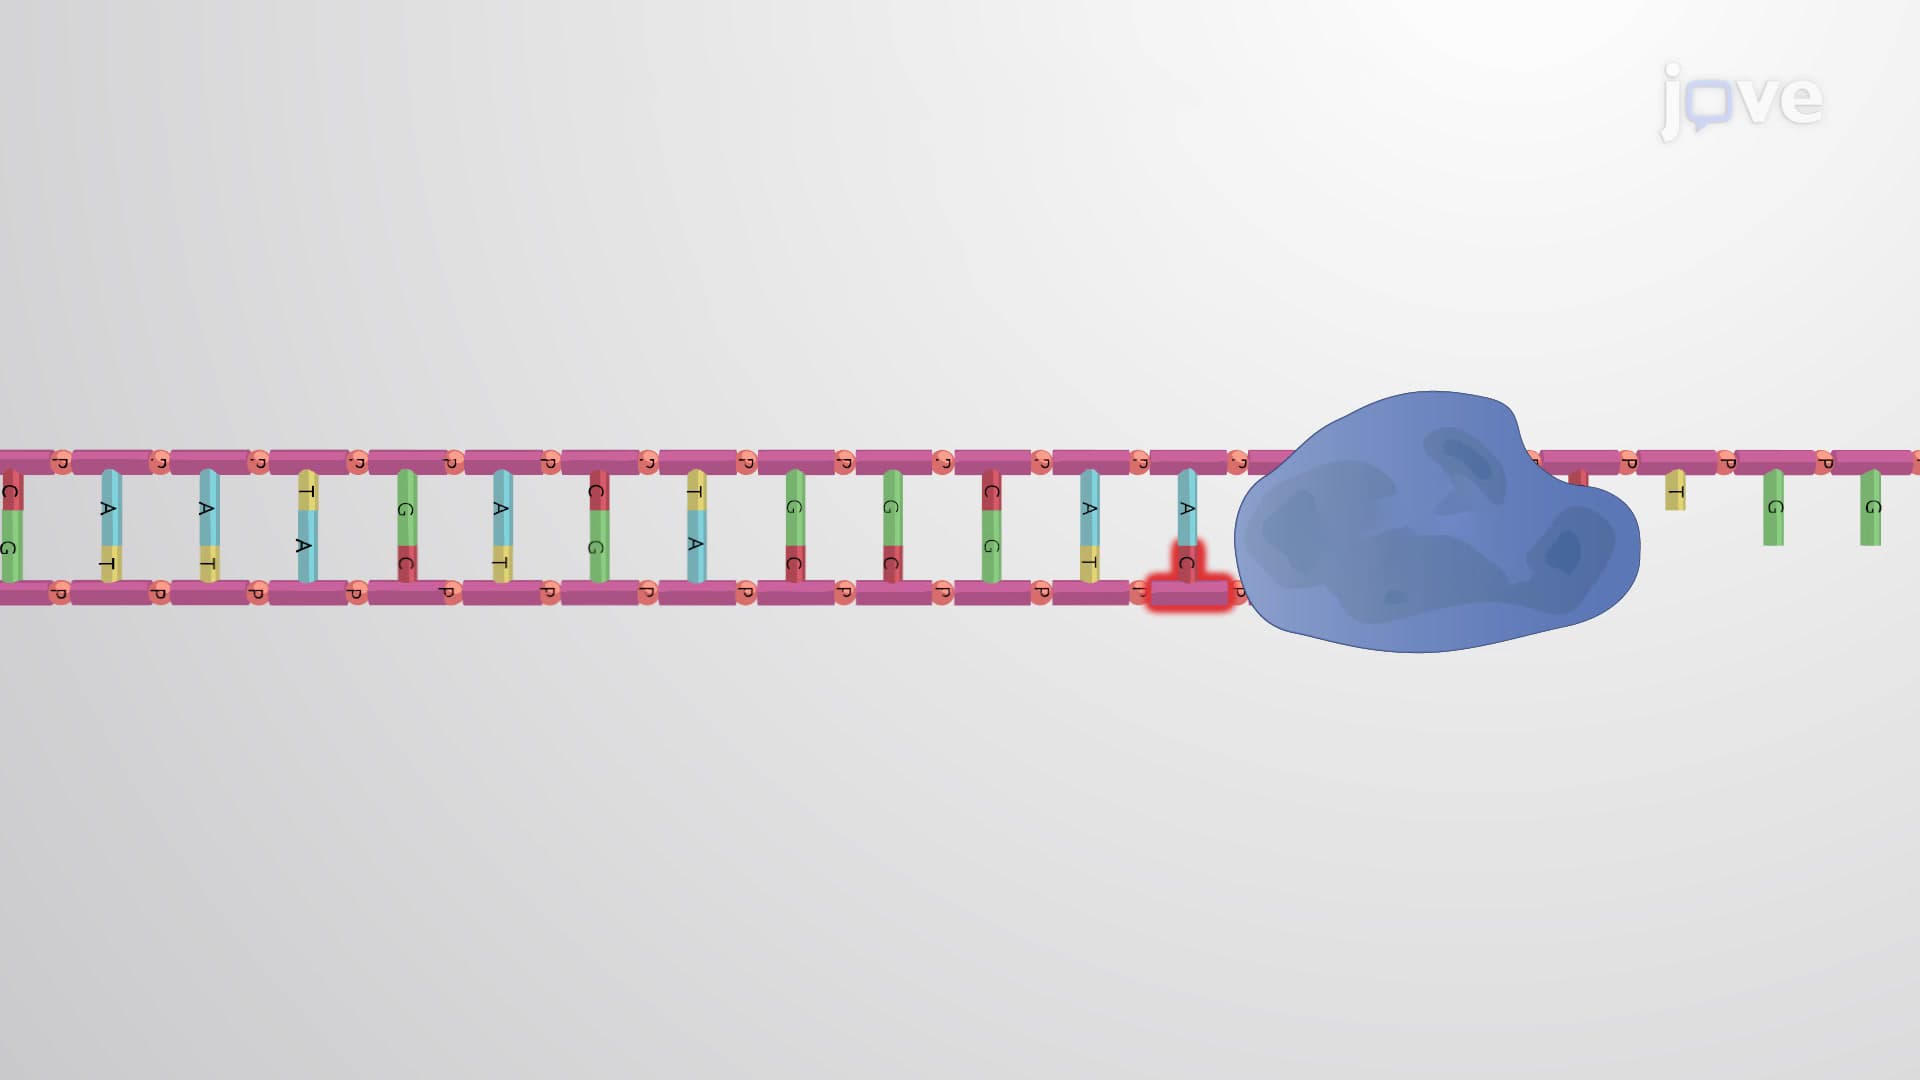
DNA replication is a well-evolved process that copies millions of base pairs with high fidelity during each cell division. Occasionally a wrong base or a long stretch of wrong bases may get added to the daughter strands. If the errors are left unchecked, cells might accumulate several mutations that might endanger their survival. Therefore, the copying errors are checked and repaired at three levels.
4.1K
 01:05
Maxam-Gilbert Sequencing
10.8K
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
10.8K
 03:00
Next-generation Sequencing
86.8K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
86.8K