JUNIPER:从规模上的下一代测序数据中重建传输事件
Ivan Specht1, Gage K Moreno1, Taylor Brock-Fisher1,2
1The Broad Institute of MIT and Harvard, Cambridge, MA 02142, USA.
Research square
|April 8, 2025

概括
我们开发了JUNIPER,这是一个可扩展的工具,通过分析病原体遗传数据来重建疾病传播网络. 朱尼珀提高了对病原体传播的理解,并有助于在疫情期间进行有针对性的控制.
科学领域:
- 流行病学 流行病学
- 计算生物学 计算生物学
- 基因组学就是基因组学.
背景情况:
- 了解病原体传播动态对于有效的疾病控制至关重要.
- 传输重建的现有工具面临着可扩展性和方法上的局限性.
研究的目的:
- 开发一个高度可扩展的工具,JUNIPER (植物遗传学和流行病学重建的联合底层网络推断),用于重建病原体传播网络.
- 将宿主内变异和不完整的抽样纳入爆发重建模型.
主要方法:
- 使用下一代测序数据开发了主机内部变异频率的统计模型.
- 集成宿主内部变异与人口水平的进化和传播模型.
- 能够同时推断族系和传输树,并实现算法并行化以实现可扩展性.
主要成果:
- 在超过16万个SARS-CoV-2基因组上验证了宿主内部变异模型.
- 证明了JUNIPER在牛H5N1 (1500多例) 和人类COVID-19 (13000多例) 大规模数据集上的实用性.
- 在加利福尼亚州量化了H5N1传染率的升高,并确定了高可信度传染事件;显示了疫苗接种在减少SARS-CoV-2传染方面的有效性.
结论:
- JUNIPER克服了现有工具的计算和方法限制.
- 为大规模的病原体传播研究提供了强大的框架.
- 通过提供对疾病爆发的关键见解,促进有针对性的控制措施.
更多相关视频
 10:41
Leveraging CyVerse Resources for De Novo Comparative Transcriptomics of Underserved Non-model Organisms
Published on: May 9, 2017
9.2K
 06:59
Using Next Generation Sequencing to Identify Mutations Associated with Repair of a CAS9-induced Double Strand Break Near the CD4 Promoter
Published on: March 31, 2022
2.4K
相关概念视频
 03:00
Next-generation Sequencing
86.3K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
86.3K
 01:57
Sanger Sequencing
751.4K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
751.4K
 03:36
Genome Annotation and Assembly
18.7K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
18.7K
 01:05
Maxam-Gilbert Sequencing
10.6K
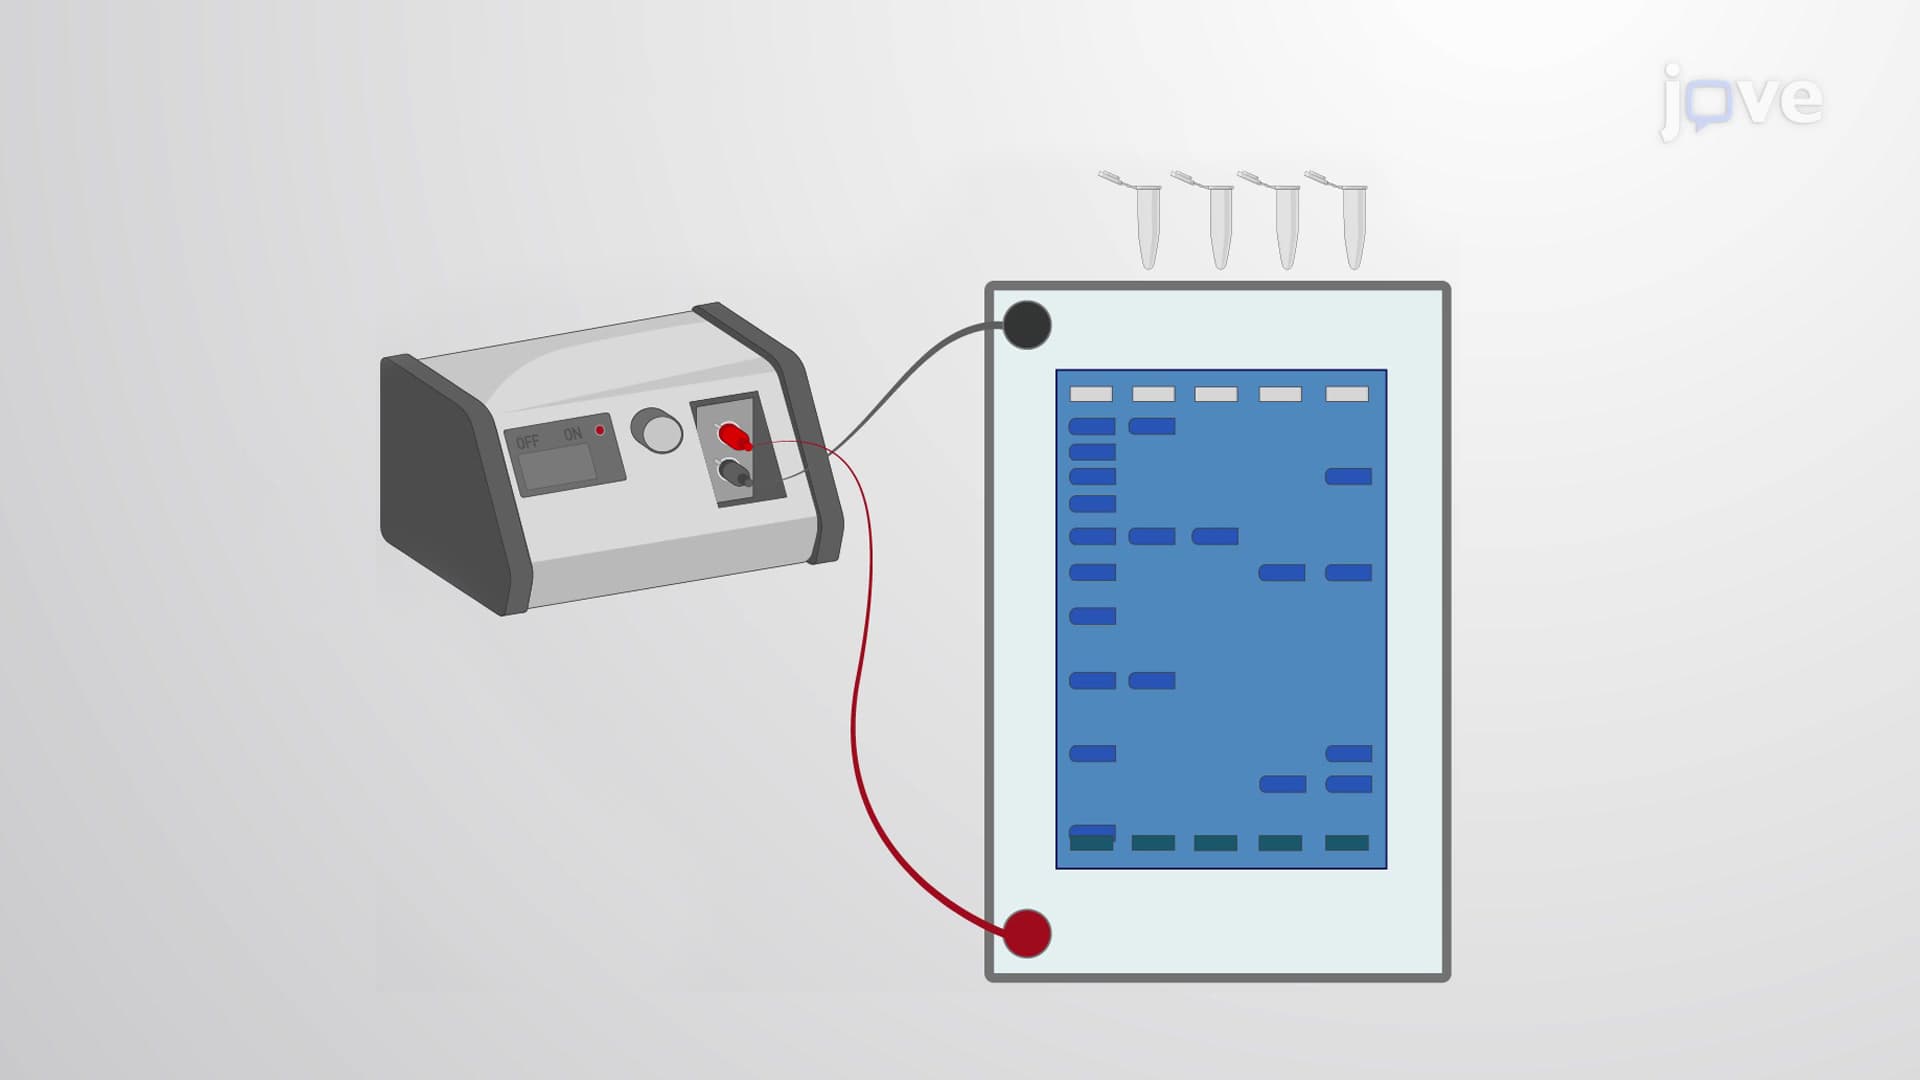
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
10.6K
 02:54
Evolutionary Relationships through Genome Comparisons
5.6K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
5.6K
 03:21
RNA-seq
9.7K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.7K