构建一个全面的图书馆的重复序列,以注释的基因组
Delphine Giraud1, Nathalie Choisne2, Marilyne Summo3,4
1UR AGAP Corse, INRAE, Institut Agro, CIRAD, University of Montpellier, San Giuliano, F-20230, France. delphine.giraud79@gmail.com.
BMC genomic data
|April 17, 2025

概括
这项研究使用多种生物信息学工具全面注释类基因组中的重复序列. 开发的参考库有助于理解类物种的基因组进化和基因调节.
科学领域:
- 基因组学就是基因组学.
- 生物信息学是一种生物信息学.
- 植物科学 植物科学
背景情况:
- 对重复序列的全面注释对于理解基因组动态和基因调节至关重要.
- 目前的类基因组注释是不完整的,因为它依赖于有限的生物信息学工具,专注于可转换的元素.
研究的目的:
- 为类基因组中的所有重复序列开发一个全面的注释方法.
- 为类和相关物种创建一个重复序列的参考库.
- 促进进一步研究重复元素在类植物进化和基因调节中的作用.
主要方法:
- 综合补充的重复寻找程序 (REPET,CAULIFINDER,TAREAN) 进行.
- 开发了一种细粒度注释方法,用于祖先的类物种 (C. medica,C. micrantha,C. reticulata,C. maxima).
- 开发了一种更快的注释方法,以丰富其他类和Aurantioideae物种的数据集.
主要成果:
- 创建了3091个共识序列的参考库,其中94.5%是可转移的元素.
- 首次 (160个序列) 描述了类植物内源性calimovirids的多样性.
- 识别了10颗卫星的DNA.
结论:
- 多重重复检测方法可以实现全面的果基因组注释.
- 该参考库改善了在属物种化和品种形成期间对重复序列动态的理解.
- 该 RepeatLoc Citrus 网页界面有助于研究重复元素在基因组结构和基因调节中的作用.
更多相关视频
 13:03
Robust DNA Isolation and High-throughput Sequencing Library Construction for Herbarium Specimens
Published on: March 8, 2018
10.5K
 10:40
Comprehensive Workflow for the Genome-wide Identification and Expression Meta-analysis of the ATL E3 Ubiquitin Ligase Gene Family in Grapevine
Published on: December 22, 2017
10.4K
相关概念视频
 03:36
Genome Annotation and Assembly
18.7K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
18.7K
 03:21
RNA-seq
9.7K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.7K
 01:05
Maxam-Gilbert Sequencing
10.5K
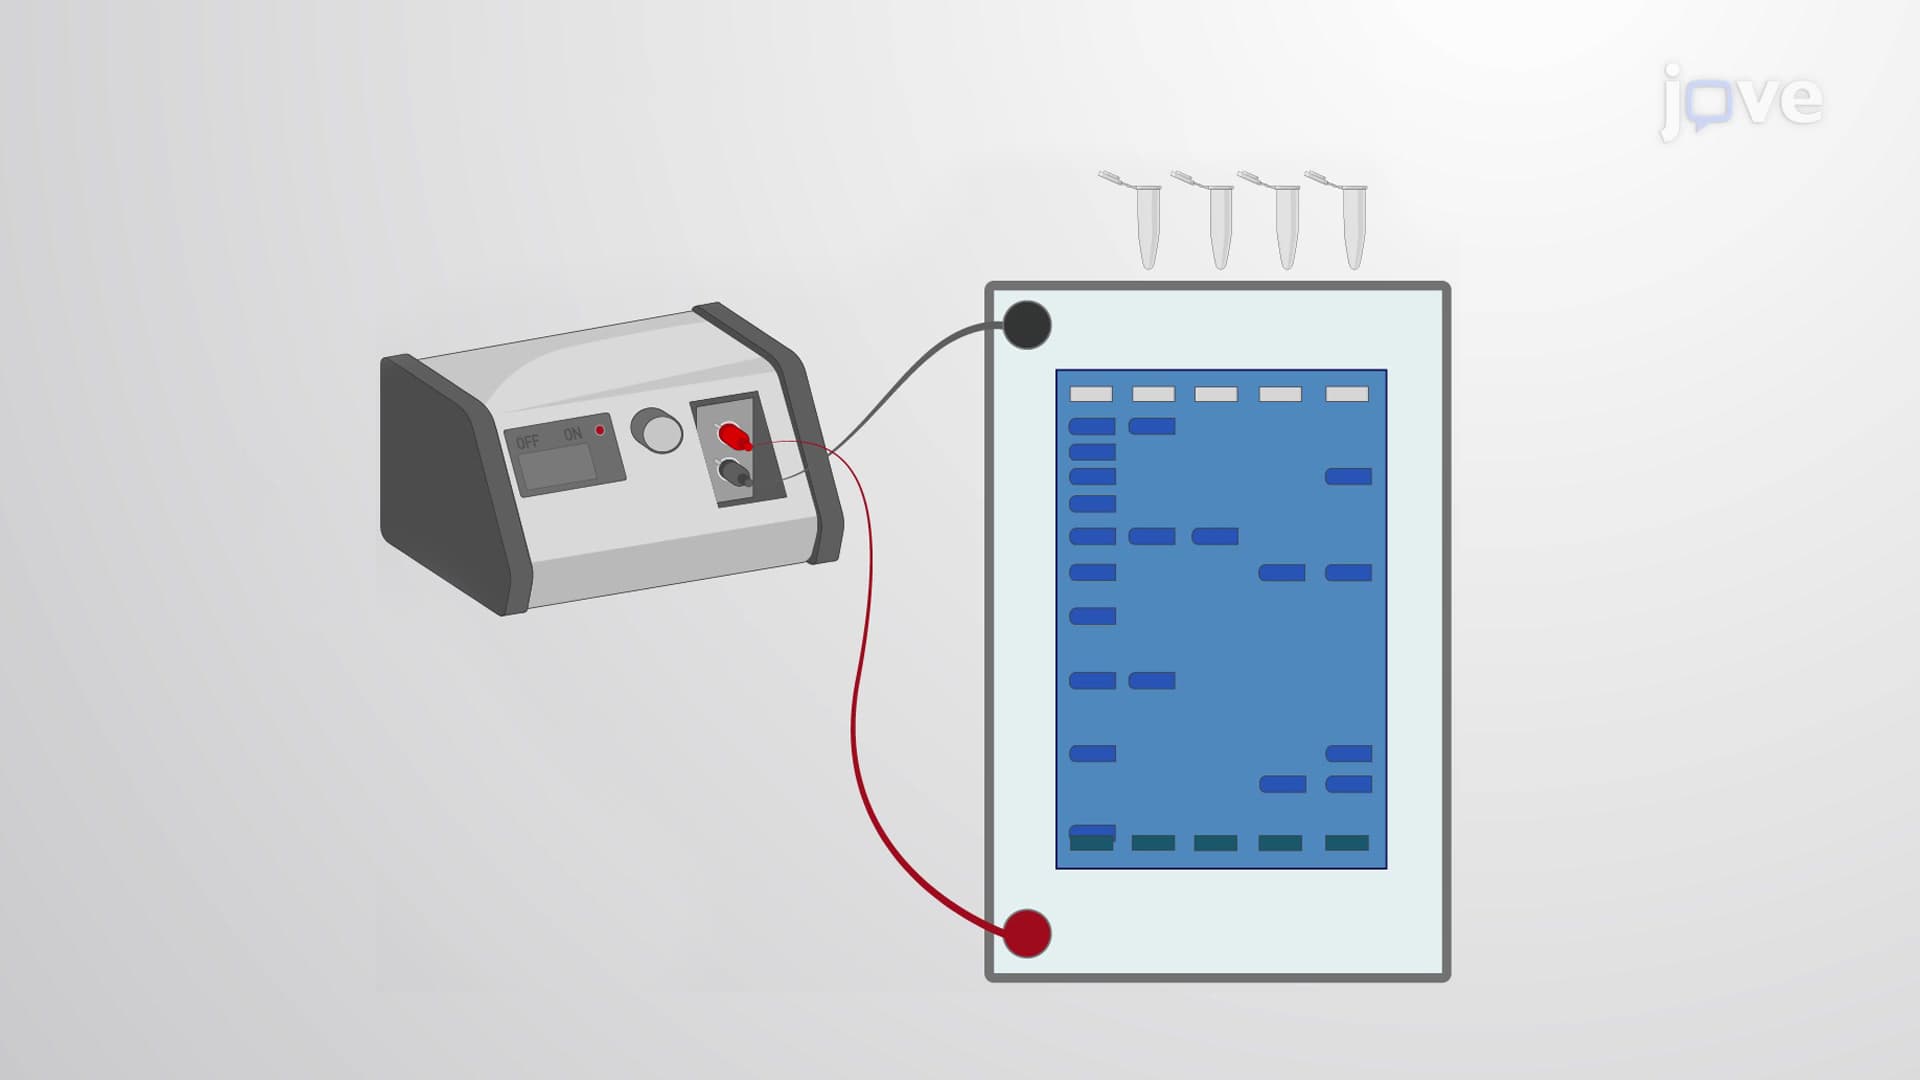
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
10.5K
 02:51
Multi-species Conserved Sequences
3.8K
Next-generation sequencing technologies have created large genomic databases of a variety of animals and plants. Ever since the human genome project was completed, scientists studied the genome of primates, mammals, and other phylogenetically distant living beings. Such large-scale studies have provided new insights into the evolutionary relationship between organisms.
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
3.8K
 01:57
Sanger Sequencing
751.2K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
751.2K
 02:35
RACE - Rapid Amplification of cDNA Ends
6.2K
Rapid Amplification of cDNA Ends, or RACE, is one of the most effective methods to obtain a full-length cDNA from an mRNA sequence between a known internal region to the unknown sequence at the 5’ or 3’ end. The unknown region is cloned in the cDNA by a gene-specific primer that binds the known end, and a hybrid primer that attaches a predefined anchor sequence to the unknown end of the cDNA. The sequence in between is amplified by PCR with an anchor primer and a gene-specific...
6.2K