探索DFT热化学对铁素模型复合体[CH3S4Fe2IIIS2H]的质子化状态的高灵敏度
Victor P Vysotskiy1, Ulf Ryde1
1Department of Computational Chemistry, Lund University, Chemical Centre, SE-22100 Lund, Sweden.
The Journal of chemical physics
|April 22, 2025

概括
铁硫复合体的精确热化学需要仔细选择密度函数理论 (DFT) 方法. r2SCAN,B3LYP和B97-1函数,以及密度校正的DFT和随机相近似,显示出可靠计算的希望.
科学领域:
- 计算化学的计算化学
- 量子化学 是一个量子化学.
- 生物物理化学 生物物理化学
背景情况:
- 密度函数理论 (DFT) 对3D过渡金属复合物的热化学对于函数中确切的交换组件非常敏感.
- 不准确的DFT方法可能会导致相对能量的显著误差 (数百kJ/mol),特别是在铁硫复合体的质子化状态.
研究的目的:
- 评估各种DFT函数的准确性,以计算 [2Fe-2S] 铁素模型复合体 ([CH3S4Fe2IIIS2H]-) 的质子化异构体的相对能量.
- 为了将DFT结果与高层次的ab initio方法比较,如合集群和量子蒙特卡罗.
主要方法:
- 使用各种DFT函数 (r2SCAN12-D4,B3LYP-D4,B97-1-D3(OP),TPSSh,O3LYP) 来计算[CH3S4Fe2IIIS2H]-复合物的四个质子化异构体的相对能量.
- 采用高层次的ab initio方法:无相辅助场量子蒙特卡罗 (AFQMC) 和合集群与单个和双重和扰动三重 (CCSD(T)).
- 通过使用r2SCAN函数在KS-CCSD电子密度和混合函数的直接随机相近似 (dRPA) 上进行密度校正的DFT.
主要成果:
- 与AFQMC和CCSDT基准相比,r2SCAN12-D4,B3LYP-D4和B97-1-D3 (OP) DFT方法表现最好.与AFQMC和CCSDT基准相比,r2SCAN12-D4,B3LYP-D4和B97-1-D3 (OP) DFT方法表现最好.
- 用r2SCAN函数进行密度校正的DFT提供了可靠的相对能量.
- 使用TPSSh,O3LYP和r2SCAN12混合函数的直接随机相近似 (dRPA) 计算也产生了准确的结果.
结论:
- 特定的DFT函数 (r2SCAN,B3LYP,B97-1) 和密度校正DFT和DRPA等方法为铁硫提供可靠的热化学.
- 这些发现指导了研究金属蛋白活性位点和相关无机复合物的适当计算方法的选择.
更多相关视频
相关概念视频
 01:22
EDTA: Chemistry and Properties
1.6K
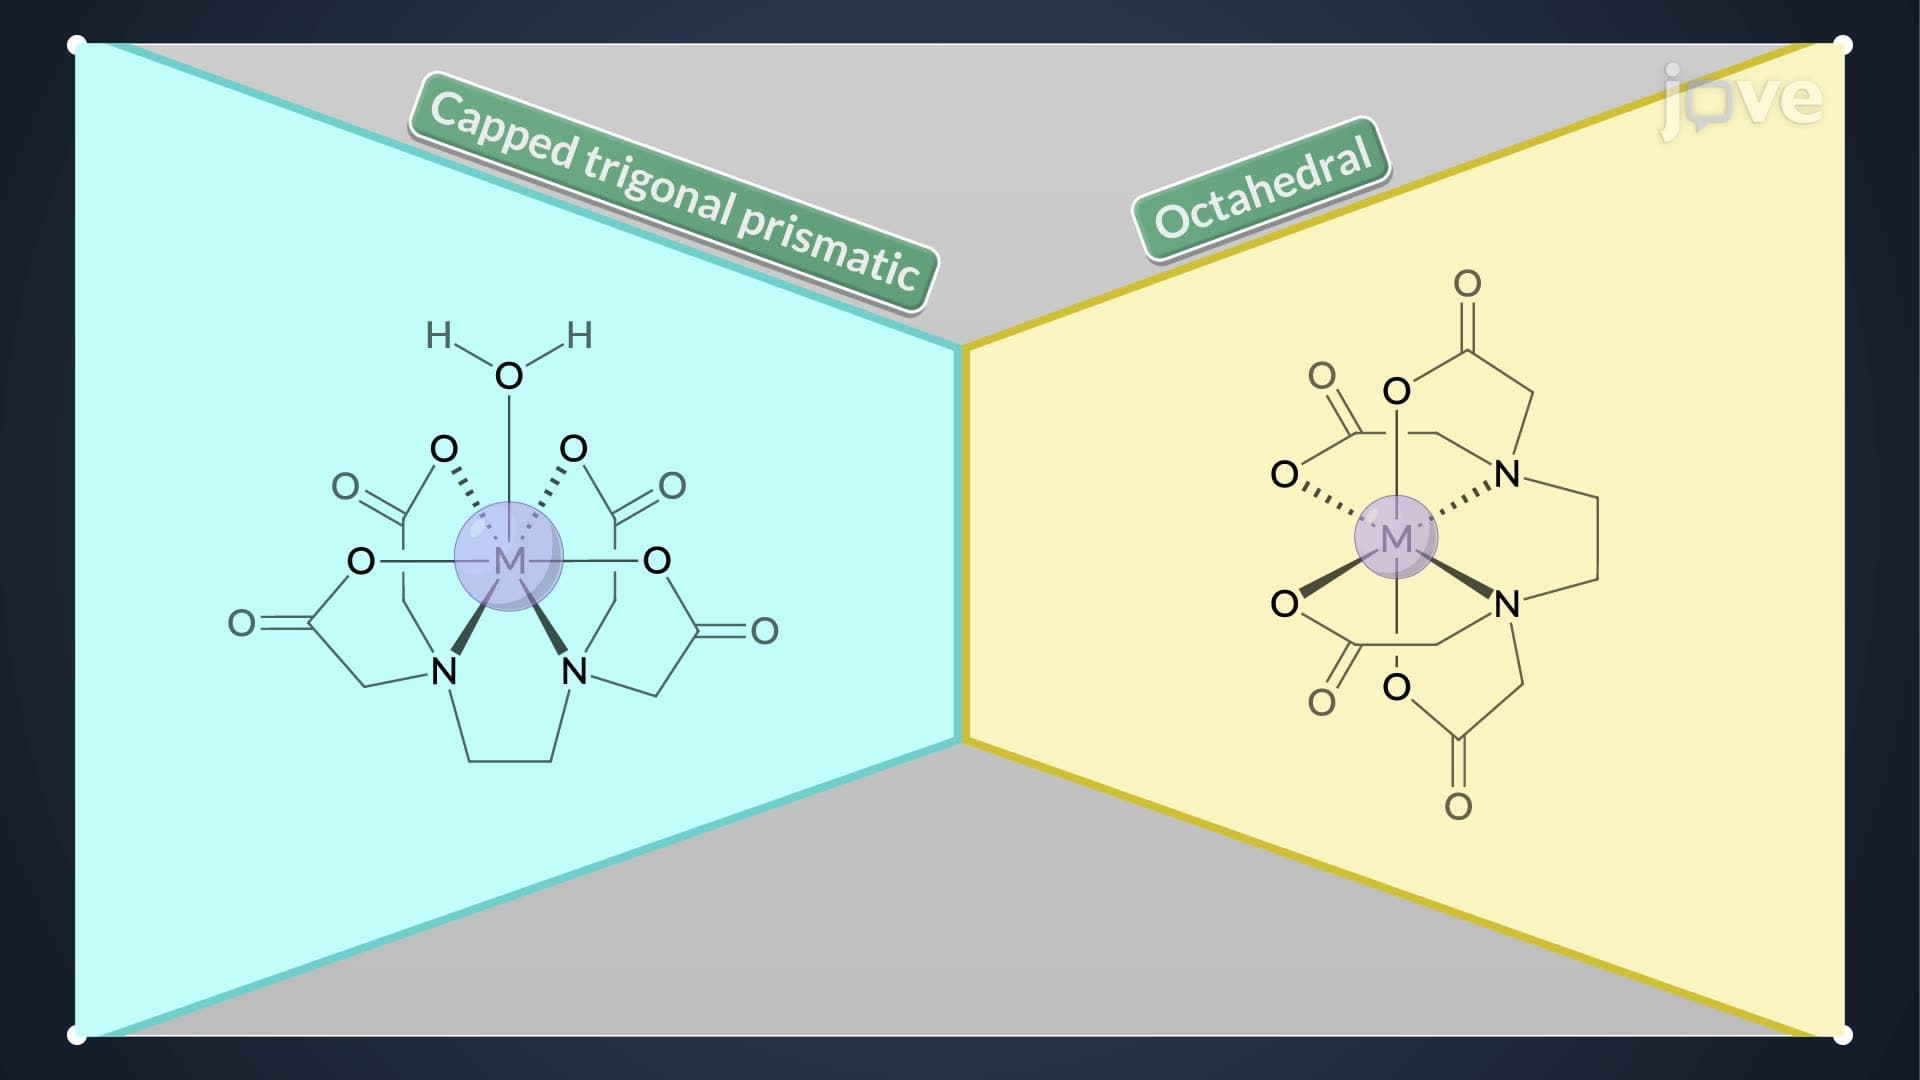
Polydentate ligands are most widely used in complexometric titrations because they form more stable complexes with the metal ions than mono- or bidentate ligands due to the chelate effect. Examples of polydentate ligands are ethylenediaminetetraacetic acid (EDTA), crown ethers, and cryptands. The most important feature of optimal polydentate ligands is the ability to form 1:1 complexes in a single-step process. Amino carboxylic acid derivatives are frequently used as complexing agents. EDTA is...
1.6K
 02:58
Crystal Field Theory - Octahedral Complexes
25.7K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
25.7K
 00:48
¹H NMR of Labile Protons: Deuterium (²H) Substitution
819
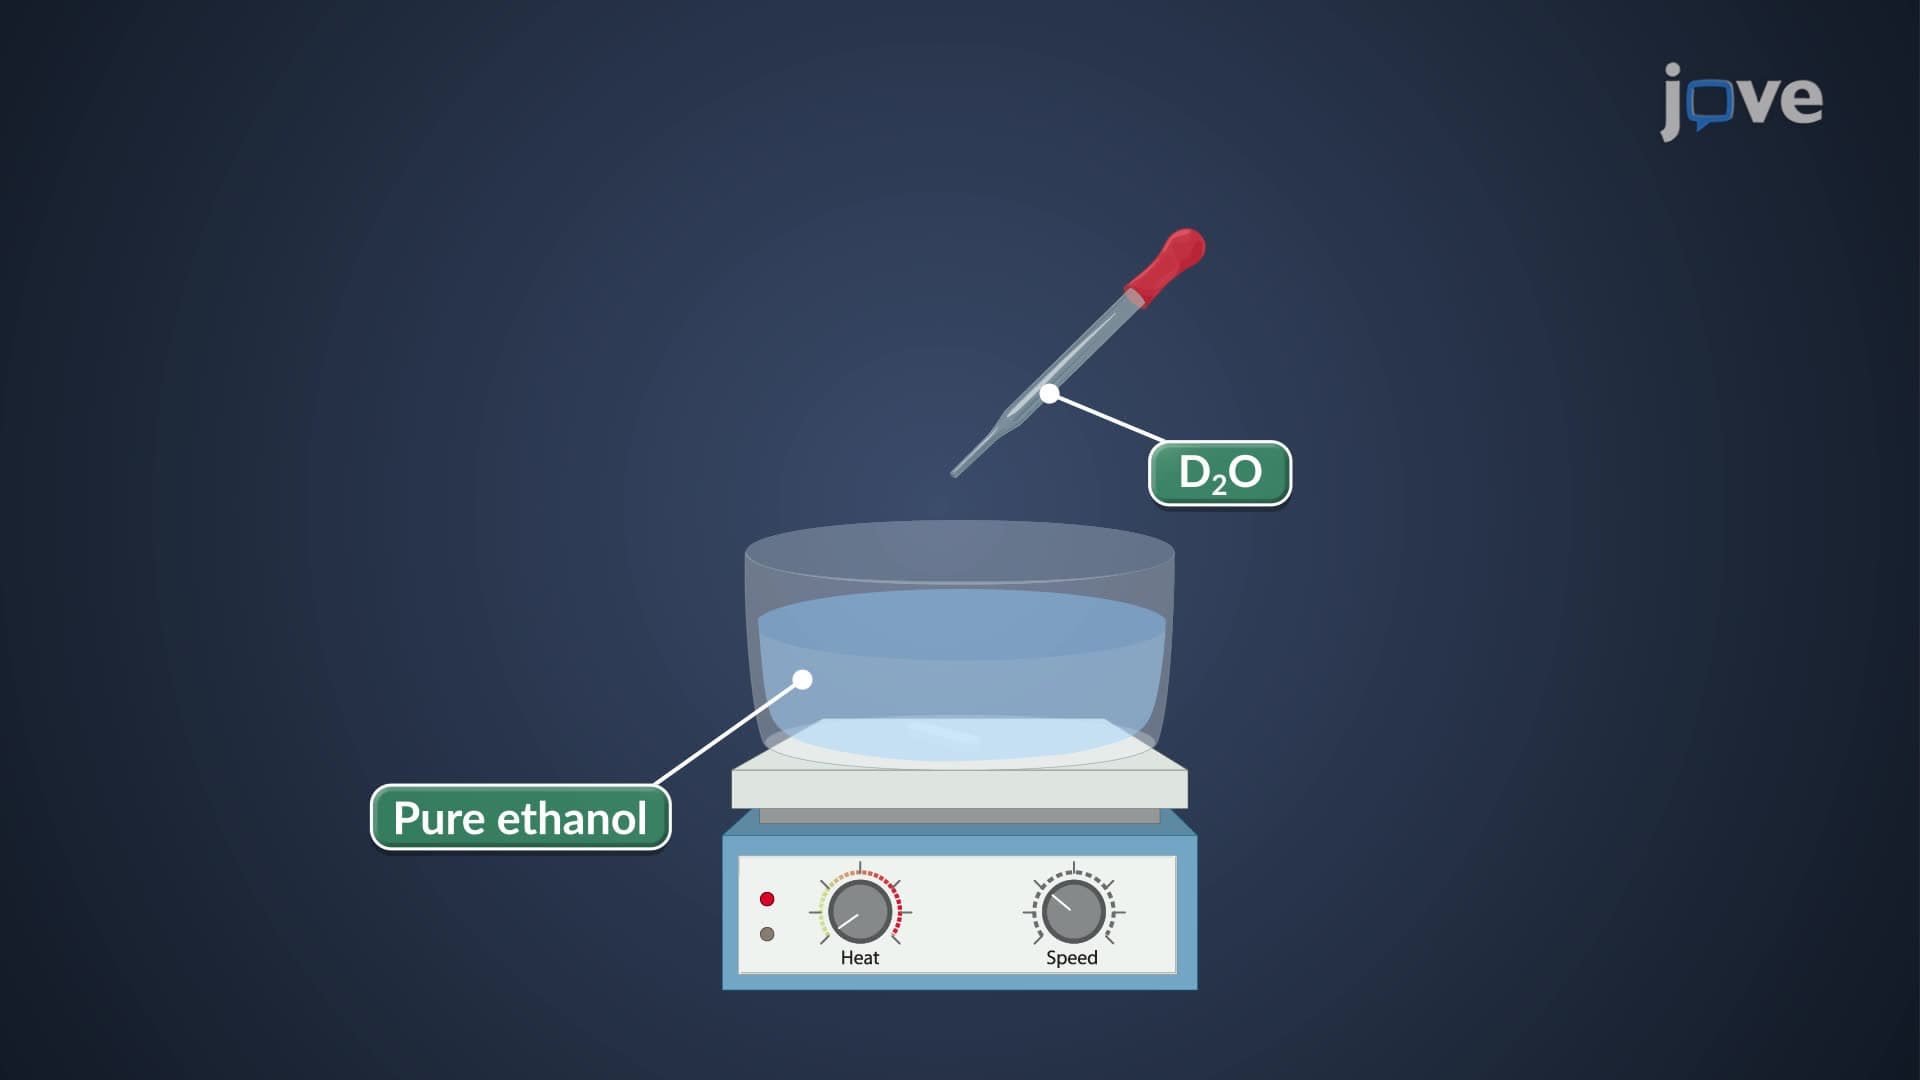
This lesson illustrates the role of deuterium substitution in simplifying the NMR spectrum of compounds comprising labile protons. One method employed is the use of deuterium. Amongst the three isotopes of hydrogen, deuterium (2H) has a nucleus composed of one proton and one neutron. When the D2O solvent is added to a pure dry ethanol solution, its labile proton is substituted with deuterium.
819
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
949
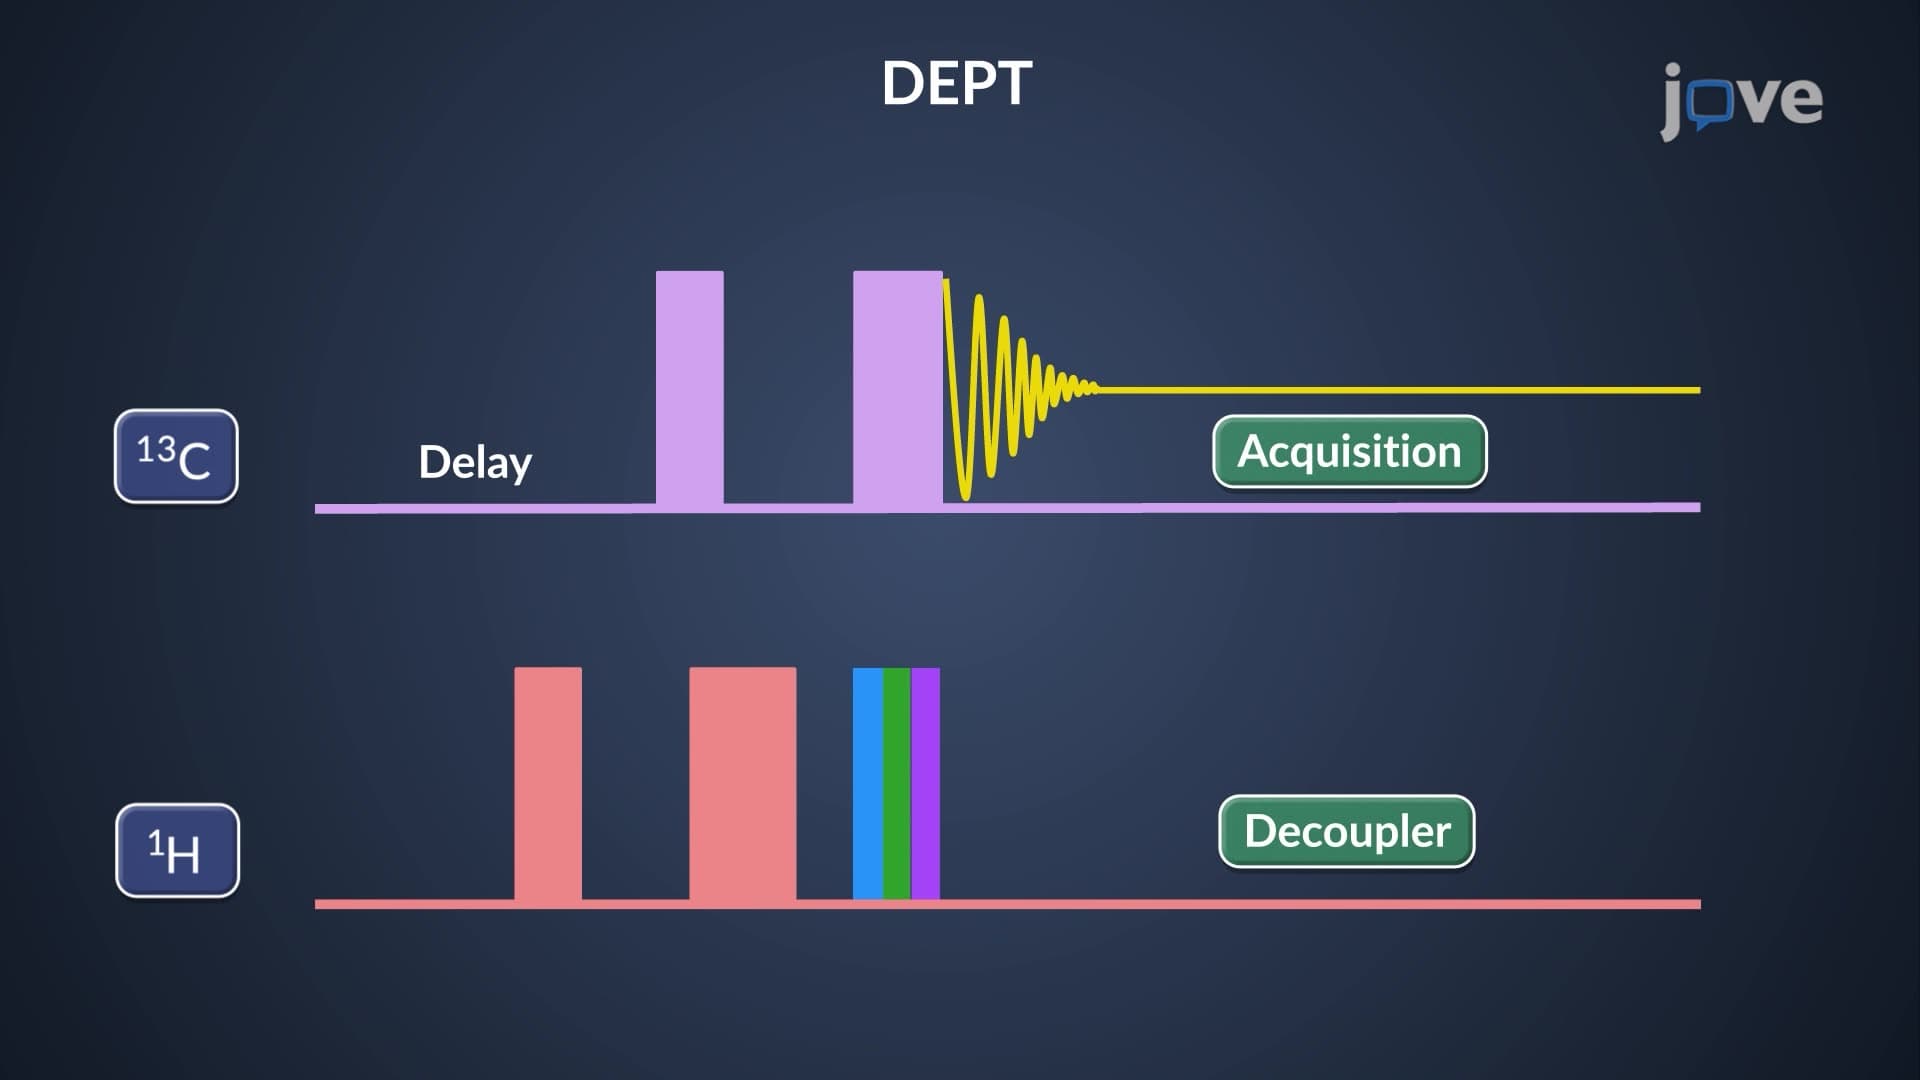
When proton-coupled carbon-13 spectra are simplified by a broadband proton decoupling technique, structural information about the coupled protons is lost. Distortionless enhancement by polarization transfer (DEPT) is a technique that provides information on the number of hydrogens attached to each carbon in a molecule. While the DEPT experiment utilizes complex pulse sequences, the pulse delay and flip angle are specifically manipulated. The resulting signals have different phases depending on...
949
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
40.7K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
40.7K
 01:30
Ladder Diagrams: Redox Equilibria
405
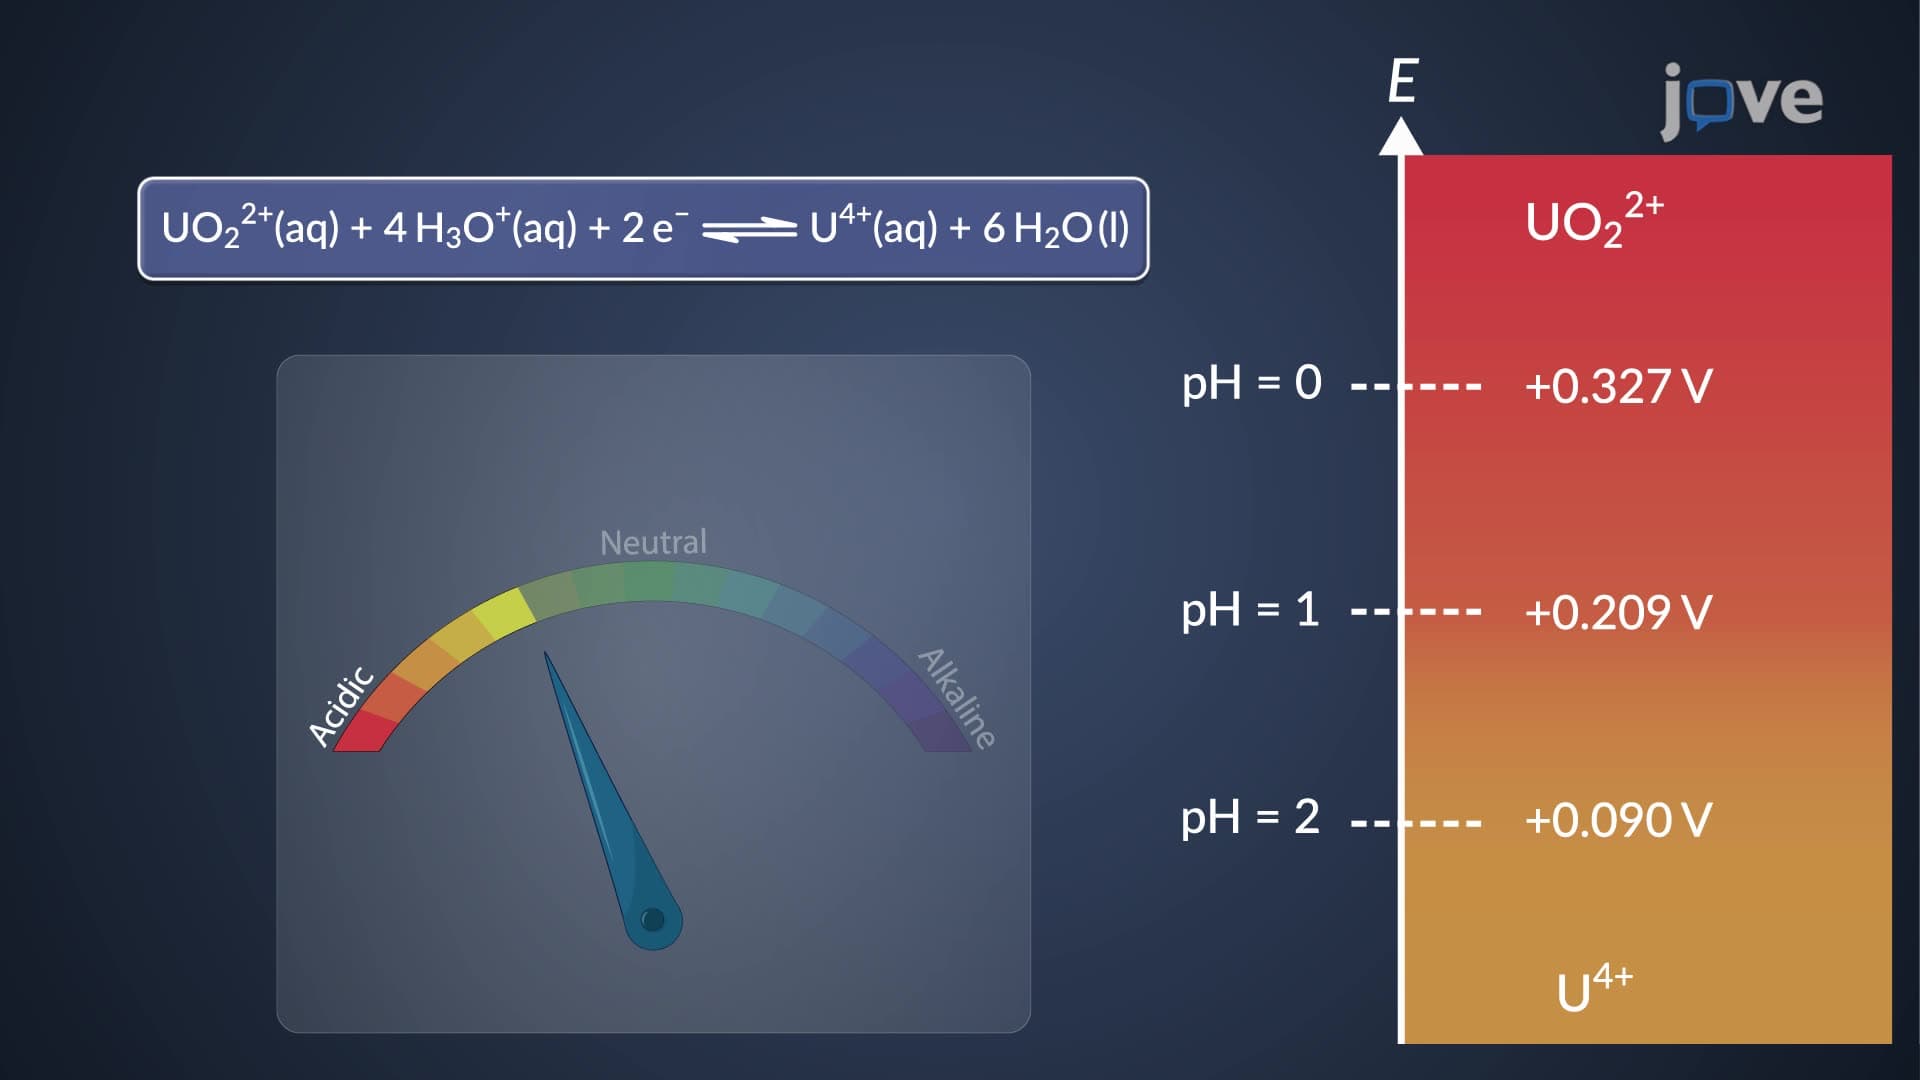
Ladder diagrams are useful tools for understanding redox equilibrium reactions, especially the effects of concentration changes on the electrochemical potential of the reaction. The vertical axis in the redox ladder diagrams represents the electrochemical potential, E. The area of predominance is demarcated using the Nernst equation.
Consider the Fe3+/Fe2+ half-reaction, which has a standard-state potential of +0.771 V. At potentials more positive than +0.771 V, Fe3+ predominates, whereas Fe2+...
Consider the Fe3+/Fe2+ half-reaction, which has a standard-state potential of +0.771 V. At potentials more positive than +0.771 V, Fe3+ predominates, whereas Fe2+...
405