
在SARS-CoV-2的测序研究中报告的与患者相关的元数据:范围审查和文献识别分析协议
Karen O'Connor1, Davy Weissenbacher2, Amir Elyaderani3
1Department of Biostatistics, Epidemiology, and Informatics, Perelman School of Medicine, University of Pennsylvania, Philadelphia, PA, United States.
JMIR research protocols
|April 22, 2025

概括
本研究评估了SARS-CoV-2基因组测序研究中的患者元数据. 结果将突出差距,并建议改善更好的传染病研究.
科学领域:
- 基因组流行病学基因组流行病学
- 病毒学 病毒学
- 生物信息学是一种生物信息学.
背景情况:
- 史无前例的SARS-CoV-2测序工作存在,在GISAID和GenBank等公共数据库中有数以百万计的记录.
- 基因组流行病学将遗传数据与患者信息联系起来,以了解传播和疾病的影响.
- 在SARS-CoV-2测序研究中,与患者相关的元数据很少,其质量在很大程度上尚未探索.
研究的目的:
- 量化评估SARS-CoV-2全基因组测序研究中患者报告的元数据的范围和质量.
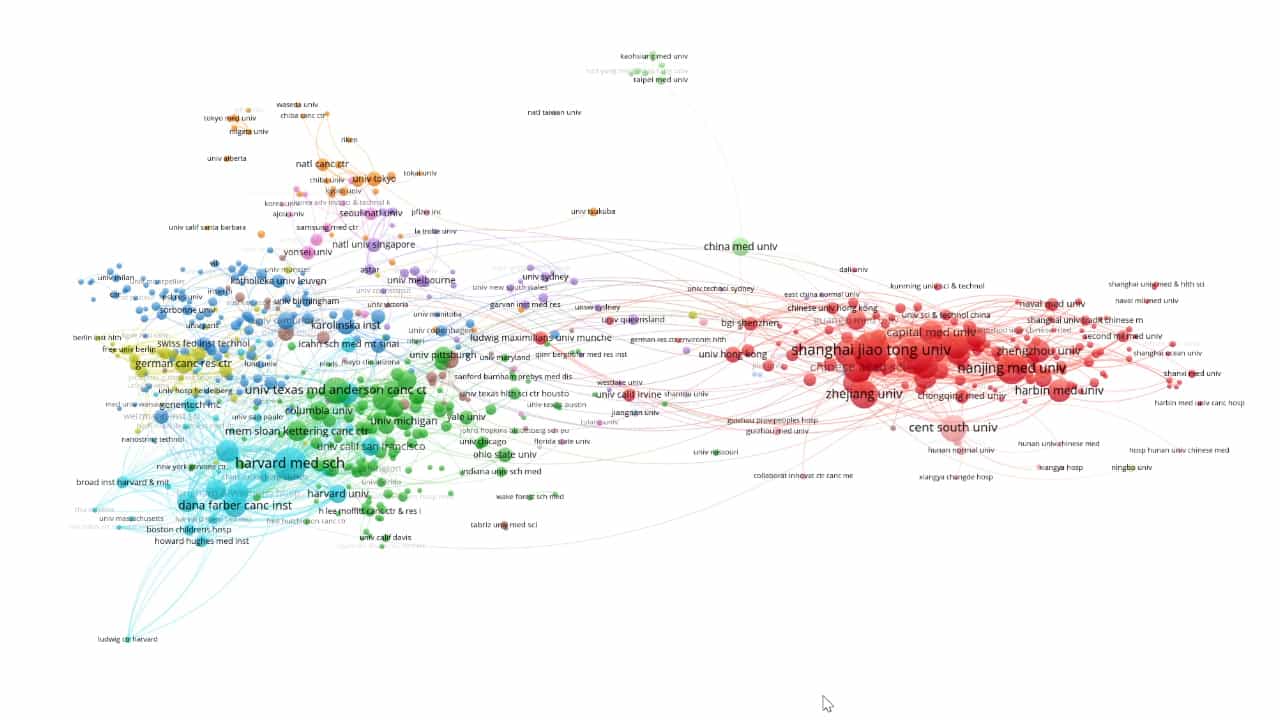
- 对与SARS-CoV-2基因组数据相关的出版模式进行文献计量分析.
- 评估机器学习分类器在识别相关研究的准确性,以便进行范围审查.
主要方法:
- 利用NIH的LitCovid集合进行序列沉积论文的自动分类.
- 在MEDLINE和PubMed Central中进行独立搜索以验证.
- 使用Covidence提取数据来合成和总结患者元数据的可用性.
- 对作者归属,引用指标和关键词进行文献计量分析.
主要成果:
- 开发并应用于LitCovid出版物的分类模型,直到2023年2月.
- 截至2024年8月的论文处理将于2024年9月开始.
- 文献搜索和选于2024年10月开始;结果将通过表格和图表进行叙述.
结论:
- 范围审查将报告SARS-CoV-2基因组研究中的患者元数据的范围和类型,并确定报告漏洞.
- 将提出建议,以提高患者元数据报告的质量和一致性.
- 图书统计分析将揭示元数据报告的趋势,可能为未来的传染病研究提供信息.
相关概念视频
 01:57
Sanger Sequencing
752.0K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
752.0K
 01:05
Single Nucleotide Polymorphisms-SNPs
13.8K
A single nucleotide polymorphism or SNP is a single nucleotide variation at a specific genomic position in a large population. It is the most prevalent type of sequence variation found in the human genome. Point mutations that occur in more than 1% of the population qualify as SNPs. These are present once every 1000 nucleotides on an average in the human genome. Replacement of a purine with another purine (A/G) or a pyrimidine with another pyrimidine (C/T) is known as a transition. In contrast,...
13.8K
 03:21
RNA-seq
9.7K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
9.7K
 03:00
Next-generation Sequencing
86.7K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
86.7K
 01:05
Maxam-Gilbert Sequencing
10.8K
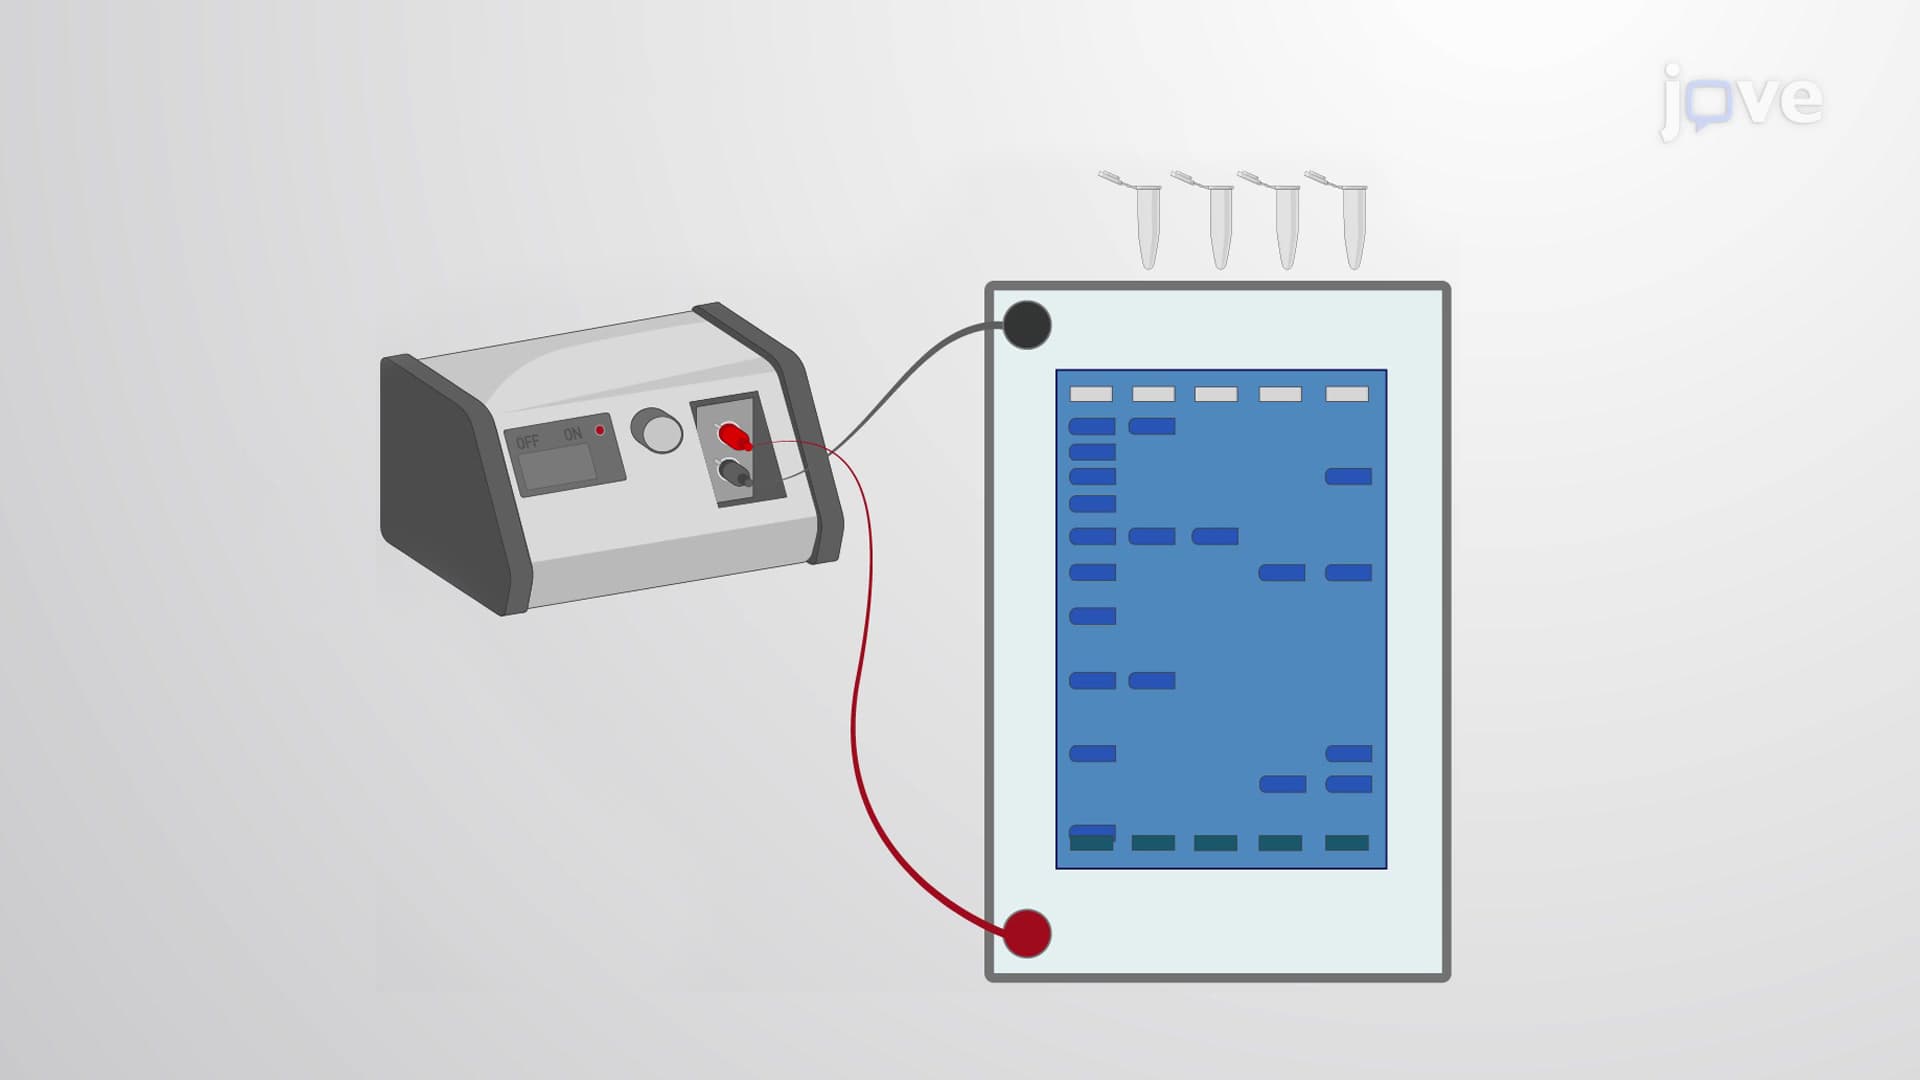
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
10.8K