

使用平面波/伪电位密度函数理论和机器学习的有机分子的光辐射光谱学:用于孤立分子,分子聚合物和有机薄膜的综合性和预测性计算协议
Francesco Porcelli1, Francesco Filippone1, Emanuela Colasante2
1Istituto di Struttura della Materia, Consiglio Nazionale delle Ricerche (ISM-CNR), Strada Provinciale 35d/9, Montelibretti 00010, Italy.
The Journal of chemical physics
|June 23, 2025

概括
我们使用平面波/伪电位密度函数理论 (PW-DFT) 开发了一个计算协议,准确预测孤立分子和分子膜的X射线光辐射光谱 (XPS),从而实现详细的电子结构分析.
科学领域:
- 计算化学的计算化学
- 频谱学是一种光谱学.
- 材料科学 材料科学 材料科学
背景情况:
- 光发射光谱学提供了对单分子水平的分子电子和结构性质的洞察.
- 准确的计算方法对于解释实验光辐射数据和分配光谱特征至关重要.
- 现有的方法在预测复杂分子的特定地点核心电离结合能方面面临挑战.
研究的目的:
- 开发和验证一个强大的计算协议,用于预测孤立分子的X射线光辐射光谱 (XPS).
- 评估各种密度函数和XPS预测计算方法的性能.
- 将协议的适用范围扩展到分子聚合物和薄膜,并探索价值光发射.
主要方法:
- 在 ΔSCF 框架内开发了一个基于平面波/伪潜在密度函数理论 (PW-DFT) 的计算协议.
- 评估了使用半局部和混合密度函数 (PBE,B3LYP,HSE,BH&HLYP) 和运动方程合集群的协议.
- 应用该方法来预测核心 (C1s,N1s,O1s) 和不同分子类的价值电离结合能.
主要成果:
- PW-DFT/ΔSCF协议准确地预测了XPS光谱的分离分子,包括芳香,异芳香,亚利法化合物,药物和生物分子.
- 证明了高精度和稳定性,即使使用半局密度函数近似.
- 该协议显示适用于分子聚合物,薄膜和价值光发射谱的预测.
结论:
- 开发的计算协议为解释和预测孤立分子和相关系统的XPS光谱提供了可靠的工具.
- 该研究验证了PW-DFT用于准确的核心级绑定能量计算的使用,从而促进实验数据分析.
- 提供了一个公共存储库,包含计算资源和机器学习数据集,以促进可重现性和进一步研究.
相关概念视频
 01:14
Molecular Spectroscopy: Absorption and Emission
3.4K
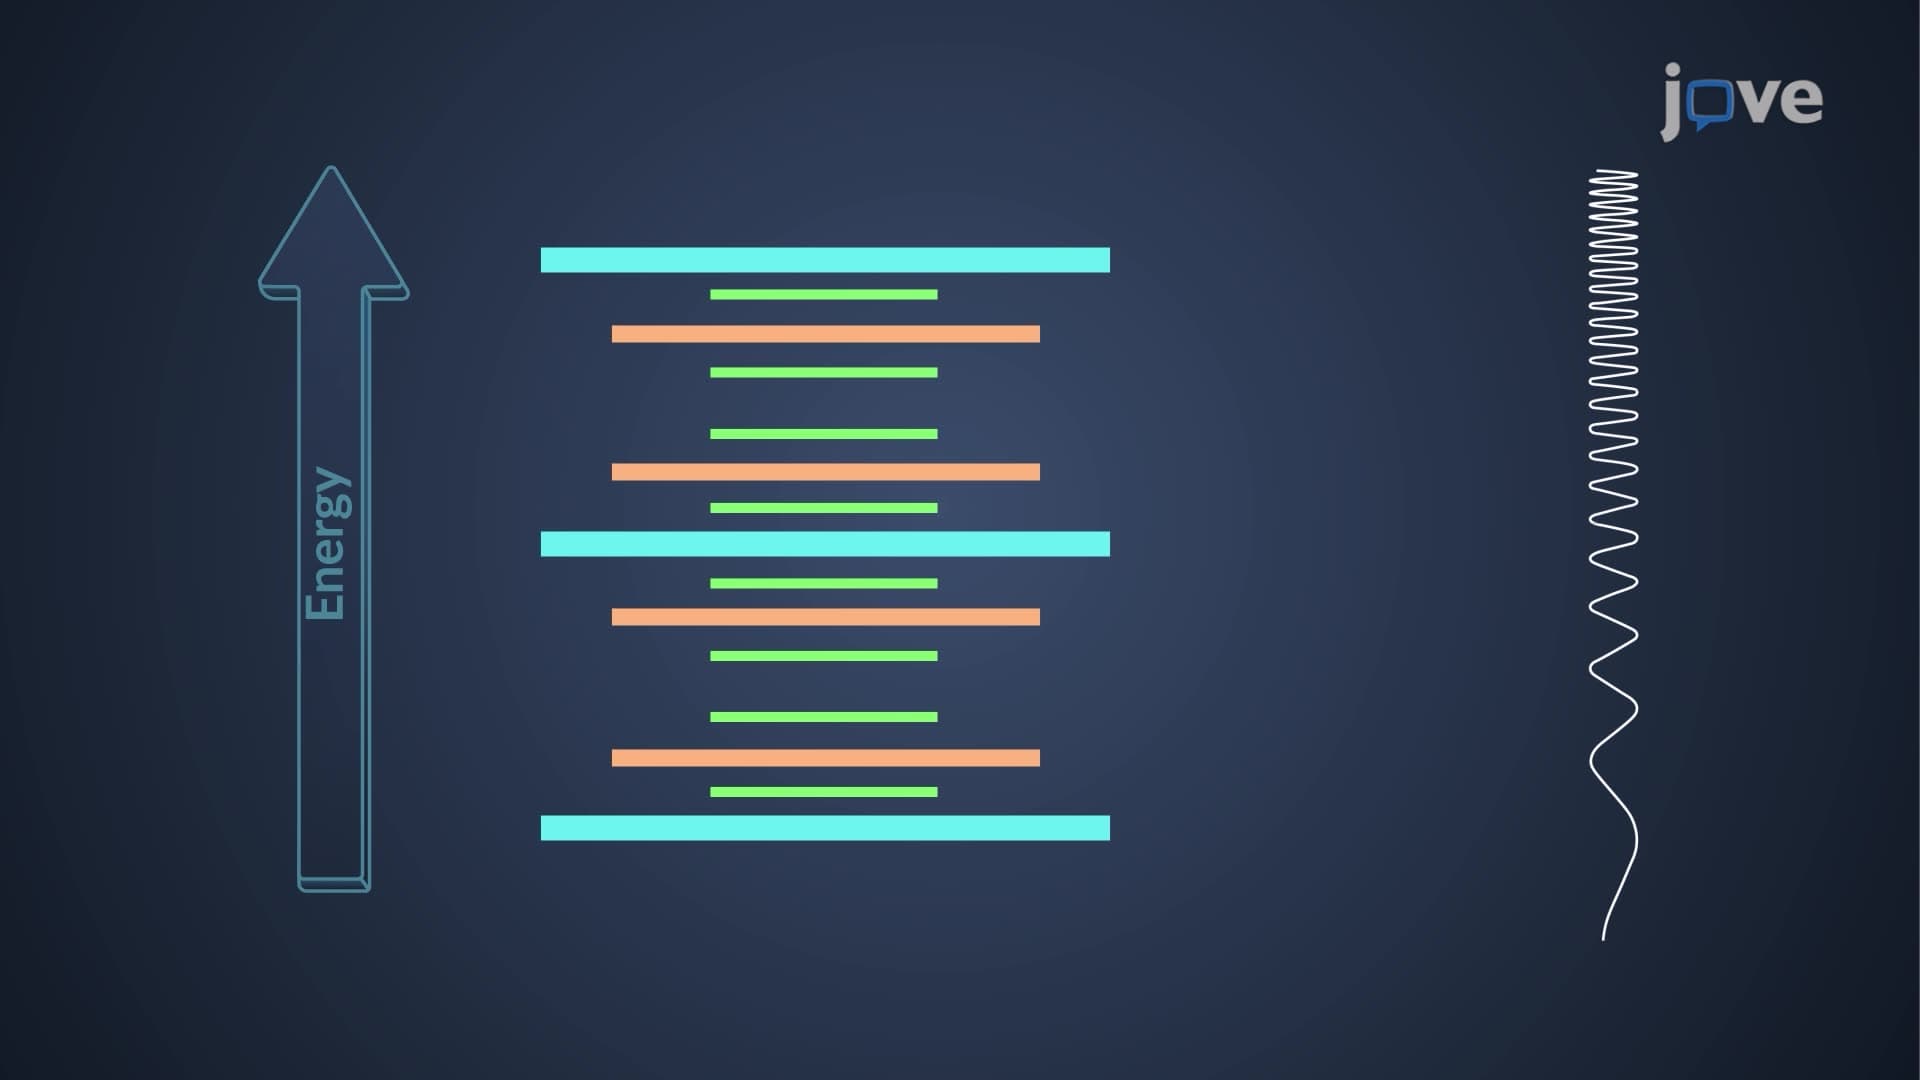
Molecules possess discrete energy levels called quantum states. Unlike atoms, which have simpler energy levels, molecules possess additional rotational and vibrational energy levels. Each energy level is separated by an energy gap, with the gaps between adjacent electronic, vibrational, and rotational levels varying significantly. The three types of energy levels in a diatomic molecule are shown in Figure 1.
3.4K
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
1.8K
In Ultraviolet–Visible (UV–Vis) spectroscopy, the absorption of electromagnetic radiation is used to probe the electronic structure of molecules. This technique provides insights into molecular electronic transitions, particularly the movement of electrons between different molecular orbitals. Radiation is absorbed if the energy of the electromagnetic radiation passing through the molecule is precisely equal to the energy difference between the excited and ground states. During this...
1.8K
 01:17
Electron Paramagnetic Resonance (EPR) Spectroscopy: Organic Radicals
2.7K
Ideally, an unpaired electron shows a single peak in the EPR spectrum due to the transition between the two spin energy states. However, coupling interactions can occur between the spins of the unpaired electron and any neighboring spin-active nuclei. This hyperfine coupling results in hyperfine splitting, where the EPR signal is split into multiplets. The signals split into 2nI + 1 peaks, where n is the number of equivalent nuclei and I is the nuclear spin. These splitting patterns provide...
2.7K
 01:32
UV–Vis Spectroscopy of Conjugated Systems
7.4K
Organic compounds with conjugated double bonds show strong absorption features in the UV–visible region of the electromagnetic spectrum attributed to π → π* electronic excitations. Generally, a UV–vis absorption spectrum is recorded as a plot of absorbance vs wavelength. The wavelength of maximum absorbance, which manifests as a peak in the absorption spectrum, is denoted as λmax.
One of the factors influencing λmax is the extent...
One of the factors influencing λmax is the extent...
7.4K
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
1.7K
A covalently bonded heteronuclear diatomic molecule can be modeled as two vibrating masses connected by a spring. The vibrational frequency of the bond can be expressed using an equation derived from Hooke's law, which describes how the force applied to stretch or compress a spring is proportional to the displacement of the spring. In this case, the atoms behave like masses, and the bond acts like a spring.
According to Hooke's law, the vibrational frequency is directly proportional to...
According to Hooke's law, the vibrational frequency is directly proportional to...
1.7K
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
25.7K
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given...
25.7K