

[AnCp3] (其中An = Th-Cf) 复合体中的电子结构,共价性和磁性异质性:从第一原理计算中的见解
Ibtesham Tarannum1, Prem Prakash Sahu1,2, Shruti Moorthy1
1Computational Inorganic Chemistry Group, Department of Chemistry, Indian Institute of Technology Hyderabad, Kandi, Sangareddy, Telangana, 502284, India.
Chemistry, an Asian journal
|June 24, 2025

概括
这项研究揭示了活性化物-连接物结合和磁性特性在活性化物系列中的变化. 它突出了早期动因体中的6d-共性向更重元素中的5f-共性的转变,这对于理解它们的行为至关重要.
科学领域:
- 无机化学 无机化学
- 计算化学计算化学
- 量子化学 是一个量子化学.
背景情况:
- 乙胺化学是一个快速发展的领域,人们越来越感兴趣了解乙胺 - 干相互作用.
- 关键性质,如共价性和磁性行为是这项研究的核心.
- 活性化物复合物的电子结构决定了它们的化学和物理性质.
研究的目的:
- 为了研究九个[AnCp3]复合物的电子结构和磁性异构性 (An = Th(III) -Cf(III)).
- 阐明动因酸-联体共价性质的性质,重点关注5f和6d轨道贡献之间的相互作用.
- 为了了解驱动行为系列共价性趋势的因素.
主要方法:
- 尺度相对论密度函数理论 (SR-DFT) 用于电子结构计算.
- 用完整的活跃空间自相一致场 (CASSCF) 方法来分析磁性.
- 分子轨道 (MO),自然人群分析 (NPA),自然局部分子轨道 (NLMO) 和ab initio联结体场理论 (AILFT) 用于详细分析.
主要成果:
- 计算预测了电子配置,从的6d1到-加利福尼亚类似物的5fn.
- 密度函数理论 (DFT) 表明,在早期的动因体中,从主导的6d-共性过渡到后来的动因体中主导的5f-共性.
- CASSCF的计算准确地复制了实验中的g移和磁感应,证实了5f-联结体共价性的意义.
结论:
- 这项研究强调了5f和6d轨道贡献在动因化物-连接体结合中的关键作用.
- 在较重的活性化物中观察到向能量驱动的协同价值的转变,影响了结合趋势.
- 这些发现强调了5f-联结体共价性在确定这些动因化物复合物的磁性特性方面的重要性.
关键词:
结合的结合 结合的结合CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF CASSCF相对应的共价性在 DFT 方面,它是最重要的.F 元素是指 F 元素的元素.磁性异构性是一种磁性异构性.

相关概念视频
 02:42
Valence Bond Theory
9.7K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
9.7K
 02:58
Crystal Field Theory - Octahedral Complexes
28.0K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
28.0K
 03:02
Colors and Magnetism
12.4K
Color in Coordination Complexes
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
12.4K
 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
1.1K
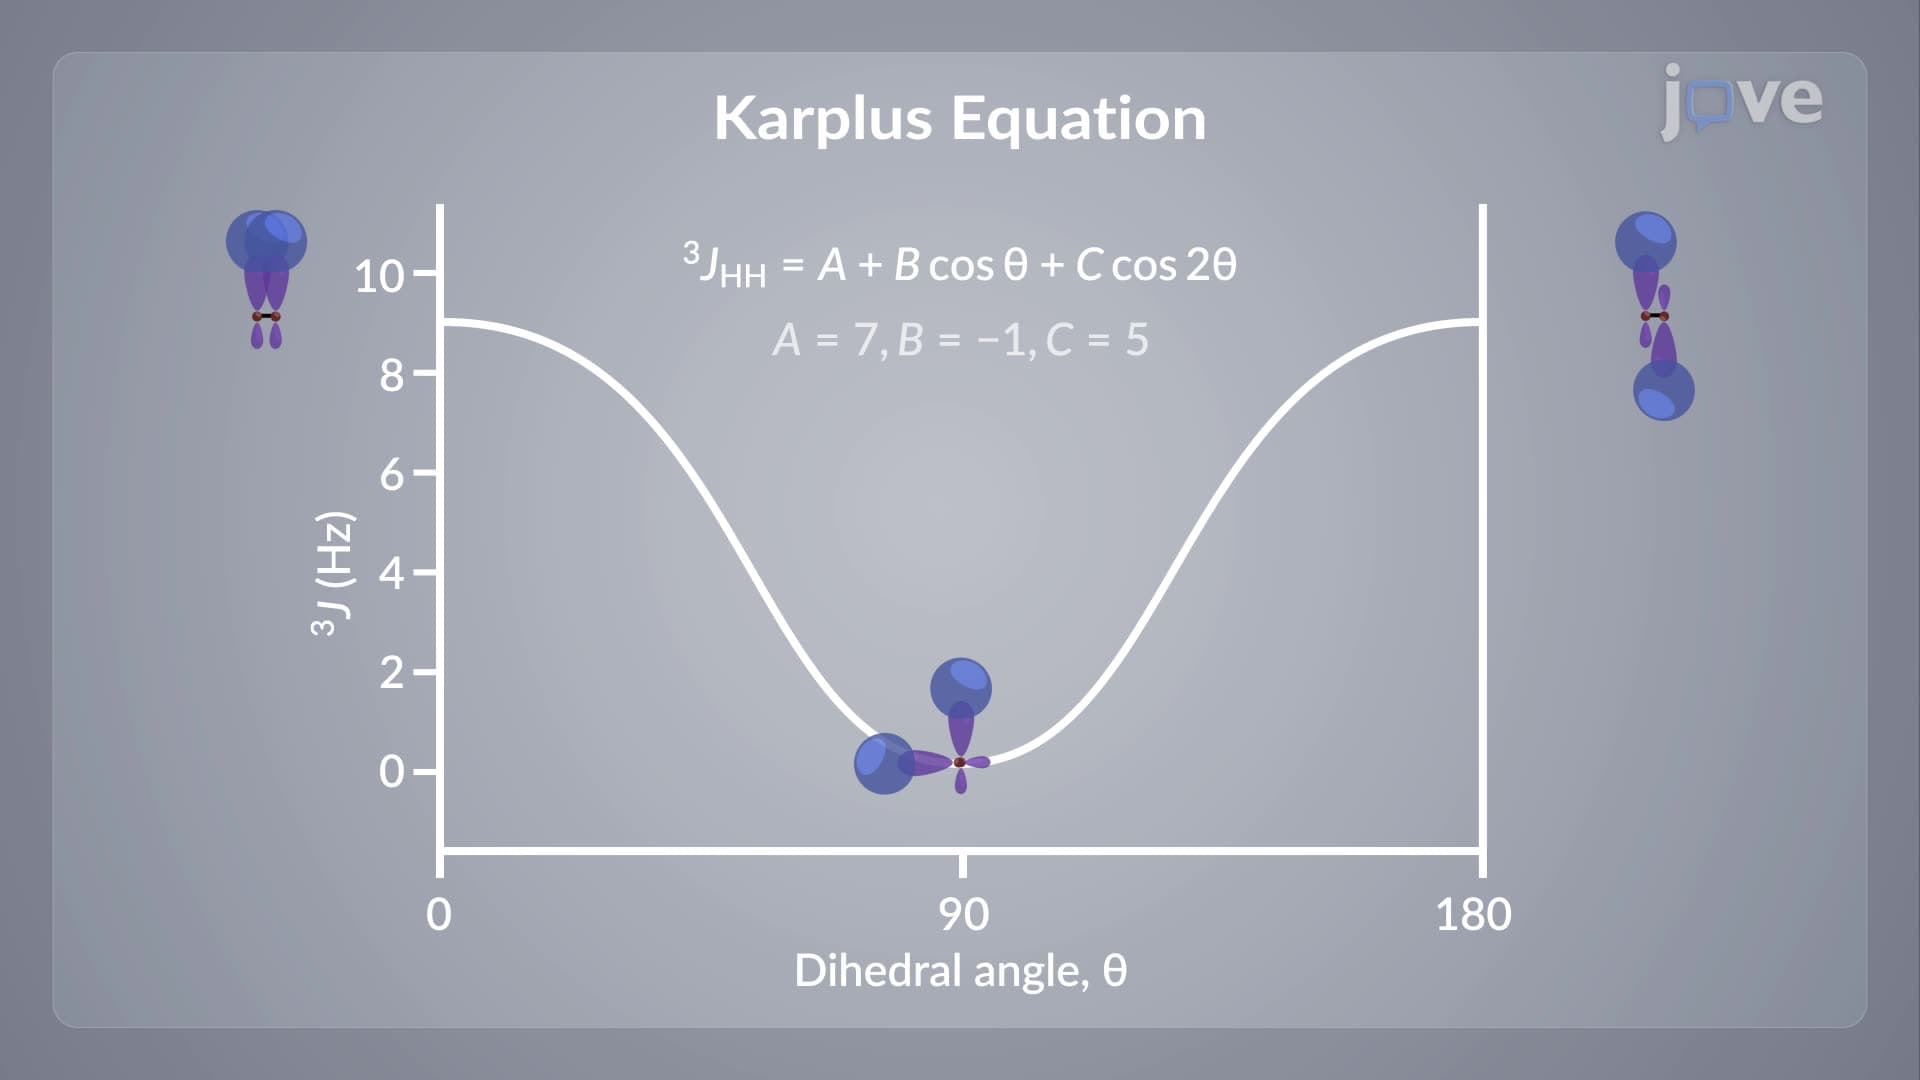
Vicinal or three-bond coupling is commonly observed between protons attached to adjacent carbons. Here, nuclear spin information is primarily transferred via electron spin interactions between adjacent C‑H bond orbitals. This generally favors the antiparallel arrangement of spins, so 3J values are usually positive.
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
1.1K
 02:28
Electronic Structure of Atoms
24.6K
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
24.6K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
44.8K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
44.8K