相关实验视频
Updated: Sep 17, 2025
05:22
Analyzing Multifactorial RNA-Seq Experiments with DiCoExpress
Published on: July 29, 2022
3.7K
传说:在多模式转录组测序数据中识别共表达的基因
Tao Deng1,2, Mengqian Huang3, Kaichen Xu3
1School of Data Science, The Chinese University of Hong Kong, Shenzhen (CUHK-Shenzhen), Shenzhen 518172, China.
Genomics, proteomics & bioinformatics
|July 1, 2025

概括
传奇集成了单细胞RNA测序 (scRNA-seq) 和空间解析的转录组 (SRT) 数据,以识别共同表达的基因组. 这种新的方法揭示了基因协同功能和空间模式,增强了生物学见解.
科学领域:
- 基因组学就是基因组学.
- 计算生物学 计算生物学
- 系统生物学 系统生物学
背景情况:
- 基因共同表达分析对于理解生物功能和疾病机制至关重要.
- 现有的方法通常独立分析单细胞RNA测序 (scRNA-seq) 或空间解析的转录组 (SRT) 数据,可能缺少集成的协同功能信号.
- 需要利用scRNA-seq和SRT数据进行全面的基因共表达分析的方法.
研究的目的:
- 引入LEGEND (多模共同表达的GEN finDer),一种用于整合scRNA-seq和SRT数据的新型计算方法.
- 在细胞类型和组织域级别上识别共同表达的基因组,捕获细微的模式.
- 为了证明LEGEND在探索基因协同功能,空间模式和与疾病相关的基因相互作用方面的实用性.
主要方法:
- 在LEGEND中使用层次聚类算法来整合scRNA-seq和SRT数据.
- 该算法旨在最大限度地提高集群内的冗余性和集群之间的互补性.
- 丰富和协同功能分析用于验证已识别的基因集群的生物相关性.
主要成果:
- 通过整合多式转录基因数据,LEGEND成功地识别了生物相关的共同表达的基因组.
- 该方法揭示了细微的基因共同表达模式和跨细胞类型和组织领域的空间连贯性.
- 传奇在探索特定环境的基因功能,基因相互作用的转变以及提高scRNA-seq和SRT数据的注释精度方面具有实用性.
结论:
- 传奇提供了一个强大的方法来整合scRNA-seq和SRT数据,以发现协同功能基因.
- 该方法提升了我们对基因协同表达,空间转录组学和生物和病理背景中的基因功能的理解.
- 通过分析集成的转录基因数据,LEGEND促进了发现新型基因功能和与疾病相关的基因交叉通话.
更多相关视频
相关概念视频
 02:24
Ribosome Profiling
3.6K
Ribosome profiling or ribo-sequencing is a deep sequencing technique that produces a snapshot of active translation in a cell. It selectively sequences the mRNAs protected by ribosomes to get an insight into a cell’s translation landscape at any given point in time.
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique...
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique...
3.6K
 03:21
RNA-seq
10.4K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
10.4K
 02:34
DNA Microarrays
18.6K
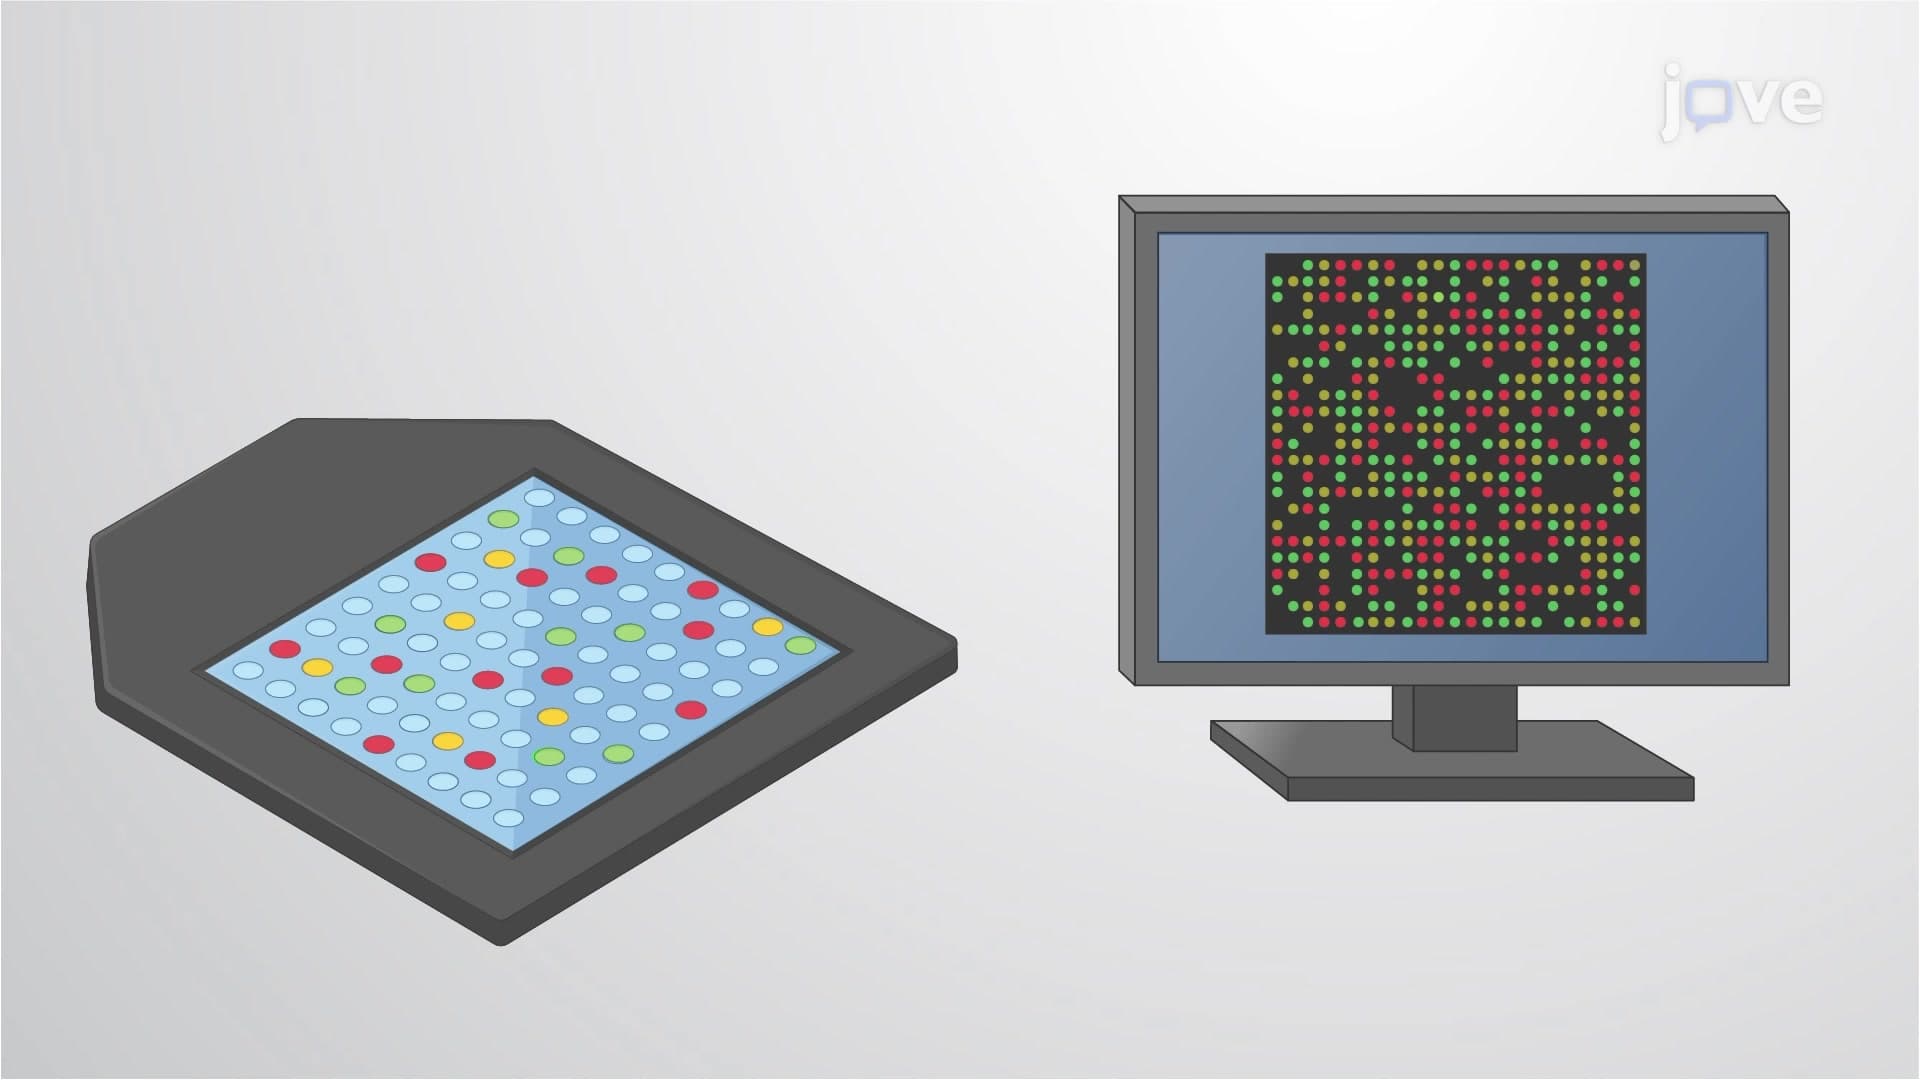
Microarrays are high-throughput and relatively inexpensive assays that can be automated to analyze large quantities of data at a time. They are used in genome-wide studies to compare gene or protein expression under two varied conditions, such as healthy and diseased states. Microarrays consist of glass or silica slides on which probe molecules are covalently attached through surface functionalization. Most commonly, the slides are prepared through the chemisorption of silanes to silica...
18.6K
 01:23
Regulation of Expression at Multiple Steps
1.0K
The gene expression in cells is regulated at different stages: (i) transcription, (ii) RNA processing, (iii) RNA localization, and (iv) translation. Transcriptional regulation is mediated by regulatory proteins such as transcription factors, activators, or repressors—these control gene expression by initiating or inhibiting the transcription of genes. Once a precursor or pre-mRNA is produced, it undergoes post-transcriptional modification, including 5' capping, splicing, and the...
1.0K