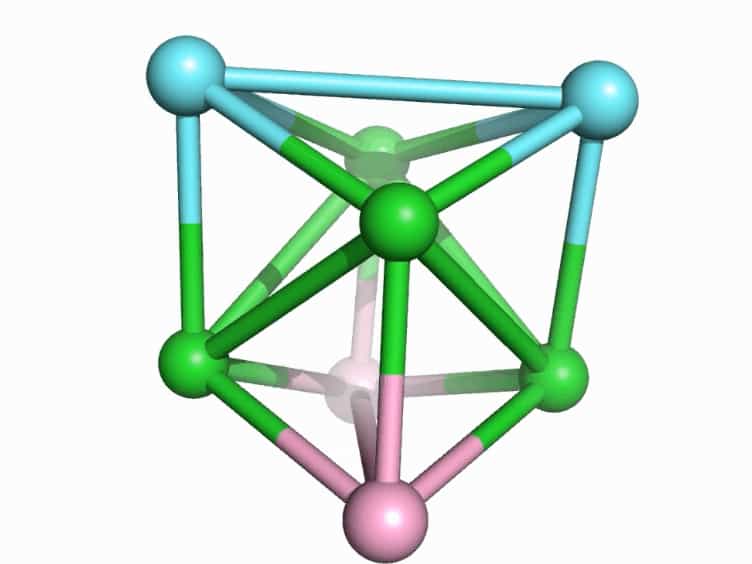

[Au11(BINAP) 4X2的溶剂依赖:X=Cl或Br]+集群电子和光学特性
Daniel J Heintzelman1, Jane A Knappenberger1, Kenneth L Knappenberger1,2
1Department of Chemistry, Pennsylvania State University, University Park, Pennsylvania 16802, United States.
The journal of physical chemistry. A
|July 10, 2025

概括
溶剂相互作用显著影响黄金纳米集群属性. 调整集群-溶剂键会改变电子和光学特征,这对于原子精确的纳米集群至关重要.
科学领域:
- 纳米材料 化学 化学
- 物理化学 物理化学
- 频谱学是一种光谱学.
背景情况:
- 原子精确的金纳米集群表现出独特的光物理特性.
- 溶解相互作用对于理解纳米集群行为至关重要.
- 单层保护的金纳米集群 ([Au11(BINAP) 4X2]+) 是研究集群溶剂效应的模型系统.
研究的目的:
- 为了研究金纳米集群中复杂的集群-溶剂相互作用.
- 为了确定溶剂同一性对[Au11(BINAP) 4X2]+的光物理性质的影响.
- 阐明溶解在调节纳米集团电子和光学特性中的作用.
主要方法:
- 紫外线可见 (UV-vis) 吸收光谱学.紫外线可见 (UV-vis) 吸收光谱学.
- 光发光光谱学. 光发光光谱学.
- 5秒短暂吸收 (fs-TA) 光谱学.
- 质子核磁共振 (1H NMR) 光谱学.
主要成果:
- 蛋白溶剂增加了[Au11(BINAP) 4X2]+纳米集群的刚性,由缩小的紫外线光谱峰值证明.
- 光发光的排放配置文件可以通过调节集群-溶剂相互作用来调整.
- fs-TA实验揭示了不同的辐射衰变路径,受溶剂相互作用强度的影响,乙醇比butanol具有更稳定的结合.
- 1H NMR证实了[Au11(BINAP) 4Cl2]+和乙醇之间的键.
结论:
- 集群溶剂键是可调的,在确定原子精确纳米集群的电子和光学性质方面发挥着至关重要的作用.
- 溶解显著影响金纳米集群的光物理行为.
- 了解和控制集群-溶剂相互作用是设计具有特定功能的纳米集群的关键.
相关概念视频
 02:42
Valence Bond Theory
9.7K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
9.7K
 03:02
Colors and Magnetism
12.3K
Color in Coordination Complexes
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
12.3K
 02:58
Crystal Field Theory - Octahedral Complexes
28.0K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
28.0K
 04:01
VSEPR Theory and the Effect of Lone Pairs
44.1K
Effect of Lone Pairs of Electrons on Molecule Geometry
44.1K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
44.8K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
44.8K
 03:02
The Aufbau Principle and Hund's Rule
62.4K
To determine the electron configuration for any particular atom, we can build the structures in the order of atomic numbers. Beginning with hydrogen, and continuing across the periods of the periodic table, we add one proton at a time to the nucleus and one electron to the proper subshell until we have described the electron configurations of all the elements. This procedure is called the aufbau principle, from the German word aufbau (“to build up”). Each added electron occupies the...
62.4K