
预测人体血中未结合的per-和多基基物质的分数:评估转移学习作为对稀缺数据问题的算法解决方案
Gabriel Sinclair1, Nathaniel Charest1, Barbara A Wetmore1
1Center for Computational Toxicology and Exposure, Office of Research and Development, U.S. Environmental Protection Agency, Research Triangle Park, North Carolina 27711, United States.
Journal of chemical information and modeling
|July 22, 2025

概括
我们探索了机器学习来预测血 (Fup) 中的未结合分数,这对于毒动力学模型至关重要. 转移学习改善了小化学品家族的预测,但在稀疏的PFAS数据方面遇到了困难,突出了需要更多实验性表征的需求.
科学领域:
- 毒理学和化学信息学
- 生理学基础的毒动力学 (PBTK) 建模
- 机器学习在化学中的应用.
背景情况:
- 血中未结合的部分 (Fup) 对PBTK模型至关重要,表明生物可用性和生物积累潜力.
- 准确的Fup预测具有挑战性,特别是在数据稀疏的化学品类,如PFAS.
- 现有的定量结构-活性关系 (QSAR) 模型需要针对特定化学品家族进行改进.
研究的目的:
- 开发和评估用于增强Fup QSAR建模的新型机器学习方法.
- 对跨化学空间的Fup预测进行转移学习策略的有效性进行调查.
- 对全球随机森林模型进行PFAS数据微调的深度学习模型的性能评估.
主要方法:
- 使用深度学习模型,在广泛的化学库上进行训练,并在一个小的PFAS数据集上进行微调.
- 采用转移学习方法,在不同的化学空间中适应模型.
- 将微调模型的性能与先前建立的全球随机森林模型进行了比较.
- 分析了PFAS结构空间,以确定影响Fup的关键特征.
主要成果:
- 转移学习在对其他小型化学品家族进行微调时,显示了更高的统计性能.
- 全球随机森林模型由于数据稀疏和不平衡,对PFAS更具竞争力.
- 该研究发现了强大的PFAS Fup建模目前数据可用性的局限性.
- 为扩大知识库,制定了未来实验性特征的建议.
结论:
- 虽然转移学习在数据更丰富的家庭中显示出Fup建模的希望,但它面临着像PFAS这样稀疏的数据集的挑战.
- 鉴于数据限制,目前的全球模型是PFAS Fup预测中最具竞争力的.
- 扩大PFAS的实验数据对于提高先进机器学习方法的可行性至关重要,包括本地和转移学习.
相关概念视频
 01:29
Drug Distribution: Plasma Protein Binding
6.7K
Drugs predominantly attach to plasma proteins, with only a small percentage remaining unbound. The unbound portion can be calculated as one minus the bound fraction. Acidic drugs form large, inactive complexes by reversibly binding to plasma albumin, which prevents them from diffusing across biological barriers. These drug-protein complexes act as reservoirs for the drugs. As the concentration of unbound drugs decreases, these complexes quickly dissociate to release the free drug, maintaining...
6.7K
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
92
Physiological models with protein binding in pharmacokinetics offer a sophisticated approach to understanding drug disposition. These models consider drug-protein interactions, enabling them to effectively predict drug concentrations in different organs and tissues. This precision aids in accurate drug dosing, providing a significant advantage over conventional models. A key process within these models is equilibration, which ensures that drug concentrations achieve a steady state within the...
92
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
83
Drug transporters are critical in drug absorption, distribution, and excretion processes. They should be included in physiological-based pharmacokinetic (PBPK) models, which help predict human drug disposition. However, predicting this is challenging during drug development, especially when liver transport is involved. However, with a realistic representation of body transport processes, an accurate model may be possible.
A recent model describes pravastatin's hepatobiliary excretion,...
A recent model describes pravastatin's hepatobiliary excretion,...
83
 00:54
Noncompartmental Analysis: Miscellaneous Pharmacokinetic Parameters
197
The noncompartmental approach is a widely used method in pharmacokinetics to assess drugs' behaviors in the body. It considers several factors, including clearance, bioavailability, and total volume of distribution.
One key aspect of the noncompartmental approach is determining a drug's total clearance. This can be done by dividing the drug dose by the area under the concentration-time curve from zero to infinity. The area under the concentration-time curve represents the drug's...
One key aspect of the noncompartmental approach is determining a drug's total clearance. This can be done by dividing the drug dose by the area under the concentration-time curve from zero to infinity. The area under the concentration-time curve represents the drug's...
197
 01:09
Hepatic Drug Clearance: Effect of Protein Binding
286
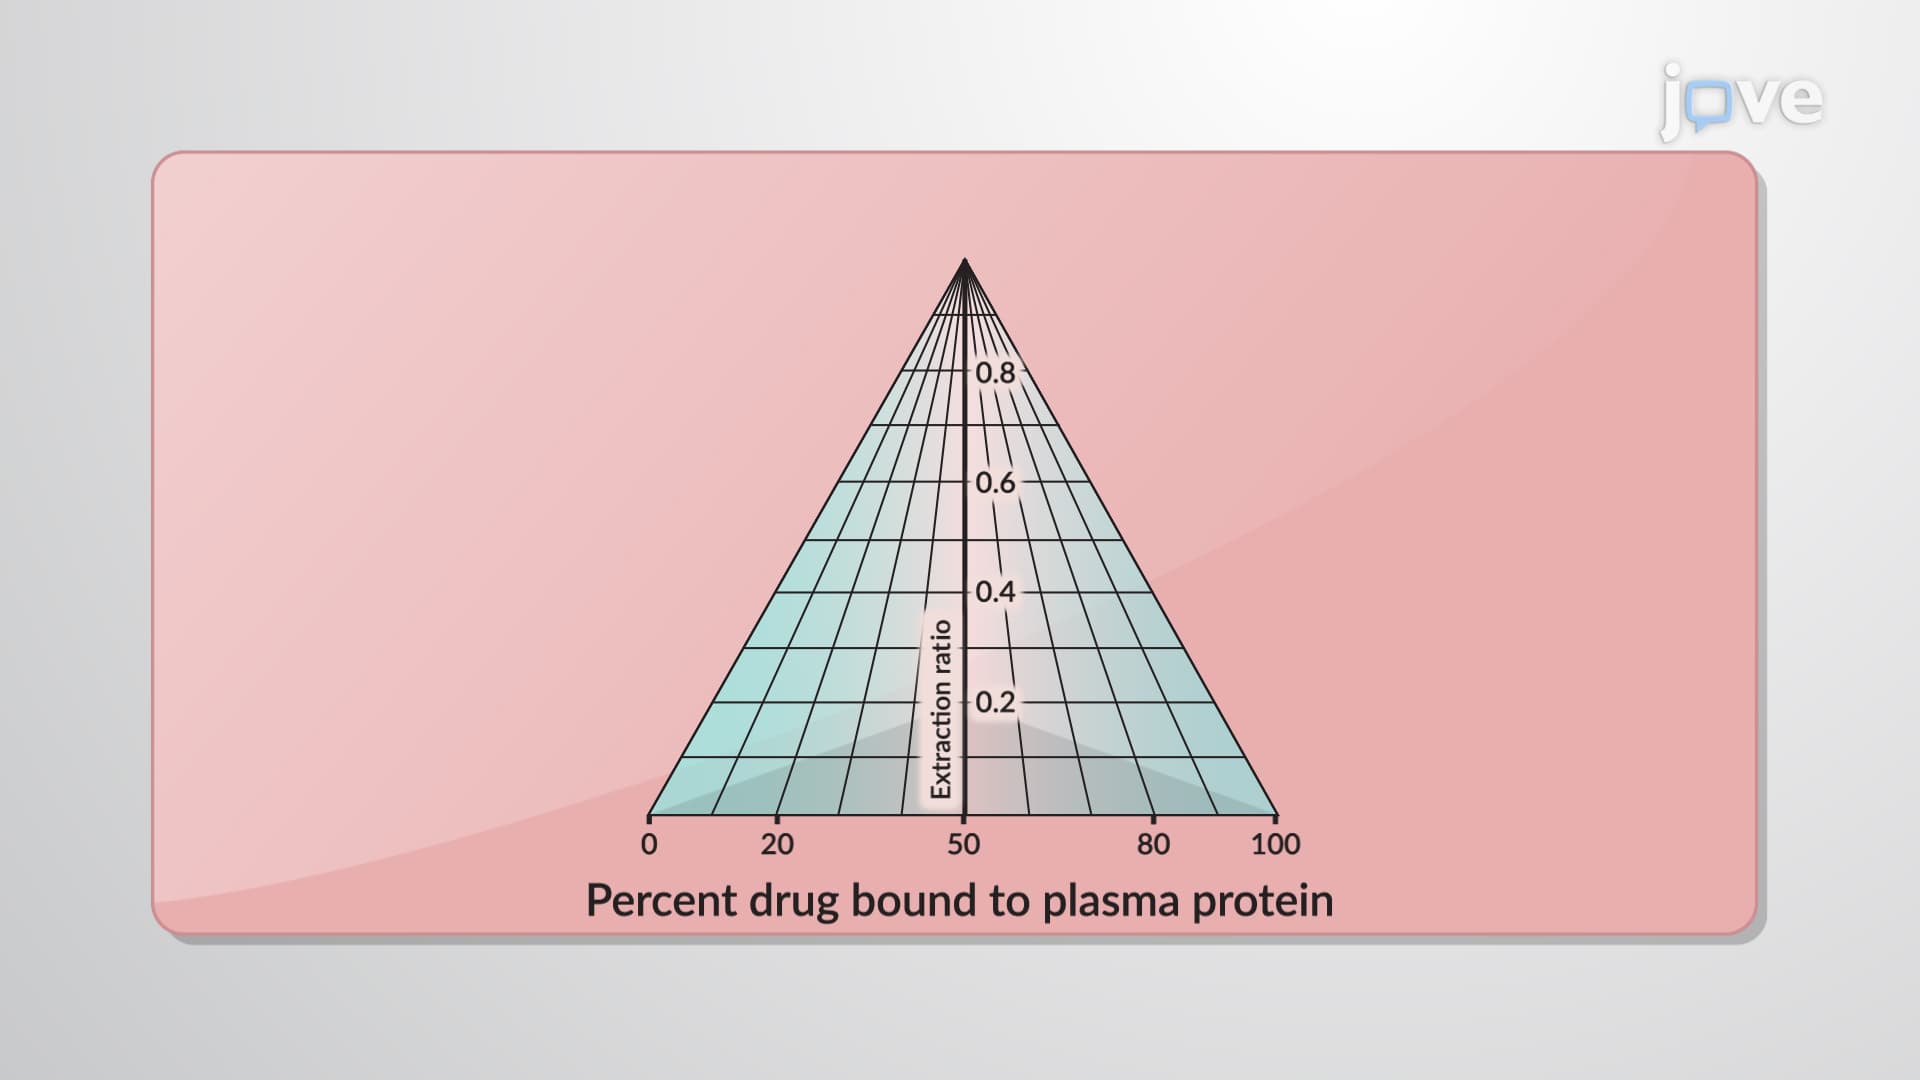
Hepatic clearance is influenced by protein binding based on the drug's extraction ratio. Drugs with high extraction ratios are considered flow-limited and remain unaffected by protein binding during hepatic clearance. On the other hand, drugs with low extraction ratios may be impacted by plasma protein binding, although the extent of this influence depends on the fraction of the drug bound.
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
286
 01:23
Drug Concentrations: Measurements
603
Drug concentration is the quantity of a drug present in a biological sample. Measuring drug amounts in biological samples allows the clinician to understand how a drug is absorbed, distributed, metabolized, and excreted. Samples can be obtained through invasive or non-invasive methods. Invasive techniques involve surgical or parenteral interventions to gather blood, cerebrospinal fluid, or tissue biopsy. Conversely, non-invasive approaches provide samples like urine, feces, and saliva.
Plasma...
Plasma...
603