相关实验视频
Updated: Sep 14, 2025
10:34
Ultra-long Read Sequencing for Whole Genomic DNA Analysis
Published on: March 15, 2019
23.1K
cONcat:从长的牛津纳米孔读取连接碎片的计算重建
Alexander J Petri1,2, Mai Thi-Huyen Nguyen3, Anjali Rajwar3
1Department of Mathematics, Science for Life Laboratory, Stockholm University, Stockholm, Sweden.
PloS one
|July 24, 2025

概括
我们开发了cONcat,这是一个生物信息学工具,用于分析来自组合图书馆的DNA测序读取. cONcat精确地识别了来自牛津纳米孔测序数据的DNA片段组成,尽管它的错误率.
科学领域:
- 生物信息学是一种生物信息学.
- 分子生物学分子生物学
- 基因组学就是基因组学.
背景情况:
- 合成组合DNA库对于蛋白质工程和研究DNA-蛋白质相互作用至关重要.
- 高通量测序,包括牛津纳米孔测序 (ONT),用于质量控制和分析选择实验.
- 目前的生物信息学工具很难从易出错的ONT读取中准确地重建DNA库组成.
研究的目的:
- 开发一种生物信息算法,用于在ONT测序读取中准确识别连接的DNA片段.
- 解决现有工具在分析使用ONT技术生成的复杂DNA库方面的局限性.
主要方法:
- 介绍了cONcat,一种使用基于编辑距离的递归覆盖方法的算法.
- 将已知的DNA片段与ONT测序读数相匹配,以确定库的组成.
- 使用模拟和实验数据与短DNA片段 (<20bp) 的验证.
主要成果:
- cONcat准确地识别了在ONT读取中连接的DNA片段的组成和顺序.
- 该算法表现良好,尽管存在固有的序列错误和ONT数据特征的短片长.
- 在ONT读数开始时的高错误率对有信心的序列覆盖构成了挑战.
结论:
- cONcat提供了一个强大的解决方案,用于分析使用ONT测序的组合DNA库.
- 可能需要实验策略来缓解ONT读取启动错误带来的挑战.
- 这些发现突出了ONT测序用于DNA库分析的潜力和局限性.
相关概念视频
 03:36
Genome Annotation and Assembly
19.3K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
19.3K
 03:21
RNA-seq
10.4K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
10.4K
 01:05
Maxam-Gilbert Sequencing
11.5K
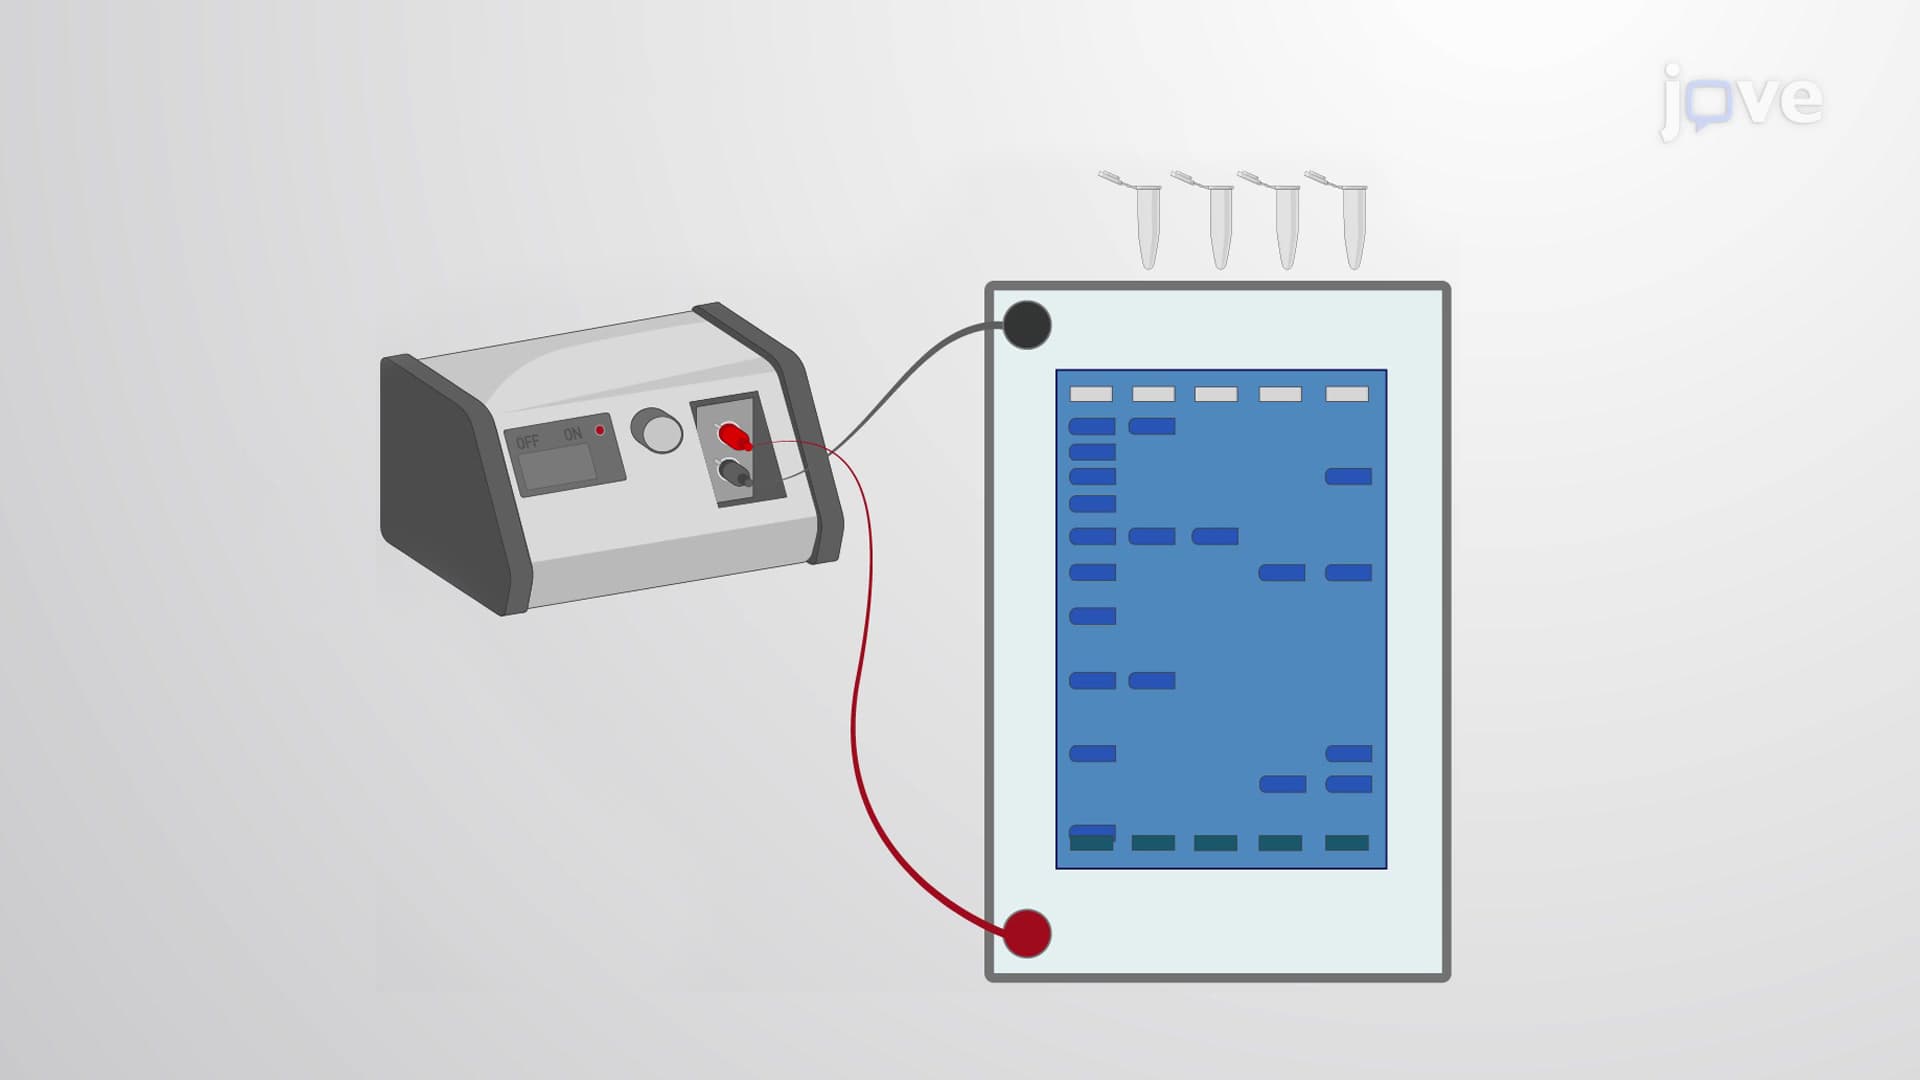
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
11.5K
 03:00
Next-generation Sequencing
92.6K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
92.6K
 01:57
Sanger Sequencing
757.4K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
757.4K