

仅对结合亲和力变化的序列预测:用于抗体工程的强大和可解释的模型
Chen Liu1, Mingchen Li1, Yang Tan1
1School of Information Science and Engineering, East China University of Science and Technology, Shanghai 200237, China.
Bioinformatics (Oxford, England)
|August 12, 2025

概括
一个深度学习模型ProtAttBA仅使用序列数据预测了抗体-抗原结合亲和力变化. 这为抗体工程的传统方法提供了快速,经济高效的替代方案.
科学领域:
- 生物化学 生物化学
- 计算生物学 计算生物学
- 免疫学 免疫学 免疫学
背景情况:
- 增强抗体-抗原结合亲和力对于抗体工程至关重要.
- 评估结合亲和力的传统实验方法昂贵且耗时.
- 当前的深度学习模型通常需要复杂的结构数据,而这些数据并不总是可用.
研究的目的:
- 开发一种深度学习模型,ProtAttBA,用于预测抗体-抗原结合亲和力变化.
- 创建一个仅依赖序列信息的模型,克服结构依赖方法的局限性.
- 为抗体工程提供一个快速且具有成本效益的计算工具.
主要方法:
- ProtAttBA利用一个预训练阶段来学习蛋白质序列模式.
- 监督训练阶段使用标记的抗体-抗原复合数据.
- 一个基于交叉注意力的回归器被训练来预测结合亲和力变化.
主要成果:
- 与基于序列和结构的方法相比,ProtAttBA在开放基准上取得了竞争性表现.
- 该模型表现出强度,特别是在不确定的复杂结构中.
- 注意机制提供了可解释性,识别了影响结合亲和力的关键残留物.
结论:
- ProtAttBA为抗体工程提供了一个快速,具有成本效益的计算工具.
- 该模型可以加速新型治疗抗体的开发.
- 基于序列的结合亲和力的预测是基于结构的方法的可行和强大的替代方案.
相关概念视频
 01:49
Conserved Binding Sites
4.4K
Many proteins’ biological role depends on their interactions with their ligands, small molecules that bind to specific locations on the protein known as ligand-binding sites. Ligand-binding sites are often conserved among homologous proteins as these sites are critical for protein function.
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
4.4K
 02:18
The Equilibrium Binding Constant and Binding Strength
13.4K
The equilibrium binding constant (Kb) quantifies the strength of a protein-ligand interaction. Kb can be calculated as follows when the reaction is at equilibrium:
13.4K
 01:58
Cooperative Allosteric Transitions
8.0K
Cooperative allosteric transitions can occur in multimeric proteins, where each subunit of the protein has its own ligand-binding site. When a ligand binds to any of these subunits, it triggers a conformational change that affects the binding sites in the other subunits; this can change the affinity of the other sites for their respective ligands. The ability of the protein to change the shape of its binding site is attributed to the presence of a mix of flexible and stable segments in the...
8.0K
 02:40
Ligand Binding Sites
13.2K
Proteins are dynamic macromolecules that carry out a wide variety of essential processes; however, the activities of most proteins depend on their interactions with other molecules or ions, known as ligands.
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
13.2K
 01:22
Protein-Drug Binding: Determination Methods
304
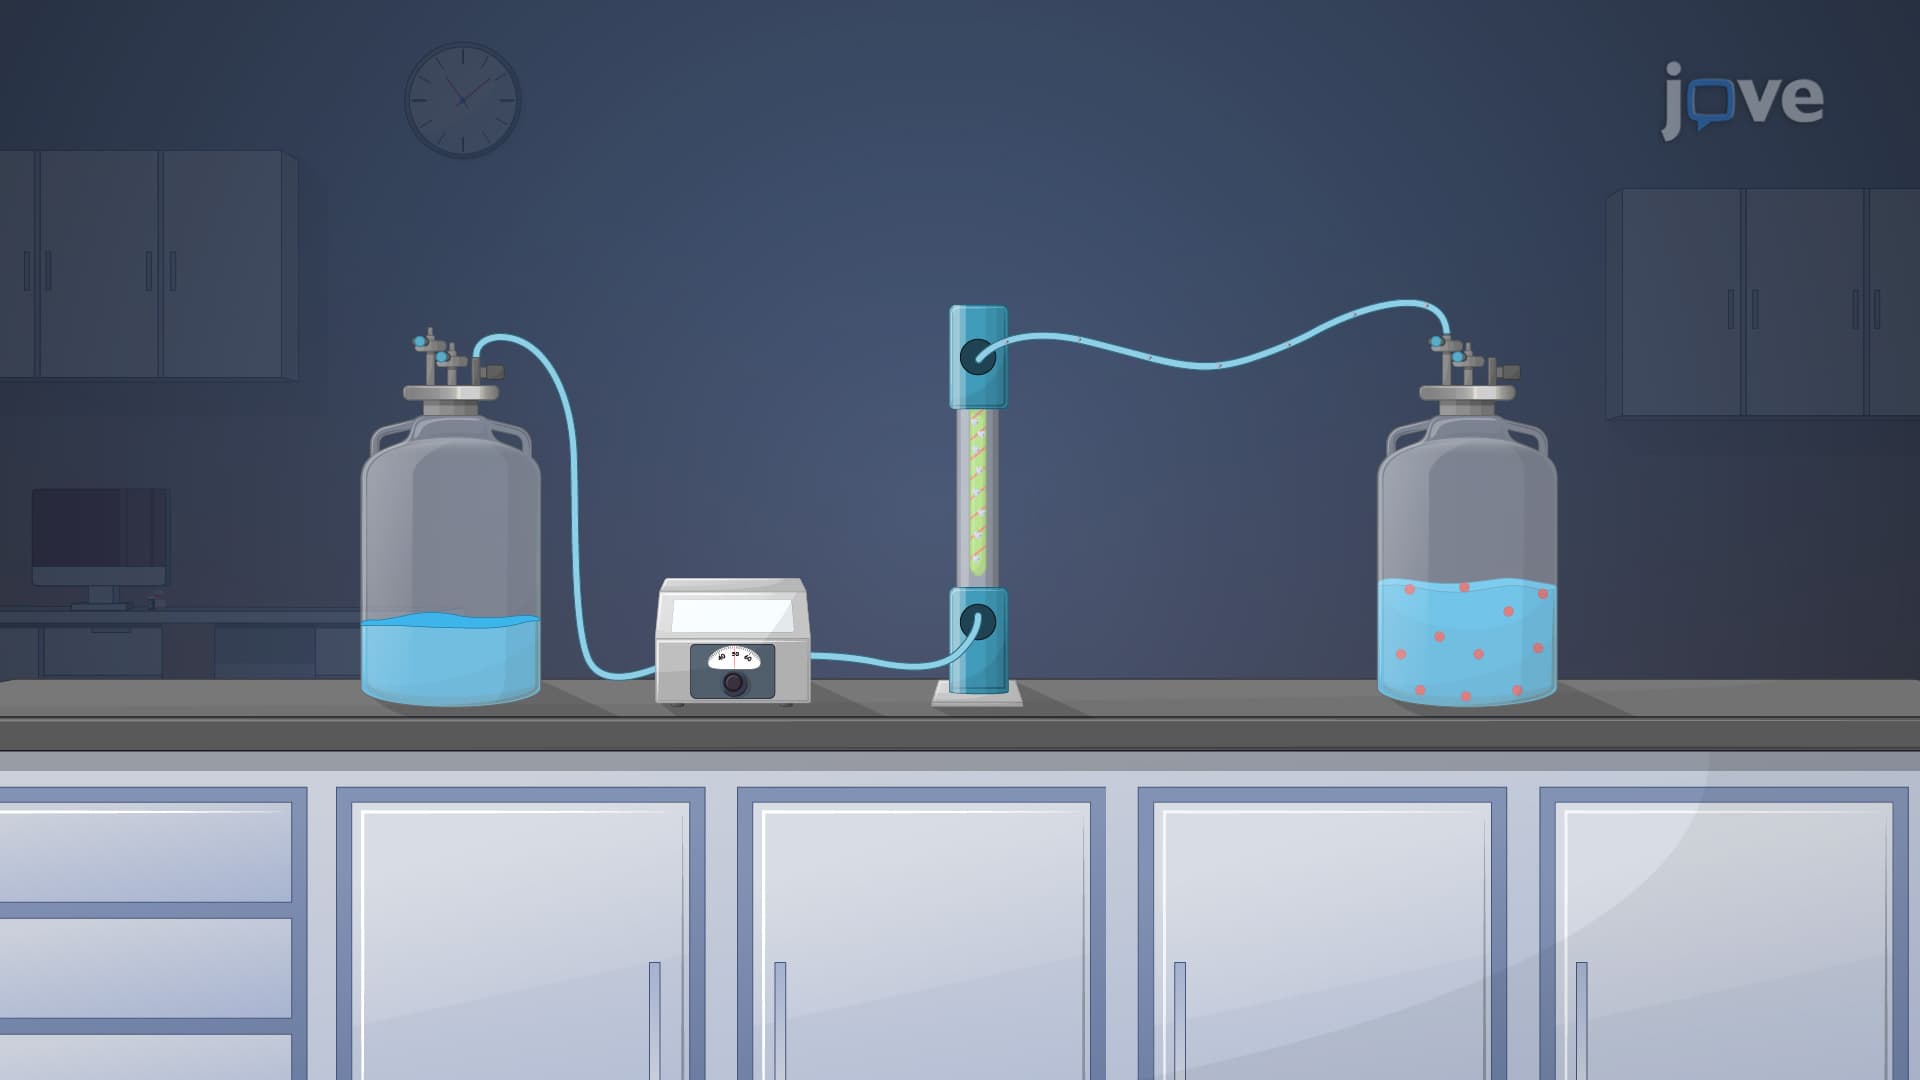
Determining protein-drug binding can be achieved through indirect and direct methods, each providing valuable insights into the interaction between proteins and drugs.
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
304