

基于激素的药解体建模和量子计算方法,用于开发hCA XII抑制剂
IEEE transactions on computational biology and bioinformatics
|August 14, 2025

概括
新的研究确定了新型化合物3和26作为碳酸无水酶XII (CA XII) 的强有力的抑制剂,这是缺氧瘤的关键标. 这些有前途的线索为癌症治疗提供了潜在的新疗法策略.
科学领域:
- 生物化学 生物化学
- 药用化学 医学化学
- 计算化学的计算化学
背景情况:
- 癌症仍然是一个重大的全球健康威胁,耐药性构成了主要挑战.
- 向特定的碳酸 anhydrase (CA) 异型,如CA XII,是癌症治疗的一个有前途的策略.
- 在各种缺氧瘤中,CA XII被认为是生物标志物,因此需要开发选择性抑制剂.
研究的目的:
- 为了发现新的碳酸无水酶XII (CA XII) 抑制剂.
- 为了超越传统的硫胺基基架.
- 为了确保对其他CA异型的选择性.
主要方法:
- 现有分子的计算选.
- 药模拟和虚拟选以识别被击中化合物.
- 在制药物动力学 (ADMET) 和分子动力学模拟中.
- 分子对接,DFT和MMGBSA计算用于稳定性和结合亲和力的评估.
主要成果:
- 一个药模型产生了43个命中分子,所有这些都通过了ADMET的预测.
- 化合物3和26通过分子对接显示出强大的结合亲和力和低能量构造 (-7.82 K/Cal和 -7.36 K/Cal).
- 分子动力学模拟表明,3和26化合物与乙胺相比具有更高的稳定性.
- 获得了针对CA XII的抑制机制的结构性见解.
结论:
- 化合物3和26代表了CA XII抑制的有希望的新候选者.
- 这些化合物为二硫胺胺提供了潜在的替代性支架.
- 对化合物3和26的进一步开发可能会导致新的抗癌疗法.
更多相关视频
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
1.5K
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
357
相关概念视频
 02:40
Ligand Binding Sites
13.2K
Proteins are dynamic macromolecules that carry out a wide variety of essential processes; however, the activities of most proteins depend on their interactions with other molecules or ions, known as ligands.
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
13.2K
 01:26
Drug Discovery: Overview
8.7K
Drug discovery is a multifaceted process involving extensive screening, testing, and optimization of lead compounds to identify potential new drugs for therapeutic use. It combines several approaches, including screening large numbers of natural products, chemical modification of known active molecules, identification of new drug targets, and rational design based on biological mechanisms and drug-receptor structure. These approaches are carried out in both academic research laboratories and...
8.7K
 01:28
Structure-Activity Relationships and Drug Design
1.0K
Drug design is a dynamic field that involves discovering and developing new medications based on specific biological targets. This process heavily relies on structure-activity relationships (SAR) and quantitative structure-activity relationships (QSAR) to guide the design and optimization of efficient drugs.
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
1.0K
 01:15
G Protein-coupled Receptors
13.4K
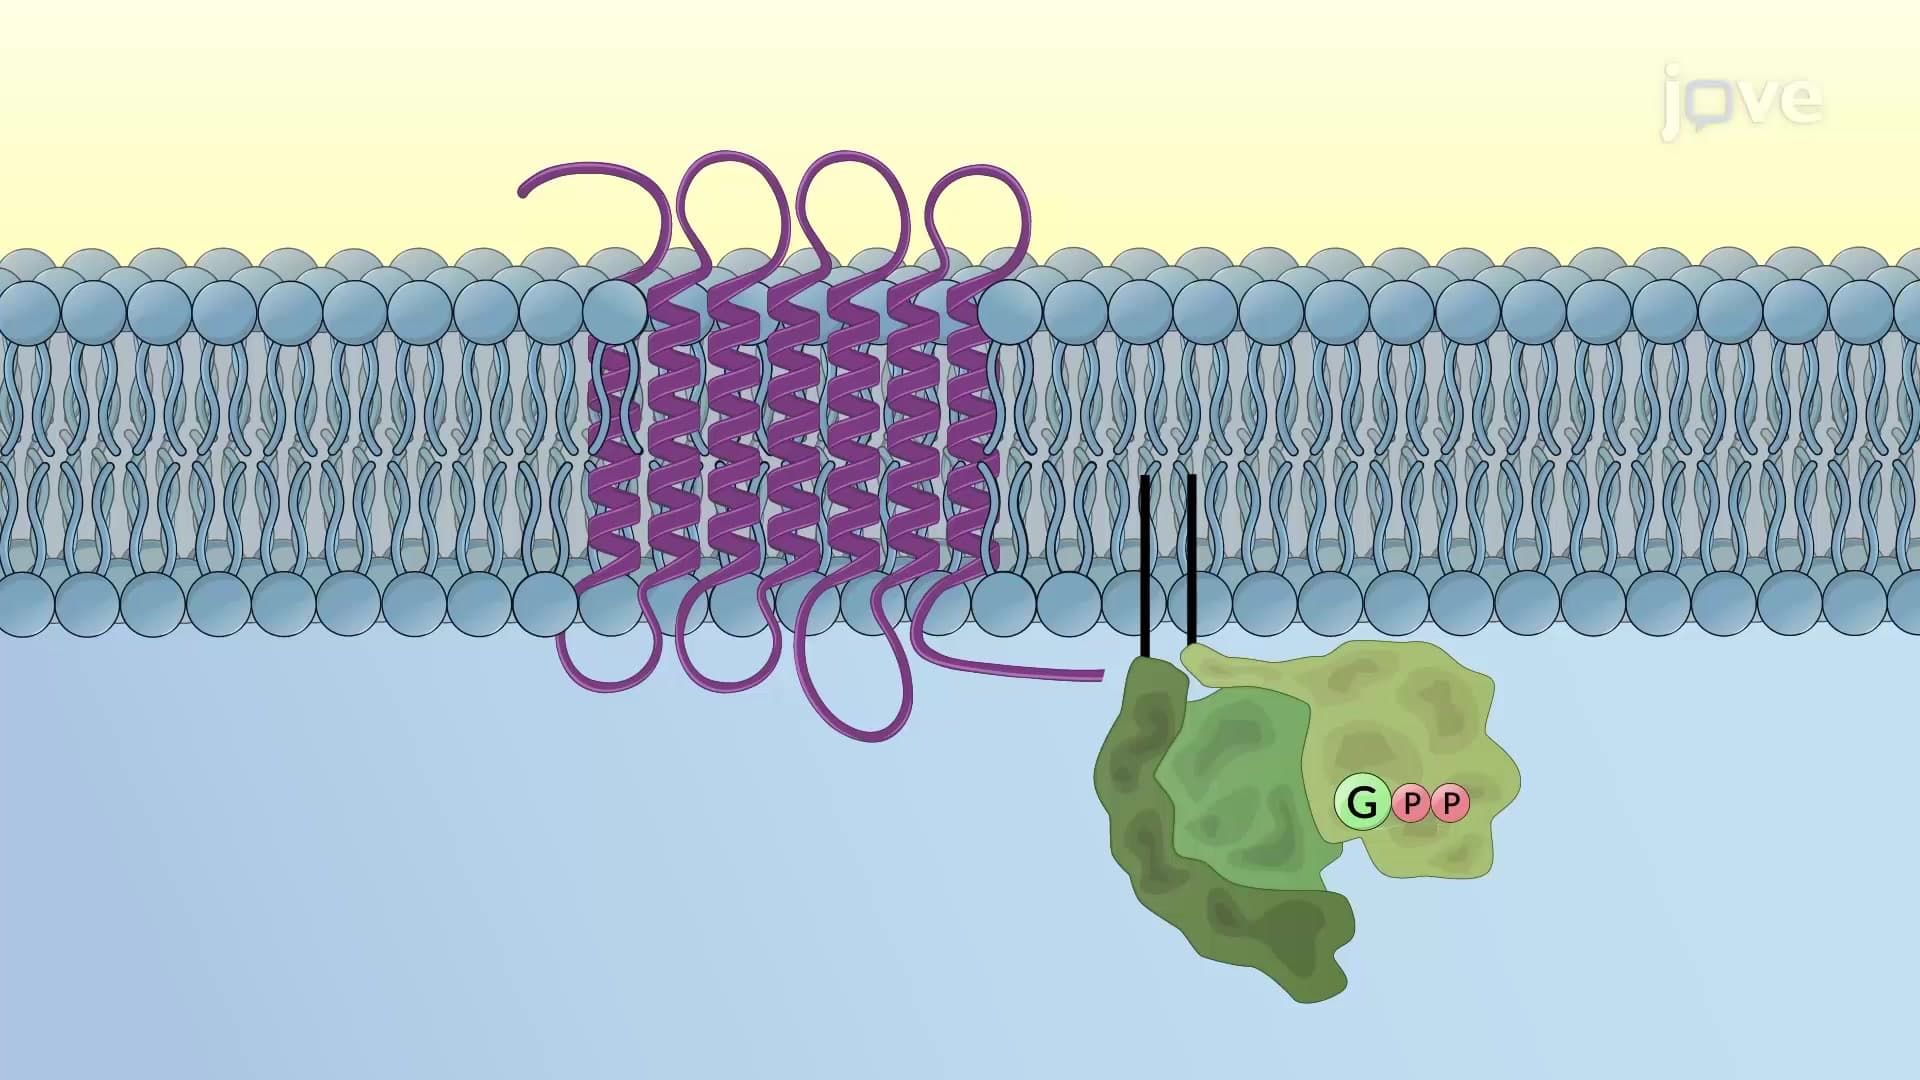
G Protein-Coupled Receptors or GPCRs are membrane-bound receptors that transiently associate with heterotrimeric G proteins and induce an appropriate response to sensory stimuli such as light, odors, hormones, cytokines, or neurotransmitters.
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
13.4K