
从测序数据中直接推断单元型
Zhen Zhang1,2,3, Bencong Zhu3, Yongyi Luo3
1The First Institute, Kunming Institute of Physics, Kunming, Yunnan Province, 650223, China.
Bioinformatics advances
|September 22, 2025

概括
这项研究引入了一个统一的概率框架,用于直接从测序读数中复原型重建. 该方法通过对测序错误进行建模,提高了复杂场景 (如多重积分和低覆盖数据) 的准确性.
科学领域:
- 基因组学就是基因组学.
- 生物信息学是一种生物信息学.
背景情况:
- 哈普洛型重建对于遗传分析至关重要,但在当前的两阶段方法 (变体调用和分阶段) 中具有挑战性,这些方法忽略了变体调用错误.
- 随着同类染色体数量的增加,哈普洛型重建的复杂性增加,这在多倍体物种或细胞混合物中很常见.
研究的目的:
- 从测序数据中开发一个统一的概率框架,用于从测序数据中直接重建单元型.
- 提高单 haplotype 推断的准确性,特别是在低覆盖率和高错误率的数据集以及多倍体样本中.
主要方法:
- 一个统一的概率框架直接利用测序读数来估计单元类型和测序错误概况.
- 建模所有被测序读取的位点,而不仅仅是变量位点,以提高错误概况估计.
主要成果:
- 与传统分阶段方法相比,拟议的方法在模拟和真实测序数据上表现出优异的性能.
- 在具有高序列错误率,低覆盖率和多重积分的场景中观察到显著的改善.
结论:
- 统一的框架提供了一个更强大,更准确的方法来复原型重建.
- 这种方法增强了哈普洛型推断的统计能力,特别是在具有挑战性的基因组环境中.
相关概念视频
 01:11
Genome-wide Association Studies-GWAS
15.3K
Genome-wide association studies or GWAS are used to identify whether common SNPs are associated with certain diseases. Suppose specific SNPs are more frequently observed in individuals with a particular disease than those without the disease. In that case, those SNPs are said to be associated with the disease. Chi-square analysis is performed to check the probability of the allele likely to be associated with the disease.
GWAS does not require the identification of the target gene involved in...
GWAS does not require the identification of the target gene involved in...
15.3K
 02:54
Evolutionary Relationships through Genome Comparisons
6.9K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
6.9K
 01:57
Sanger Sequencing
773.3K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
773.3K
 03:00
Next-generation Sequencing
97.8K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
97.8K
 01:05
Single Nucleotide Polymorphisms-SNPs
17.9K
A single nucleotide polymorphism or SNP is a single nucleotide variation at a specific genomic position in a large population. It is the most prevalent type of sequence variation found in the human genome. Point mutations that occur in more than 1% of the population qualify as SNPs. These are present once every 1000 nucleotides on an average in the human genome. Replacement of a purine with another purine (A/G) or a pyrimidine with another pyrimidine (C/T) is known as a transition. In contrast,...
17.9K
 01:05
Maxam-Gilbert Sequencing
12.6K
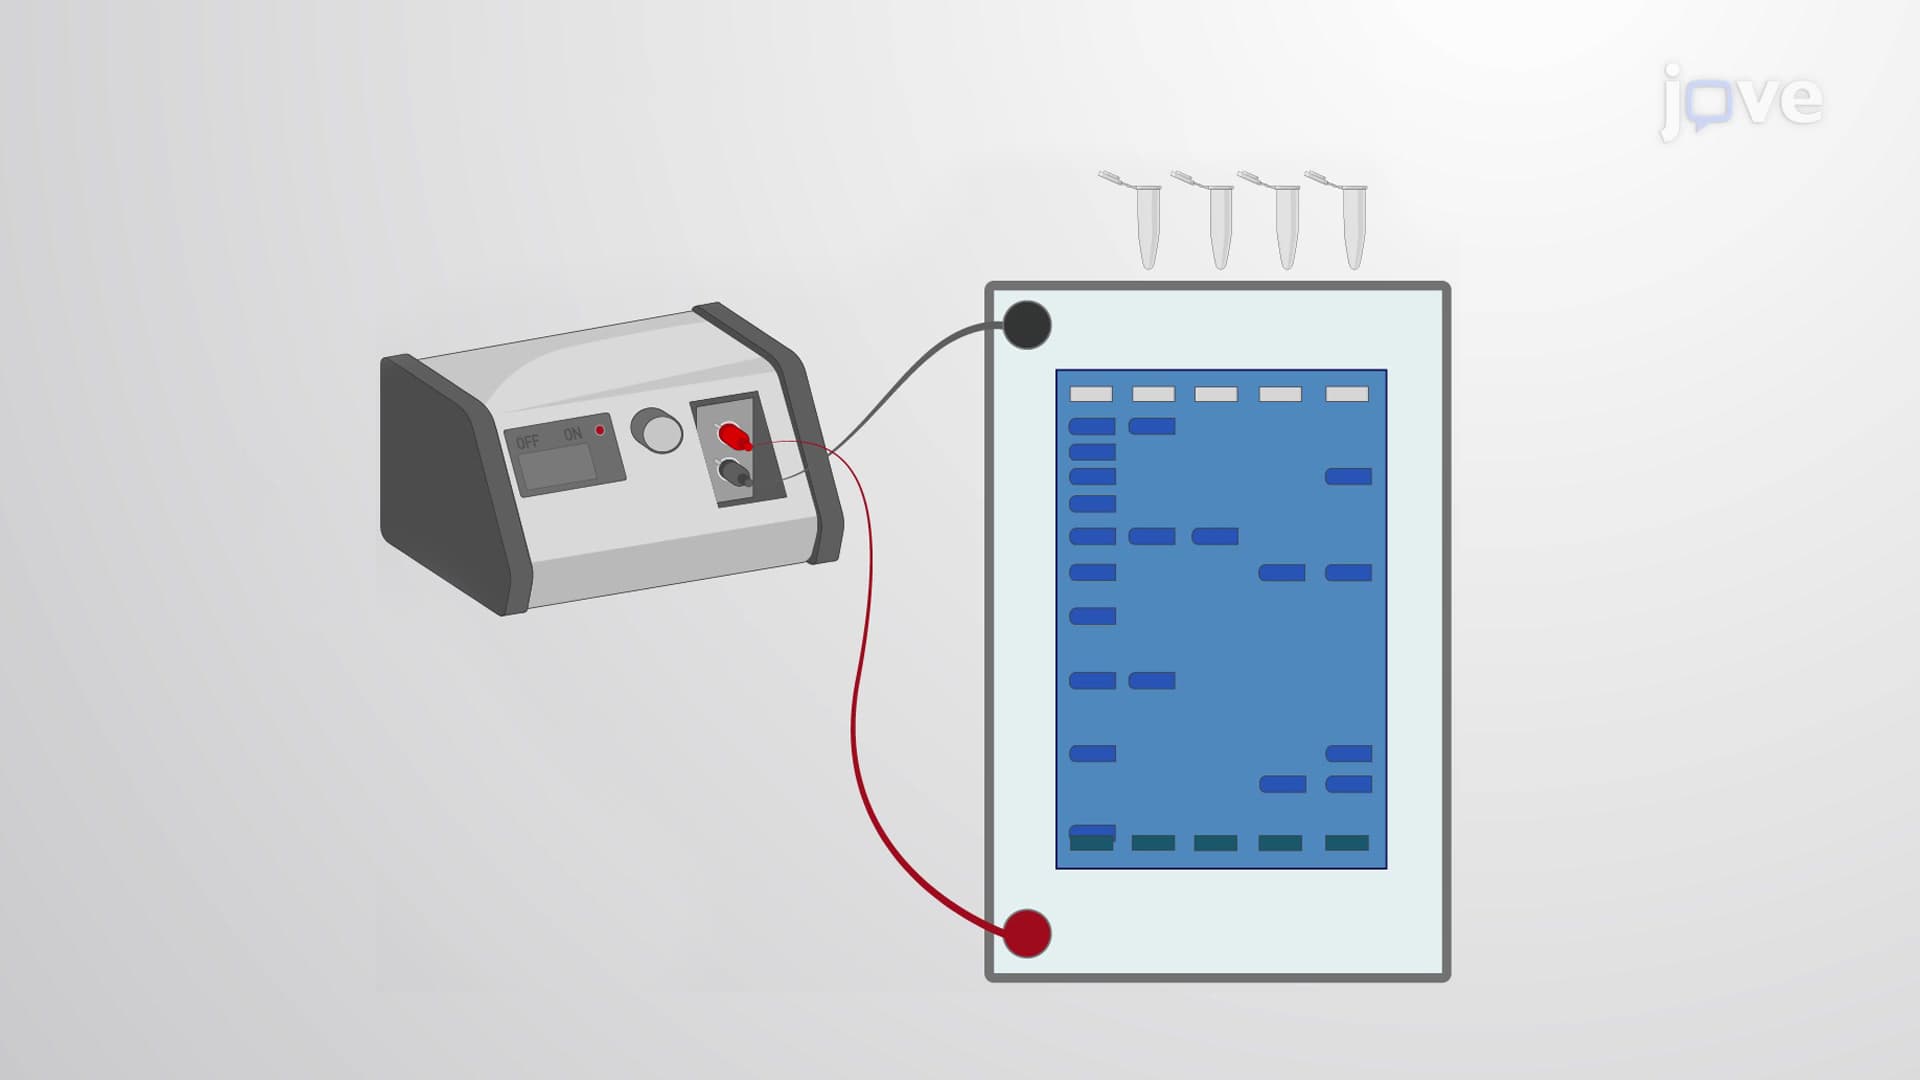
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
12.6K