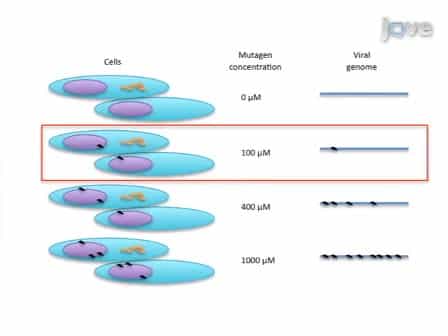
病毒基因组序列数据集显示了对特定链的替代偏差的普遍证据,这些偏差最好使用不可逆转的核酸替代模型来描述
Rita Sianga-Mete1, Penelope Hartnady1, Wimbai Caroline Mandikumba1
1Division of Computational Biology, Institute of Infectious Diseases and Molecular Medicine, Department of Integrative Biomedical Sciences, Faculty of Health Sciences, University of Cape Town, Rondebosch, South Africa.
eLife
|September 30, 2025

概括
非可逆核酸替代模型,特别是12速率模型 (NREV12),惊人地改善了病毒基因组的基因树适应性. 然而,这些模型对于常规的家族遗传推断并不必不可少,尽管通过增加非可逆性提高了准确性.
科学领域:
- 进化生物学是进化的生物学.
- 人类遗传学 是一个学科.
- 分子进化的分子进化.
背景情况:
- 遗传学树通常使用时间可逆模型推断,假设核酸对之间的对称替代率.
- 生物化学突变过程可能不对称,挑战时间可逆模型的假设.
研究的目的:
- 引入和评估两个不可逆转的核酸替代模型:双链基因组的NREV6和单链基因组的NREV12.
- 使用各种基因组数据集,比较非可逆模型与一般时间可逆 (GTR) 模型的合适性.
- 评估非可逆模型对遗传树推断准确性的影响.
主要方法:
- 开发了两个不可逆转的核酸替代模型:NREV6 (6率) 和NREV12 (12率).
- 应用概率比率和Akaike信息标准测试来比较dSRNA,dSDNA,ssDNA和ssRNA数据集中的模型匹配.
- 模拟不同程度的非可逆性 (DNRs) 用NREV12和GTR模型来测试家族遗传推断的准确性.
主要成果:
- 对于大多数dsRNA和dsDNA数据集,NREV12模型显示出比GTR和NREV6更好的匹配.
- 正如预期的那样,NREV12为大多数ssDNA和ssRNA数据集提供了显著更好的匹配.
- 增加的非可逆性提高了NREV12和GTR的族系树拓推断精度,但降低了分支长度的精度.
结论:
- 非可逆模型,特别是NREV12,为许多病毒物种的突变过程提供了改进的分析.
- 尽管在某些方面准确度有所提高,但不可逆转的模型不被认为是常规的家族遗传推断的必要条件.
- 这项研究强调了在遗传学分析中考虑模型对称性的重要性,特别是在病毒进化方面.
相关概念视频
 00:36
Viral Mutations
39.5K
A mutation is a change in the sequence of bases of DNA or RNA in a genome. Some mutations occur during replication of the genome due to errors made by the polymerase enzymes that replicate DNA or RNA. Unlike DNA polymerase, RNA polymerase is prone to errors because it is not capable of “proofreading” its work. Viruses with RNA-based genomes, like HIV, therefore accrue mutations faster than viruses with DNA-based genomes. Because mutation and recombination provide the raw material...
39.5K
 01:26
Size and Structure of Viral Genomes
606
Viral genomes exhibit remarkable diversity in size, structure, and composition, influencing their replication strategies and interactions with host cells. These genomes consist of either DNA or RNA and may be linear or circular. Additionally, they can be single-stranded or double-stranded, with each configuration affecting how the virus propagates within a host. RNA viruses, for instance, generally have smaller genomes than DNA viruses, a factor that contributes to their high mutation rates and...
606
 00:57
Viral Recombination
24.8K
Cells are sometimes infected by more than one virus at once. When two viruses disassemble to expose their genomes for replication in the same cell, similar regions of their genomes can pair together and exchange sequences in a process called recombination. Alternatively, viruses with segmented genomes can swap segments in a process called reassortment.
24.8K
 01:29
Viruses with RNA Genomes
698
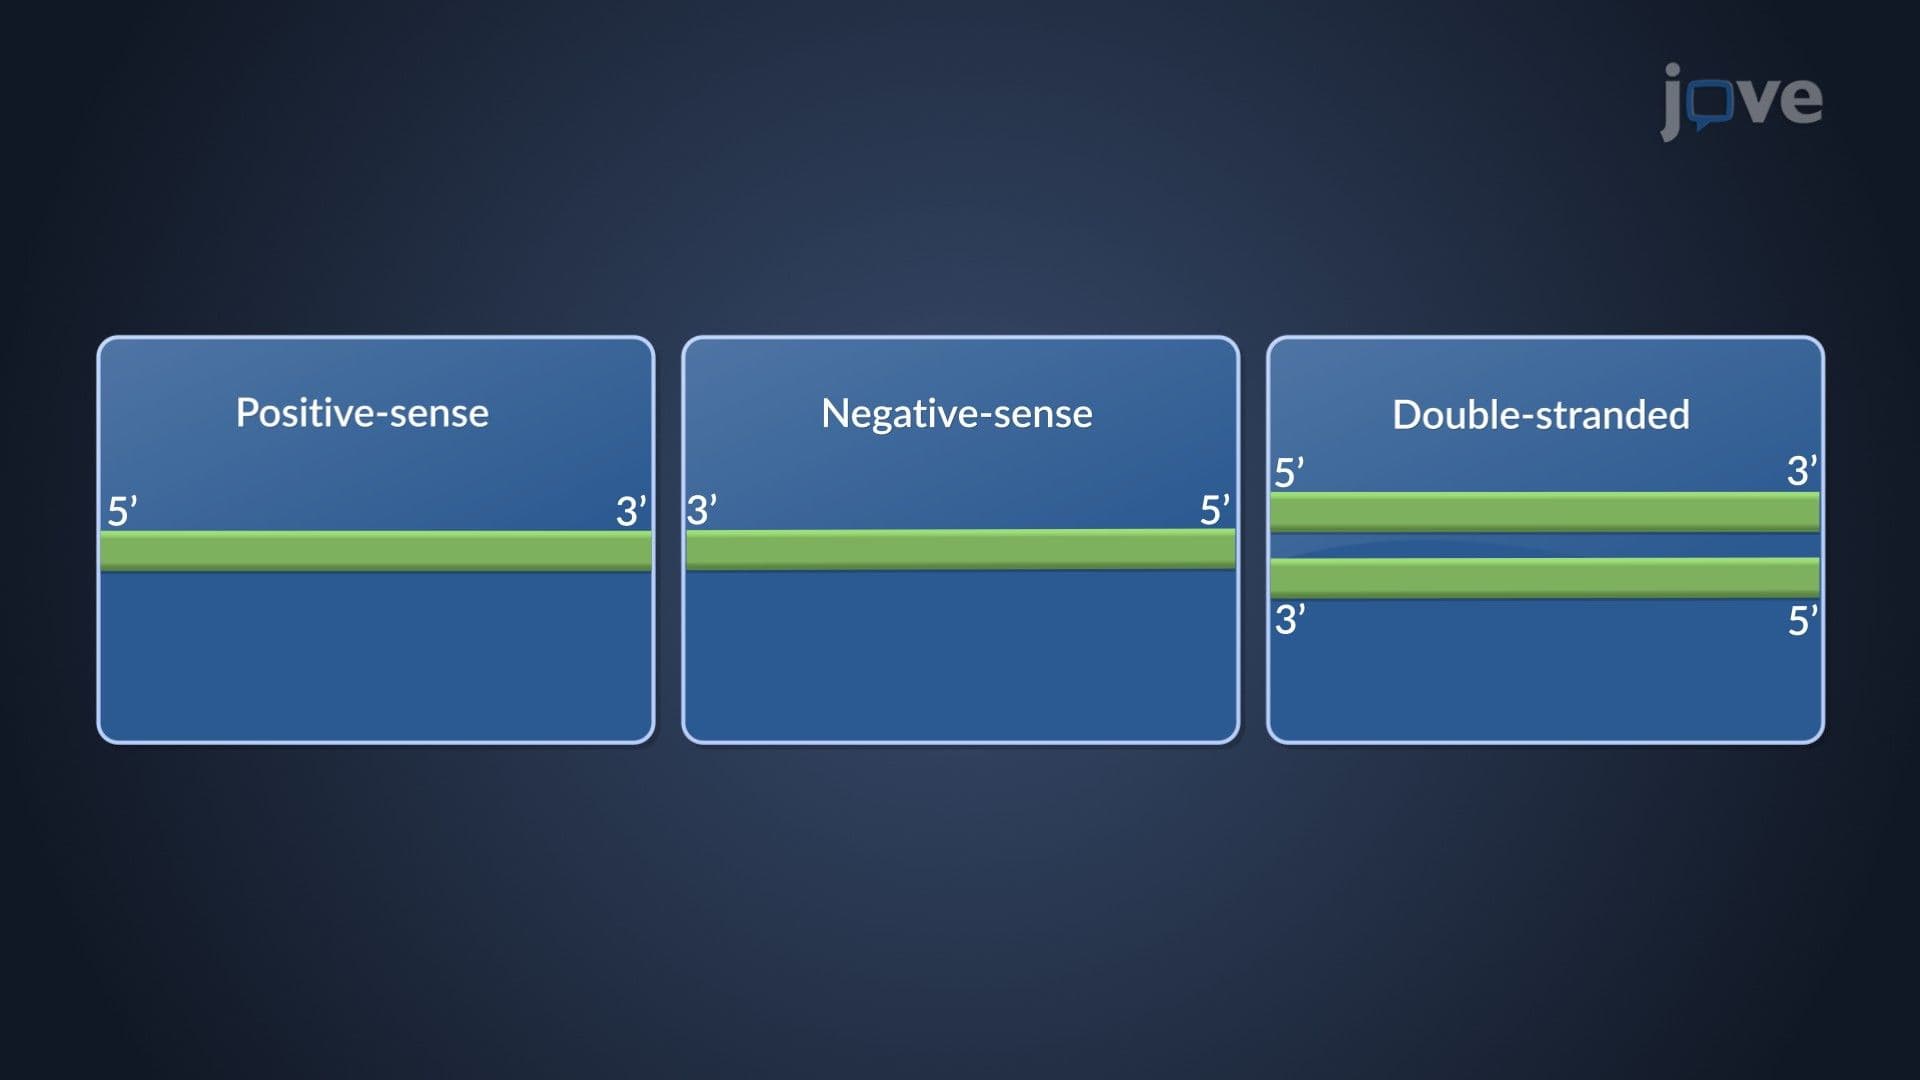
RNA viruses are categorized into positive-strand, negative-strand, or double-stranded groups based on their genomic structure and replication mechanisms. This classification dictates how they exploit host cellular machinery for protein synthesis and replication. Some RNA viruses also utilize reverse transcription as part of their life cycle, further diversifying their replication strategies.Positive-Strand RNA VirusesPositive-strand RNA viruses have genomes that function directly as messenger...
698
 02:05
Gene Evolution - Fast or Slow?
7.9K
The genomes of eukaryotes are punctuated by long stretches of sequence which do not code for proteins or RNAs. Although some of these regions do contain crucial regulatory sequences, the vast majority of this DNA serves no known function. Typically, these regions of the genome are the ones in which the fastest change, in evolutionary terms, is observed, because there is typically little to no selection pressure acting on these regions to preserve their sequences.
In contrast, regions which code...
In contrast, regions which code...
7.9K
 02:33
Retroviruses
14.5K
Retroviruses and retrotransposons both insert copies of their genetic elements into the genome of the host cell. Thus, the viral genes are passed on when the host genome is replicated or translated. A typical retroviral DNA sequence contains 3-4 genes that encode the different proteins required for its structural assembly and function as a molecular parasite. This DNA is transcribed into a single mRNA, which is very similar in structure to conventional mRNAs, i.e., it is capped at the 5’...
14.5K