
概率引导的动态聚变多任务 (PDFM) 框架用于质谱分类
Yinchu Wang1,2,3, Wei Zhang1,2,3, Zilong Liu1,2,3
1Center for Metrology Scientific Data and Energy Metrology, National Institute of Metrology, Beijing 100029, China.

本研究介绍了PDFM,这是一个用于质谱 (MS) 数据分类的新型深度学习框架. PDFM提高了准确性和稳定性,改善了生物医学和临床诊断中的罕见类别识别.
科学领域:
- 生物医学数据分析
- 计算生物学是一种计算生物学.
- 机器学习用于质谱学.
背景情况:
- 传统的深度学习模型由于确定性特征和固定的融合而与质谱 (MS) 数据扎.
- 局限性包括单任务优化,阻碍精确的MS数据分类.
研究的目的:
- 提出PDFM,一个渐进的深度学习框架,用于增强MS数据分类.
- 提高MS数据集中的精度,稳定性和罕见类别的识别.
主要方法:
- 使用变量自编码器 (VAE) 和具有动态权重 (TDW) 架构的变压器实现的 PDFM 框架.
- VAE指导注意力峰值;动态权重融合全球 (VAE) 和地方 (变压器) 特性.
- 多目标损失函数集成了分类,重建和分布对齐,以提高稳定性.
主要成果:
- 在没有批量效应的数据集上实现了4.73%的准确度增长.
- 在分类准确性方面,跨批改进了3.49%-4.66%.
- 在小样本大小中,在罕见类别中,F1得分提升了44.07%.
结论:
- PDFM为质谱数据分析提供了一种新而精确的方法.
- 该框架显示了在生物医学和临床诊断中推进翻译应用的巨大潜力.
更多相关视频
 10:37
Deep Proteome Profiling by Isobaric Labeling, Extensive Liquid Chromatography, Mass Spectrometry, and Software-assisted Quantification
Published on: November 15, 2017
 12:11
Simultaneous Affinity Enrichment of Two Post-Translational Modifications for Quantification and Site Localization
Published on: February 27, 2020
相关概念视频
 01:21
Tandem Mass Spectrometry
 01:21
Mass Spectrometry: Complex Analysis
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
 01:19
MALDI-TOF Mass Spectrometry
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:19
Mass Spectrometry: Overview
 01:16
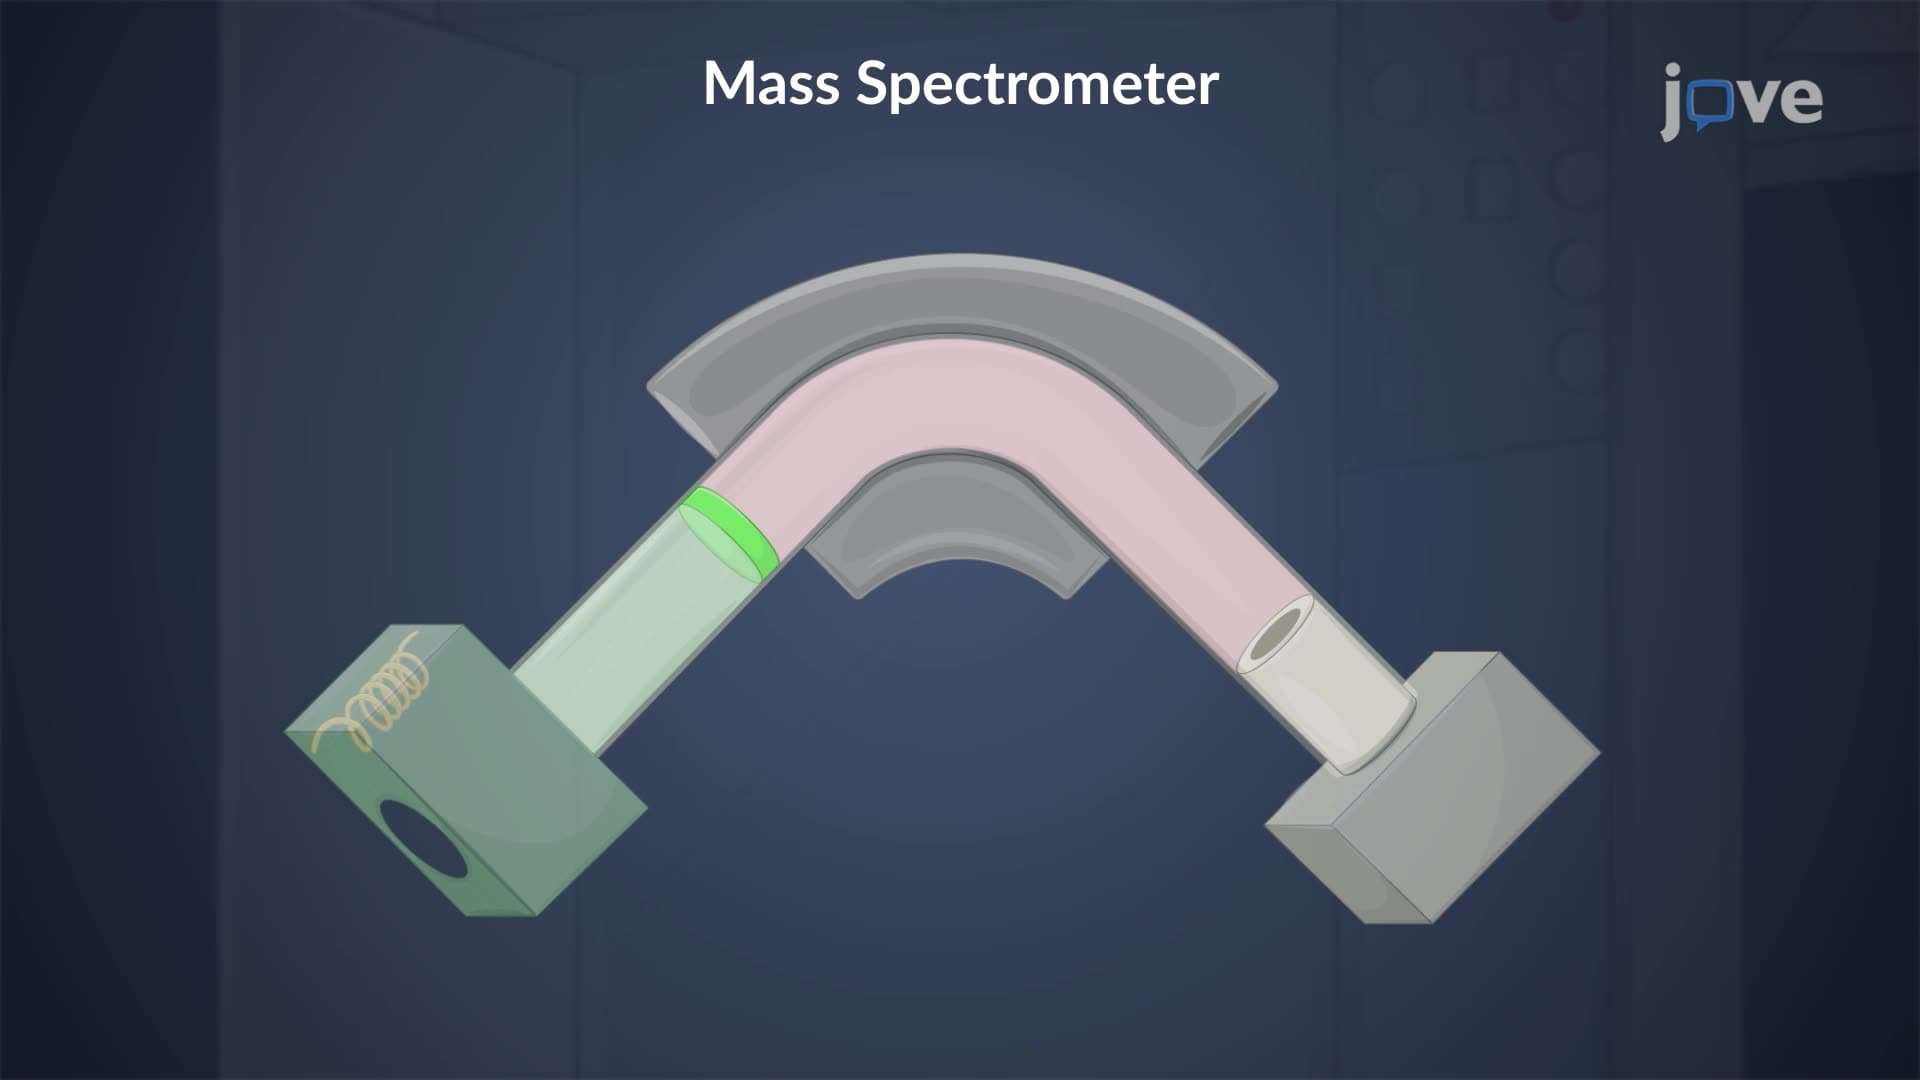
Mass Spectrometers