
分子结构,DFT计算,以及对含有1,2,3-triazole和4-bromophenyl部分的imidazo[1,2-a]pyridine衍生物的对接研究
Corneliu Cojocaru1, Mihaela Balan-Porcăraşu2, Gheorghe Roman3
1Department of Inorganic Polymers, Petru Poni Institute of Macromolecular Chemistry, 41A Grigore Ghica Vodă Alley, 700487, Iaşi, Romania. cojocaru.corneliu@icmpp.ro.
Journal of computer-aided molecular design
|October 24, 2025

概括
这项研究使用计算化学来分析一种imidazo[1,2-a]pyridine衍生物 (IPD). 理论计算与实验数据保持一致,揭示了抗癌药物开发的关键电子特性和潜在相互作用.
科学领域:
- 计算化学是一种计算化学.
- 药品化学 药品化学 是一个
- 药物发现 药物发现
背景情况:
- 伊米达佐[1,2-a]氨酸衍生物是药物化学中的重要支架.
- 了解这些化合物的电子和结构性质对于药物设计至关重要.
研究的目的:
- 在imidazo[1,2-a]pyridine衍生物IPD上进行理论研究.
- 将计算结果与实验数据进行比较 (X射线晶体学,光谱学).
- 通过分子对接模拟来探索潜在的抗癌活性.
主要方法:
- 密度函数理论 (DFT) 用于几何优化和电子属性分析.
- 分子对接模拟以研究与β-cyclodextrin和EGFR-TK的相互作用.
- 对静电电位图和质子亲和力的分析.
主要成果:
- DFT优化的几何学与实验性X射线结晶学数据有很好的一致性.
- 静电电位图确定了潜在的电友攻击地点.
- 预测imidazo[1,2-a]pyridine原子是最有可能发生质子化的地点.
- 分子对接揭示了EGFR-TK的相互作用,表明了潜在的抗癌应用.
- 在对接的复合体中的分子间相互作用涉及边境分子轨道合.
结论:
- 该研究成功地与IPD的实验数据对计算方法进行了验证.
- 理论见解为了解IPD的反应和相互作用提供了基础.
- 这些发现支持IPD作为向EGFR-TK的抗癌剂的潜力.

相关概念视频
 01:25
Basicity of Heterocyclic Aromatic Amines
6.9K
Heterocyclic amines, where the N atom is a part of an alicyclic system, are similar in basicity to alkylamines. Interestingly, the heterocyclic amine having a nitrogen atom as part of an aromatic ring has much less basicity than its corresponding alicyclic counterpart. For this reason, as presented in Figure 1, piperidine (pKb = 2.8) is significantly more basic than pyridine (pKb = 8.8).
6.9K
 01:13
Five-Membered Heterocyclic Aromatic Compounds: Overview
5.3K
Heterocyclic aromatic compounds are cyclic compounds that are aromatic and have one or more heteroatoms—atoms other than carbon, in the ring. Depending upon the number of atoms present in the ring, they can be either five or six-membered. Examples of five-membered heterocyclic aromatic compounds include pyrrole, furan, thiophene, and imidazole. Pyrrole consists of one nitrogen atom having one lone pair of electrons. Furan and thiophene have one oxygen and one sulfur heteroatom,...
5.3K
 02:27
Predicting Molecular Geometry
44.9K
VSEPR Theory for Determination of Electron Pair Geometries
44.9K
 01:18
Structure of Benzene: Molecular Orbital Model
12.0K
According to the molecular orbital (MO) model, benzene has a planar structure with a regular hexagon of six sp2 hybridized carbons. As shown in Figure 1, each carbon is bonded to three other atoms with C–C–C and H–C–C bond angles of 120°. The C–H bond length is 109 pm, and the C–C bond length is 139 pm which is midway between the single bond length of sp3 hybridized carbons (154 pm) and sp2 hybridized carbons (133 pm).
12.0K
 01:19
Structure of Amines
3.2K
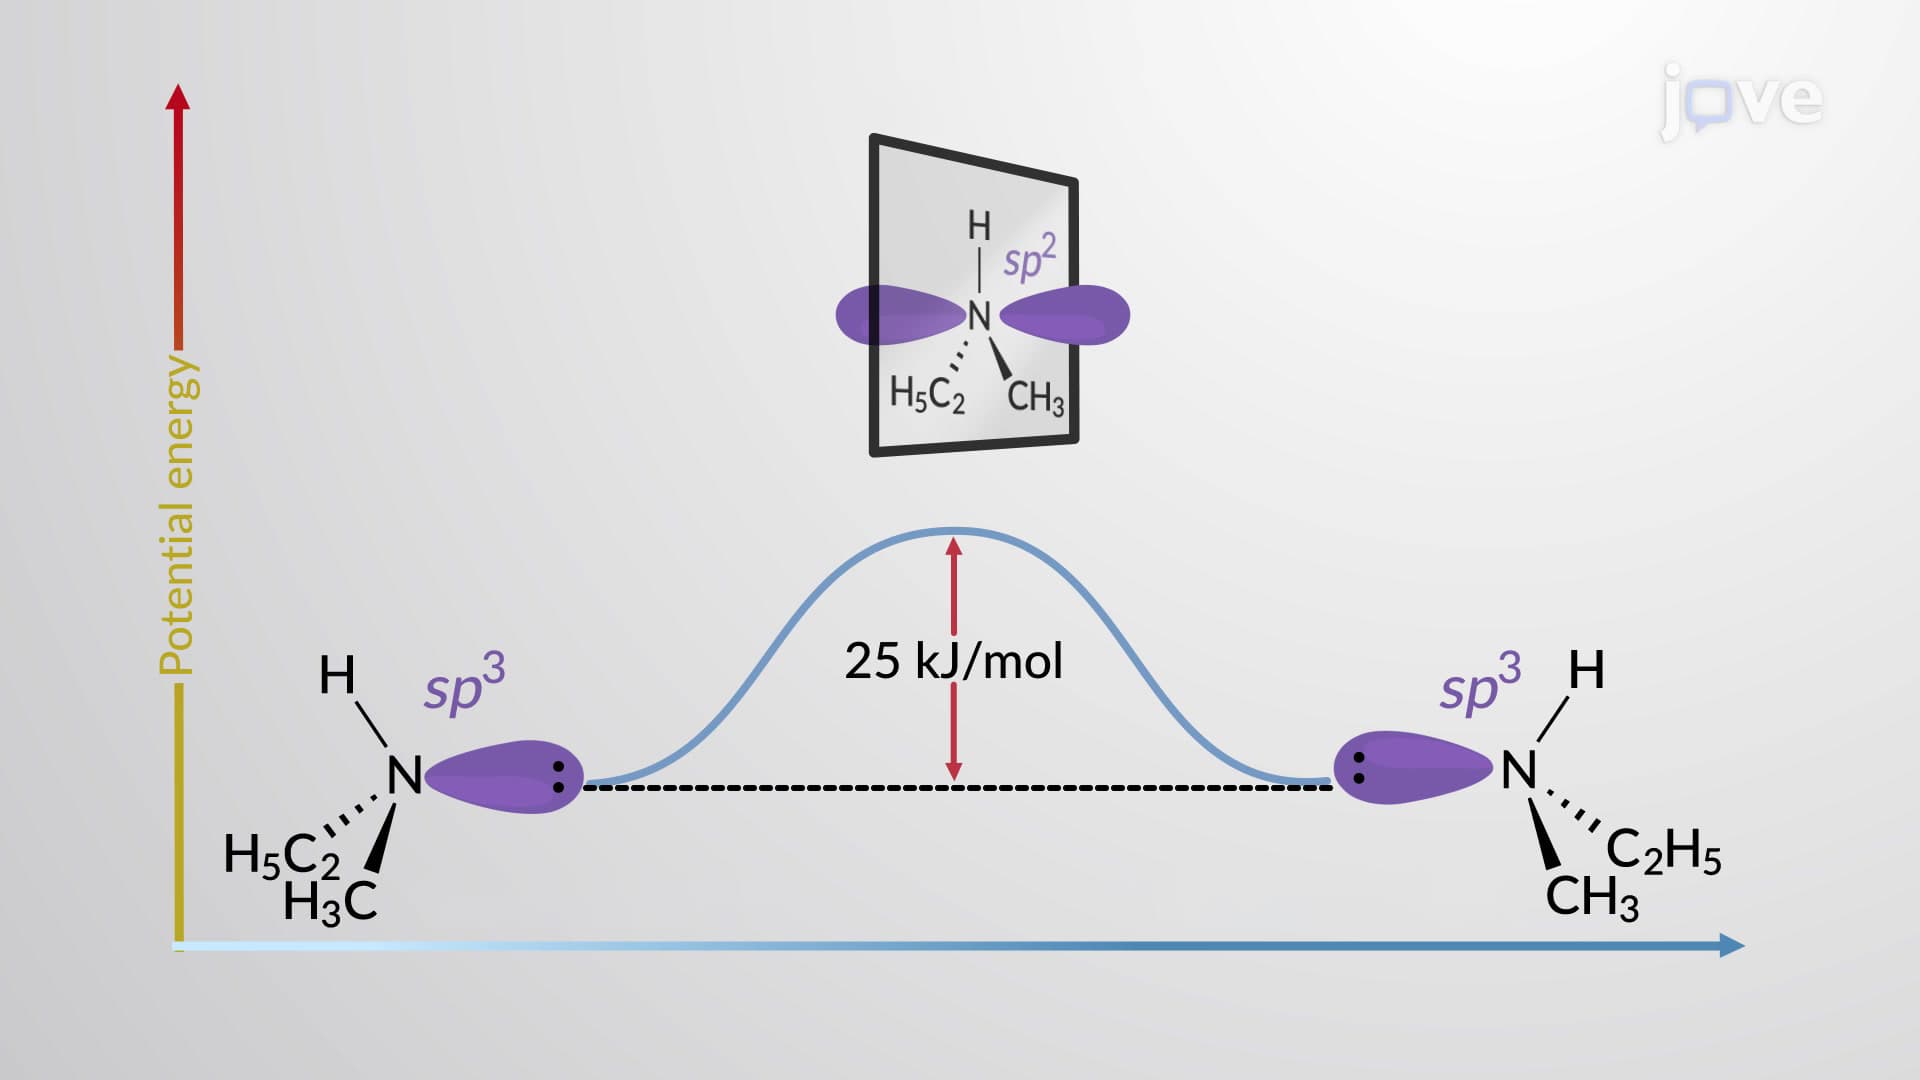
The hybridized nitrogen atom in amines possesses a lone pair of electrons and is bound to three substituents with a bond angle of around 108°, which is less than the tetrahedral angle of 109.5°. However, the C–N–H bond angle is slightly larger at 112°, with a carbon–nitrogen bond length of 147 pm. This carbon–nitrogen bond length of of amines is longer than the carbon–oxygen bond of alcohols (143 pm) but shorter than alkanes’ carbon–carbon bond (154 pm). These aspects are...
3.2K
 01:24
π Molecular Orbitals of 1,3-Butadiene
11.3K
Conjugated dienes have lower heats of hydrogenation than cumulated and isolated dienes, making them more stable. The enhanced stabilization of conjugated systems can be understood from their π molecular orbitals.
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
11.3K