
在F@O-doped PbTiO3中,自发的极化,n型半金属性,低晶格导热性和高结构稳定性
S Haider1, Bassem F Felemban2, Hafiz Tauqeer Ali2
1Department of Physics, University of Sargodha 40100 Sargodha Pakistan safdar.nazir@uos.edu.pk +92-334-9719060.
RSC advances
|November 3, 2025

概括
这项研究研究了用于自旋电子的F@O化PbTiO3矿氧化物. 兴奋剂诱导了半金属铁磁行为,显示了高级记忆和能源应用的前景.
科学领域:
- 材料科学 材料科学 材料科学
- 凝聚物质物理学 凝聚物质物理学
- 计算化学的计算化学
背景情况:
- 铁电和铁磁材料对于自旋电子设备至关重要.
- PbTiO3矿是这种应用的有希望的候选物.
- 了解兴奋剂效应是优化材料性能的关键.
研究的目的:
- 为了研究原始和F@O-化PbTiO3矿的结构,电子和磁性特性.
- 探索这些材料在螺旋电子和能量转换应用中的潜力.
- 为了确定药物结构的稳定性和关键性能指标.
主要方法:
- 使用ab initio计算来研究原始的PbTiO3矿氧化物和F@O-doped矿氧化物.
- 热力学和机械稳定性通过形成和弹性系数进行评估.
- 计算了电子带结构,磁性属性和功绩数字.
主要成果:
- 纯净的PbTiO3具有高自发偏振 (88μC cm-2),并且是一种非磁性绝缘体.
- 在PbO层中F@O注会诱导n型半金属铁磁行为,其磁矩为1.0μB.
- 合结构显示出显著的自发两极分化和旋转小数通道中的巨大能量差距,确保了半金属性.
结论:
- 用F@O合的PbTiO3矿具有出色的结构稳定性,高自发极化和半金属铁磁性质.
- 这些材料是先进的自旋电子设备的有希望的候选者,包括多态随机访问内存和数据存储.
- 这些发现表明,由于有利的导热率和优点数字,能量转换应用的潜力很大.
相关概念视频
 02:42
Valence Bond Theory
11.2K
Coordination compounds and complexes exhibit different colors, geometries, and magnetic behavior, depending on the metal atom/ion and ligands from which they are composed. In an attempt to explain the bonding and structure of coordination complexes, Linus Pauling proposed the valence bond theory, or VBT, using the concepts of hybridization and the overlapping of the atomic orbitals. According to VBT, the central metal atom or ion (Lewis acid) hybridizes to provide empty orbitals of suitable...
11.2K
 02:48
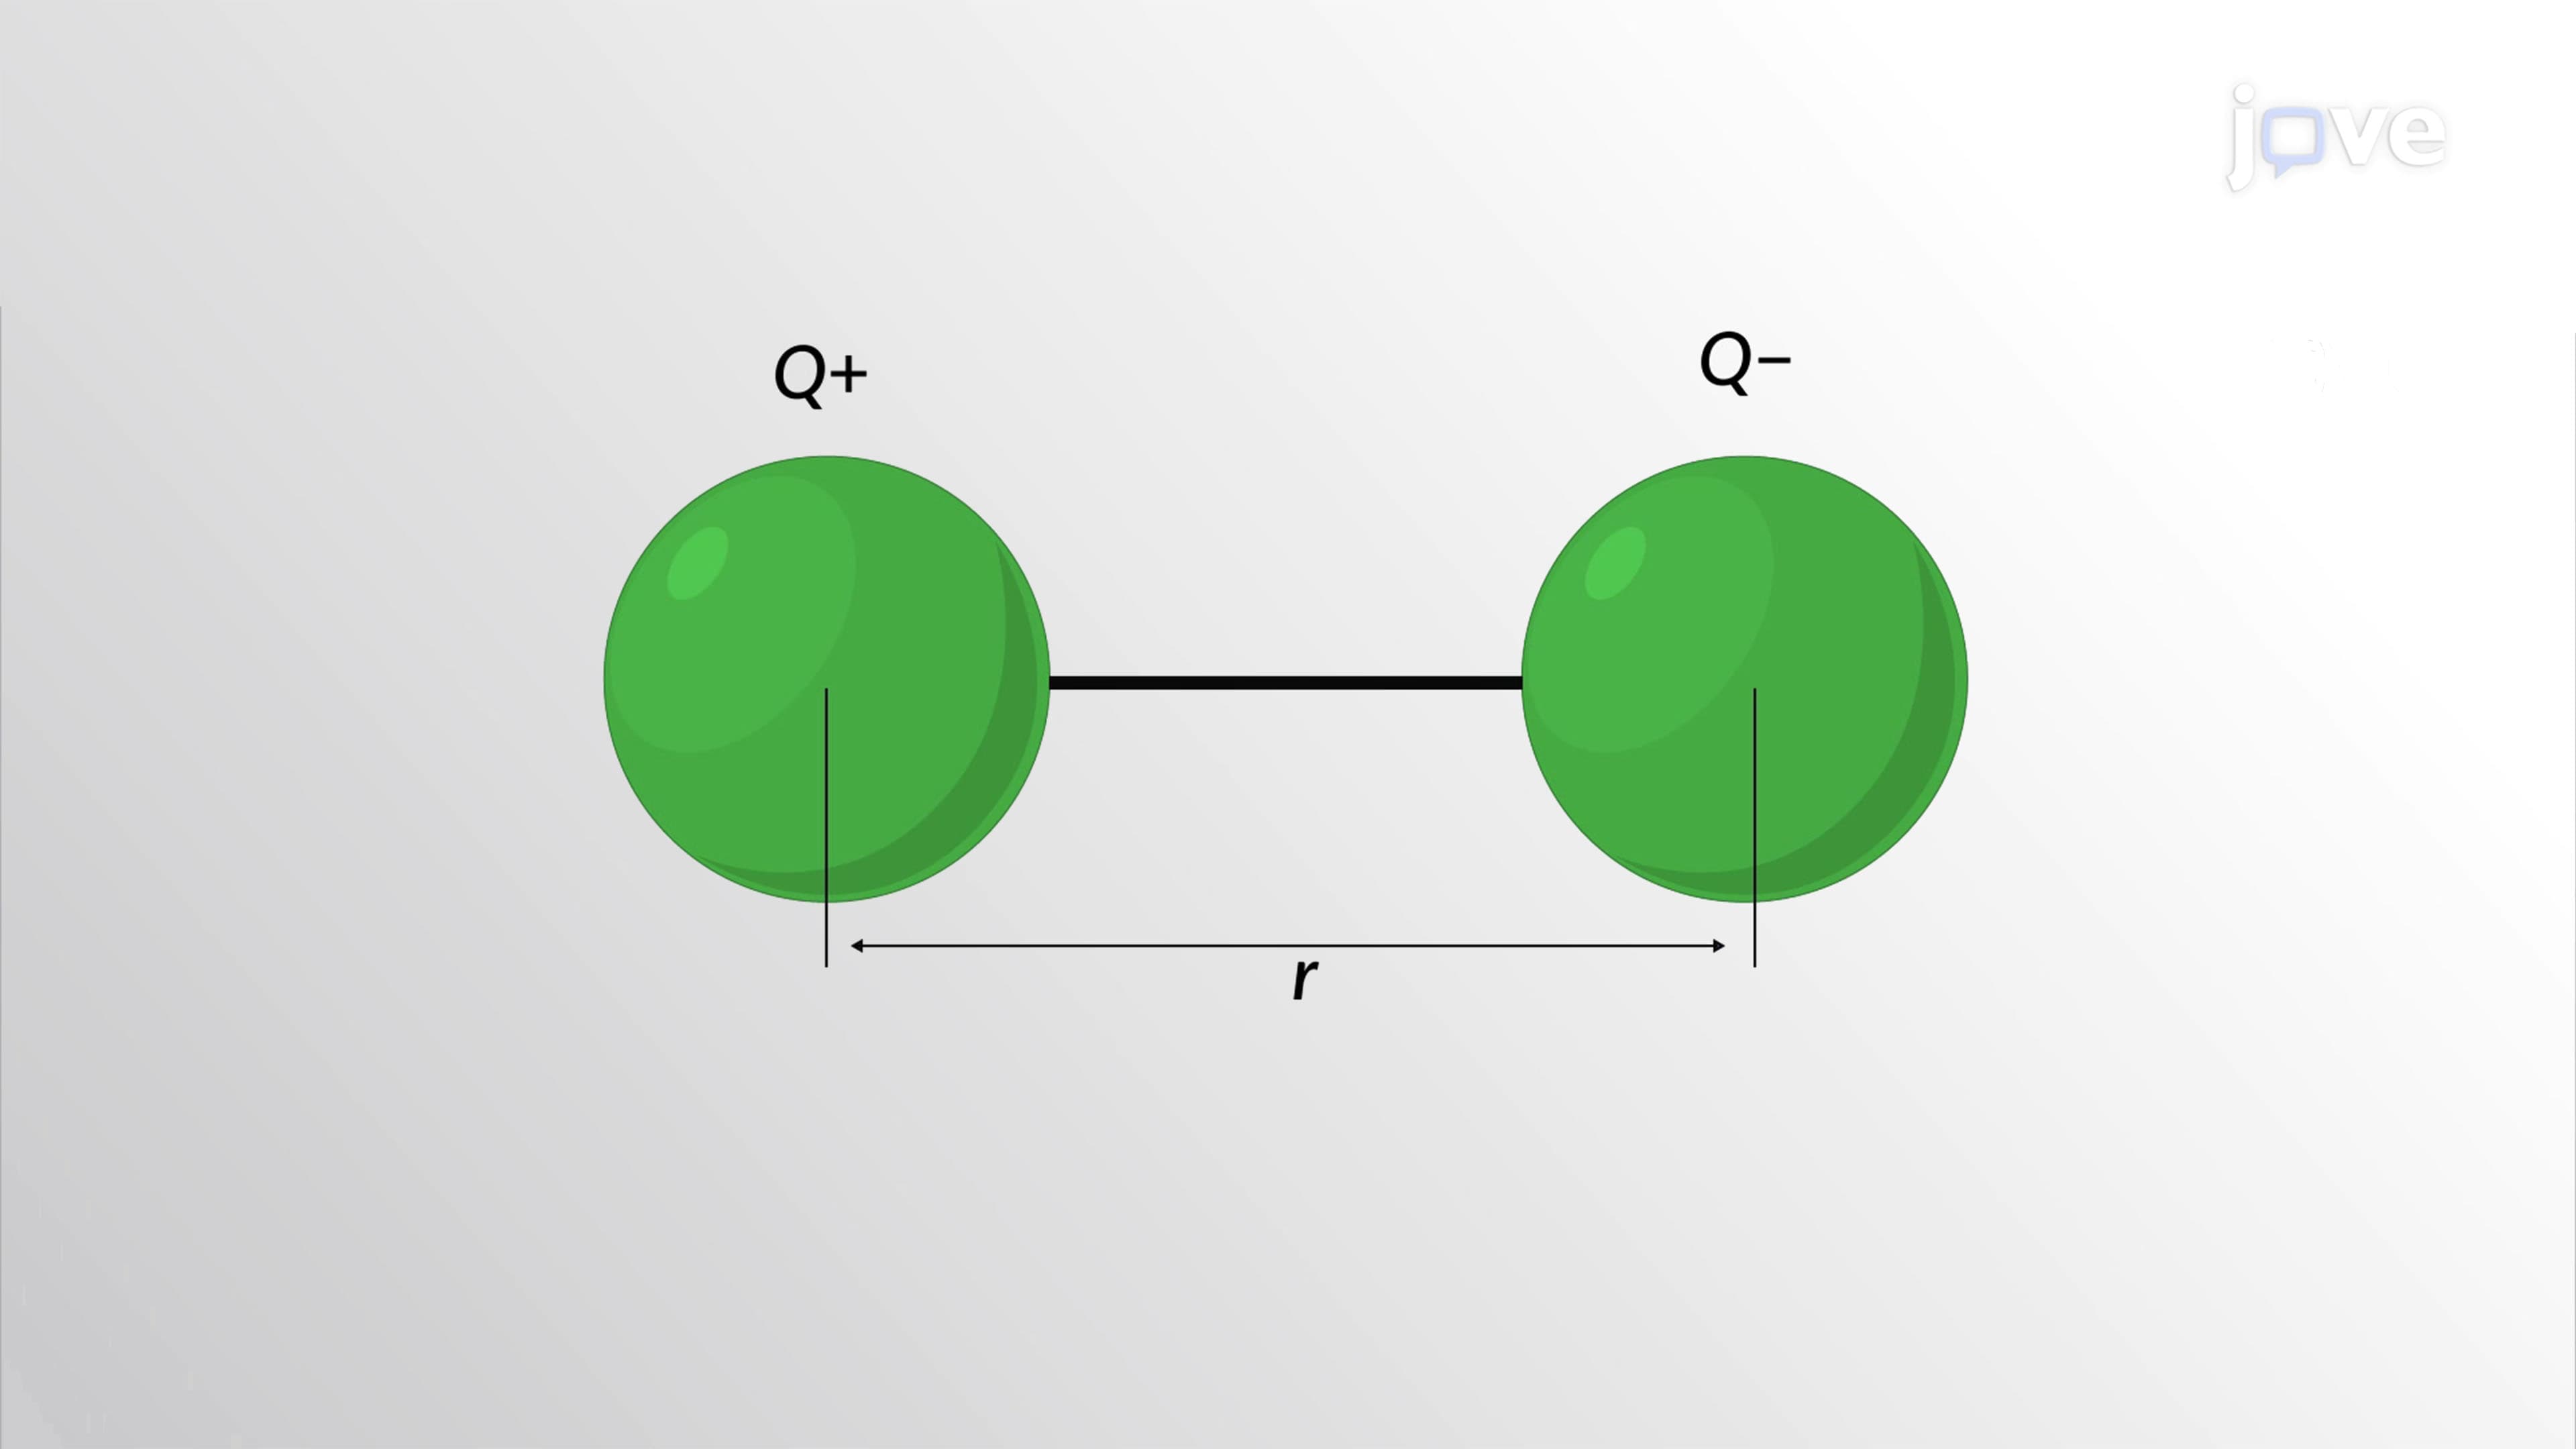
Bond Polarity, Dipole Moment, and Percent Ionic Character
34.7K
Bond Polarity
34.7K
 03:37
Molecular Shape and Polarity
73.6K
Dipole Moment of a Molecule
73.6K
 02:58
Crystal Field Theory - Octahedral Complexes
30.5K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
30.5K
 03:24
Hybridization of Atomic Orbitals I
65.3K
The mathematical expression known as the wave function, ψ, contains information about each orbital and the wavelike properties of electrons in an isolated atom. When atoms are bound together in a molecule, the wave functions combine to produce new mathematical descriptions that have different shapes. This process of combining the wave functions for atomic orbitals is called hybridization and is mathematically accomplished by the linear combination of atomic orbitals. The new orbitals that...
65.3K
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
48.1K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
48.1K