相关实验视频
Updated: Jul 5, 2026
06:51
A Patient-Derived Xenograft Model for Venous Malformation
Published on: June 15, 2020
5.7K
患有前立酶缺乏症的儿科患者的波尔图-鼻状血管疾病:一个病例报告
Melissa Castro1, Christian Martinez2, Carole Brathwaite3
1Division of Pediatric Gastroenterology, Hepatology, and Nutrition Vanderbilt University Medical Center Nashville Tennessee USA.
JPGN reports
|November 17, 2025

概括
甲酶缺乏症 (PD) 可以导致肝脏问题,如门性高血压. 这一案例突出显示了PD与波多鼻状血管疾病之间的潜在联系,并提出了新的研究途径.
科学领域:
- 遗传学 遗传学 是一个
- 肝病学 肝病学是一种肝病学.
- 儿科 儿科 儿科
背景情况:
- 益酶缺乏症 (PD) 是一种罕见的遗传疾病,影响着原体代谢.
- 肝脏参与PD的特征不清楚,病例报告有限.
- 非肝硬化门高血压是一种可能出现的严重并发症.
研究的目的:
- 在患有前立酶缺乏症的儿童中呈现早期发病的非肝性门高血压病例.
- 探索益利达酶缺乏症和波尔托-鼻状血管疾病 (PSVD) 之间的潜在关联.
- 为了研究益利达酶在血管改造和肝脏病理学中的作用.
主要方法:
- 一个儿科患者的临床病例介绍.
- 诊断工作包括对酸酶-D基因变异的基因测试和生物化学测试.
- 肝活检分析揭示了鼻状扩张和结节性.
- 怀疑血管病变时使用托西利祖马布治疗.
主要成果:
- 这位患者出现了新生儿血液溶解性贫血症,肝壮症,后来患上了食道静脉.
- 遗传和生物化学测试证实了益利达酶缺乏.
- 肝脏活检的结果与波托-鼻状血管疾病一致,特别是结节性再生性增生症.
- 托西利祖马布治疗导致临床改善.
结论:
- 这一案例表明,利达酶缺乏症和波尔托-鼻状血管疾病之间可能存在联系.
- 普罗利达酶可能在血管重塑中发挥作用,有助于PD中的肝脏病理.
- 早期诊断和识别PD相关的肝脏并发症对于管理至关重要.
相关概念视频
 01:35
Pedigree Analysis
Overview
 01:22
Lysosomal Hydrolases
Lysosomes are the site for the degradation of macromolecules and biological polymers released during membrane trafficking events such as secretory, endocytic, autophagic, and phagocytic pathways. The membrane-enclosed area of the lysosome, called the lumen, contains hydrolytic enzymes active in an acidic environment. These acid hydrolases are functional at a pH between 4.5 and 5 and are involved in cellular processes such as cell signaling, energy metabolism, restoration of the plasma membrane,...
 01:27
Protein Import into the Peroxisomes
Cells contain membrane-bound organelles called peroxisomes that oxidize organic molecules by transferring hydrogen atoms to oxygen, producing hydrogen peroxide. Peroxisomes enzymatically convert the released hydrogen peroxide into water and oxygen.
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:20
Inborn Errors of Metabolism
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
 01:24
Pharmacokinetics in Pediatric Patients: Drug Metabolism
In pediatric care, understanding the nuances of hepatic drug metabolism is crucial, as it significantly differs from that of adults. This divergence is primarily due to the developmental stage of drug-metabolizing enzymes, which affects how medications are processed in the body. In neonates, for instance, the activity of Phase I enzymes—critical for the initial breakdown of drugs—is markedly reduced, functioning at just 20–40% of the levels seen in adults. This reduction poses a challenge in...
 01:22
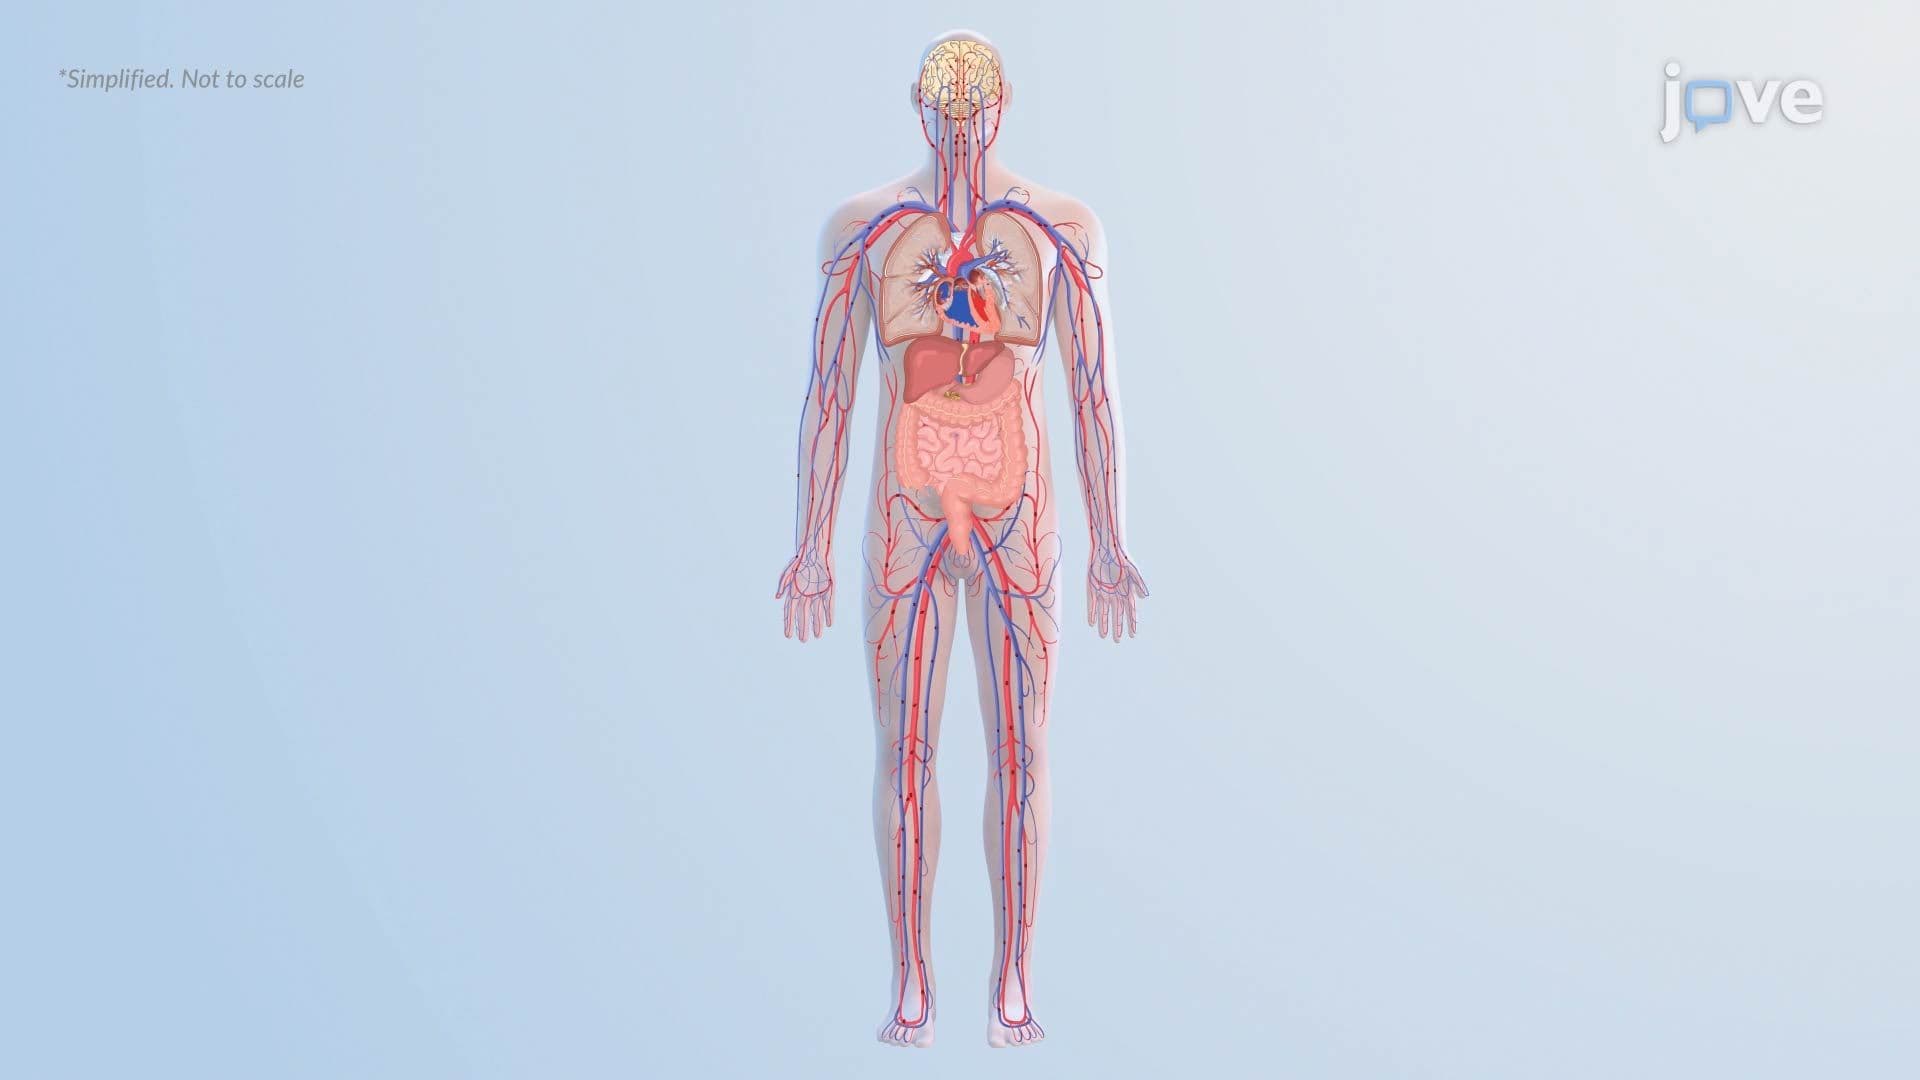
Portal Hypertension
Portal hypertension is an increase in blood pressure within the portal venous system. Normally, this pressure is less than 5 mmHg. It is considered clinically significant when it rises above 10 mmHg. At this threshold, complications from altered blood flow and venous congestion emerge.EtiologyPortal hypertension arises from conditions that impede blood flow through the liver. The most common cause is cirrhosis, in which chronic liver injury leads to fibrotic scarring. This fibrosis narrows or...