
了解由几何相位效应引起的电子结构方法中的故障
Eirik F Kjønstad1, Henrik Koch1
1Department of Chemistry, Norwegian University of Science and Technology, 7491 Trondheim, Norway.
The Journal of chemical physics
|November 17, 2025

概括
由电子波函数信号变化产生的几何相位效应,显著影响电子结构计算. 忽视这个阶段可能会导致近似的错误,导致不准确的基本状态预测.
科学领域:
- 量子化学 是一个量子化学.
- 理论化学 理论化学
- 计算化学的计算化学
背景情况:
- 几何相位效应影响基于核配置的电子波函数.
- 它对电子结构近似的影响在很大程度上被忽视了.
- 圆交点是电子状态退化的关键点.
研究的目的:
- 为了证明几何相效应对近似电子结构方法的显著影响.
- 分析几何相对圆交点附近波函数行为的后果.
- 为了解释计算化学方法的失败,由于忽视了这种相位效应.
主要方法:
- 电子哈密尔顿数和波函数的理论分析.
- 关于圆交点周围波函数行为的数学证明.
- 数字模拟来证明波函数参数的影响.
主要成果:
- 几何相位需要波函数组件在形交叉点消失,除非交叉点一致.
- 如果不考虑几何相位,则会导致波函数参数的不连续性.
- 数字证据显示,结合集群和扰乱理论中的崩.
结论:
- 几何相位效应对于精确的电子结构计算至关重要.
- 估计必须考虑这个阶段,以避免趋同问题,并预测正确的基本状态.
- 该效应的全球性需要在整个核配置空间进行仔细考虑.
更多相关视频


相关概念视频
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
48.0K
Tetrahedral Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
48.0K
 02:27
Predicting Molecular Geometry
44.6K
VSEPR Theory for Determination of Electron Pair Geometries
44.6K
 02:58
Crystal Field Theory - Octahedral Complexes
30.5K
Crystal Field Theory
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
30.5K
 04:01
VSEPR Theory and the Effect of Lone Pairs
52.1K
Effect of Lone Pairs of Electrons on Molecule Geometry
52.1K
 02:52
VSEPR Theory and the Basic Shapes
82.7K
Overview of VSEPR Theory
82.7K
 02:37
VSEPR Theory
13.9K
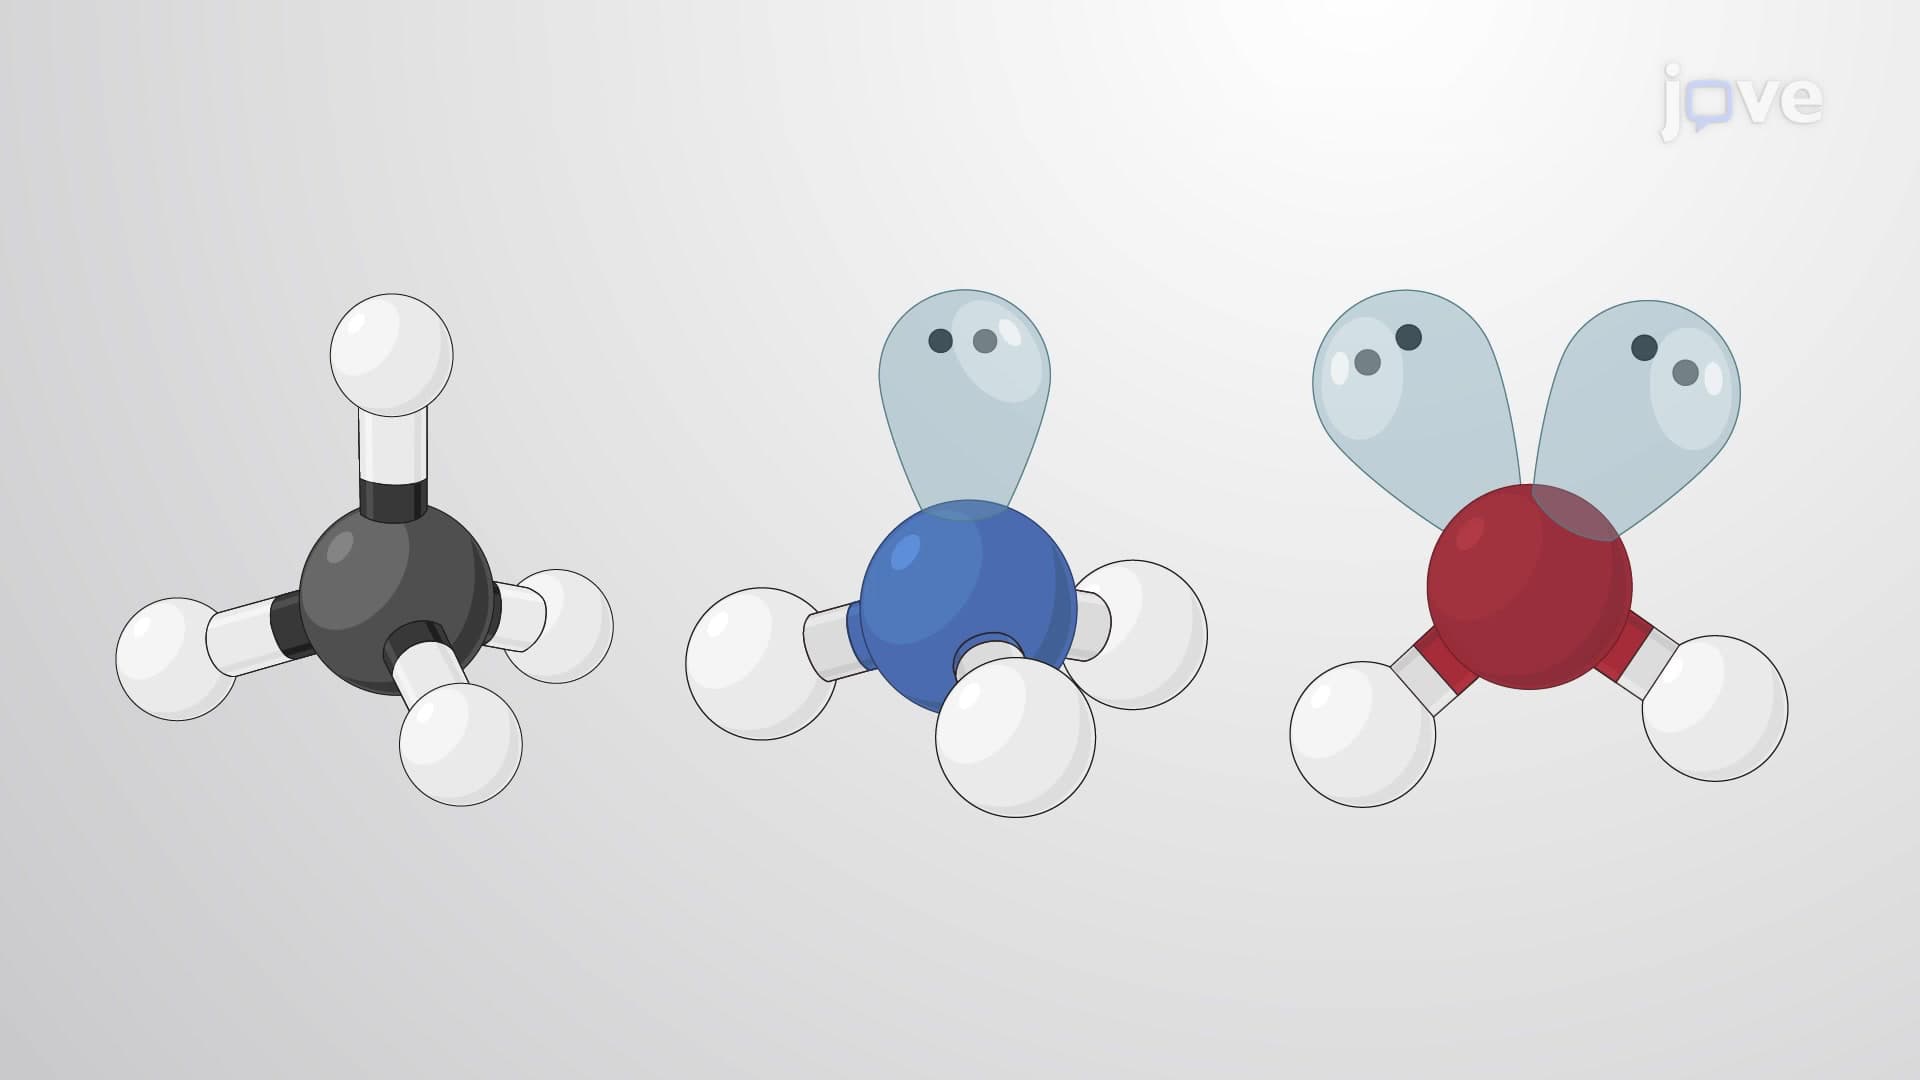
Valence shell electron-pair repulsion theory (VSEPR theory) enables us to predict the molecular structure around a central atom from an examination of the number of bonds and lone electron pairs in its Lewis structure. The VSEPR model assumes that electron pairs in the valence shell of a central atom will adopt an arrangement that minimizes repulsions between these electron pairs by maximizing the distance between them. The electrons in the valence shell of a central atom form either bonding...
13.9K