药理动力学证据支持在患有蛋白C缺乏症的患者中皮下使用蛋白C缩剂
Zhaoyang Li1, Inmaculada C Sorribes2, Jennifer Schneider2
1Takeda Development Center Americas, Inc., Cambridge, Massachusetts, United States.
TH open : companion journal to thrombosis and haemostasis
|December 12, 2025

概括
皮下 (SC) 蛋白C缩剂的使用显示了严重先天性蛋白C缺乏症 (SCPCD) 的有希望的药理动力学 (PK) 数据. 模型模拟支持各种SC剂量方案,表明快速治疗度可能需要高负载剂量.
科学领域:
- 药理动力学和药理动力学
- 血液学 血液学 血液学
- 药物开发 药物开发
背景情况:
- 蛋白C缩剂 (Ceprotin®) 已批准用于静脉注射 (IV) 治疗严重先天性蛋白C缺乏症 (SCPCD).
- 皮下 (SC) 给药提供了潜在的益处,特别是对于儿科和新生儿患者,但需要药理动力学数据来优化剂量.
- 目前SC使用的蛋白C缩物是经验性的,缺乏可靠的PK数据.
研究的目的:
- 在患有SCPCD的患者中,描述SC蛋白C度的群体药动力学 (PopPK) 概况.
- 在SCPCD患者中提供支持SC蛋白C度剂量优化的数据.
- 通过模拟来评估不同的SC剂量方案的疗效.
主要方法:
- 为SC蛋白C缩物开发了一种PopPK模型,调整了以前建立的IV模型.
- 模拟使用三阶段 (基于静脉注射方案) 和一阶段 (基于临床实践) 剂量方案进行.
- 目标最大 (Cmax) 和最低 (Ctrough) 度分别为100 IU/dL和25 IU/dL.
主要成果:
- 波普克模型证明了稳定性,并准确地描述了SC蛋白C缩PK数据.
- 模拟预测,在三阶段情景中,6-9%的患者和在单阶段情景中5-45%的患者在第一剂后会达到Cmax>100 IU/dL.
- 在稳定状态下,预计超过83%的患者在所有模拟剂量场景中保持Ctrough>25 IU/dL.
结论:
- 基于模型的模拟提供了证据,支持在SCPCD患者中使用各种SC剂量方案的蛋白C缩物.
- 在不同的年龄组中,可以在急性或预防性环境中使用SC施用,以目标蛋白C活性水平为指导.
- 一个高的初始加载剂量可能是必要的,以快速达到治疗度.
相关概念视频
 01:09
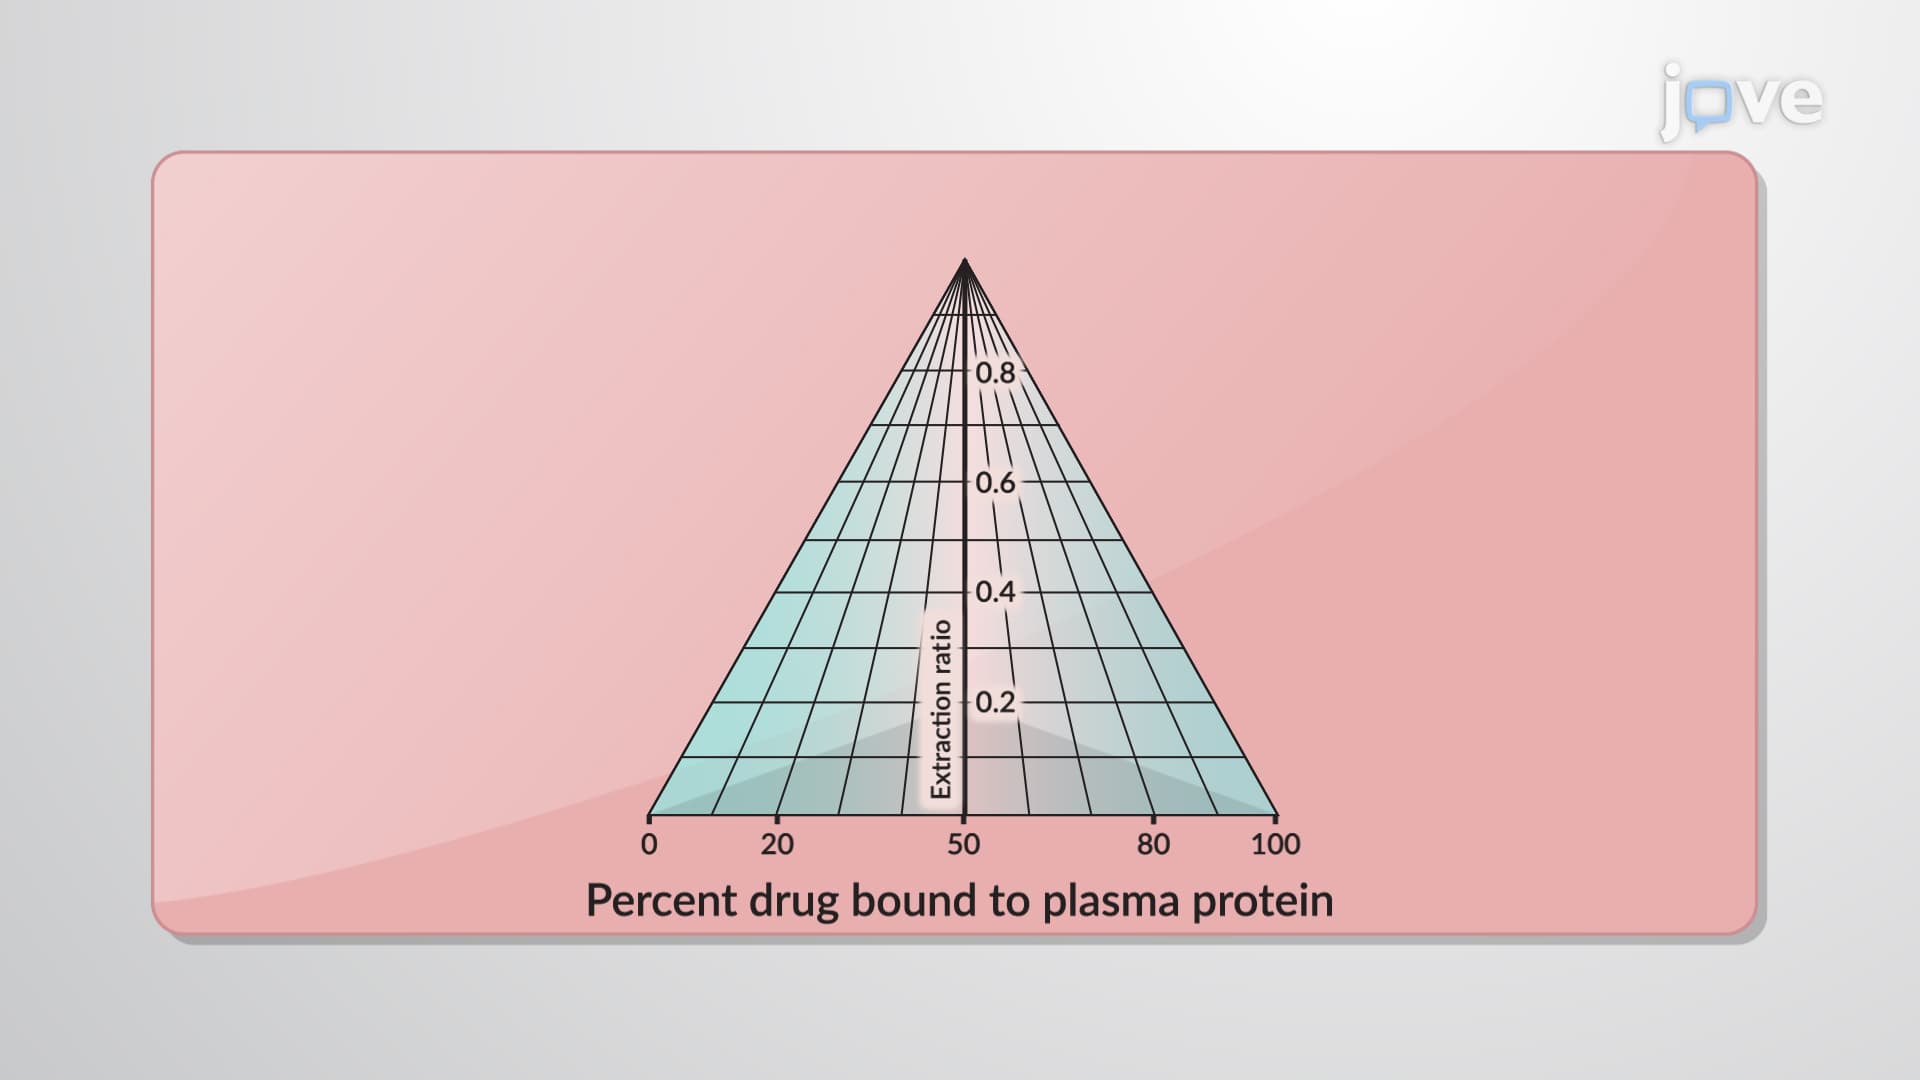
Hepatic Drug Clearance: Effect of Protein Binding
494
Hepatic clearance is influenced by protein binding based on the drug's extraction ratio. Drugs with high extraction ratios are considered flow-limited and remain unaffected by protein binding during hepatic clearance. On the other hand, drugs with low extraction ratios may be impacted by plasma protein binding, although the extent of this influence depends on the fraction of the drug bound.
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
494
 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
560
When a drug follows nonlinear pharmacokinetics, its bioavailability, the amount of the drug that reaches the systemic circulation, can change with different doses. This is due to the presence of a saturable pathway. The pathway becomes saturated as the drug concentration increases, decreasing the absorption rate. Consequently, the drug's bioavailability may be lower than expected at higher doses.
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
560
 01:16
Protein-Drug Binding: Mechanism and Kinetics
1.6K
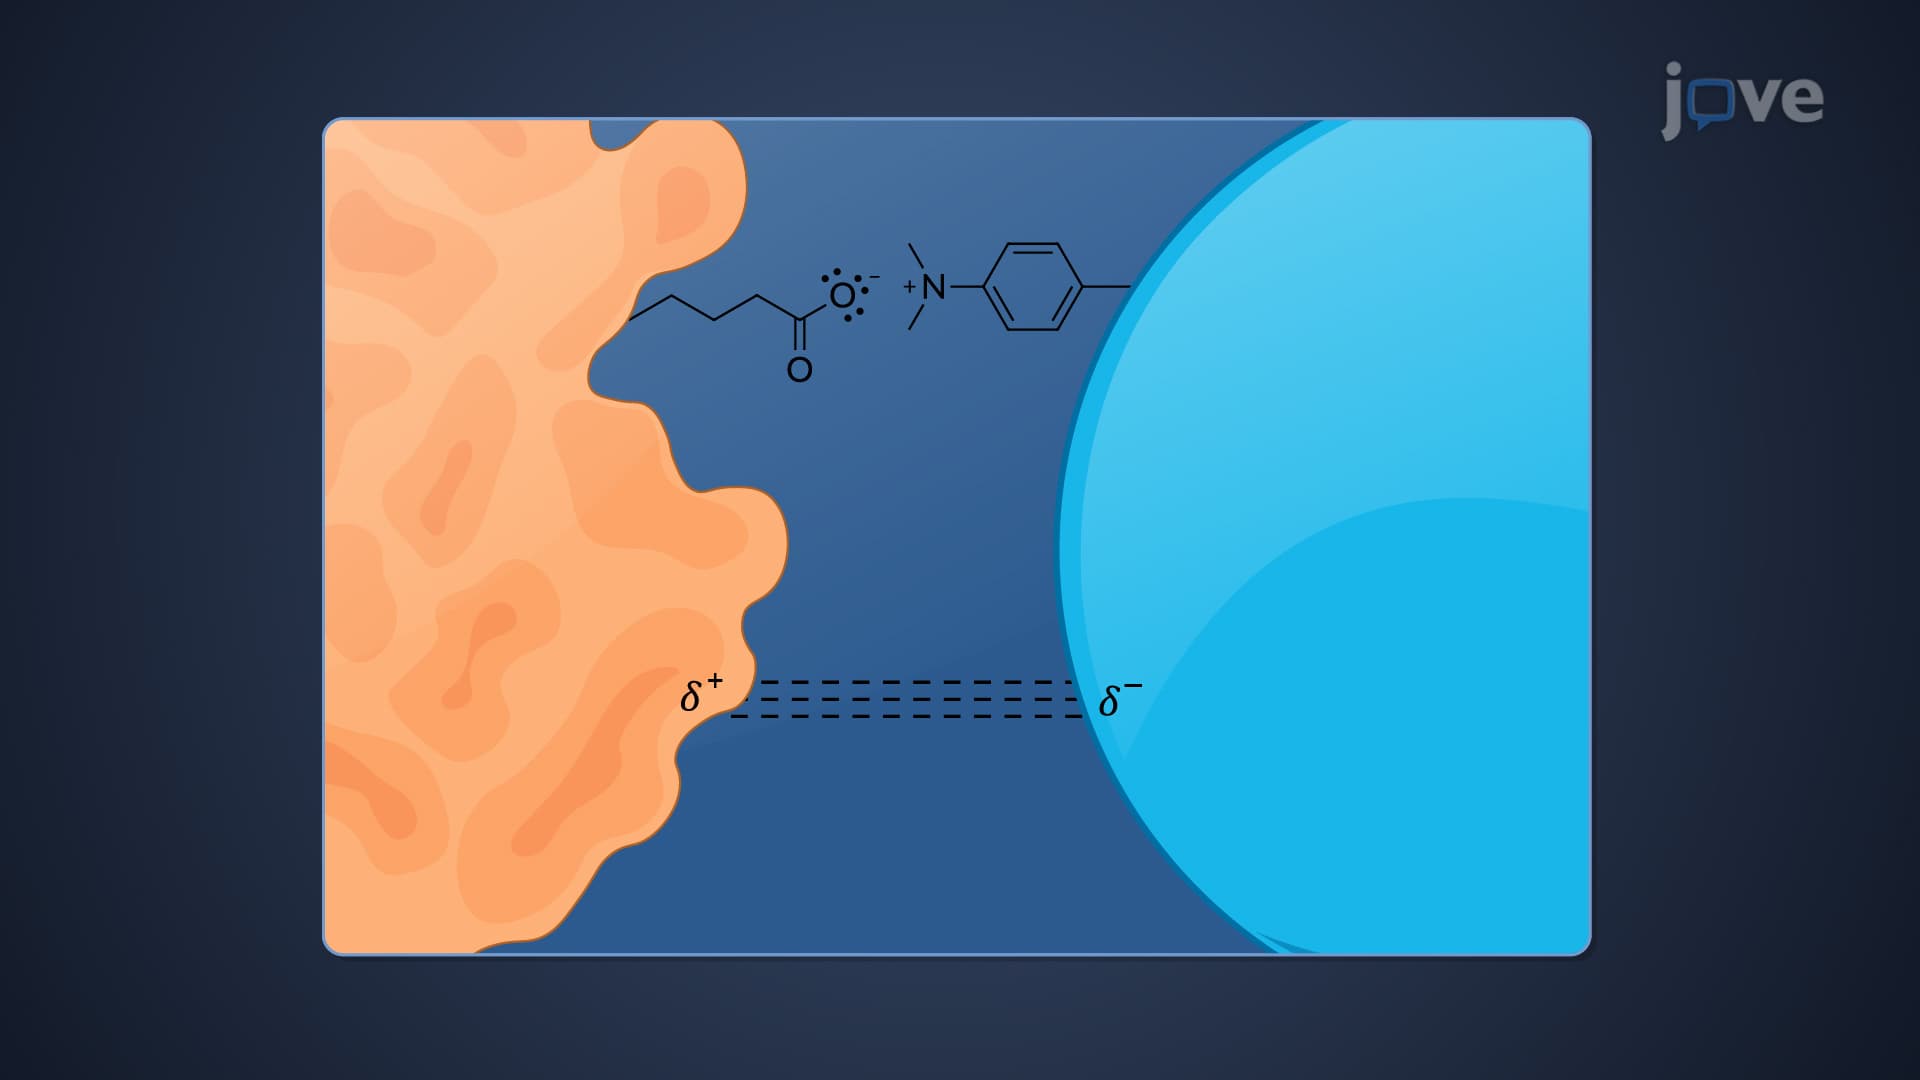
Protein-drug binding refers to the interaction between drugs and proteins within the body. This binding process can occur intracellularly, involving drug interactions with enzymes or receptors within cells, or extracellularly, involving plasma proteins in the blood.
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
1.6K
 01:23
Drug Concentrations: Measurements
1.0K
Drug concentration is the quantity of a drug present in a biological sample. Measuring drug amounts in biological samples allows the clinician to understand how a drug is absorbed, distributed, metabolized, and excreted. Samples can be obtained through invasive or non-invasive methods. Invasive techniques involve surgical or parenteral interventions to gather blood, cerebrospinal fluid, or tissue biopsy. Conversely, non-invasive approaches provide samples like urine, feces, and saliva.
Plasma...
Plasma...
1.0K
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
196
Physiological models with protein binding in pharmacokinetics offer a sophisticated approach to understanding drug disposition. These models consider drug-protein interactions, enabling them to effectively predict drug concentrations in different organs and tissues. This precision aids in accurate drug dosing, providing a significant advantage over conventional models. A key process within these models is equilibration, which ensures that drug concentrations achieve a steady state within the...
196
 01:29
Drug Distribution: Plasma Protein Binding
8.2K
Drugs predominantly attach to plasma proteins, with only a small percentage remaining unbound. The unbound portion can be calculated as one minus the bound fraction. Acidic drugs form large, inactive complexes by reversibly binding to plasma albumin, which prevents them from diffusing across biological barriers. These drug-protein complexes act as reservoirs for the drugs. As the concentration of unbound drugs decreases, these complexes quickly dissociate to release the free drug, maintaining...
8.2K