
解开新的基于的π-联系统:从最佳调整的远程纠正DFT函数,PBC-DFT方法和Oxa[n]Helicenes兴奋状态研究的洞察力
Rahul Kumar1, Shreyansh Singh1, Jayati Sarkar1
1Department of Chemical Engineering, Indian Institute of Technology Delhi, New Delhi, India.
Journal of computational chemistry
|December 29, 2025

概括
这项研究对密度功能理论 (DFT) 函数进行了基准测试,用于计算基于的烯中的最高占用分子轨道-最低不占用分子轨道 (HOMO-LUMO) 能量差距. 这些发现为设计新型有机电子和光子材料提供了准确的预测.
科学领域:
- 计算化学计算化学
- 材料科学 材料科学 材料科学
- 有机电子 有机电子
背景情况:
- 烯是具有独特螺旋结构的多环芳香化合物.
- 融合 ([n]FH) 提供可调节的光电子特性.
- 精确计算最高占成的分子轨道-最低不占用的分子轨道 (HOMO-LUMO) 能量差距对于预测材料行为至关重要.
研究的目的:
- 建立一个可靠的方法来计算HOMO-LUMO基于的基基 ([n]FH) 的能量差距.
- 将16个密度函数理论 (DFT) 函数与高级合集群计算进行基准测试.
- 为了研究[n]FH的二分化电阻和光电子特性.
主要方法:
- 使用了16个DFT函数来计算HOMO-LUMO差距.
- 使用CCSD的基本差距作为基准值进行基准测试.
- 在DCM溶剂中进行了二元化研究和分子动力学 (MD) 模拟.
- 在气体和溶剂阶段进行的DFT计算,包括周期边界条件 (PBC-DFT).
主要成果:
- [n]FH表现出与S-,Se-和Te-类似物相比蓝色移动的HOMO-LUMO差距.
- [n]FH在中性和阴离子状态下表现出高抗二聚化能力,即使是对抗离子.
- 最佳调整的函数 (LC-ωPBE,LC-BLYP, ωB97XD) 提供了精确的溶剂中的HOMO-LUMO间隙,与CAM-B3LYP.
- 基于的基离子体表现出强烈的红外吸收,类似于S-模拟物.
结论:
- 基准研究能够准确地预测氧[n]基因的HOMO-LUMO差距.
- 最佳调整的DFT功能为这些系统提供可靠的激发状态特性.
- 基于的烯是电子和光子学中先进有机材料的有希望的候选者.

相关概念视频
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
2.2K
The absorption of UV–visible light by conjugated systems causes the promotion of an electron from the ground state to the excited state. Consequently, photochemical electrocyclic reactions proceed via the excited-state HOMO rather than the ground-state HOMO. Since the ground- and excited-state HOMOs have different symmetries, the stereochemical outcome of electrocyclic reactions depends on the mode of activation; i.e., thermal or photochemical.
Selection Rules: Photochemical Activation
Selection Rules: Photochemical Activation
2.2K
 01:24
π Molecular Orbitals of 1,3-Butadiene
11.2K
Conjugated dienes have lower heats of hydrogenation than cumulated and isolated dienes, making them more stable. The enhanced stabilization of conjugated systems can be understood from their π molecular orbitals.
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
11.2K
 01:13
Five-Membered Heterocyclic Aromatic Compounds: Overview
5.2K
Heterocyclic aromatic compounds are cyclic compounds that are aromatic and have one or more heteroatoms—atoms other than carbon, in the ring. Depending upon the number of atoms present in the ring, they can be either five or six-membered. Examples of five-membered heterocyclic aromatic compounds include pyrrole, furan, thiophene, and imidazole. Pyrrole consists of one nitrogen atom having one lone pair of electrons. Furan and thiophene have one oxygen and one sulfur heteroatom,...
5.2K
 01:18
Frost Circles for Different Conjugated Systems
3.5K
The inscribed polygon method is consistent with Hückel’s 4n + 2 rule and helps to learn whether the given cyclic compound is aromatic or not. The compound is stable and aromatic if every bonding molecular orbital (MO) is completely filled with a pair of electrons. However, if the non-bonding or antibonding orbitals are filled with electrons, the compound is unstable and not aromatic. Consider the Frost circle diagrams for cycloalkenes containing 4 to 8 carbons.
3.5K
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
28.0K
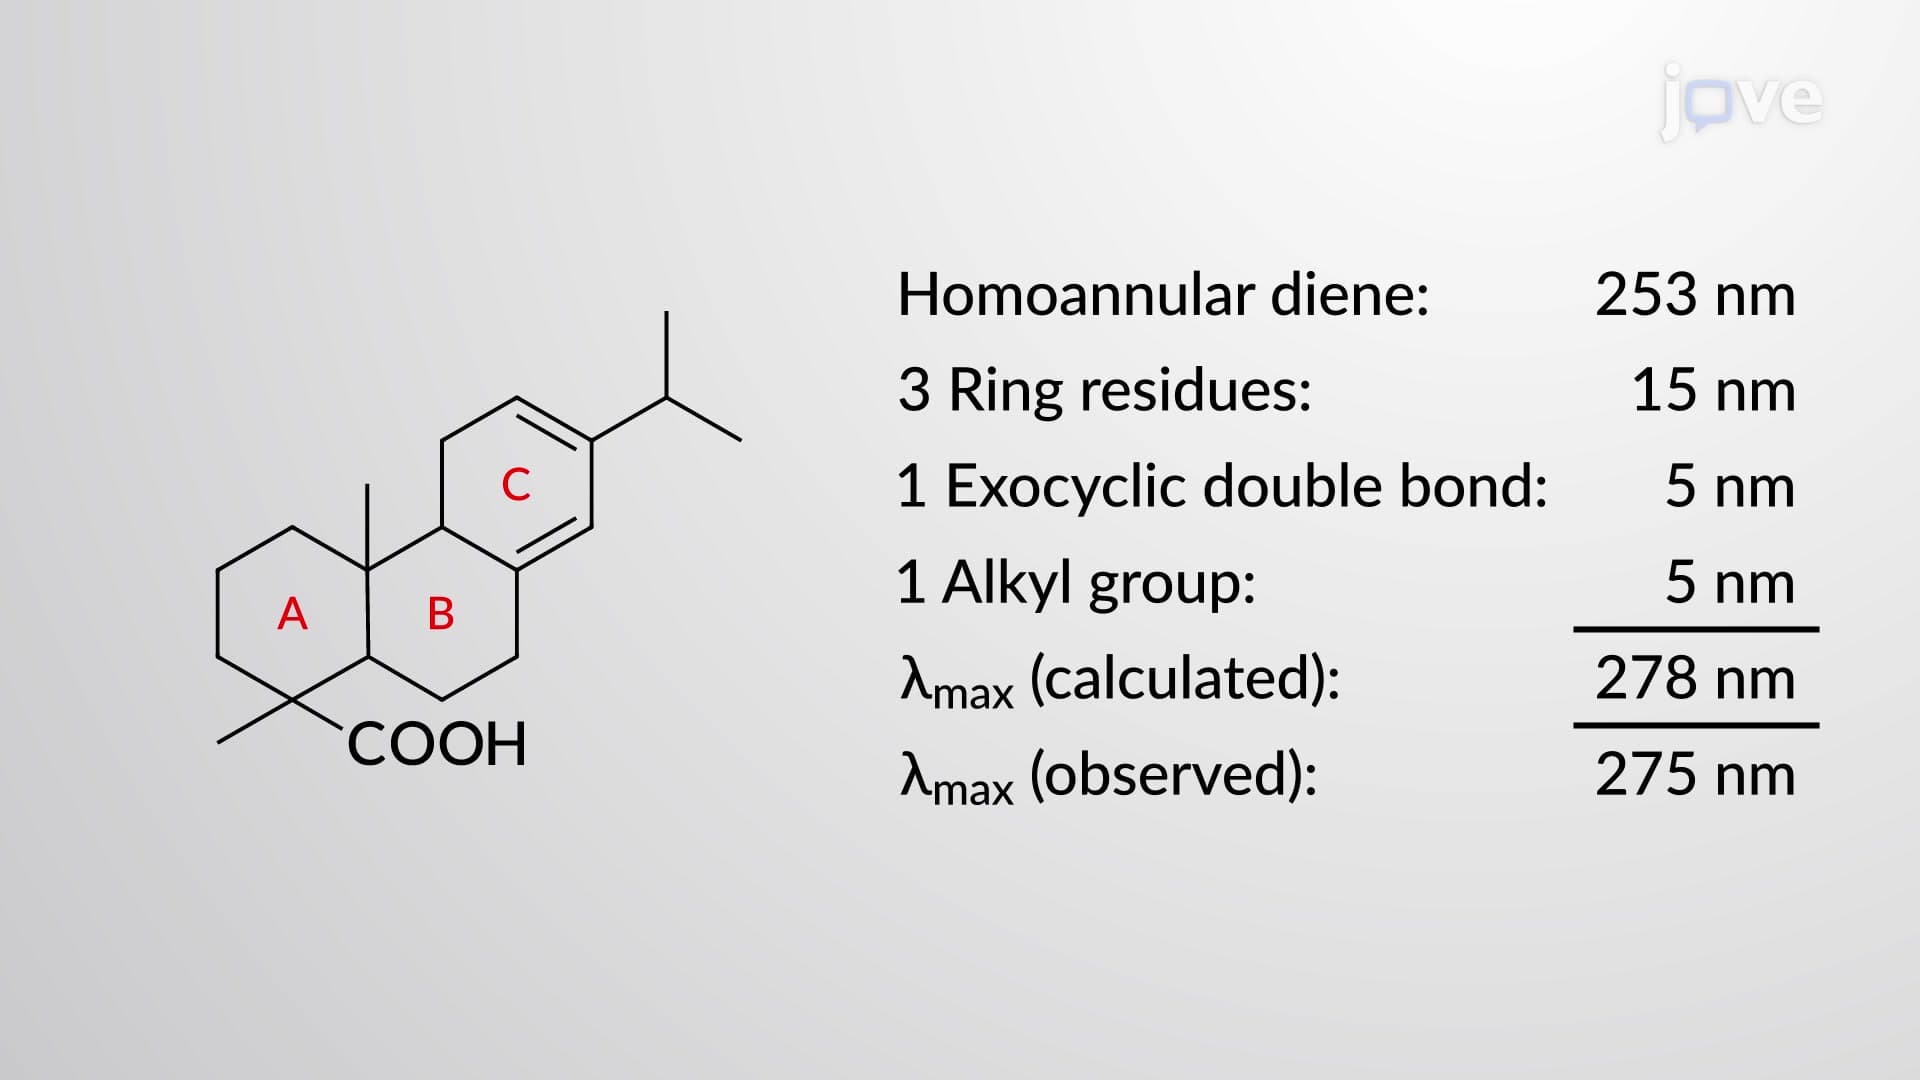
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the...
28.0K
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
3.6K
Cycloheptatriene is a neutral monocyclic unsaturated hydrocarbon that consists of an odd number of carbon atoms and an intervening sp3 carbon in the ring. The three double bonds in the ring correspond to 6 π electrons, which is a Huckel number, and therefore satisfies the criteria of 4n + 2 π electrons. However, the intervening sp3 carbon disrupts the continuous overlap of p orbitals. As a result, cycloheptatriene is not aromatic.
Removing one hydrogen from the intervening CH2 group...
Removing one hydrogen from the intervening CH2 group...
3.6K